Related Experiment Video
Updated: Aug 16, 2026

Subtyping of Campylobacter jejuni ssp. doylei Isolates Using Mass Spectrometry-based PhyloProteomics MSPP
Published on: October 30, 2016
Characterization of circulating Mycoplasma hyopneumoniae variants in the Midwestern United States using
Alyssa M Betlach1, Doug Marthaler2, Eduardo Fano3
1Department of Veterinary Population Medicine, College of Veterinary Medicine, University of Minnesota, St. Paul, MN, USA; Swine Vet Center, St. Peter, MN, USA.

Abstract:
In the United States, Multiple-Locus Variable number tandem repeat Analysis (MLVA) and complete P146 gene sequence are employed to characterize Mycoplasma hyopneumoniae from clinical samples. However, a comparison of MLVA and P146 sequencing and interpretation of assay results has not been conducted. Therefore, the aim of this study was to characterize and compare M. hyopneumoniae variants detected in the Midwestern United States using MLVA and P146 gene sequencing. A total of 160 samples were analyzed for this investigation. Both molecular techniques were compared for assay sensitivity, discriminatory power, and congruence. Epidemiological relationships in clustering of M. hyopneumoniae variants were evaluated employing Principal Coordinate Analysis. Fair agreement (κ = 0.34) in assay outcome was calculated between the two techniques. Ability to obtain a VNTR type or a P146 sequence was dependent upon the relative bacterial load (i.e., Ct value) in the sample. Simpson's diversity index was higher for MLVA (D = 0.899) than for P146 sequencing (D = 0.844). High congruence for the number of tandem repeats detected in the poly-serine region of P146 was also obtained. Similar epidemiological inferences were generated from the two assays, as production flow explained most of the variation in the clustering of VNTR types and P146 sequences. Results from this study highlighted differences in sensitivity and discriminatory power between the two molecular techniques. Nevertheless, both techniques revealed a wide genetic diversity among M. hyopneumoniae variants, with similar epidemiological inferences generated. Further research utilizing whole-genome sequencing could help identify other areas within the genome that are unrepresented by MLVA and P146 sequencing, can aid characterizing M. hyopneumoniae variants.
More Related Videos
 10:33
Multi-locus Variable-number Tandem-repeat Analysis of the Fish-pathogenic Bacterium Yersinia ruckeri by Multiplex PCR and Capillary Electrophoresis
Published on: June 17, 2019
 06:04
Antigen-Capture Enzyme-Linked Immunosorbent Assay for Specific Detection of Mycoplasma pneumoniae
Published on: February 24, 2023
Related Concept Videos
 01:29
Modern Molecular Taxonomy
 01:25
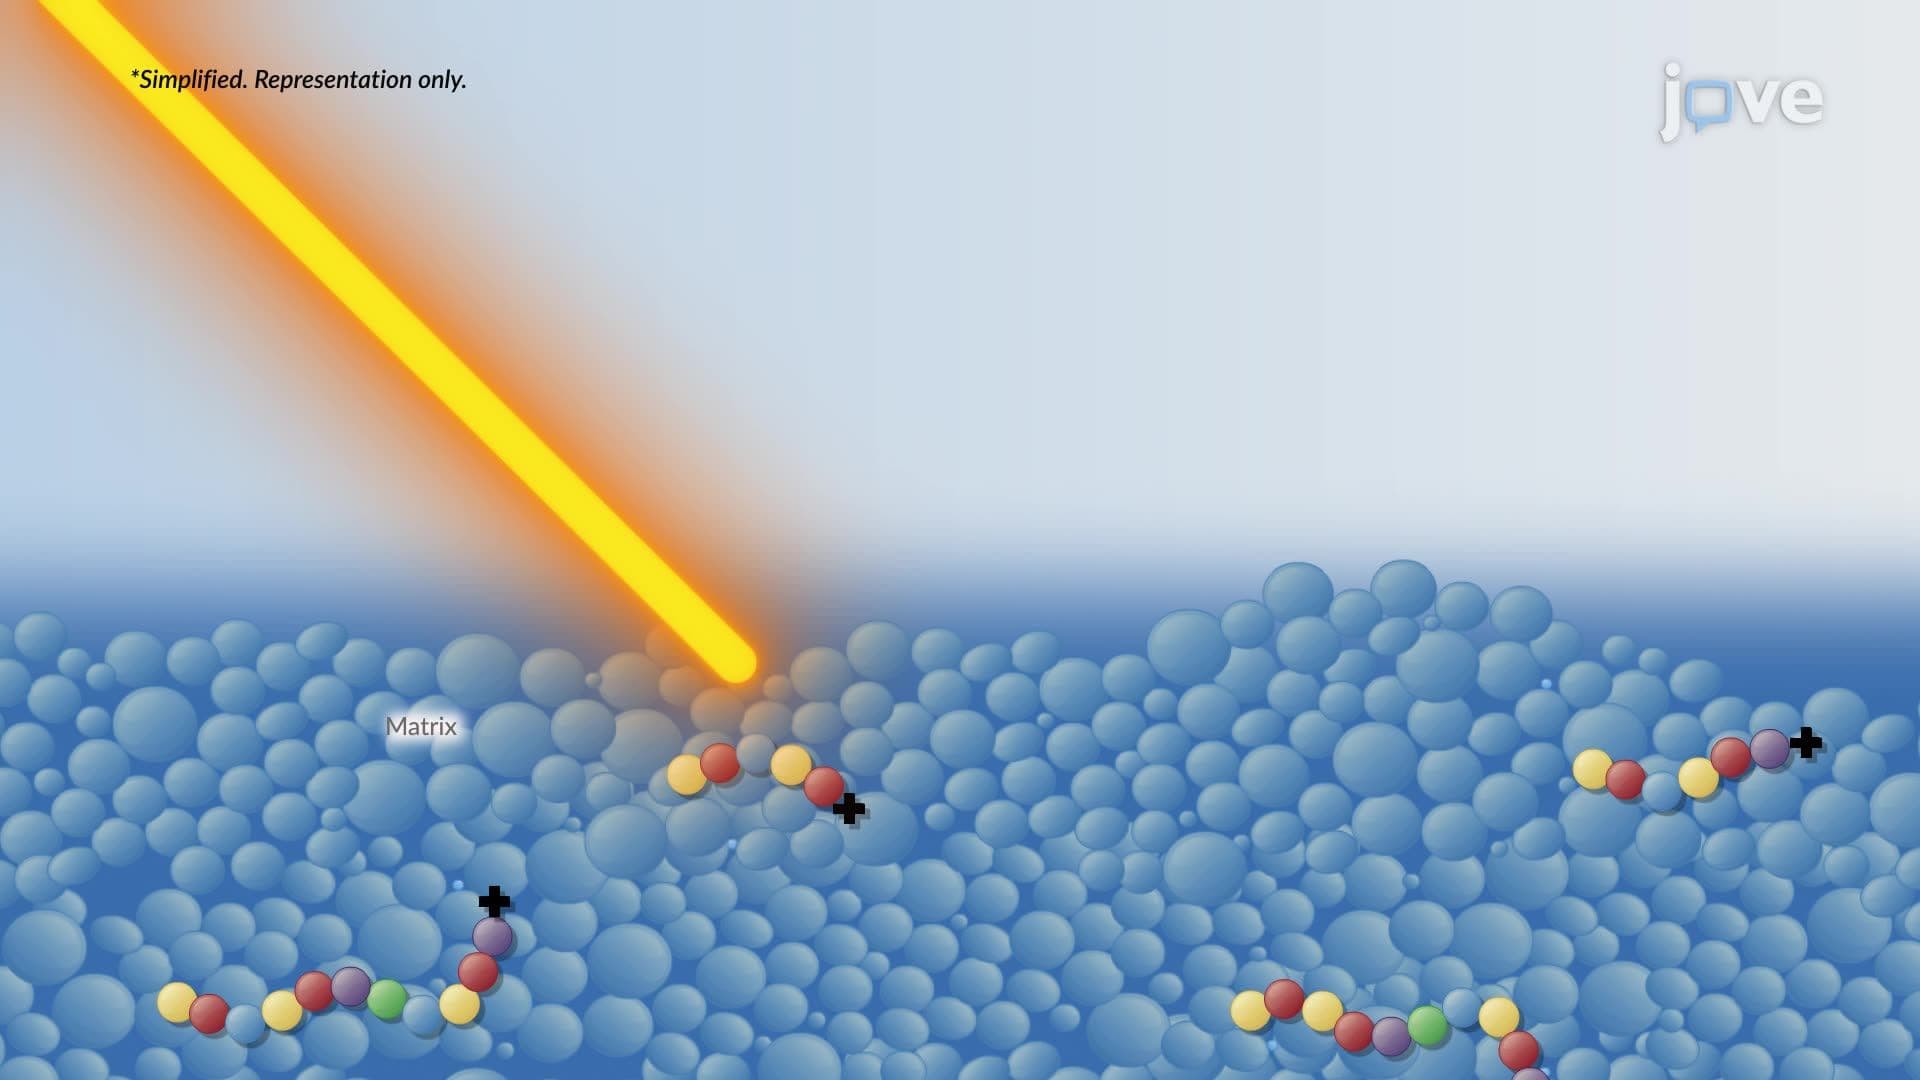
Rapid Identification of Pathogens
 01:14
Atypical Pneumonia