Related Experiment Video
Updated: Jul 14, 2026

Analysis of SAMHD1 Restriction by Flow Cytometry in Human Myeloid U937 Cells
Published on: June 13, 2021
Inhibitors of SAMHD1 Obtained from Chemical Tethering to the Guanine Antiviral Acyclovir
Matthew Egleston1, Shridhar Bhat1, A Hasan Howlader2
1Department of Pharmacology and Molecular Sciences, Johns Hopkins University School of Medicine, 725 North Wolfe Street, Baltimore, Maryland 21205, United States.

Abstract:
Sterile alpha motif histidine-aspartate domain protein 1 (SAMHD1) is an enzyme with diverse activities. Its dNTPase activity degrades all canonical dNTPs and many anticancer nucleoside drugs, while its single-stranded nucleic acid binding activity promotes DNA repair and RNA homeostasis in cells. These functions require guanine nucleotide binding to a specific allosteric site (A1) on the enzyme. We previously described how the activities of SAMHD1 could be inhibited in vitro with fragment-based inhibitor design, using dGMP as a targeting fragment for the A1 site. However, these dGMP-tethered inhibitors had poor cell permeability due to the charged guanine monophosphate group. Here, we describe a new approach where the amino form of the guanine acyclic nucleoside acyclovir (NH2-ACV) is used as the targeting fragment, allowing facile coupling to activated carboxylic acids (R-COOH), either directly or using linkers. This approach generates a neutral amide instead of charged monophosphate attachment points. High-throughput screening of a ∼375 compound carboxylic acid library identified two compounds (8, 11) with similar micromolar affinities for SAMHD1. Compound 11 was obtained by direct coupling to NH2-ACV, while compound 8 used a five-carbon linker. Both inhibitors had the same dibromonaphthol component from the carboxylic acid library screen. A crystal structure of a complex between SAMHD1 and 8, combined with computational models of bound 11, suggest how the dibromonaphthol promotes binding. The findings establish that guanine-based inhibitors targeting the A1 site do not require nucleotide or cyclic nucleoside structural elements. This guanine site targeting strategy is highly amenable to further chemical optimization.
Related Concept Videos
 01:25
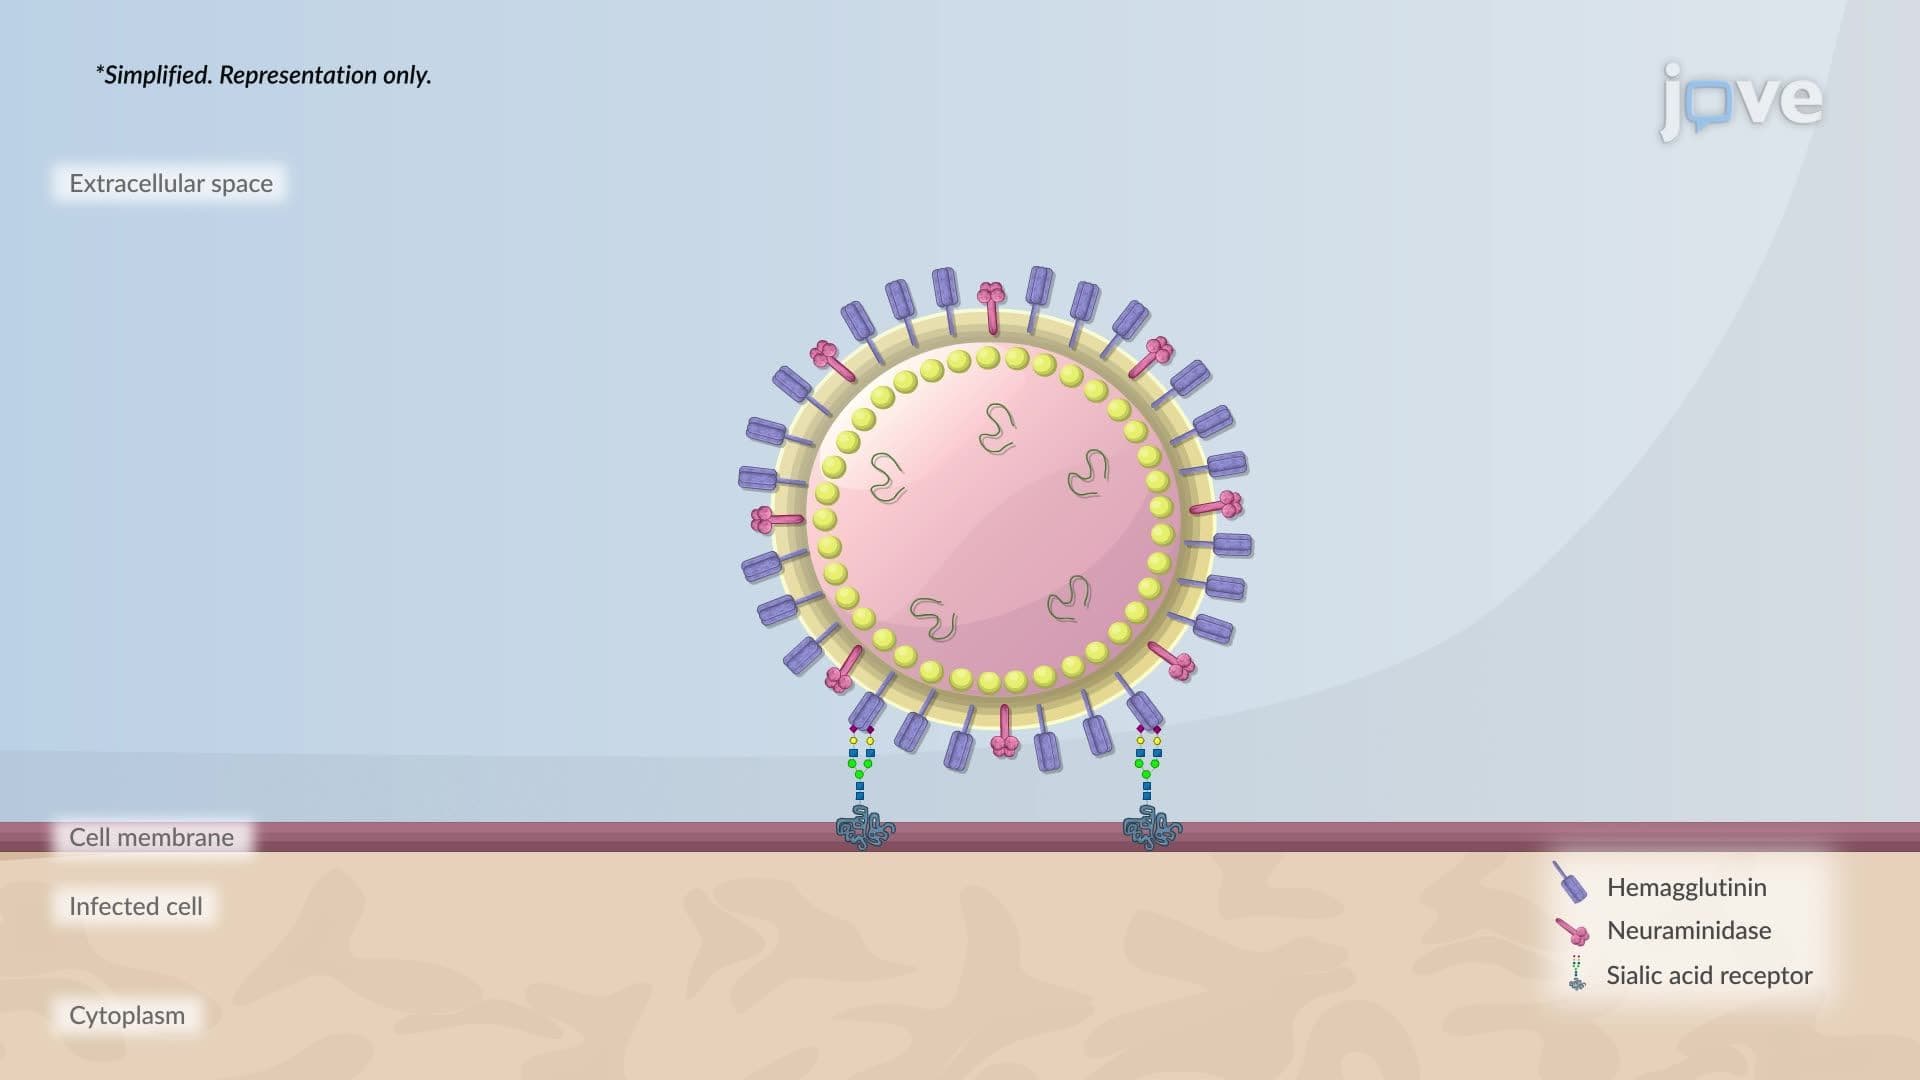
Inhibitors Of Virion Release
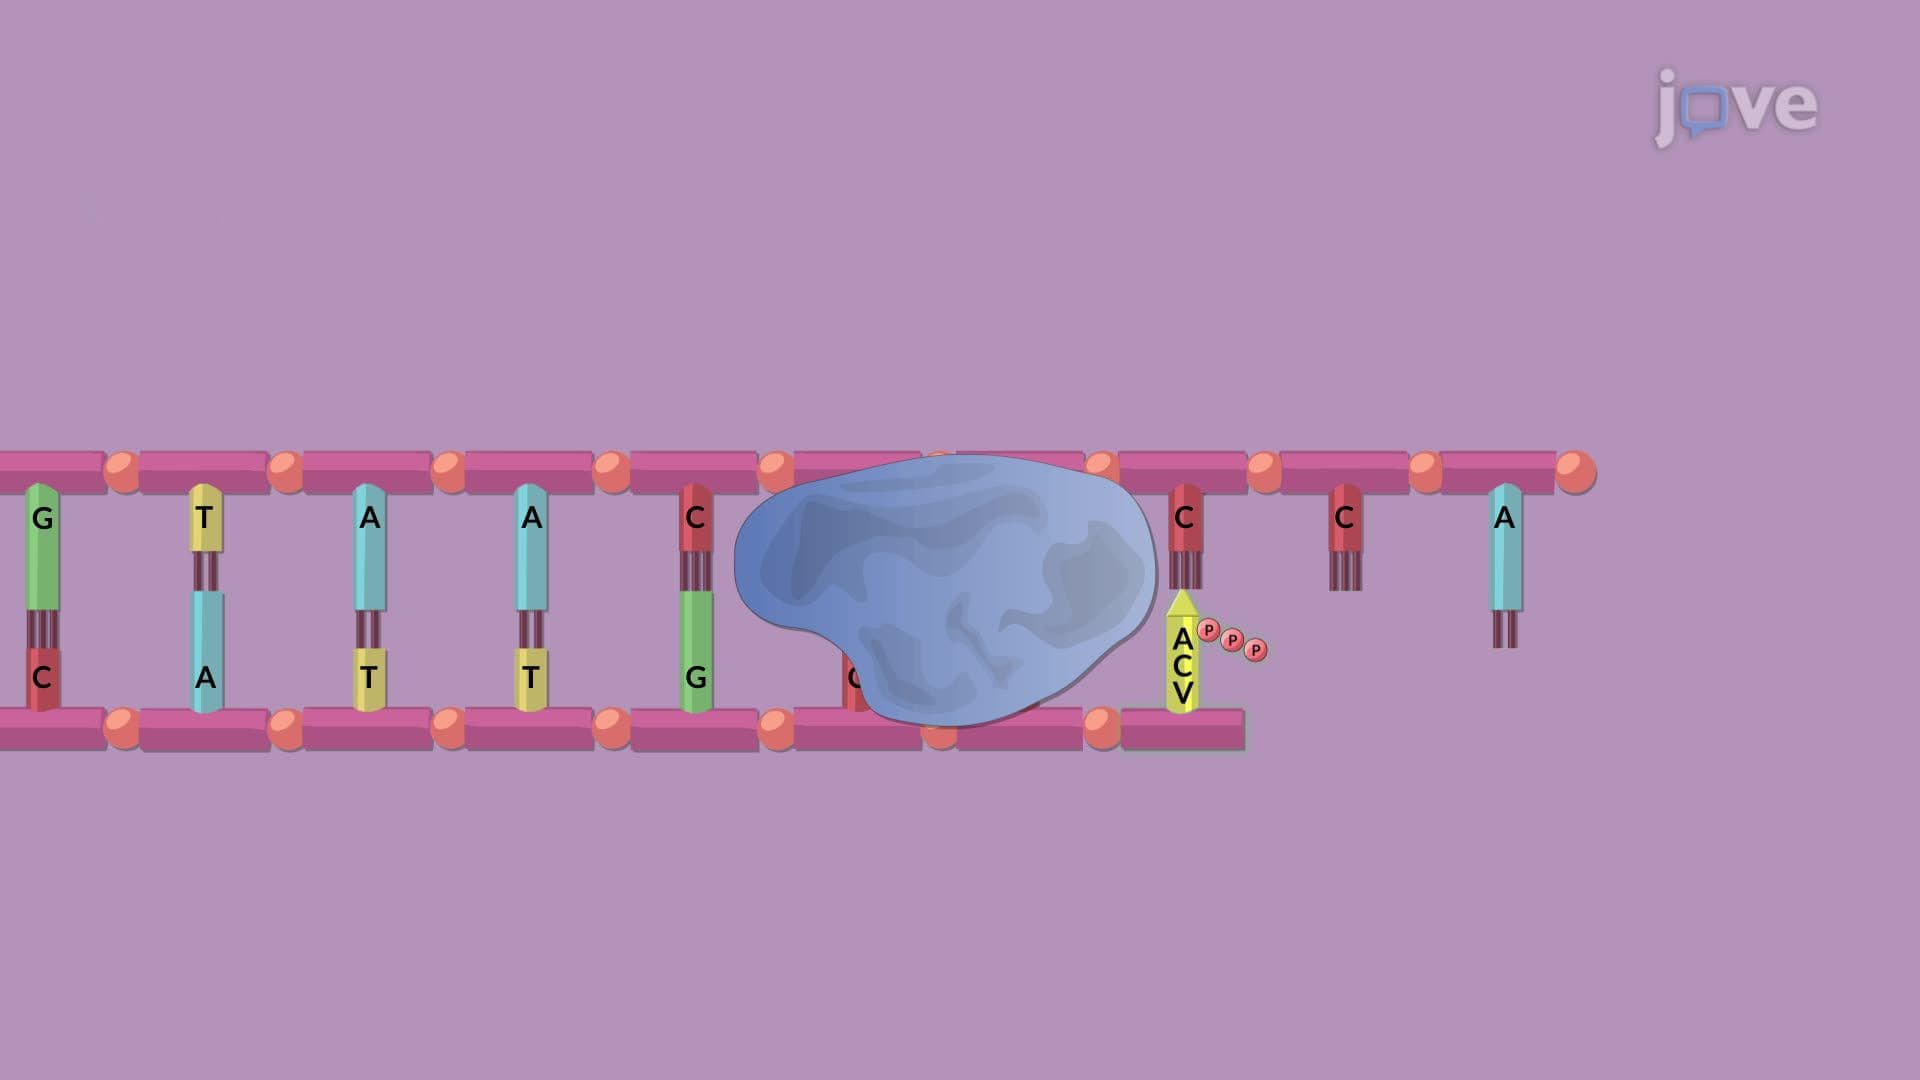 01:22
Antiviral Nucleoside Inhibitors