Related Experiment Video
Updated: May 26, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Effective Electron-Vibration Coupling by Ab Initio Methods
Maximilian F X Dorfner1, Frank Ortmann1
1TUM School of Natural Sciences, Technische Universität München, 85748 Garching b. München, Germany.

Abstract:
The description of electron-phonon coupling in materials is complex, with varying definitions of coupling constants in the literature and different theoretical approaches available. This article analyzes different levels of theory to introduce and compute these coupling constants. Within the quasi-particle picture, we derive an effective linear-coupling Hamiltonian, describing the interaction of electronic quasi-particles with vibrations. This description allows a comparison between coupling constants computed using density functional theory and higher-level quasi-particle approaches by identifying the Kohn-Sham potential as an approximation to the frequency-independent part of the self-energy. We also investigate their dependence on the exchange-correlation (XC) functional. Despite significant deviations of the Kohn-Sham eigenvalues, which arise from different XC functionals, the resulting coupling constants are remarkably similar. A comparison to quasi-particle methods, such as the well-established G0W0 approach, reveals significant quasi-particle weight renormalization. Surprisingly, however, in nearly all the considered cases, the coupling constants computed in the DFT framework are excellent approximates of the ones in the quasi-particle framework, which is traced back to a significant cancellation of competing terms. Other quasi-particle methods, such as the Outer Valence Green's Function approach and the ΔSCF method, are also included in the comparison. Moreover, we investigate the coupling of vibrations to excitonic excitations and find, by comparison to time-dependent density functional theory and extended multiconfiguration quasi-degenerate second-order perturbation theory, that knowing the underlying electron- and hole-vibration couplings is sufficient to accurately determine the exciton-vibration coupling constants in the studied cases.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 01:27
π Electron Effects on Chemical Shift: Overview
 01:22
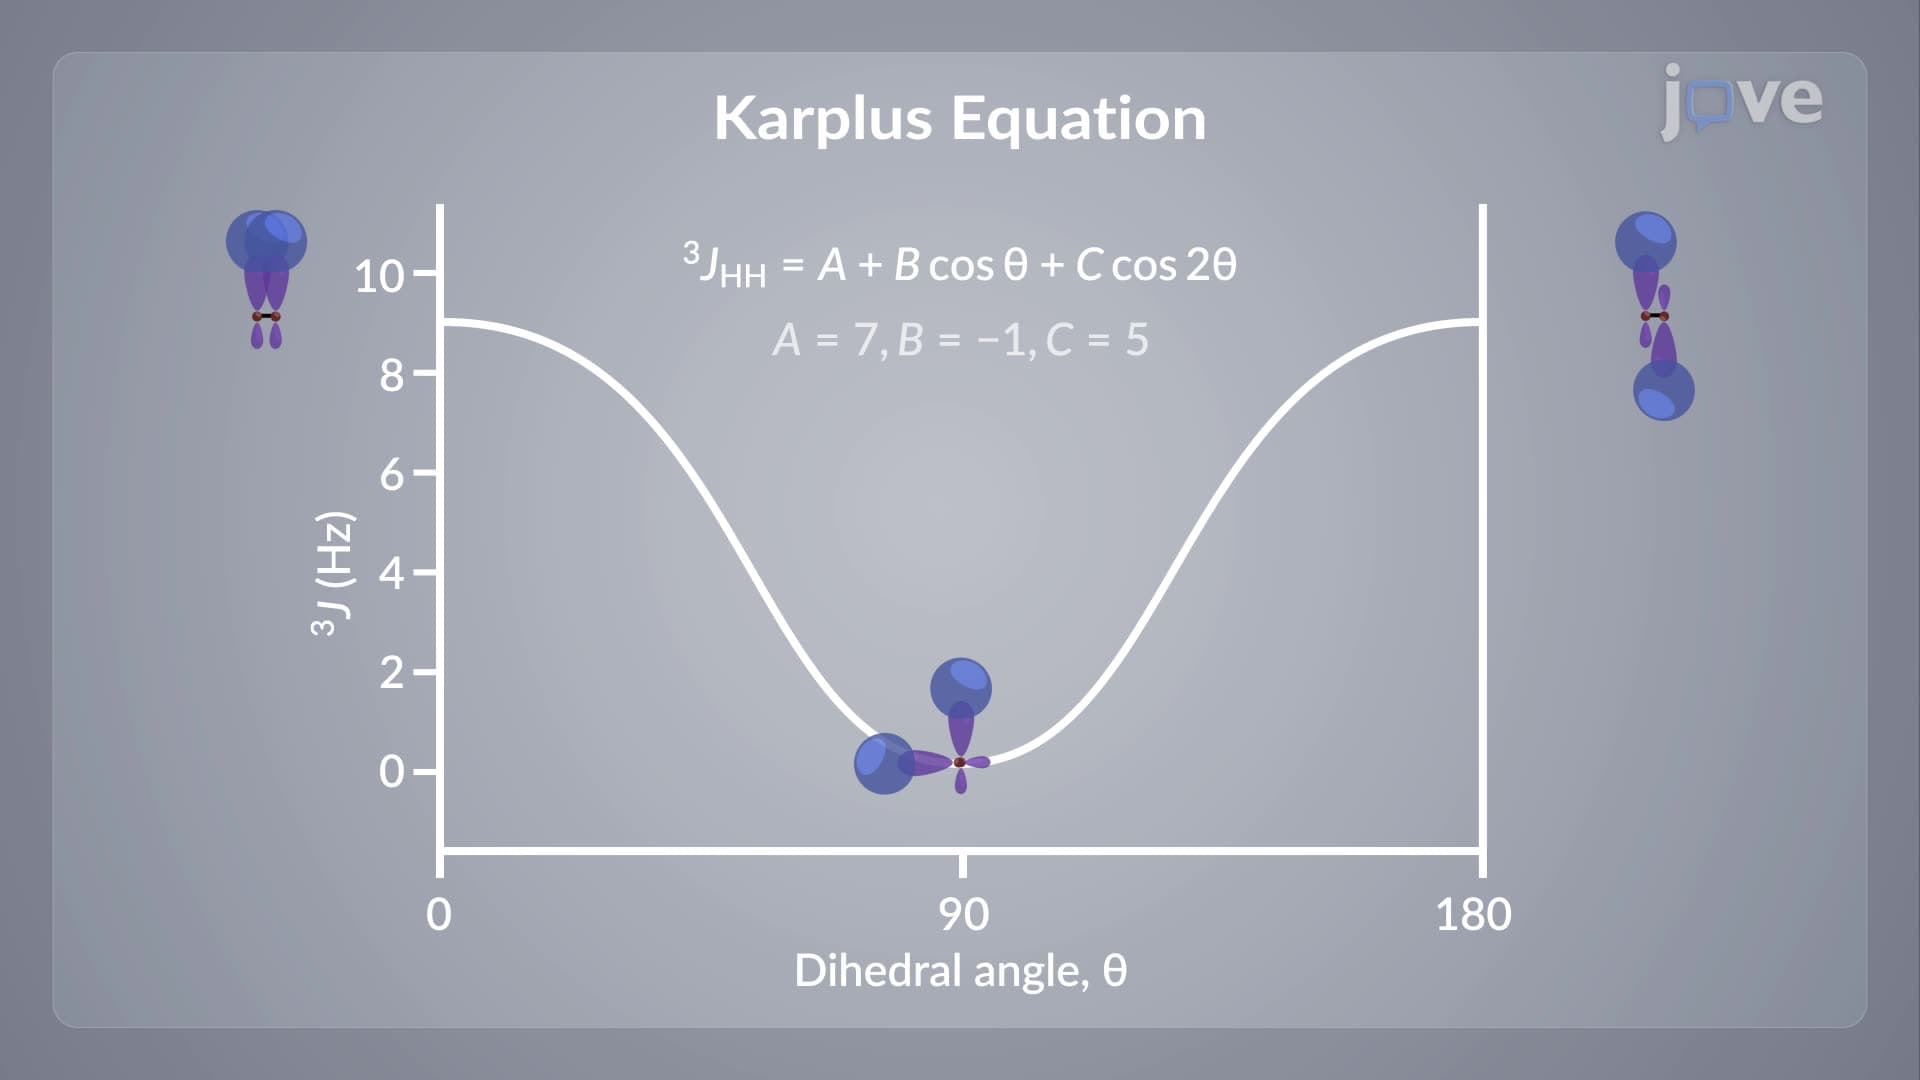
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:17
Spin–Spin Coupling: One-Bond Coupling
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 01:18
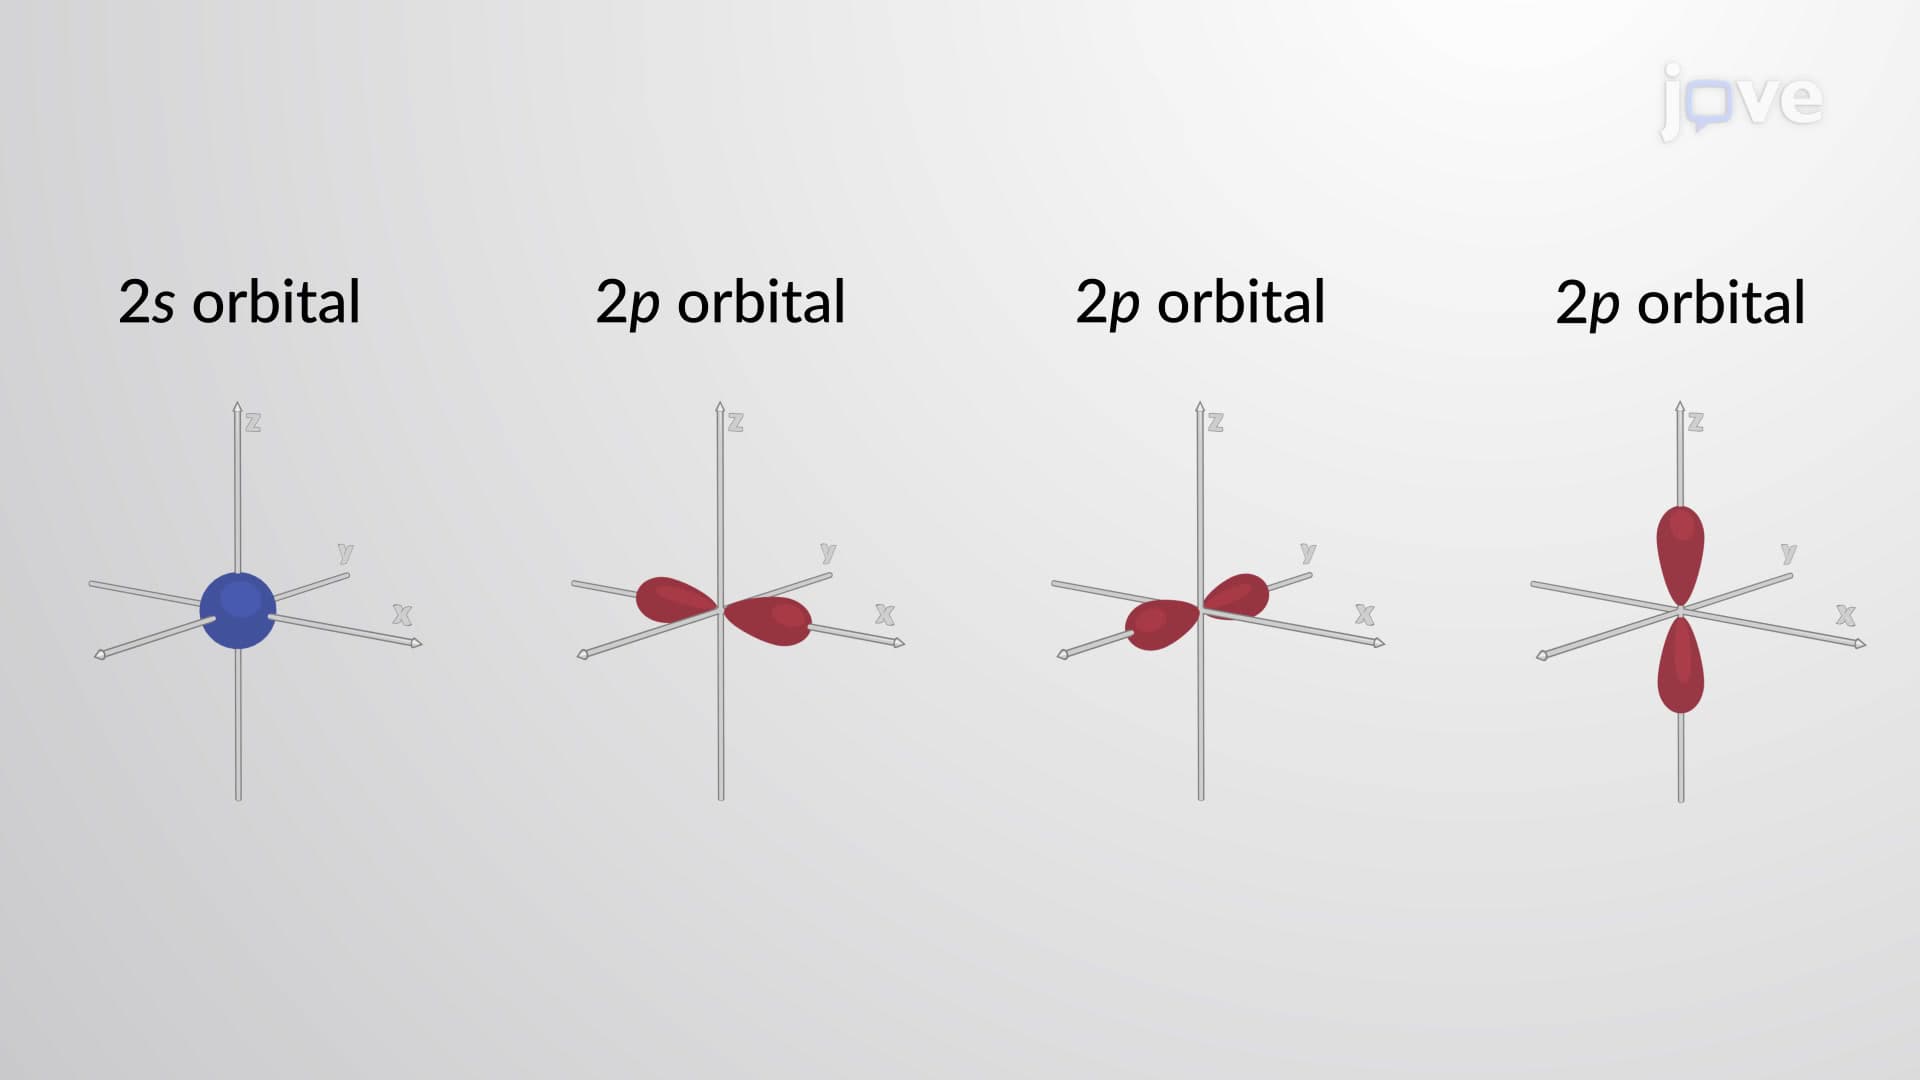
Electron Orbital Model
The first shell is closest to the nucleus, and it has only one subshell with a single spherical orbital called the...