Related Experiment Video
Updated: May 21, 2025

Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Local hybrid alternatives to the orbital density approximation reduce the orbital dependence of self-interaction
1Department of Chemistry and Biochemistry, Texas Christian University, 2800 S. University Dr., Fort Worth, Texas 7629, USA.

Abstract:
This work presents local hybrid alternatives to the orbital density approximation employed in self-interaction corrected density functional theory (SIC-DFT) and extended for use in DFT-corrected correlated wavefunction approaches (CAS-DFT). When combined with standard approximate density functionals, the orbital density approximation leaves SIC-DFT energies strongly dependent on unitary transforms among occupied orbitals and leaves CAS-DFT energies overbound. The alternatives presented here reduce both errors. The orbital density approximation and the local hybrid alternatives are shown to approximate an underlying nondiagonal exchange-correlation hole. A preliminary extension is presented to active-virtual correlation. These results motivate exploration of local hybrid concepts in SIC-DFT and CAS-DFT.
More Related Videos


Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 02:38
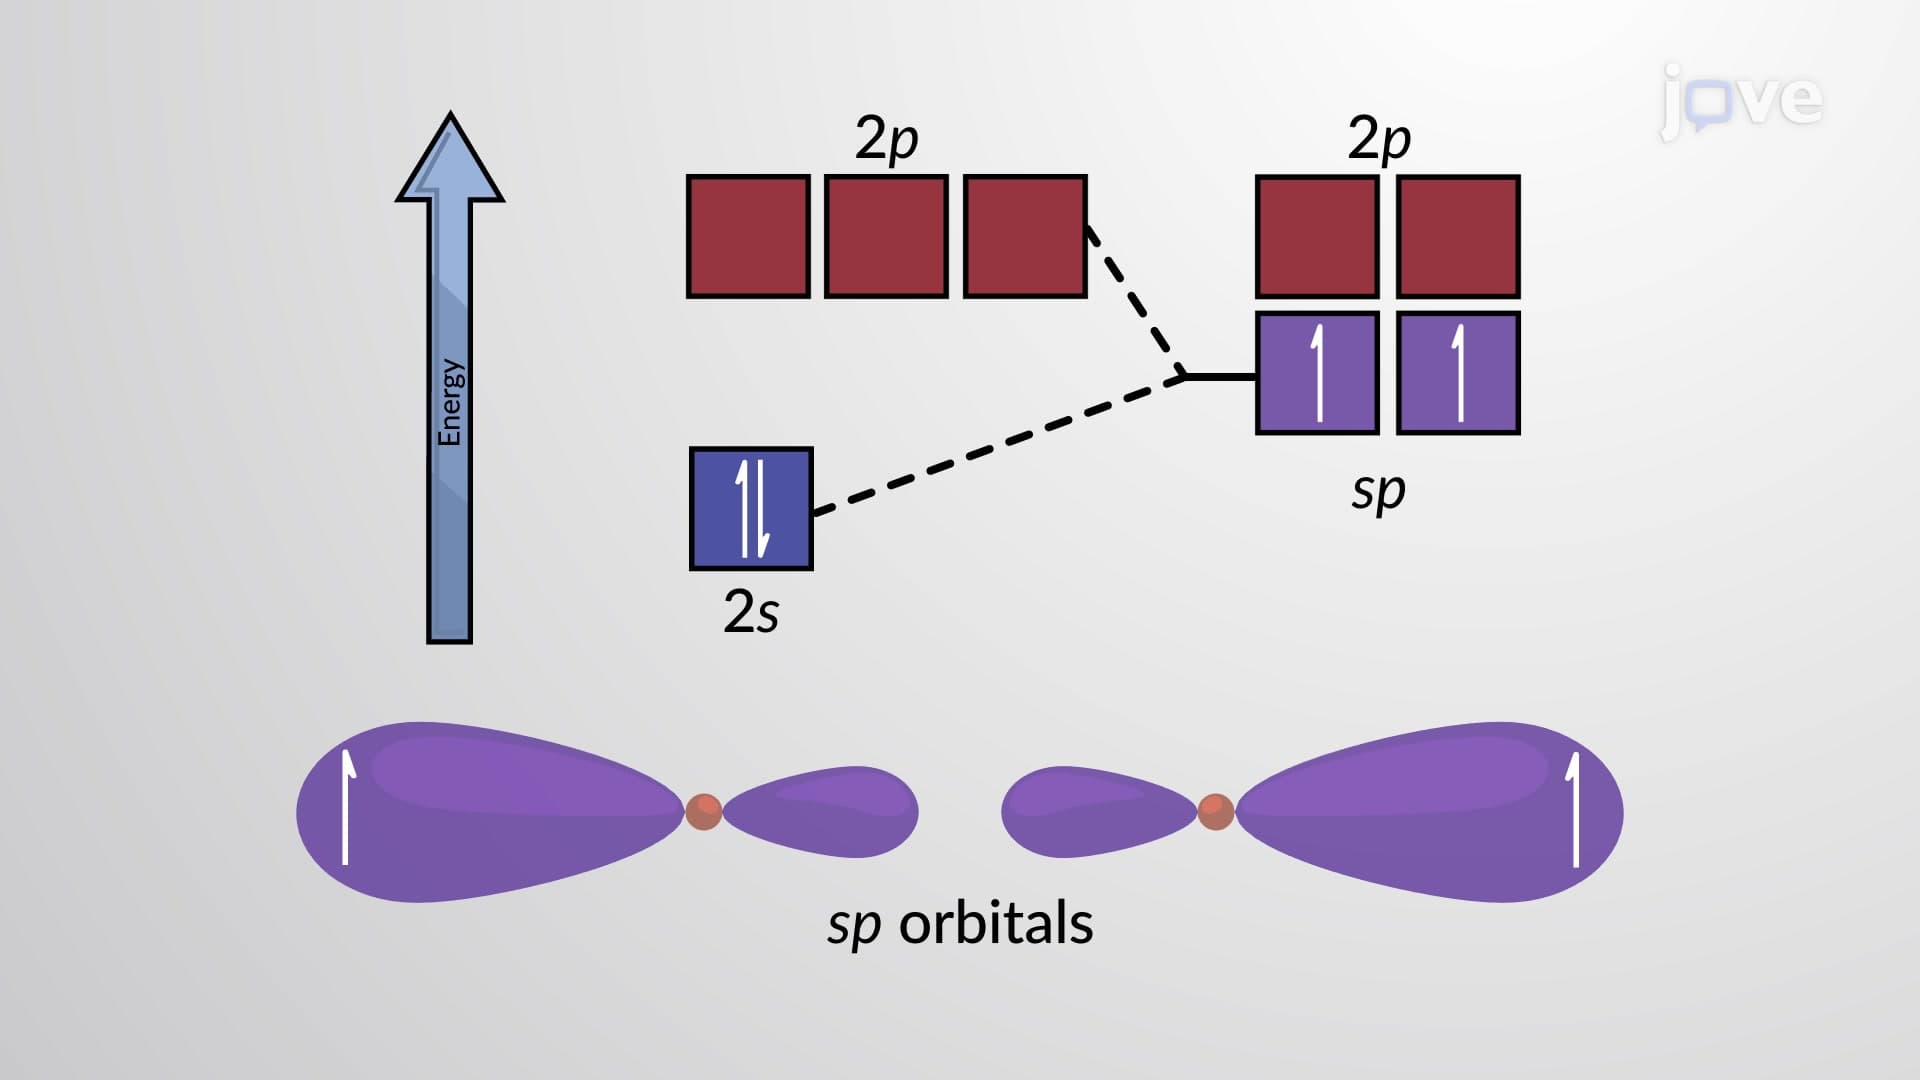
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:35
Hybridization of Atomic Orbitals II
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding