Related Experiment Video
Updated: May 21, 2025

An In Vitro Approach to Study Mitochondrial Dysfunction: A Cybrid Model
Published on: March 9, 2022
Natural History of Patients With Mitochondrial ATPase Deficiency Due to Pathogenic Variants of MT-ATP6 and MT-ATP8
Sara Carli1, Anna Levarlet1, Daria Diodato2
1Department of Medical and Surgical Sciences, Alma Mater Studiorum, University of Bologna, Italy.

Background And Objectives:
The mitochondrial DNA (mtDNA) genes MT-ATP6 and MT-ATP8 encode for subunits α and 8 (A6L) of the adenosine triphosphate synthase complex. Pathogenetic variants in MT-ATP6/8 cause incurable mitochondrial syndromes encompassing a wide spectrum of clinical features including ataxia, motor and language developmental delay, deafness, retinitis pigmentosa, and Leigh pattern in brain MRI. Typically, higher levels of mtDNA variants lead to more severe symptomatology although even individuals with similar mtDNA mutational loads exhibit high clinical variability. Hence, the establishment of potential therapeutics is currently challenging. In this article, we present an international multicenter study designed to provide a retrospective natural history of patients with MT-ATP6/8 deficiency and to identify primary and secondary end points for future clinical trials.
Methods:
Clinical, biochemical, and molecular genetics data of patients with genetically confirmed MT-ATP6/8 defects were collected and analyzed from Italian, German, US, and Spain national reference centers through ethical committee-approved mitochondrial patients' national registries or local programs.
Results:
A cohort of 111 patients, 98 unreported, were analyzed (55 male, 56 female). Patients had infantile-onset disease (<1 year) in 44% of cases, pediatric-onset (≥1 year and ≤12 years) in 36%, and late-onset (>12 years) in 20%. Kaplan-Meier analysis showed a significant difference (p value = 0.0349) in the survival of infantile and pediatric patients compared with adult patients, although only 8% of patients were not alive at the last follow-up. The CNS was the most frequently affected tissue (93%), followed by the muscle (75%), eye (46%), and heart (18%). Brain MRI showed isolated Leigh-like lesions (58%), Leigh-like lesions and cortical and/or cerebellar atrophy (15%), isolated cerebellar atrophy (10%), and other lesions (21%). At the last follow-up, 11% of patients were wheelchair-bound. Metabolic acidosis or acute deterioration complicated the clinical course in ≅55% of early-onset patients. Molecular genetics studies identified 26 pathogenic variants (6 of them novel). Reduced citrulline levels and increased alanine and lactate levels were reported in 56%, 49%, and 71% of patients, respectively, suggesting their role as potential biomarkers.
Discussion:
Our results define a more accurate classification based on the age at onset for MT-ATPase deficiency and provide fundamental clinical and biochemical data for disease management.
Insights
This study details the natural history of mitochondrial ATPase (MT-ATP6/8) deficiency, revealing age-at-onset impacts survival and identifying potential biomarkers for better disease management and future clinical trials.
Area of Science:
- Genetics and Molecular Biology
- Neurology
- Biochemistry
Background:
- Mitochondrial DNA (mtDNA) variants in MT-ATP6/8 genes cause severe, incurable mitochondrial syndromes.
- Clinical variability is high, complicating therapeutic development.
- Understanding natural history is crucial for identifying clinical trial endpoints.
Purpose of the Study:
- To establish the retrospective natural history of patients with MT-ATP6/8 deficiency.
- To identify primary and secondary endpoints for future clinical trials.
- To improve classification and management of MT-ATPase deficiency.
Main Methods:
- International multicenter study analyzing clinical, biochemical, and molecular genetics data.
- Data collected from national registries and local programs across Italy, Germany, US, and Spain.
- Ethical committee approval obtained for patient data analysis.
Main Results:
- 111 patients analyzed (98 unreported), with infantile-onset disease in 44%.
- Significant survival differences observed based on age at onset (p=0.0349).
- Central nervous system (93%) and muscle (75%) were most affected; potential biomarkers identified (citrulline, alanine, lactate).
Conclusions:
- Age at onset provides a more accurate classification for MT-ATPase deficiency.
- Fundamental clinical and biochemical data are established for disease management.
- Study provides a foundation for future therapeutic strategies and clinical trials.
More Related Videos
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase COX/SDH Double-labeling Histochemistry
Published on: November 23, 2011
 07:24
Genotyping Single Nucleotide Polymorphisms in the Mitochondrial Genome by Pyrosequencing
Published on: February 10, 2023
Related Concept Videos
 01:48
ATP Synthase: Mechanism
 01:18
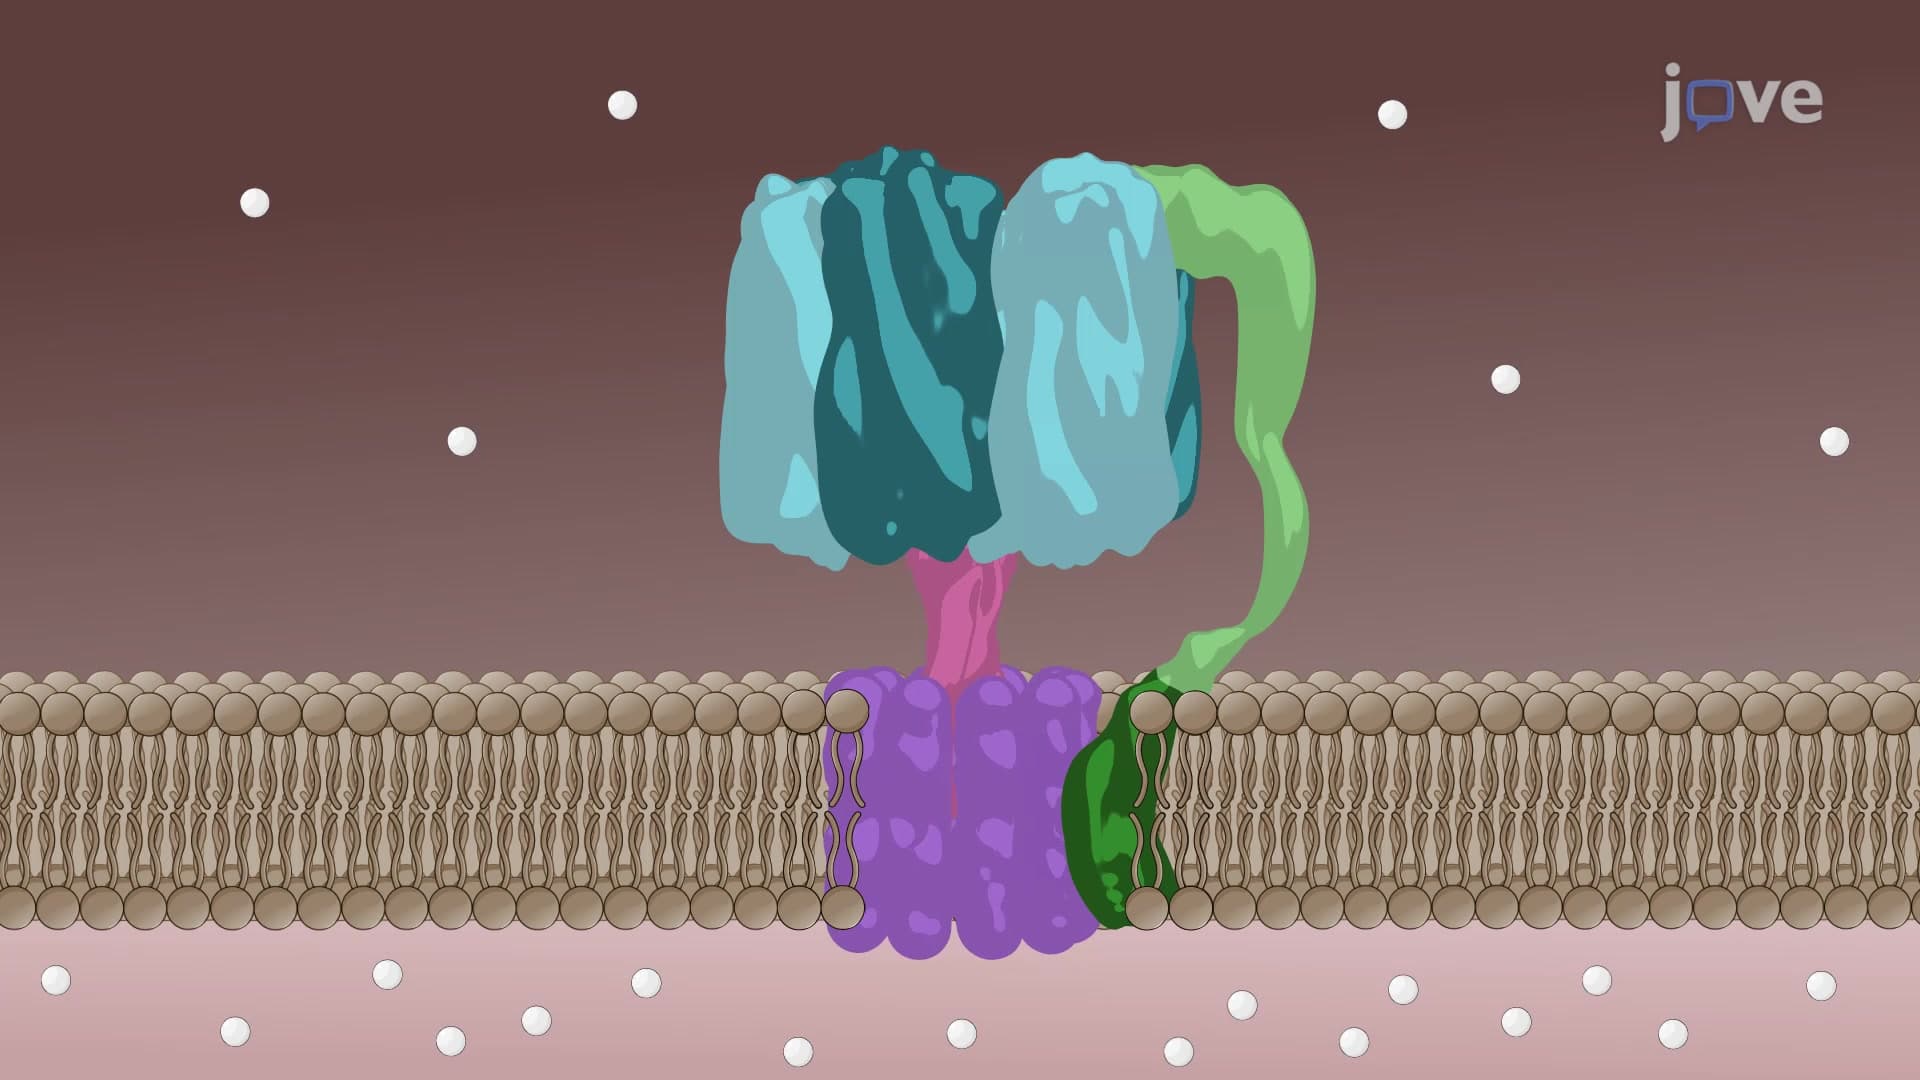
ATP Synthase: Structure
 01:32
Chemiosmosis
Electron Transport Chain
The electron transport chain involves a series of protein complexes on the inner mitochondrial membrane that undergo a series of redox reactions. At the end of this chain, the electrons...
 02:59
Animal Mitochondrial Genetics
 01:42
The ADP/ATP Carrier Protein
 01:28
The Inner Mitochondrial Membrane