Related Experiment Video
Updated: Jul 21, 2026

A Method for Screening and Validation of Resistant Mutations Against Kinase Inhibitors
Published on: December 7, 2014
Computational insights of deucravacitinib's selectivity for TYK2 pseudokinase vs. JAK kinase domain via molecular
Manish Ramchandani1, Amit Kumar Goyal1
1Department of Pharmacy, School of Chemical Sciences, Central University of Rajasthan, Ajmer, India.

Abstract:
Deucravacitinib (Sotyktu) stands out as a novel and highly specific oral inhibitor targeting tyrosine kinase 2 (TYK2). Its mechanism of action involves an allosteric binding, to catalytically inactive pseudokinase domain of TYK2, this stabilizes an inhibitory contact between the catalytic and regulatory domains. This inhibition of Janus kinase (JAK) is associated with suppression of cytokine signaling using diverse molecules defining wide importance in current research. In our recent investigation, we examined the selectivity of the TYK2JH2 inhibitor, deucravacitinib, against four JAK kinases (JAK1, JAK2, JAK3, TYK2) and TYK2 pseudokinases utilizing a merged approach involving molecular docking, molecular dynamics analysis (300 ns), and binding free energy calculation through the molecular mechanics Poisson - Boltzmann surface area (MM-PBSA) scheme. The results obtained indicate that deucravacitinib effectively interacts with the ATP-binding site of four JAK kinases and TYK2 pseudokinase through hydrogen bond formation, electrostatic attraction, and notably, van der Waals interaction. We found the calculated binding affinity demonstrates a reduction in the TYK2JH2-deucravacitinib complex due to an increased favorable intermolecular electrostatic contribution. Consequently, deucravacitinib exhibits greater selectivity for the TYK2 pseudokinase domain compared to the other four JAKs. Moreover, the interaction with DPG motif residues and the hinge region contributed to the stabilization of deucravacitinib through robustly formed hydrogen bonds. The interaction with the hydrophobic catalytic region caused the ATP-binding site to adopt a closed conformation, thereby minimizing protein movement at the glycine loop of the JAK pseudokinase protein. In summary, our study holds significant potential for informing the strategic design of TYK2 inhibitors with enhanced affinity.
Related Concept Videos
 01:26
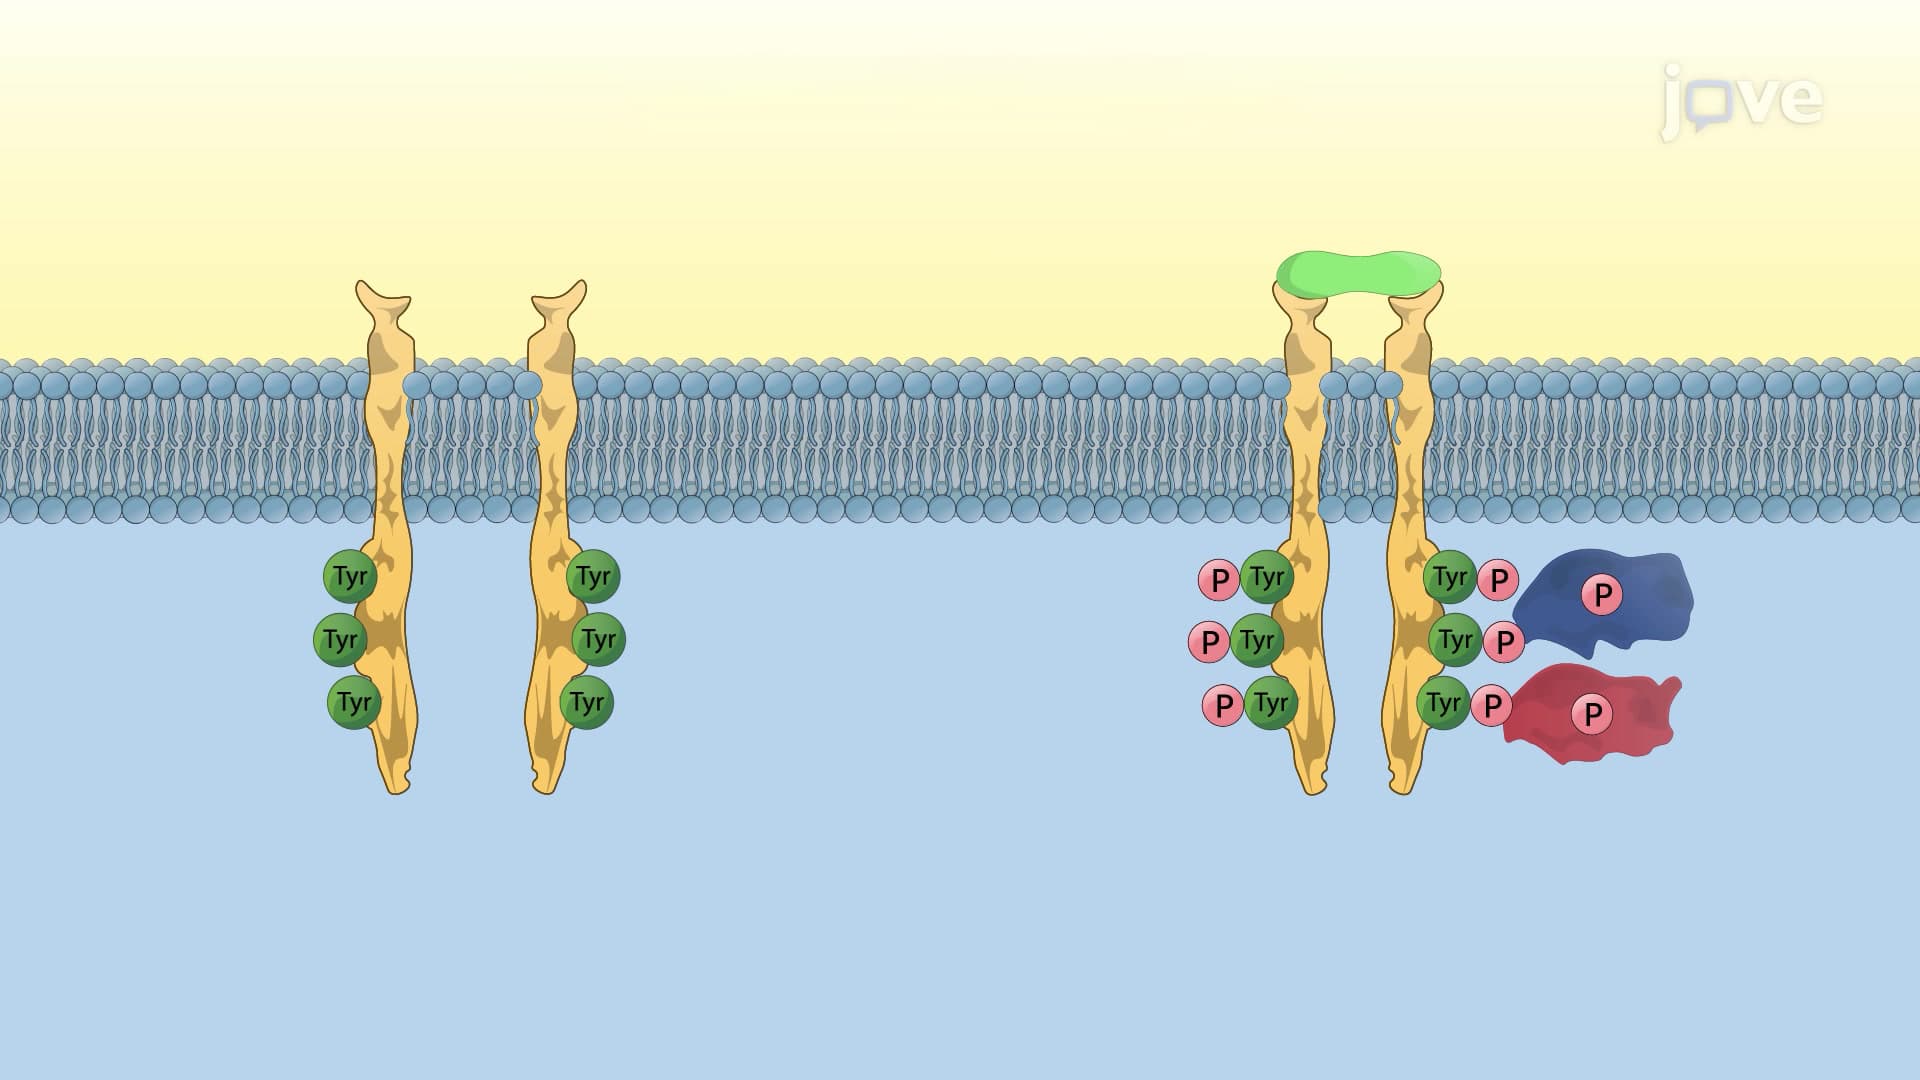
Receptor Tyrosine Kinases
 01:16
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:22
Protein-Drug Binding: Determination Methods
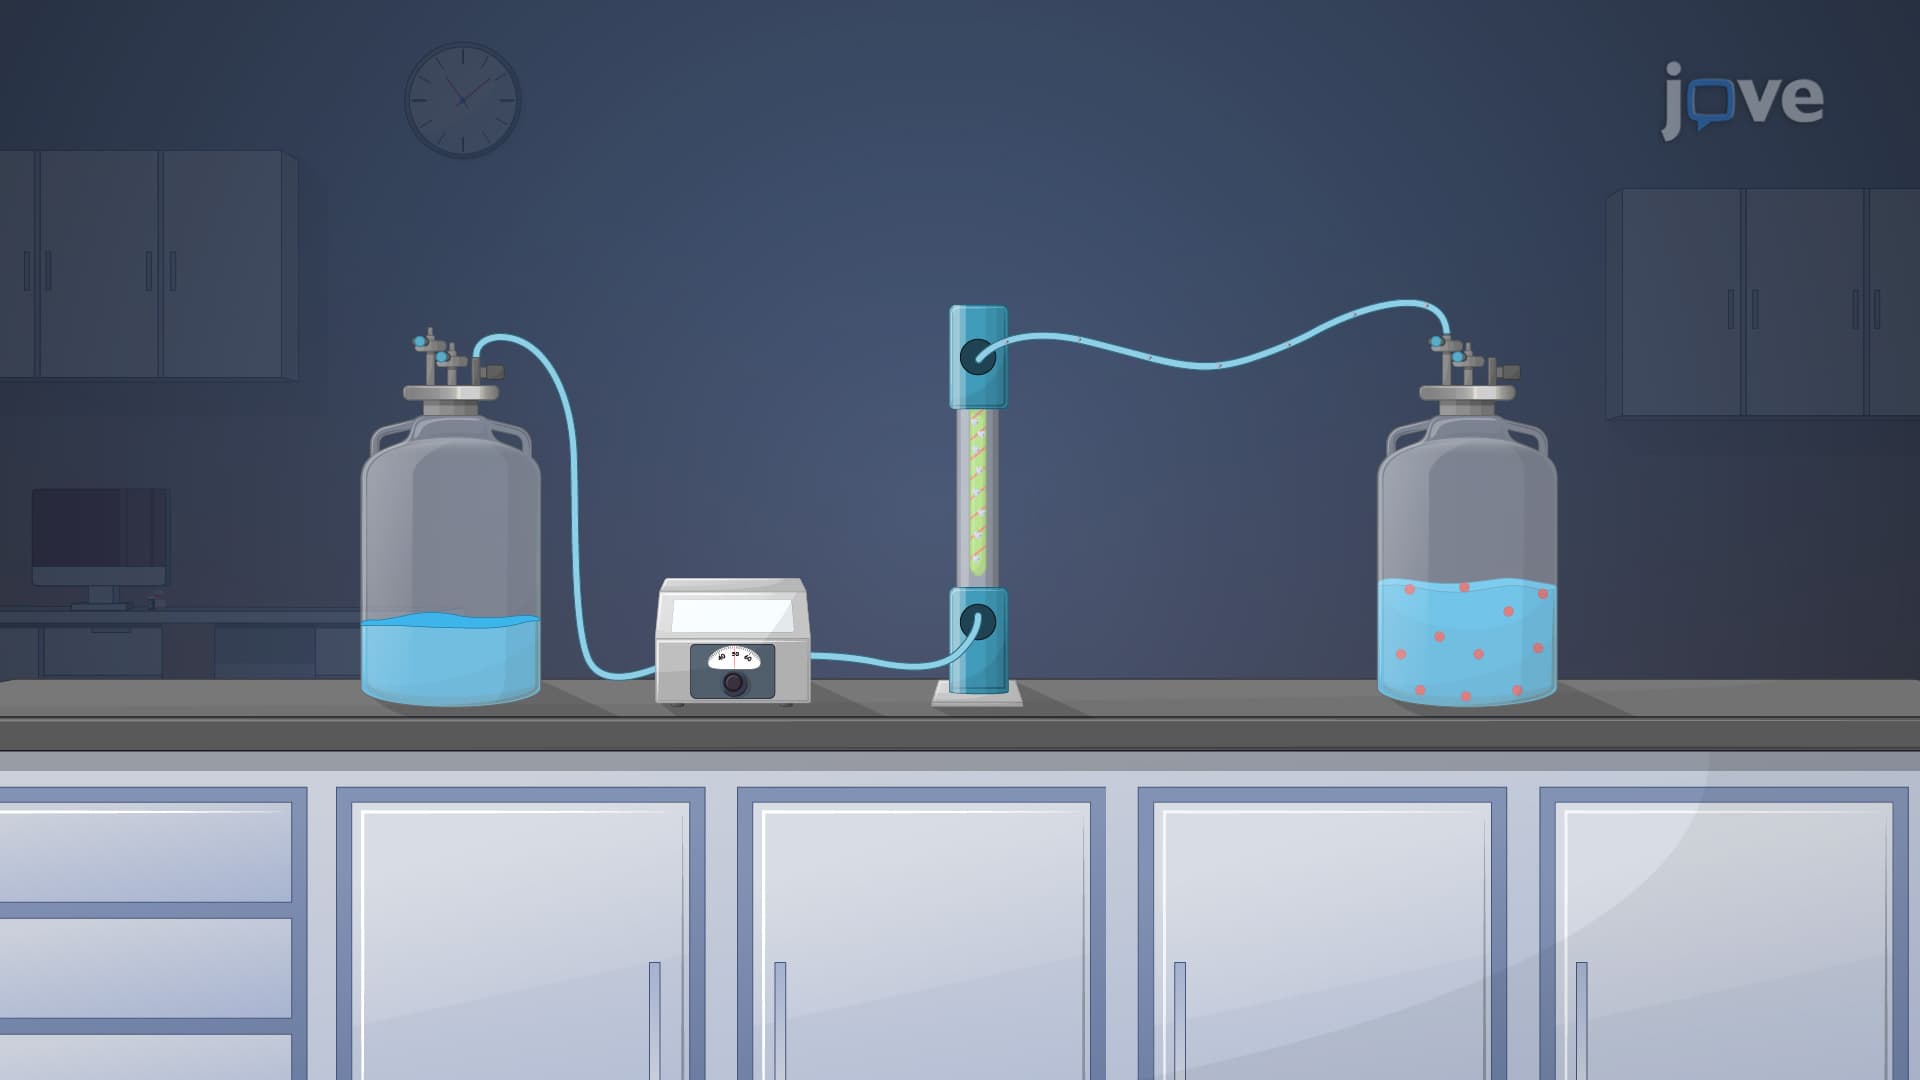
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:26
Pharmacokinetic Models: Comparison and Selection Criterion
Physiological models take a detailed approach by considering specific molecular processes. They can predict drug distribution, metabolism, and elimination changes, providing a comprehensive understanding of how drugs interact with the body.
 01:27
Pharmacodynamic Models: Overview
 01:15
Impact of Pharmacokinetic–Pharmacodynamic Models: Regulatory Decisions