Related Experiment Video
Updated: May 10, 2025

Monitoring Stub1-Mediated Pexophagy
Published on: May 12, 2023
Mitochondria and Peroxisome Crosstalk in Peroxisome Biogenesis Disorder 8A Caused by a Rare Variant in PEX16 Gene
Mohamad Wehbe1, Rudy N Zalzal1, Riyad El-Khoury2
1Faculty of Medicine, American University of Beirut, Beirut, Lebanon.

Abstract:
Peroxisome biogenesis disorder 8A is a rare autosomal recessive disorder caused by mutations in the PEX16 gene. We report the clinical, biochemical, and molecular features of a patient harboring the homozygous NM_004813.4: c.526C>T, p.(Arg176*) mutation in PEX16 associated with mitochondrial dysfunction. This newborn presented with microcephaly, encephalopathy, hypotonia, failure to thrive, hepatomegaly, and abnormal retinal pigmentation. He had elevated plasma very long-chain fatty acids. Skeletal muscle biopsy revealed significant mitochondrial depletion with deficiencies of the respiratory chain Complexes I-IV, with significant reductions in cytochrome c oxidase and citrate synthase activity. The peroxisome biogenesis disorder 8A was confirmed by whole genome sequencing. This is the first case delineating the association of mitochondrial dysfunction with peroxisome biogenesis disorder 8A caused by the above mutation. Further studies are needed to elucidate the underlying pathophysiological mechanisms of mitochondria and peroxisome crosstalk.
Insights
This study details a rare peroxisome biogenesis disorder 8A case linked to PEX16 gene mutation and severe mitochondrial dysfunction. The findings highlight a novel association between these two conditions in affected newborns.
Area of Science:
- Genetics
- Biochemistry
- Neurology
Background:
- Peroxisome biogenesis disorder 8A is a rare autosomal recessive condition.
- It is caused by mutations in the PEX16 gene.
- Mitochondrial dysfunction is a critical cellular process.
Purpose of the Study:
- To report the clinical, biochemical, and molecular characteristics of a patient with PEX16 mutation.
- To investigate the association between peroxisome biogenesis disorder 8A and mitochondrial dysfunction.
- To confirm the diagnosis using whole genome sequencing.
Main Methods:
- Clinical examination and biochemical assays (plasma very long-chain fatty acids).
- Skeletal muscle biopsy to assess mitochondrial morphology and respiratory chain enzyme activity.
- Whole genome sequencing for molecular confirmation.
Main Results:
- A newborn presented with microcephaly, encephalopathy, hypotonia, failure to thrive, hepatomegaly, and retinal abnormalities.
- Elevated plasma very long-chain fatty acids were detected.
- Muscle biopsy showed mitochondrial depletion and deficiencies in respiratory chain complexes I-IV.
Conclusions:
- The study confirms peroxisome biogenesis disorder 8A in a patient with a novel PEX16 mutation (c.526C>T, p.(Arg176*)).
- This is the first reported case associating this specific PEX16 mutation with significant mitochondrial dysfunction.
- Further research is required to understand the crosstalk between mitochondria and peroxisomes in this disorder.
Related Concept Videos
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:24
Peroxisomes
 01:30
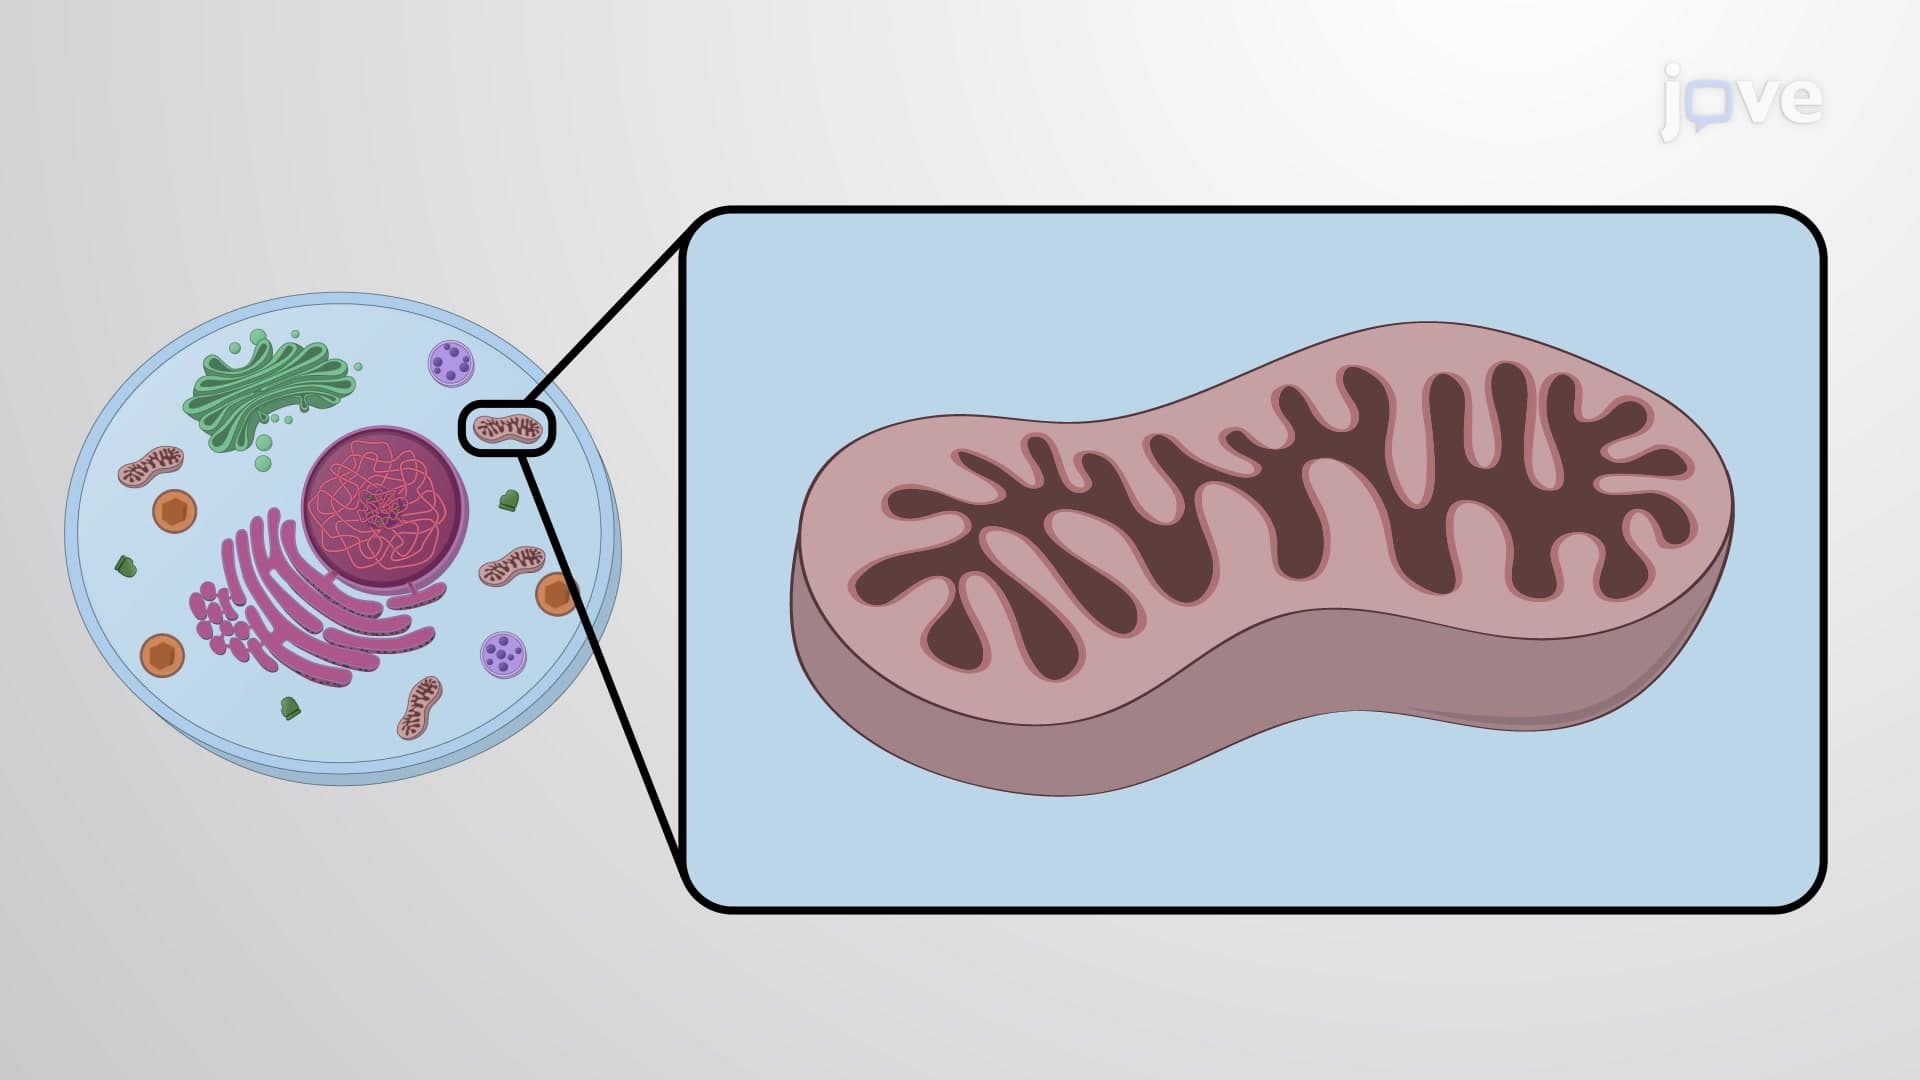
Peroxisomes and Mitochondria
The peroxisome is a single membrane-bound cellular organelle that can perform several different functions, including lipid metabolism and chemical detoxification. The enzymes within...
 01:19
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:20
Inborn Errors of Metabolism
 02:59
Animal Mitochondrial Genetics