Related Experiment Video
Updated: Jun 13, 2026

Low Molecular Weight Protein Enrichment on Mesoporous Silica Thin Films for Biomarker Discovery
Published on: April 17, 2012
Combining Mass Spectrometry with Machine Learning to Identify Novel Protein Signatures: The Example of Multisystem
Jeisac Guzmán Rivera1, Haiyan Zheng2, Benjamin Richlin3
1Public Health Research Institute, Rutgers New Jersey Medical School, Rutgers Biomedical and Health Sciences, Newark, NJ.

Objectives:
We demonstrate an approach that integrates biomarker analysis with machine learning to identify protein signatures, using the example of SARS-CoV-2-induced Multisystem Inflammatory Syndrome in Children (MIS-C).
Methods:
We used plasma samples collected from subjects diagnosed with MIS-C and compared them first to controls with asymptomatic/mild SARS-CoV-2 infection and then to controls with pneumonia or Kawasaki disease. We used mass spectrometry to identify proteins. Support vector machine (SVM) algorithm-based classification schemes were used to analyze protein pathways. We assessed diagnostic accuracy using internal and external cross-validation.
Results:
Proteomic analysis of a training dataset containing MIS-C (N=17), and asymptomatic/mild SARS-CoV-2 infected control samples (N=20) identified 643 proteins, of which 101 were differentially expressed. Plasma proteins associated with inflammation and coagulation increased and those associated with lipid metabolism decreased in MIS-C relative to controls. The SVM machine learning algorithm identified a three-protein model (ORM1, AZGP1, SERPINA3) that achieved 90.0% specificity, 88.2% sensitivity, and 93.5% area under the curve (AUC) distinguishing MIS-C from controls in the training set. Performance was retained in the validation dataset utilizing MIS-C (N=17) and asymptomatic/mild SARS-CoV-2 infected control samples (N=10) (90.0% specificity, 84.2% sensitivity, 87.4% AUC). We next replicated our approach to compare MIS-C with similarly presenting syndromes, such as pneumonia (N=17) and Kawasaki Disease (N=13) and found a distinct three-protein signature (VWF, SERPINA3, and FCGBP) that accurately distinguished MIS-C from the other conditions (97.5% specificity, 89.5% sensitivity, 95.6% AUC). We also developed a software tool that may be used to evaluate other protein pathway signatures using our data.
Conclusions:
We used MIS-C, a novel hyperinflammatory illness, to demonstrate that the use of mass spectrometry to identify candidate plasma proteins followed by machine learning, specifically SVM, is an efficient strategy for identifying and evaluating biomarker signatures for disease classification.
More Related Videos
 10:01
Combining Chemical Cross-linking and Mass Spectrometry of Intact Protein Complexes to Study the Architecture of Multi-subunit Protein Assemblies
Published on: November 28, 2017
 00:07
Quantification of Protein Interaction Network Dynamics using Multiplexed Co-Immunoprecipitation
Published on: August 21, 2019
Related Concept Videos
 01:19
Mass Spectrometry: Overview
 01:19
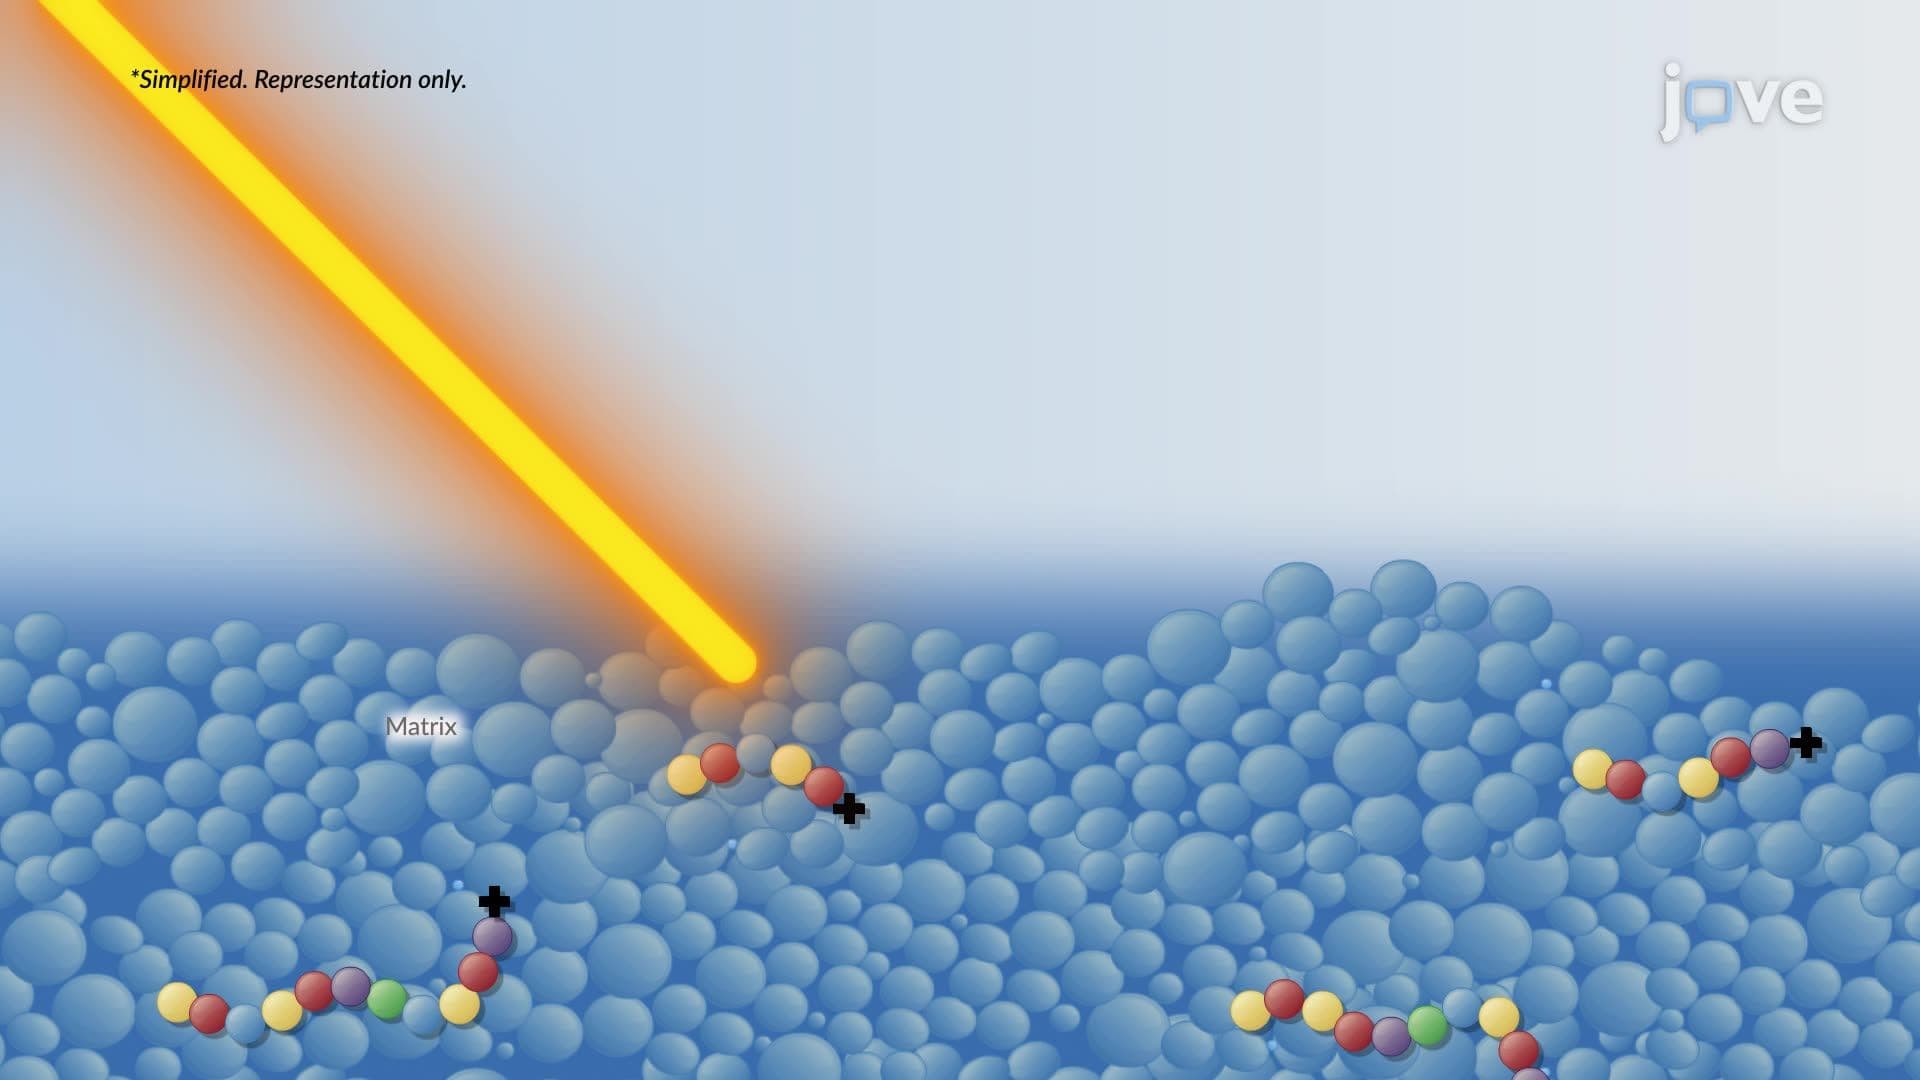
MALDI-TOF Mass Spectrometry
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:21
Mass Spectrometry: Complex Analysis
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
 01:21
Tandem Mass Spectrometry
 01:25
Rapid Identification of Pathogens