Related Experiment Video
Updated: Sep 20, 2025

Synthesis and Bioconjugation of Thiol-Reactive Reagents for the Creation of Site-Selectively Modified Immunoconjugates
Published on: March 6, 2019
The Impact of Conjugation Mode and Site on Tubulysin Antibody-Drug-Conjugate Efficacy and Stability
Sayumi Yamazoe1, Qinqin Cheng1, Srikanth Kotapati1
1Bristol Myers Squibb, Research and Development, Redwood City, California, 94063, United States.

Abstract:
Antibody-drug conjugates (ADCs) represent a prominent class of biotherapeutics engineered to selectively deliver cytotoxic payloads to tumors, thereby facilitating targeted cell killing. While first-generation ADCs, created by conjugating payloads to surface-accessible lysine or hinge-cysteine residues, have achieved clinical success, several site-specific ADCs with defined drug-to-antibody ratios are currently under clinical investigation. Herein, the efficacy, stability, and pharmacokinetics of ADCs generated by attaching the drug linker to surface-exposed lysine residues, hinge-cysteine residues, and the C'E loop in the CH2 domain (mediated by bacterial transglutaminase) using a tubulysin payload are compared. In N87 xenograft mice, the order of efficacy is C'E loop > hinge-cysteine > lysine-conjugated ADCs. Among the three ADCs evaluated, the site-specific ADC demonstrates superior in vivo stability (minimal payload-linker deconjugation and limited payload metabolism/deacetylation) and favorable pharmacokinetics (longer half-life, low clearance, high exposure). In contrast, the lysine-conjugated ADC exhibits the least stability and poorest pharmacokinetics, which directly correlate with its efficacy. Further investigation into cysteine-engineered site-specific ADCs with payloads conjugated at various sites confirms that both the conjugation chemistry and the site of conjugation significantly influence the in vivo stability and pharmacokinetics of site-specific ADCs.
Insights
Site-specific antibody-drug conjugates (ADCs) show improved efficacy and stability compared to lysine-conjugated ADCs. Conjugation site significantly impacts ADC performance, with C'E loop conjugation demonstrating superior results in vivo.
Area of Science:
- Biotechnology
- Pharmacology
- Oncology
Background:
- Antibody-drug conjugates (ADCs) are advanced biotherapeutics for targeted cancer therapy.
- Traditional ADCs use lysine or cysteine conjugation, but site-specific methods offer improved precision.
- Site-specific ADCs aim for defined drug-to-antibody ratios and enhanced therapeutic profiles.
Purpose of the Study:
- To compare the efficacy, stability, and pharmacokinetics of ADCs with different conjugation sites (lysine, hinge-cysteine, C'E loop) using a tubulysin payload.
- To evaluate the impact of conjugation chemistry and site on in vivo ADC performance.
- To identify optimal conjugation strategies for next-generation ADCs.
Main Methods:
- Generation of ADCs by conjugating a tubulysin payload to lysine residues, hinge-cysteine residues, and the C'E loop via bacterial transglutaminase.
- In vivo efficacy studies using N87 xenograft mouse models.
- Assessment of in vivo stability, including payload-linker deconjugation and payload metabolism.
- Pharmacokinetic analysis to determine half-life, clearance, and drug exposure.
Main Results:
- ADCs conjugated at the C'E loop demonstrated the highest efficacy in N87 xenograft mice, followed by hinge-cysteine and then lysine-conjugated ADCs.
- The site-specific ADC exhibited superior in vivo stability and favorable pharmacokinetics (longer half-life, low clearance, high exposure).
- Lysine-conjugated ADCs showed the least stability and poorest pharmacokinetics, correlating with lower efficacy.
Conclusions:
- The site of drug conjugation significantly influences the in vivo stability, pharmacokinetics, and overall efficacy of antibody-drug conjugates.
- Site-specific conjugation, particularly at the C'E loop, offers a promising strategy for developing more effective and stable ADCs.
- Further research into cysteine-engineered site-specific ADCs confirms the critical role of conjugation chemistry and site selection.
More Related Videos
 11:02
Genetic Encoding of a Non-Canonical Amino Acid for the Generation of Antibody-Drug Conjugates Through a Fast Bioorthogonal Reaction
Published on: September 14, 2018
 11:58
Initial Evaluation of Antibody-conjugates Modified with Viral-derived Peptides for Increasing Cellular Accumulation and Improving Tumor Targeting
Published on: March 8, 2018
Related Concept Videos
 01:14
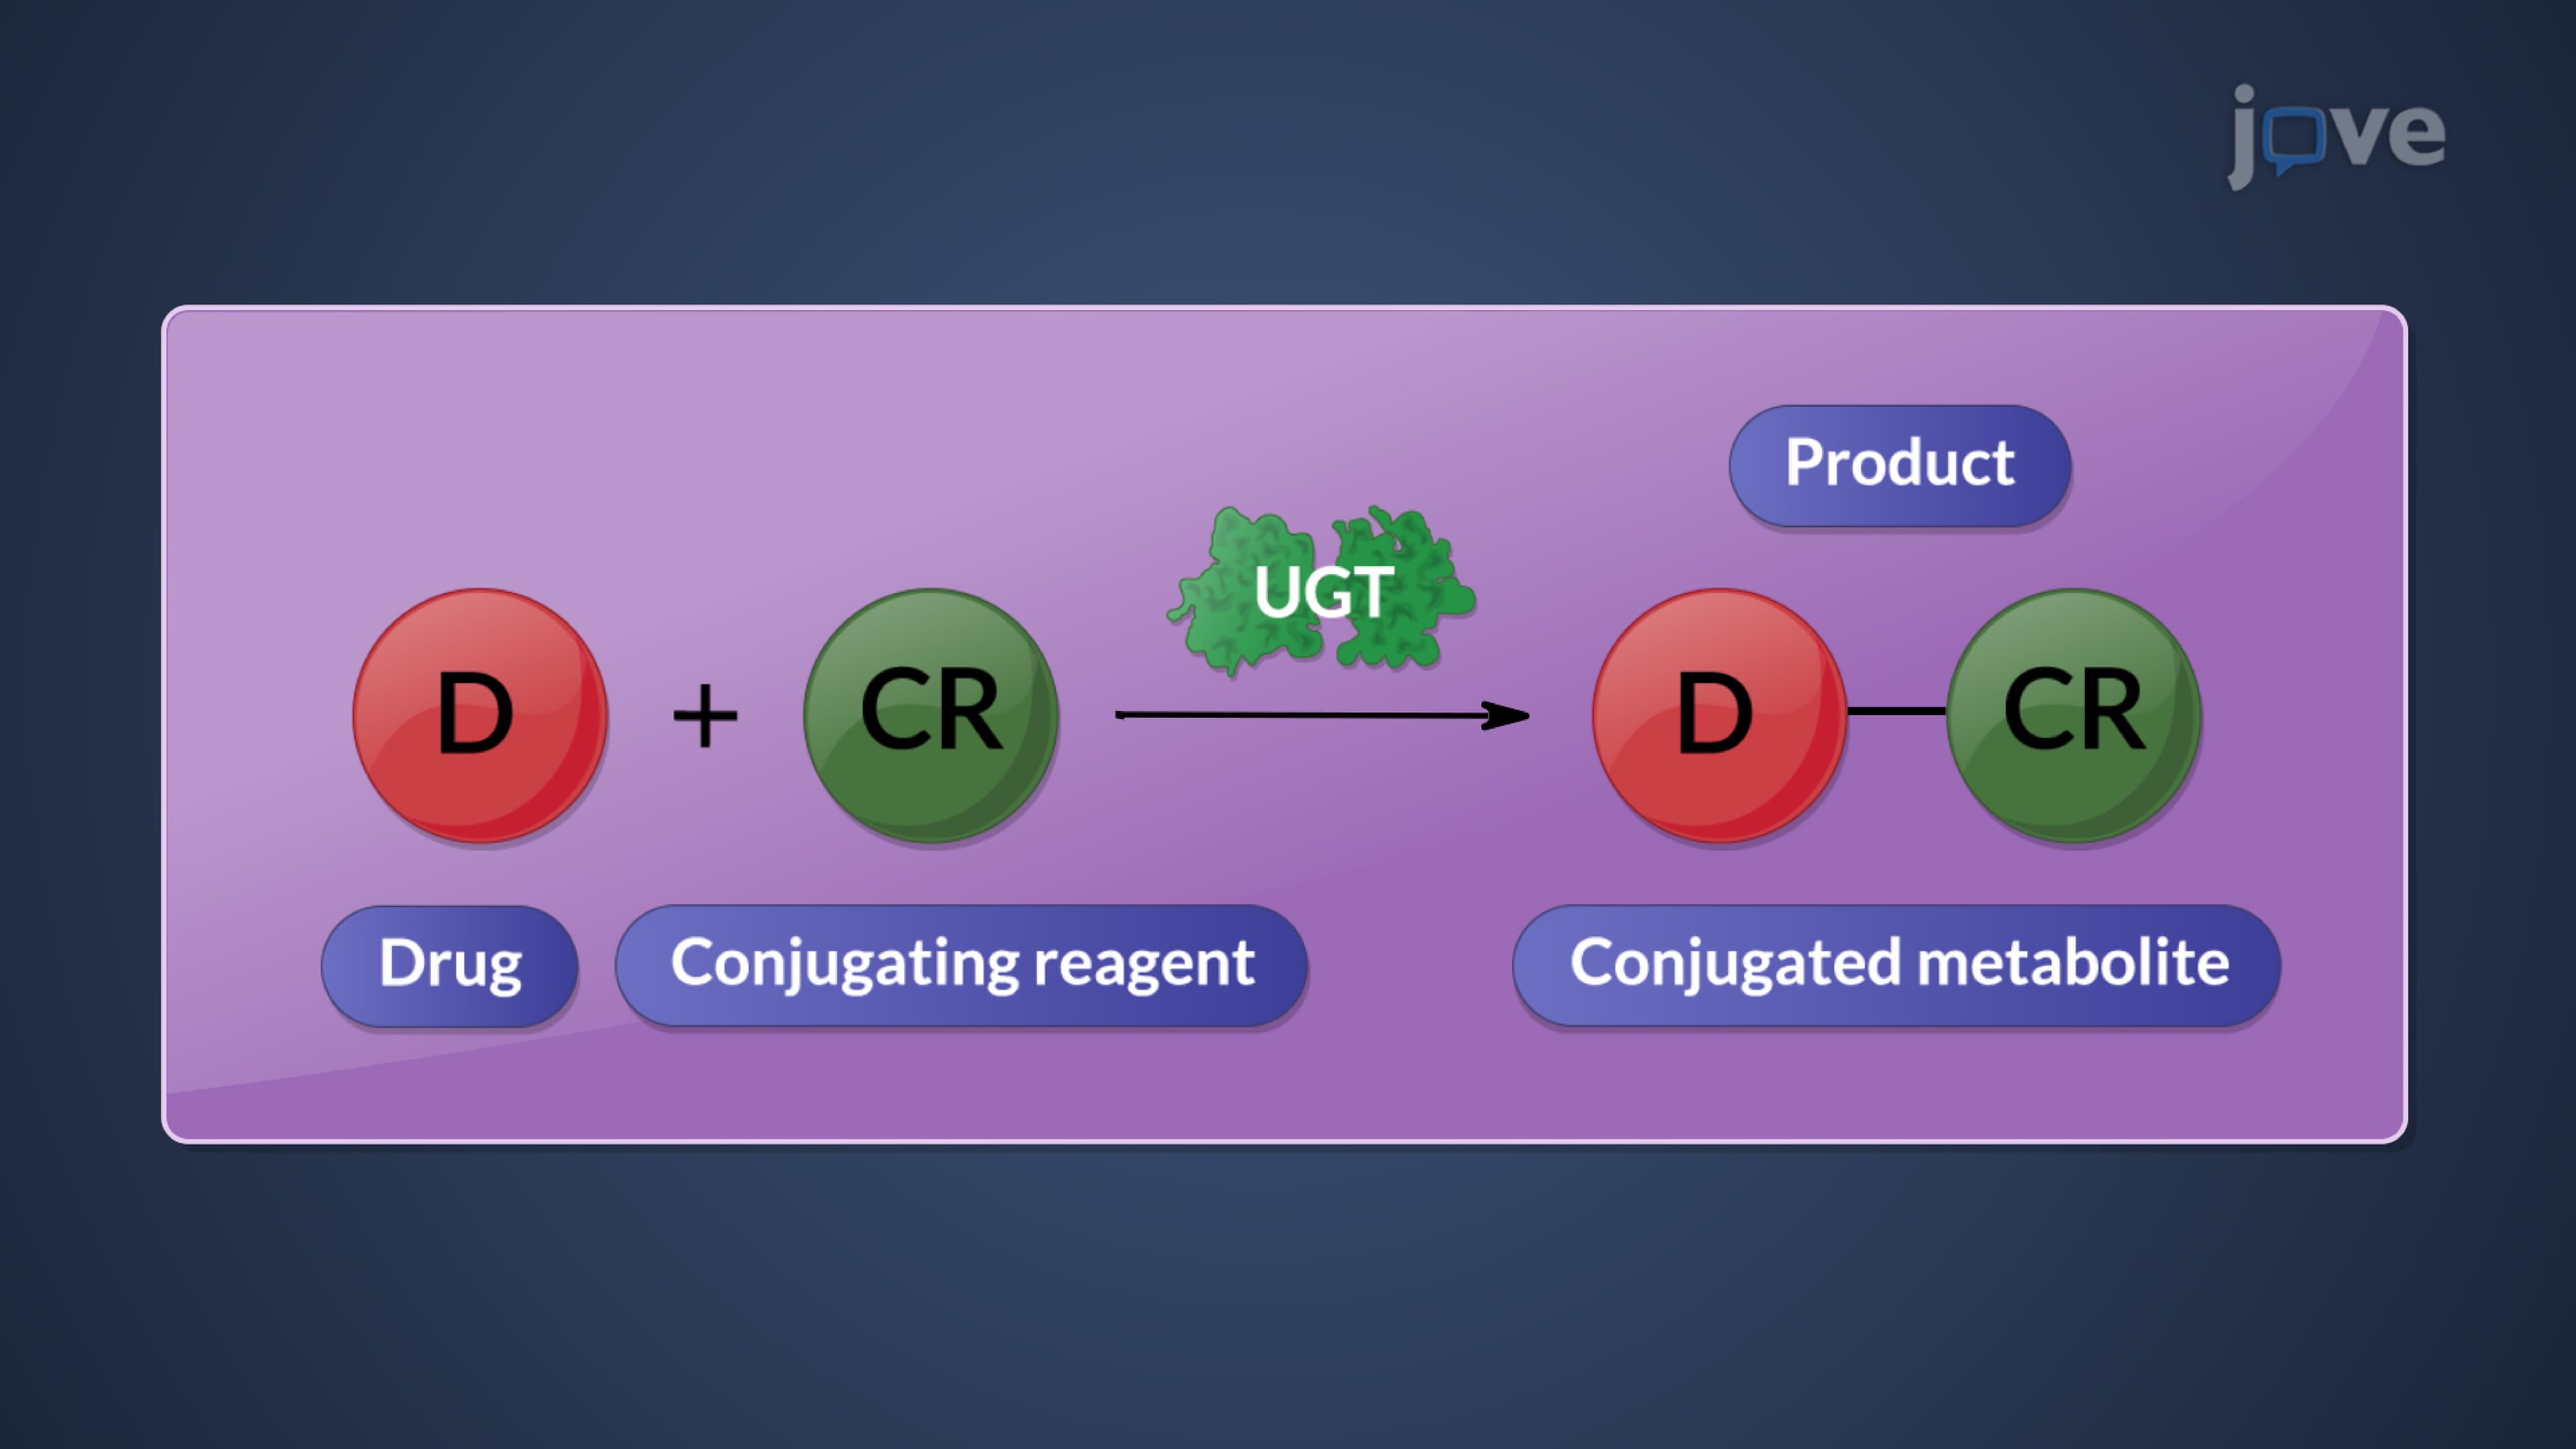
Phase II Conjugation Reactions: Overview
 01:14
Drug Metabolism: Phase II Reactions
 01:19
Phase II Reactions: Sulfation and Conjugation with α-Amino Acids
 01:20
Factors Affecting Dissolution: Drug Permeability, Stability and Stereochemistry
 01:15
Drug Elimination by Renal Route: Tubular Secretion