Related Experiment Video
Updated: Sep 18, 2025
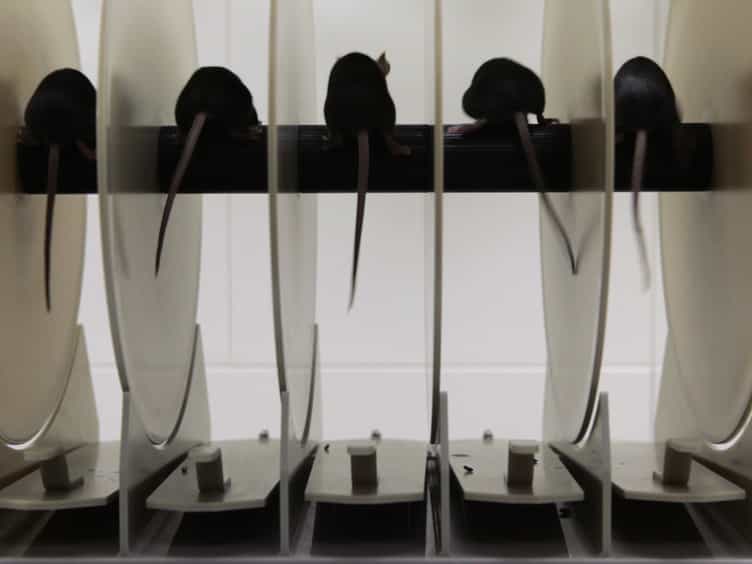

Assessing Functional Performance in the Mdx Mouse Model
Published on: March 27, 2014
Fbxl3 deletion mitigates myopathy in mdx mice through upregulation of myogenin
1Department of Cardiology, Suzhou Ninth People's Hospital, Suzhou Ninth Hospital Affiliated to Soochow University, 215200, Suzhou, PR China.

Abstract:
Duchenne muscular dystrophy (DMD) is a severe X-linked neuromuscular disorder with limited therapeutic options, highlighting the urgent need for novel treatment strategies. In this study, we investigated the role of FBXL3 in DMD pathogenesis and assessed its potential as a gene therapy target. Using mdx mice, a well-established preclinical model of DMD, we found that satellite cell-specific deletion of FBXL3 significantly improved muscle pathology and functional performance. FBXL3-deficient mdx mice exhibited increased body and muscle mass, along with enhanced grip strength and endurance capacity. Histological analyses demonstrated a marked increase in both the number and cross-sectional area of centrally nucleated fibers, indicative of enhanced regenerative activity. These changes were associated with elevated myogenin expression and reduced inflammation and fibrosis, suggesting that FBXL3 functions as a negative regulator of muscle repair. Moreover, targeted FBXL3 silencing via adeno-associated virus (AAV) delivery to the gastrocnemius muscle resulted in increased muscle mass and further upregulation of myogenin, supporting its therapeutic relevance. Together, these findings identify FBXL3 as a key modulator of muscle regeneration via repression of myogenin and provide compelling evidence for its inhibition as a promising gene therapy strategy in DMD.
More Related Videos
 14:10
Isometric and Eccentric Force Generation Assessment of Skeletal Muscles Isolated from Murine Models of Muscular Dystrophies
Published on: January 31, 2013
 06:52
Behavioral and Locomotor Measurements Using an Open Field Activity Monitoring System for Skeletal Muscle Diseases
Published on: September 29, 2014
