Related Experiment Video
Updated: Sep 18, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
On the functional dependence of transition-potential coupled cluster
Alexis Antoinette Ann Delgado1, Devin A Matthews1
1Department of Chemistry, Southern Methodist University, Dallas, Texas 75275, USA.

Abstract:
Orbital relaxation of the core region is a primary source of error in the computation of core ionization and core excitation energies. Recently, Transition-Potential Coupled Cluster (TP-CC) methods have been used to explicitly treat orbital relaxation using non-variational molecular orbitals determined by reoccupation of orbitals optimized for a fractional core occupation. The amount of fractional occupation is governed by parameter λ, and recommended values for accurate TP-CCSD and XTP-CCSD computations of carbon, nitrogen, oxygen, and fluorine K edges were previously determined. Herein, we explore the performance of several density functionals for generating the fractionally occupied orbitals used in TP-CCSD. These functionals include HF, BP86, BH&HLYP, B3LYP, M06-2X, and ωB97m-V. The fractionally occupied orbitals computed across the various functionals were subsequently employed as the initial orbitals for our TP-CCSD calculations of organic K-edge x-ray absorption and photoelectron spectra. Regardless of the functional used to generate the fractionally occupied orbitals, the TP-CCSD calculations yield accurate and comparable core ionization energies, core excitation energies, and oscillator strengths.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 01:27
¹H NMR: Long-Range Coupling
In alkenes, spin information is communicated via σ–π overlap, as seen in allylic (four-bond) and homoallylic (five-bond) couplings. These coupling interactions are stronger when the σ bond is parallel to the alkene...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:22
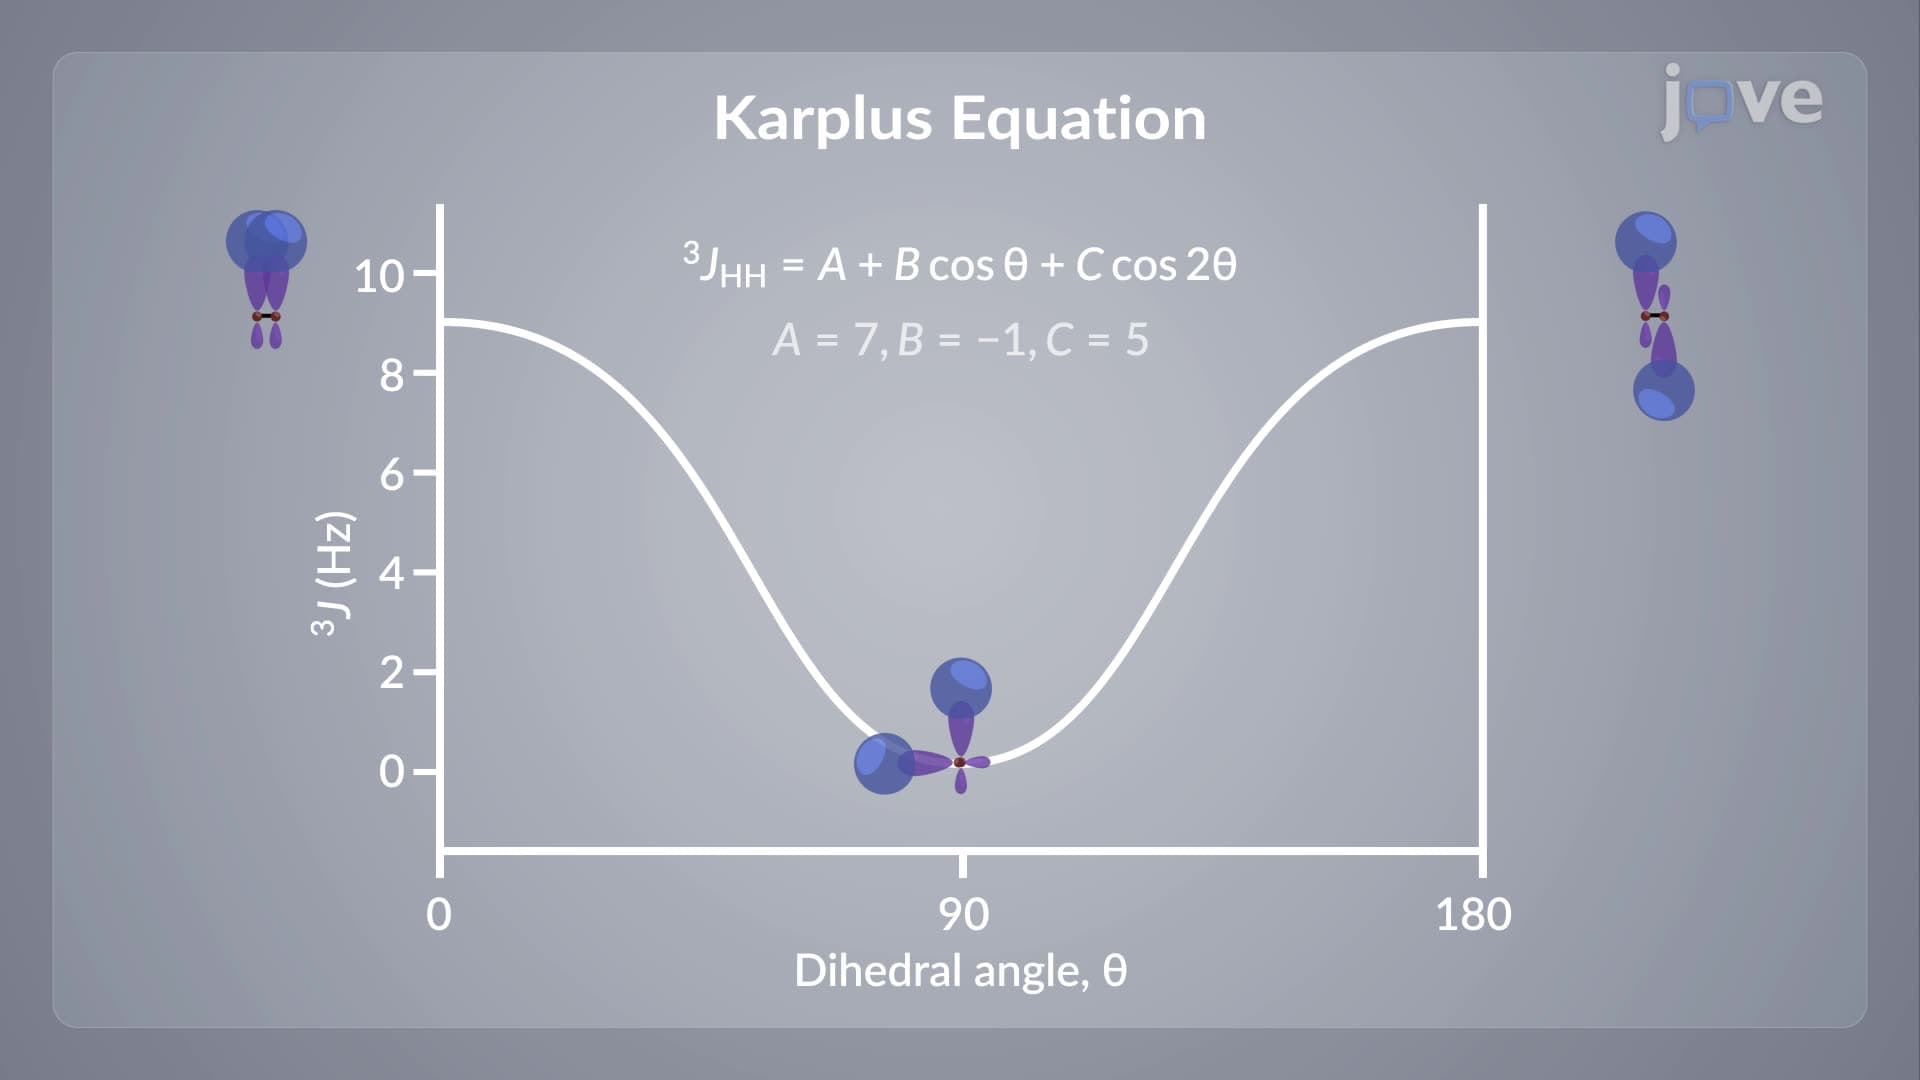
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...
 02:40
MO Theory and Covalent Bonding