Related Experiment Video
Updated: Sep 17, 2025
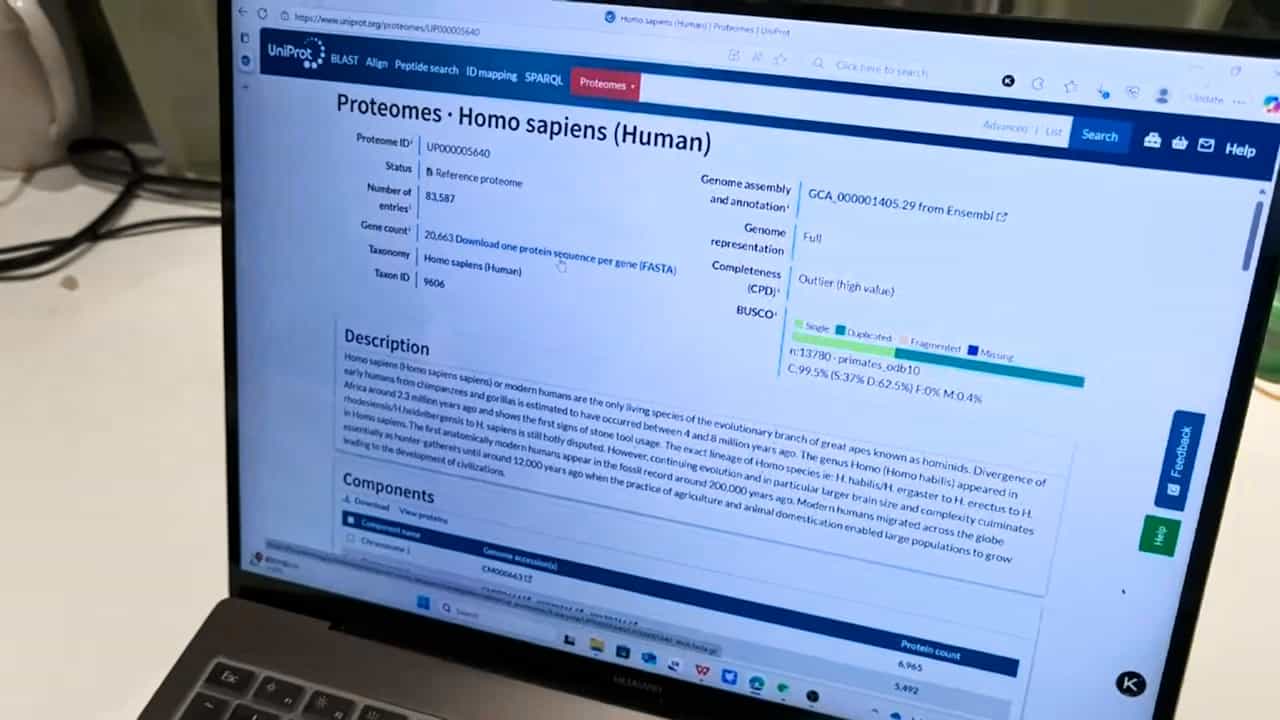

Navigating the Mass Spectrometry-Based Proteomic Data Using Free Computational Tools
Published on: August 19, 2025
Evaluation of normalization strategies for mass spectrometry-based multi-omics datasets
Chi Yen Tseng1, Jessica A Salguero2, Joshua D Breidenbach2
1Biochemistry and Biotechnology Group, Bioscience Division, Los Alamos National Laboratory, Los Alamos, NM, 84545, USA. chiyen_tseng@lanl.gov.

Introduction:
Data normalization is crucial for multi-omics integration, reducing systematic errors and maximizing the likelihood of discovering true biological variation. Most studies assess normalization for a single omics type or use datasets from separate experiments. Few address time-course data, where normalization might bias temporal differentiation. In this study, we compared common normalization methods and a machine learning approach, Systematical Error Removal using Random Forest (SERRF), using multi-omics datasets generated from the same experiment-even from the same cell lysate.
Objectives:
To develop a straightforward process to assess normalization effects and identify the most robust methods across multi-omics datasets.
Methods:
We analyzed metabolomics, lipidomics, and proteomics datasets from primary human cardiomyocytes and motor neurons exposed to acetylcholine-active compounds over time. Normalization effectiveness was evaluated based on improvement in QC features consistency and observing the change in treatment and time-related variance.
Results:
Probabilistic Quotient Normalization (PQN) and Locally Estimated Scatterplot Smoothing (LOESS) QC were identified as optimal for metabolomics and lipidomics, while PQN, Median, and LOESS normalization excelled for proteomics. These methods consistently enhanced QC feature consistency in metabolomics and lipidomics, and preserved time-related variance or treatment-related variance in proteomics, demonstrating their effectiveness and robustness. SERRF normalization, applied only to metabolomics in this study, outperformed other methods in some datasets but inadvertently masked treatment-related variance in others.
Conclusion:
Our evaluation identified PQN and LoessQC as the top methods for metabolomics and lipidomics, and PQN, Median, and Loess normalization for proteomics, in multi-omics integration in a temporal study.
More Related Videos
 08:27
Large-Scale Multi-Omics Genome-Wide Association Studies Mo-GWAS: Guidelines for Sample Preparation and Normalization
Published on: July 27, 2021
 07:34
Large Scale Non-targeted Metabolomic Profiling of Serum by Ultra Performance Liquid Chromatography-Mass Spectrometry UPLC-MS
Published on: March 14, 2013
Related Concept Videos
 01:21
Mass Spectrometry: Complex Analysis
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
 01:19
Mass Spectrometry: Overview
 01:13
Mass Analyzers: Overview