Related Experiment Video
Updated: Sep 17, 2025


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Identification of potential COVID-19 Mpro inhibitors through covalent drug docking, molecular dynamics simulation,
Mohammad Hossein Haghir Ebrahim Abadi1,2, Fatemeh Bayani1,2, Yahya Sefidbakht3,4
1Department of Cell and Molecular Biology, Faculty of Life Sciences and Biotechnology, Shahid Beheshti University, Tehran, 1983969411, Iran.

Abstract:
The viral main protease is a critical drug target due to its role in the SARS-CoV-2 life cycle. In this study, we evaluated 2,000 potential Mpro inhibitors recommended by the FragRep server, with a focus on interactions with the CYS145 residue. Six top candidates were selected for each of the 7JKV and 7TDU Mpro structures, and covalent molecular docking was performed using SeeSAR. We docked twelve ligands to various Mpro sites and performed 100 ns molecular dynamics (MD) simulations on the highest-scoring ligands: lig-14-23, lig-6-32, lig-2-3, lig-0101, lig-7612, lig-811, and lig-837. MD analysis, including Root Mean Square Deviation (RMSD), Radius of Gyration (Rg), Solvent Accessible Surface Area (SASA), and Root Mean Square Fluctuation (RMSF), confirmed the stability of the covalent complexes. Non-covalent ligands, which exhibited higher toxicity profiles, were excluded from the molecular dynamics analysis. The results indicate that lig-7612 and lig-837 form stable interactions with Mpro, suggesting their potential as drug candidates. These inhibitors exhibit pharmacological properties that could influence drug interactions and metabolism in vivo. Further safety and pharmacokinetic assessments are required for therapeutic development.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:04
Protein-protein Interfaces
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:16
Protein-Drug Binding: Mechanism and Kinetics
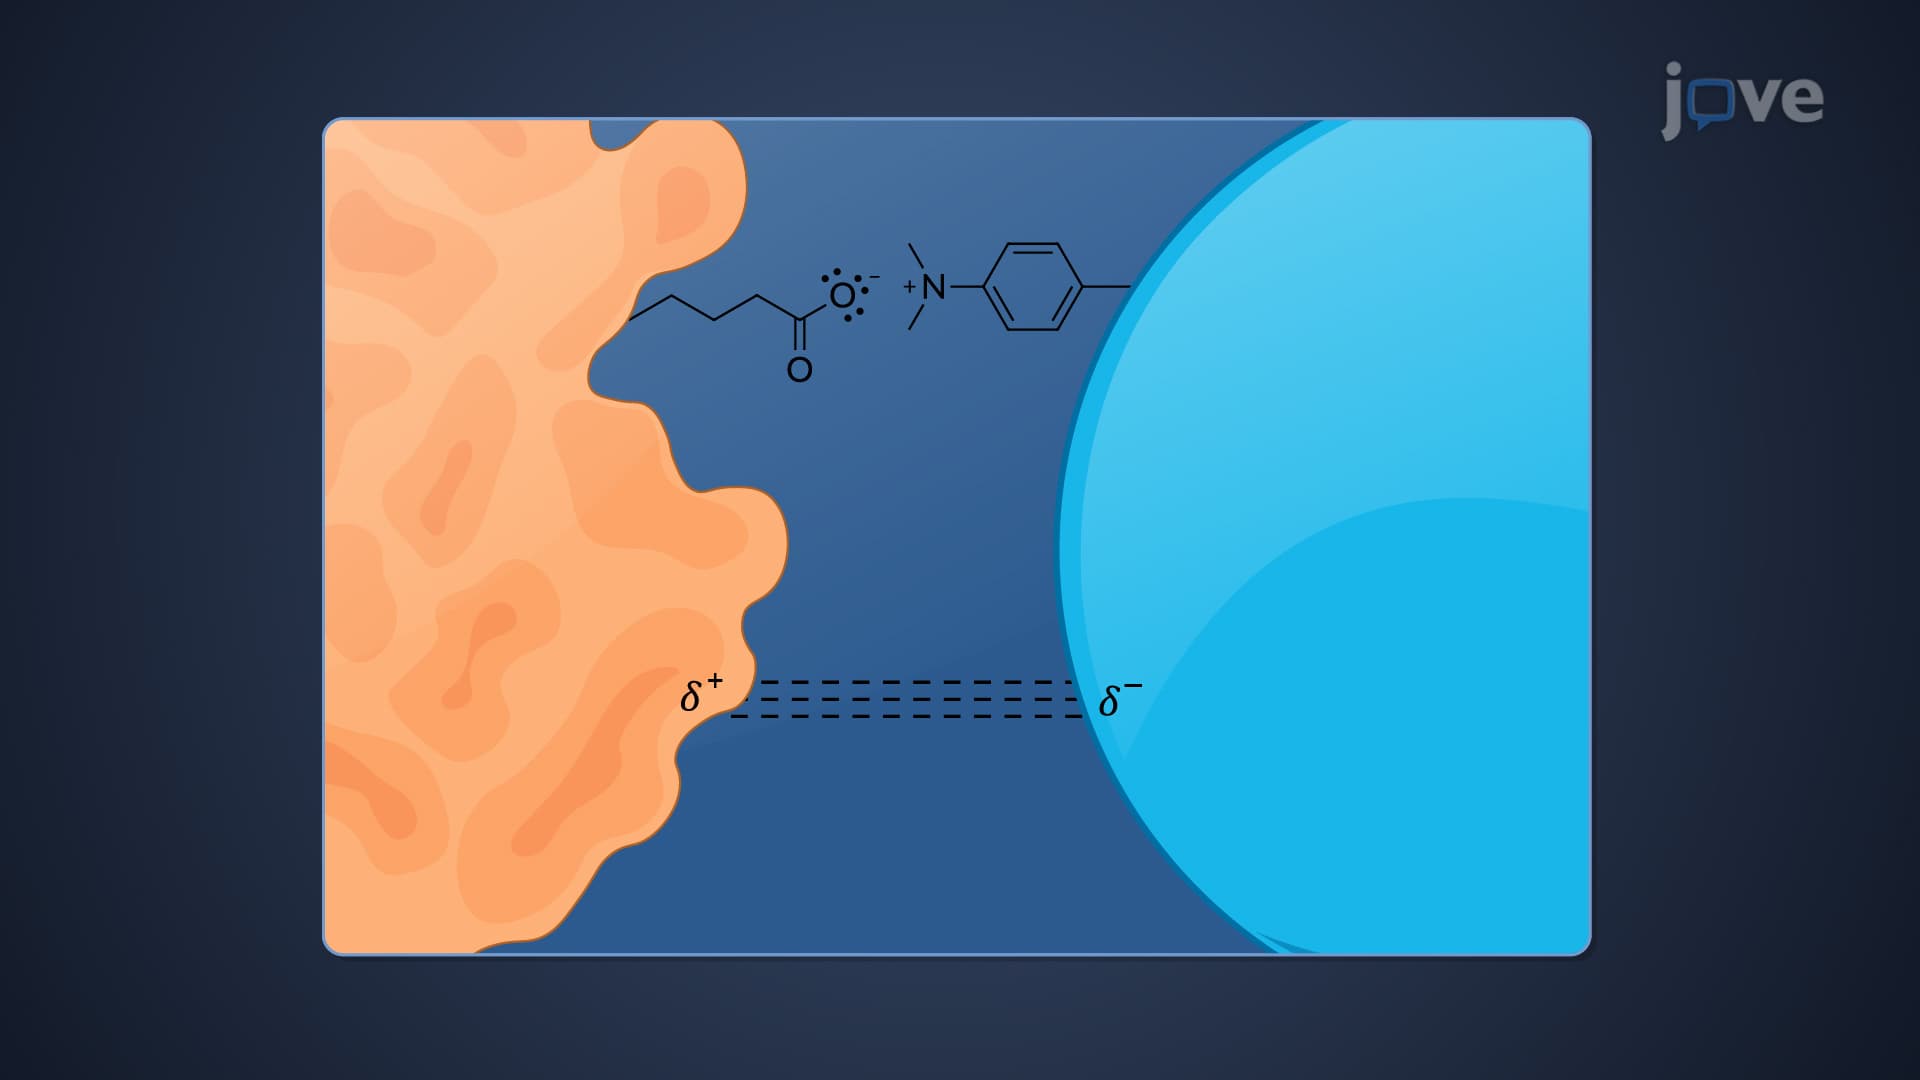
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...