Related Experiment Video
Updated: Sep 16, 2025


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based
Ahmad Fadhlurrahman Ahmad Hidayat1, Saharuddin Bin Mohamad2
1Bioinformatics Programme, Institute of Biological Sciences, Faculty of Science, Universiti Malaya.

Abstract:
The drug discovery process is a rigorous, time-consuming, and expensive operation. The computational approach in drug discovery allows researchers to prioritize the most promising compounds for further testing, which would greatly reduce the required resources, leading to an increment of the overall efficiency in the drug discovery pipelines. Structure-based drug discovery is a common approach that requires the structural information of the target protein in a three-dimensional format. However, the current limitation of most computer-aided drug discovery strategies is their inability to introduce the flexibility and dynamics of the target protein structure during the ligand-protein docking simulation. While both induced fit docking and ensemble-based docking aim to address protein flexibility in the docking procedure, the latter can provide a more comprehensive view of dynamic protein behavior by incorporating multiple conformations throughout the simulation. In this report, we demonstrate and discuss the application of a technique called ensemble-based docking analysis that indirectly introduces the target protein structure's flexibility and dynamics in the molecular docking process. The protein and ligand selected for ensemble-based docking studies were lysozyme and Flovokawain B (FB), respectively. FB has been previously reported to have binding activity with lysozyme. A molecular dynamics (MD) simulation was performed on lysozyme in the presence of water, and the total energy, root-mean-square deviation (RMSD), and root-mean-square fluctuation (RMSF) were examined. Conformation clustering was generated based on several clustering cutoff values and was chosen for additional docking analysis with FB. Cluster no 2 gives the lowest binding energy at -29.37 kJ/mol. Molecular docking images were generated to anticipate the presence of binding forces. By incorporating the structural dynamics of the protein, the ensemble-based docking approach can better capture the range of possible binding scenarios, leading to more reliable predictions of binding outcomes.
More Related Videos
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:04
Protein-protein Interfaces
 01:16
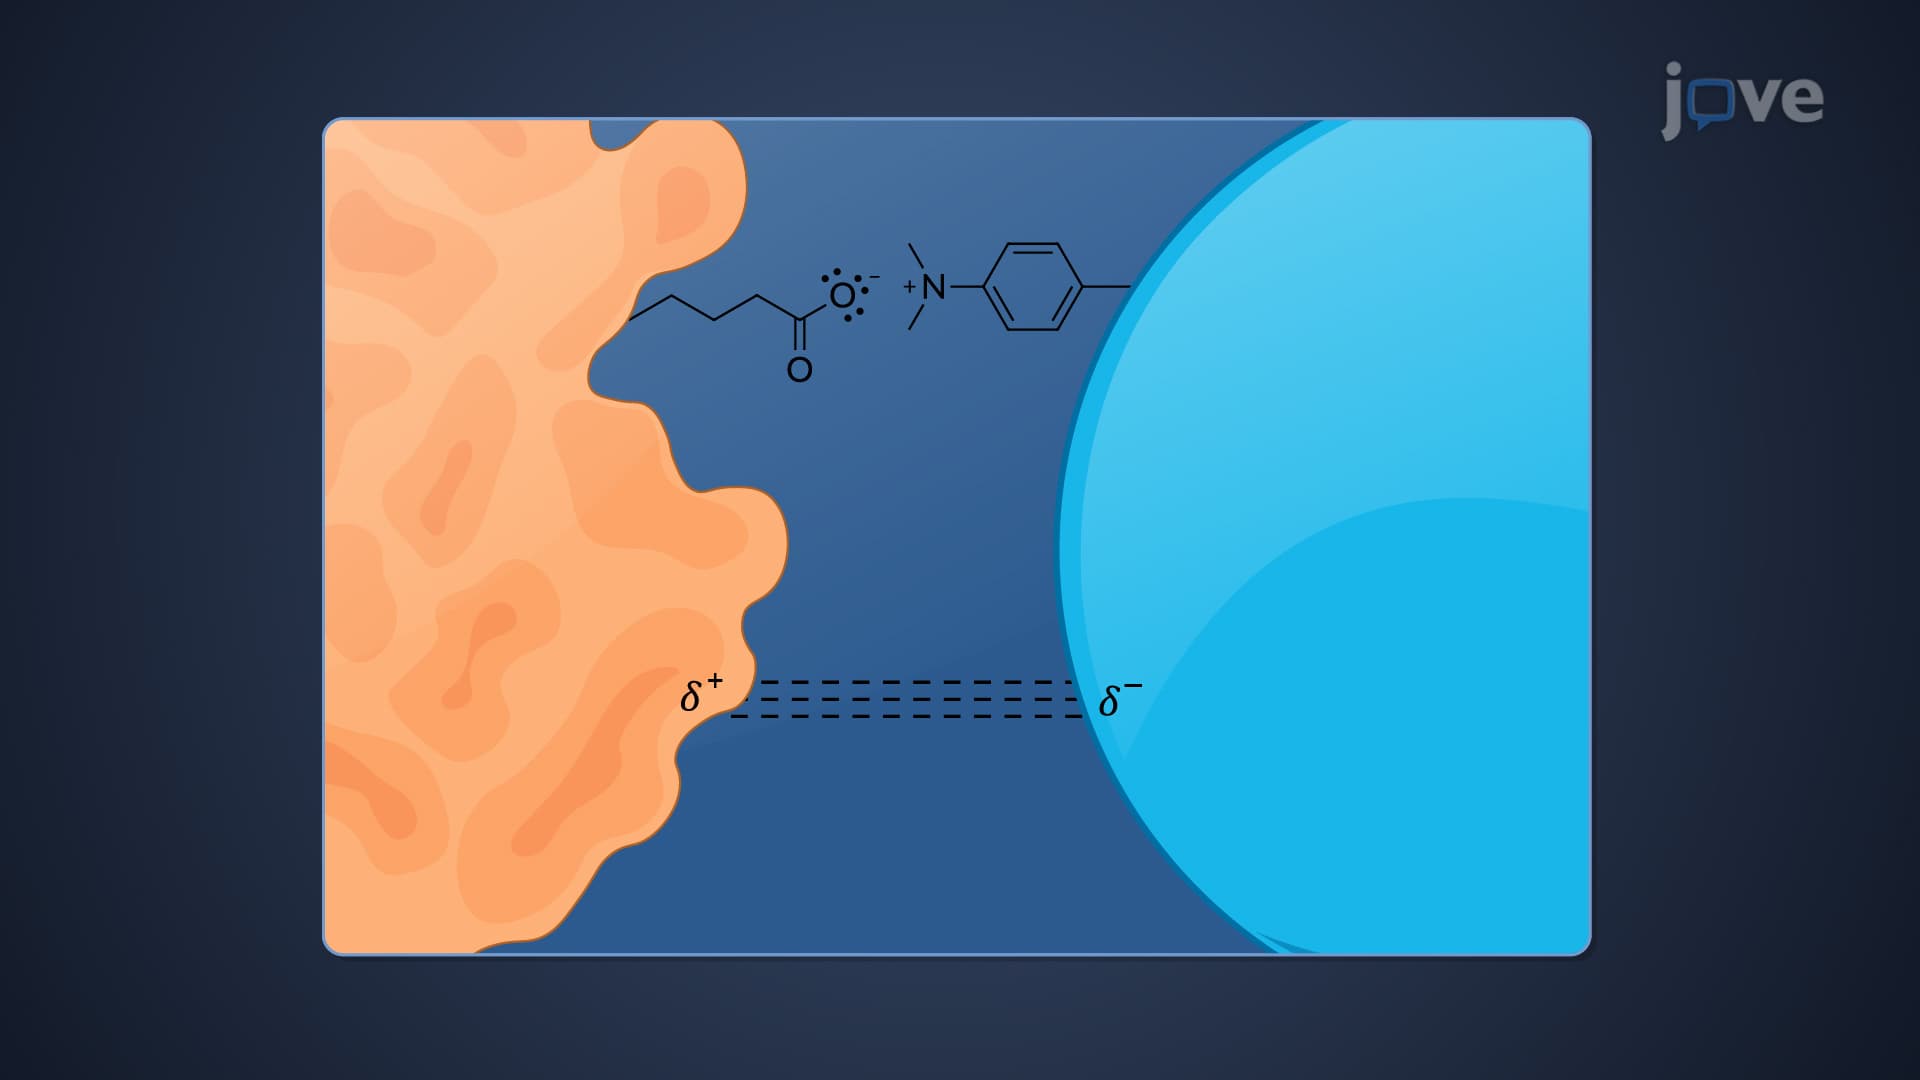
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:24
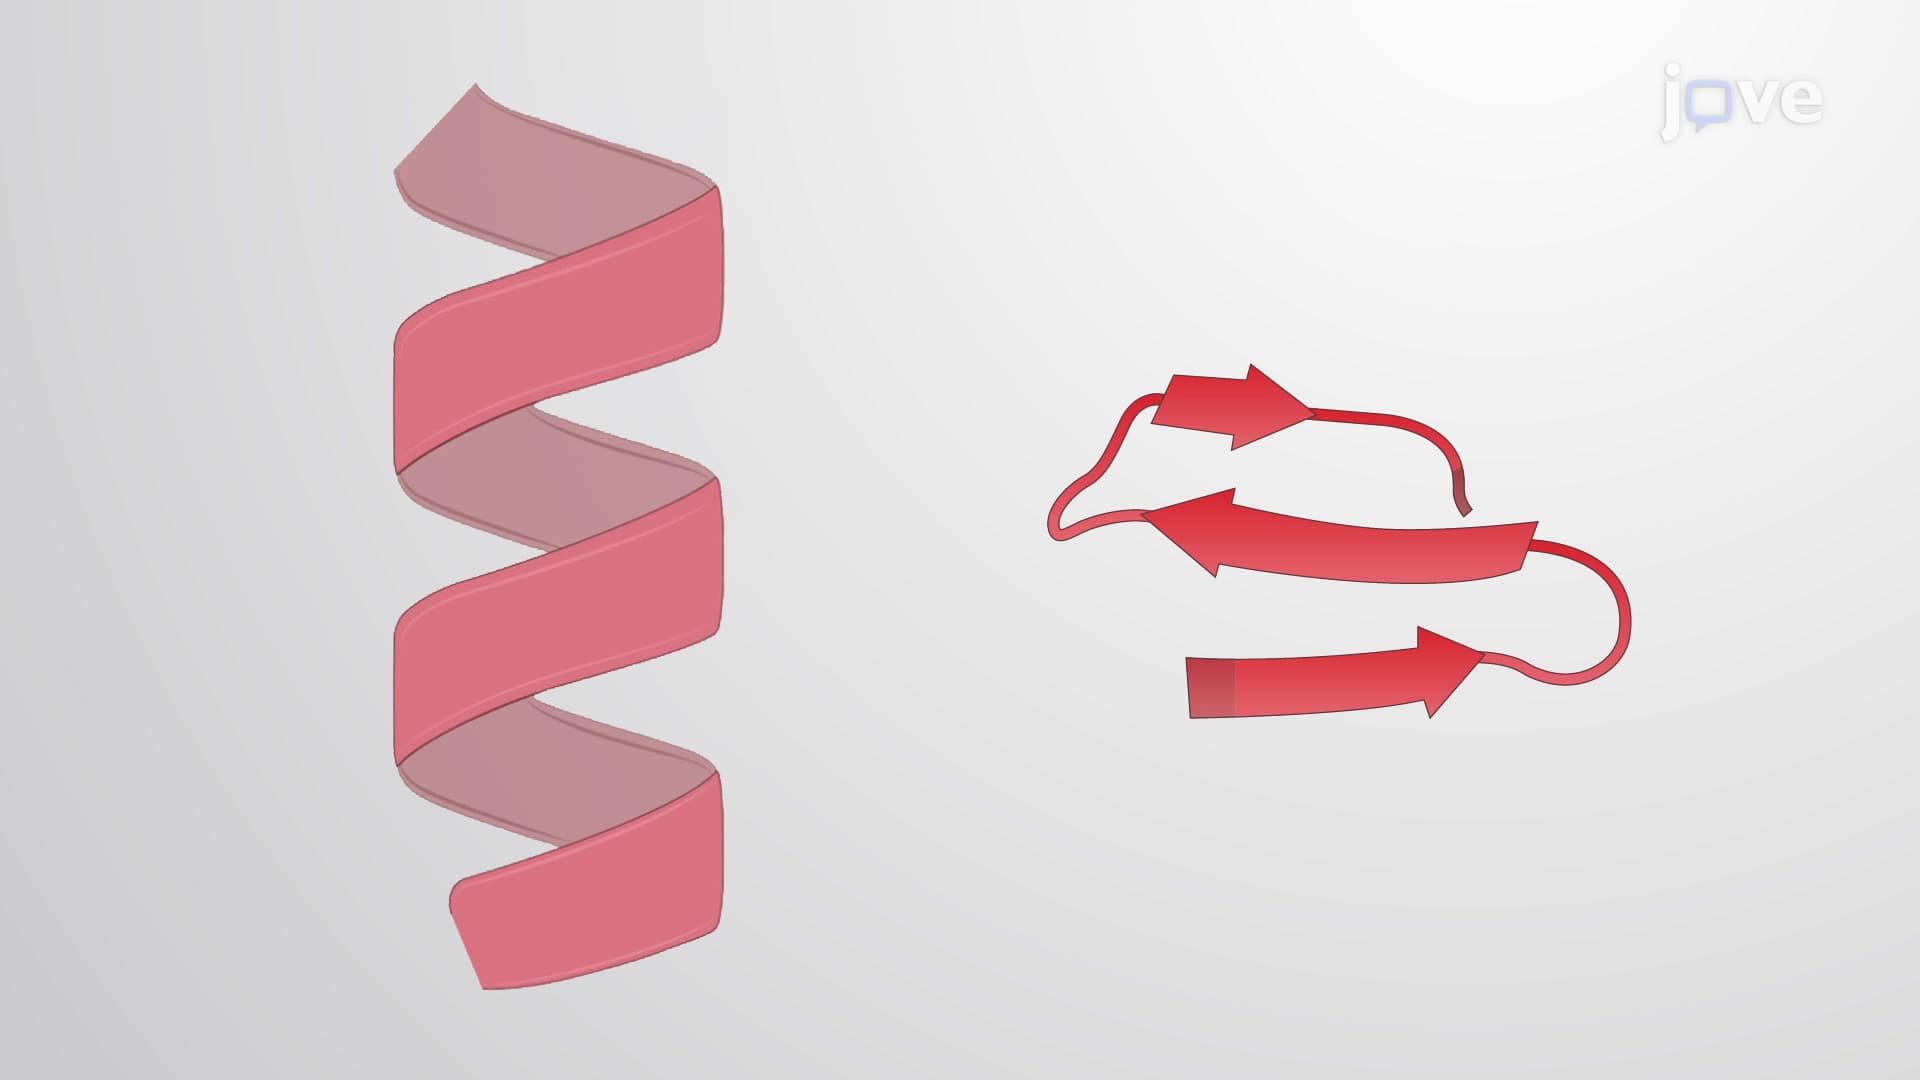
Protein Organization
The primary structure of a protein is its amino acid sequence....
 01:22
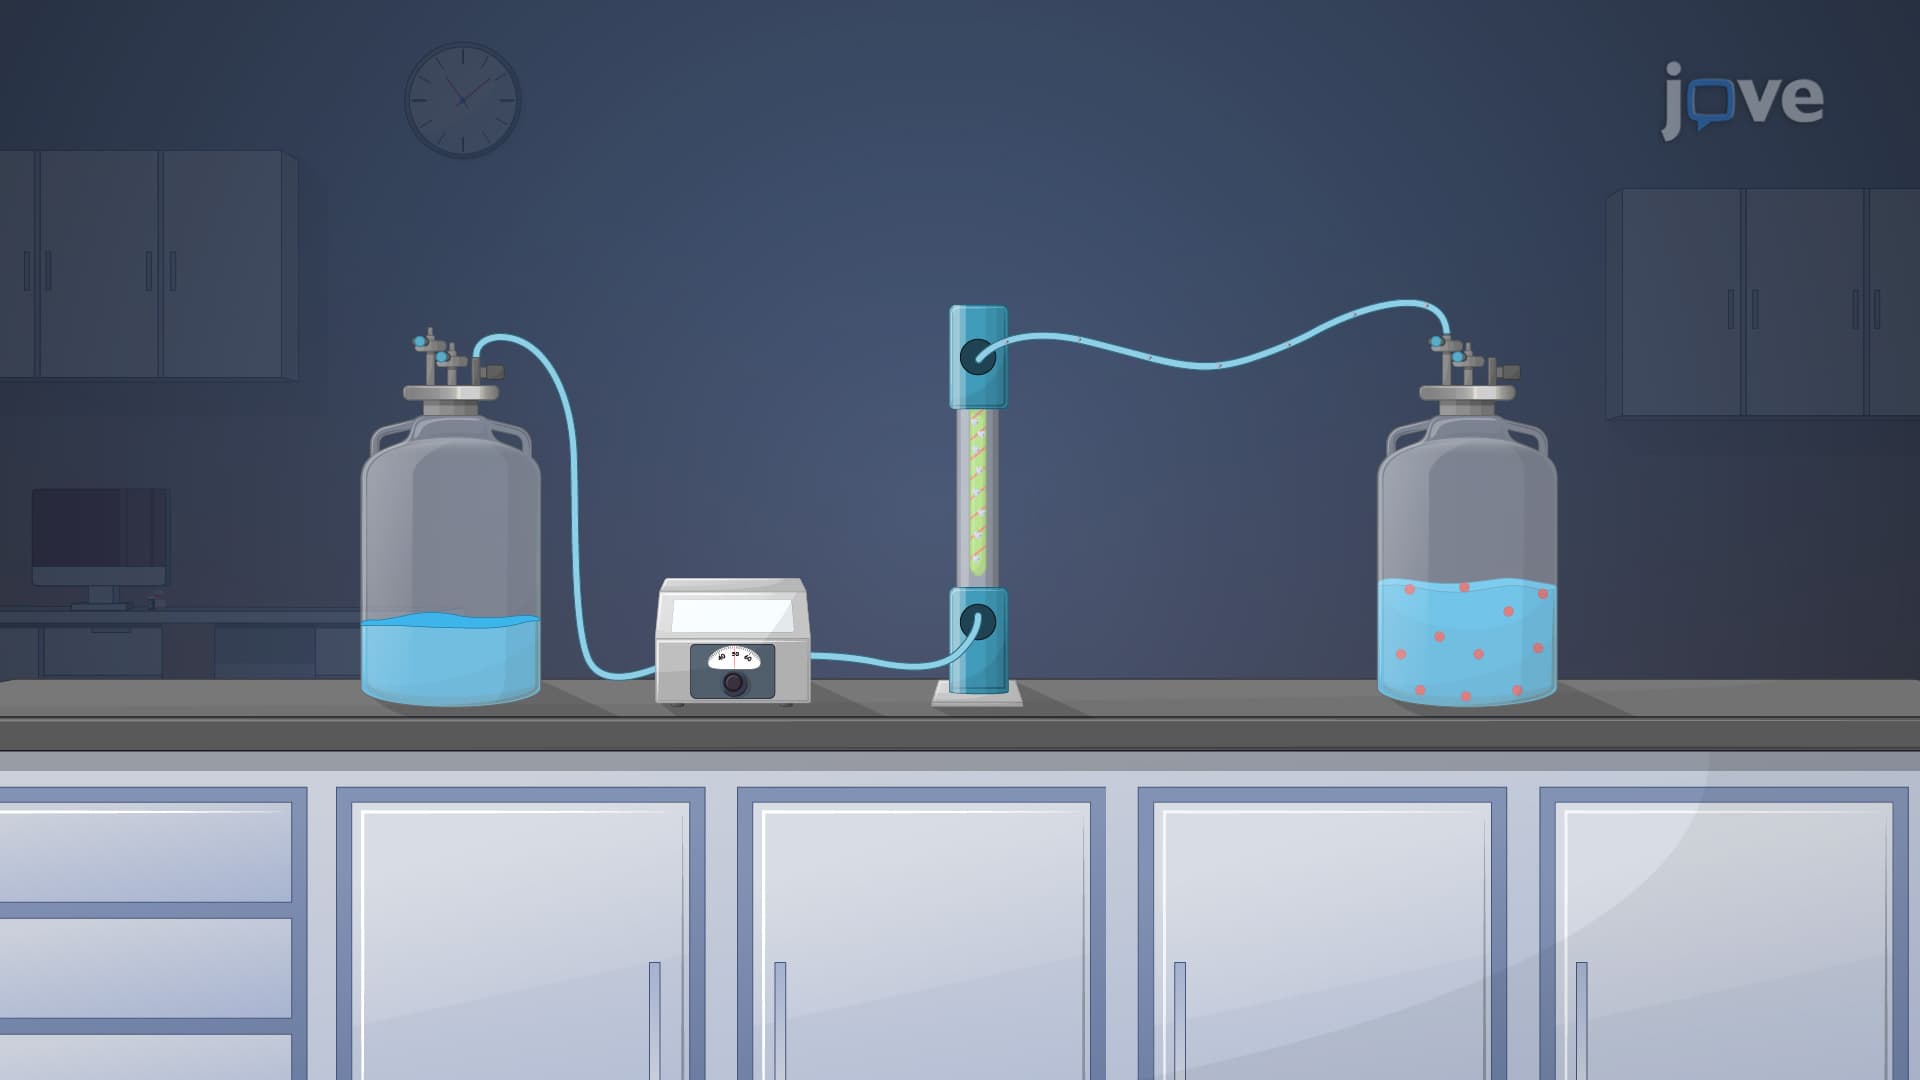
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...