Related Experiment Video
Updated: Sep 16, 2025

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Prediction of enzyme inhibition (IC50) using a combination of protein-ligand docking and semiempirical quantum
Robert C Glen1,2, Jason C Cole3, James J P Stewart4
1Department of Chemistry, University of Cambridge, Lensfield Road, Cambridge, CB21EW, UK.

Context:
The ability to predict the relative binding energies of ligands to a biological receptor would be of great value in drug discovery. However, accurately calculating the predicted binding energies is limited by the high accuracy required, by the presence of multiple minima on the potential energy surface, and by issues specific to the intrinsic properties of the binding site, such as details of the geometry of the ligand-protein complex. To address these issues, a systematic analysis of potential sources of error was carried out which resulted in a few relatively small changes being made to the MOPAC program.
Methods:
A set of 77 ligands was constructed for which experimentally determined IC50 values were available. For each of the ligands, prediction of the protein-ligand interaction energy was carried out in two distinct stages. In the first stage, the Protein-Ligand docking program GOLD was used to generate several distinct conformations of the ligand bound to a protein. The geometries of these systems were then optimised using the MOPAC program. A comparison of the relative binding energies of the ligands with the reported IC50 values showed a very poor predictive power. By partitioning the ligand set into two subsets, and eliminating six ligands that were inconsistent with the experimental results, a large increase in accuracy was obtained.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Related Concept Videos
 02:40
Ligand Binding Sites
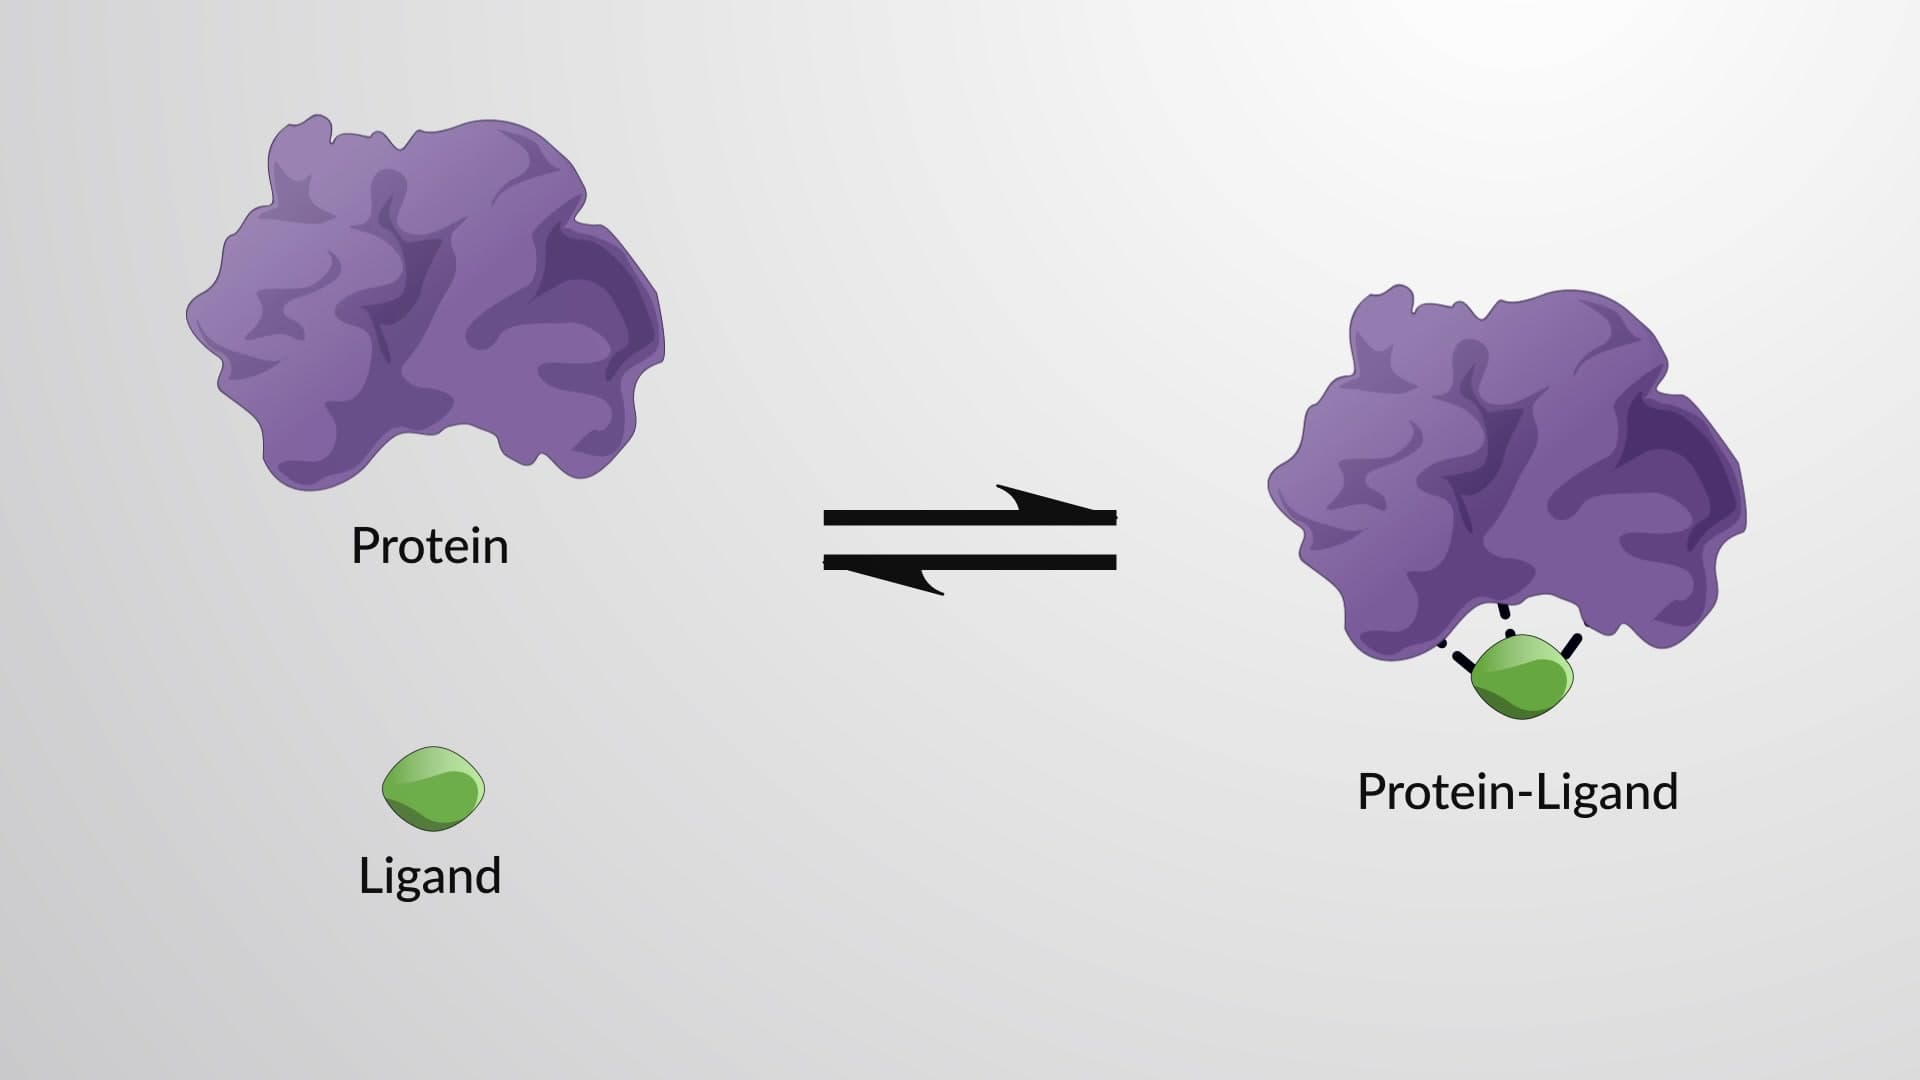
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:18
The Equilibrium Binding Constant and Binding Strength
 02:04
Protein-protein Interfaces
 01:22
Protein-Drug Binding: Determination Methods
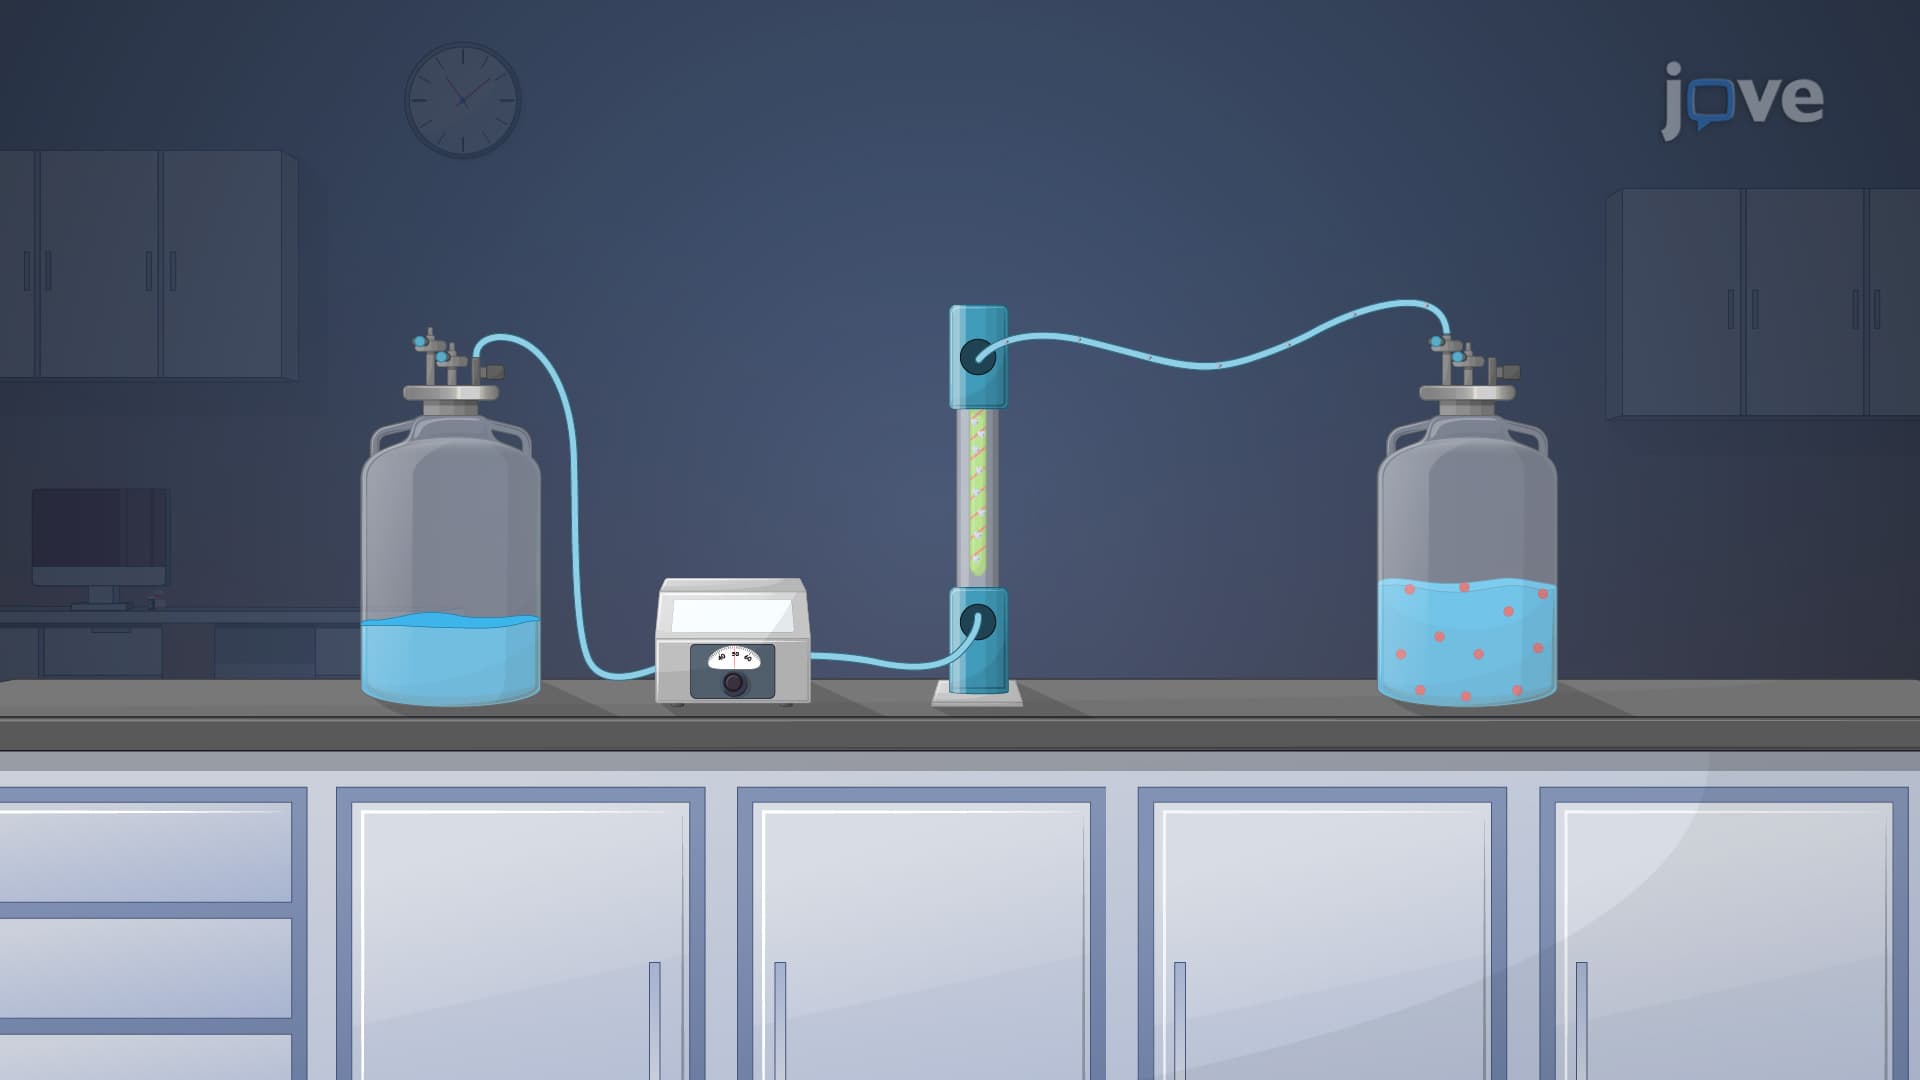
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:16
Protein-Drug Binding: Mechanism and Kinetics
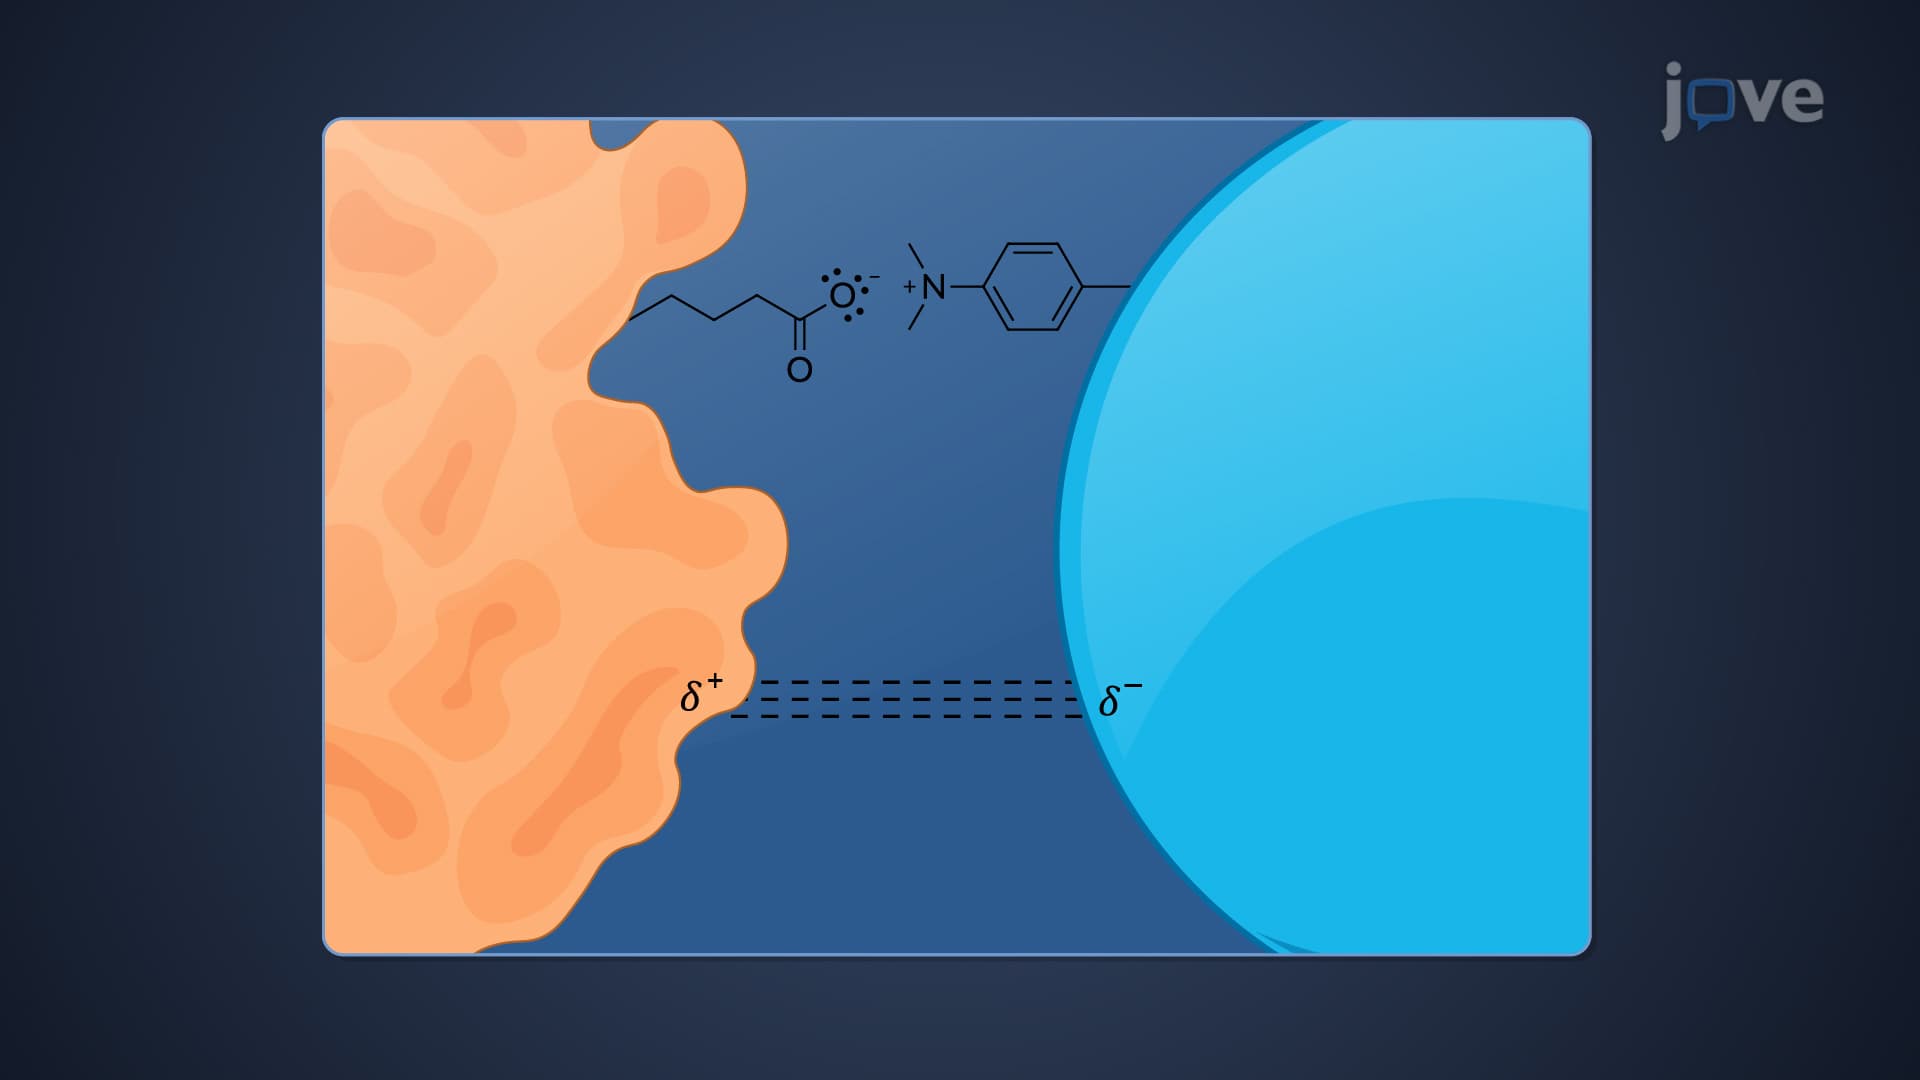
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:30
Enzyme Inhibition