Related Experiment Video
Updated: Sep 15, 2025
Optimization for Sequencing and Analysis of Degraded FFPE-RNA Samples
Published on: June 8, 2020
Strong correlation of gene counts and differentially expressed genes between a 3' RNA-Seq and an RNA hybridization
Alexander F H Haake1, Alina K Loriani Fard1, Vladimir M Jovanovic2,3
1Department of Veterinary Medicine, Institute of Veterinary Pathology, Freie Universität Berlin, Berlin, Germany.

Abstract:
Analyses of nucleic acids from archival tissues offer invaluable prospects for numerous fields of veterinary medicine, such as the study of differential gene expression in rare or historic diseases. The establishment of modern methodologies, however, raises questions regarding the comparability and reproducibility of data obtained from unlike tools. 3' RNA-Seq and direct RNA hybridization are such conceptually different approaches for high-throughput transcriptome analysis. Since both are applicable to short, partially degraded mRNA fragments, they in principle allow investigations of formalin-fixed, paraffin-embedded (FFPE) tissues that are abundantly available in pathology archives. Here, we compared the two methods in several relevant details using the RNA from the same set of 35 FFPE canine tumors as input, including sample- and gene-wise count levels, gene expression strengths and directions, as well as the overlaps of differentially expressed genes (DEGs). Both methods proved suitable for their use on archival tissues with moderately to very strong overall count correlations, as indicated by a range of Pearson and Spearman means between 0.66 and 0.87. Of note, the gene-wise count correlations depended on gene expression strength. In an entity-contrasting comparison, expression directions correlated very strongly ranging from 0.88 to 0.91, but DEGs overlapped only moderately with a Jaccard index of 0.53. Finally, we contrasted the different practically relevant aspects of the two technologies with their distinct advantages that depend on the objectives and design of the study. This comparison will guide and help to select the appropriate method and to validate and interpret the data obtained.
Related Concept Videos
 03:21
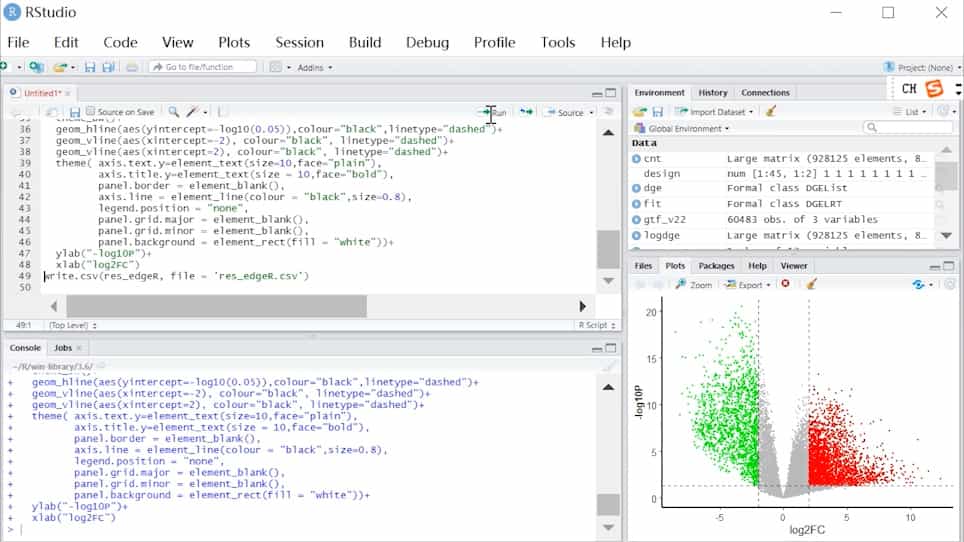
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
 02:34
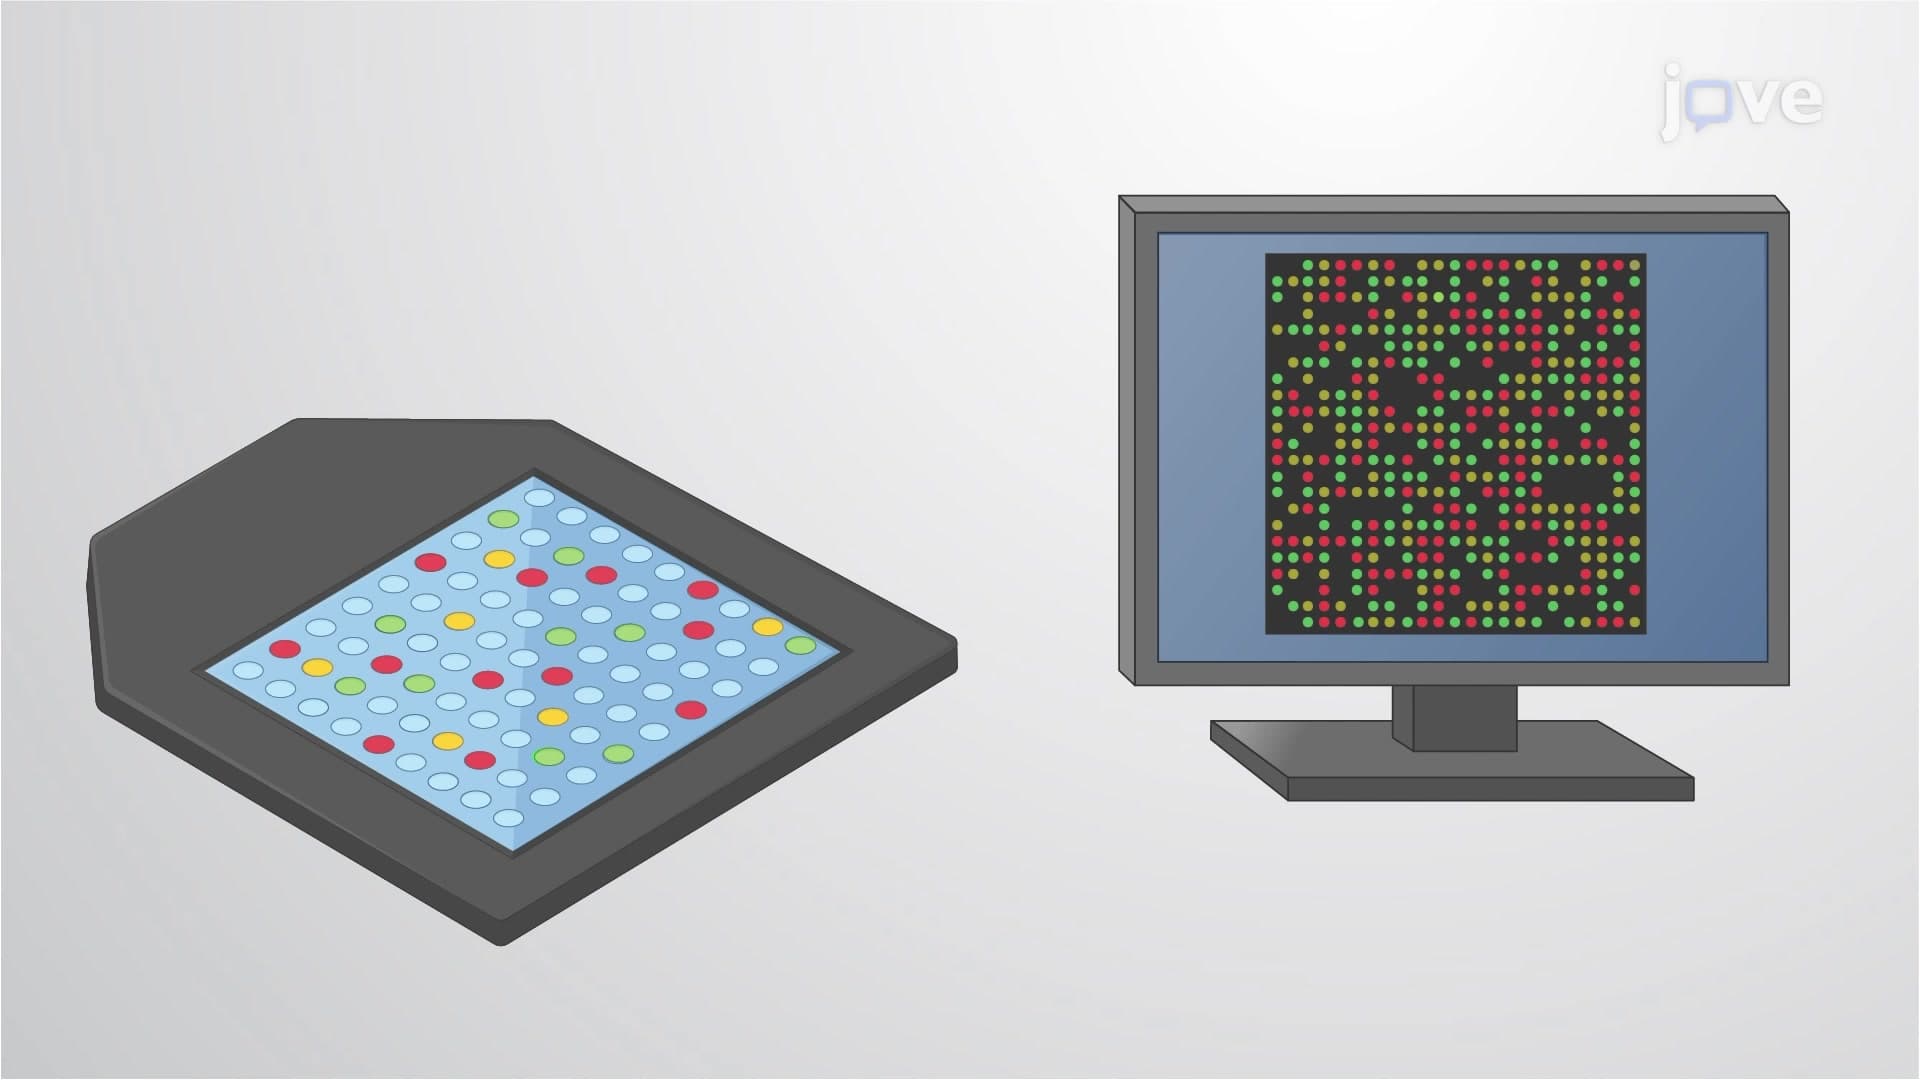
DNA Microarrays
 02:24
Ribosome Profiling
Applications of ribosome profiling
Ribosome profiling has many applications, including in vivo monitoring of translation inside a particular organ or tissue type and quantifying new protein synthesis levels.
The technique...