Related Experiment Video
Updated: Sep 15, 2025

Using En Face Immunofluorescence Staining to Observe Vascular Endothelial Cells Directly
Published on: August 20, 2019
Downregulation of LATS1/2 Drives Endothelial Senescence-Associated Stemness (SAS) and Atherothrombotic Lesion

Insights
Loss of LATS1/2 in endothelial cells triggers a senescence-associated stemness (SAS) state, promoting atherothrombosis. Targeting CD38 may offer a therapeutic strategy for vascular dysfunction and plaque instability in acute coronary syndrome patients.
Area of Science:
- Vascular Biology
- Molecular Mechanisms of Atherothrombosis
- Cellular Senescence
Background:
- Atherothrombosis, a key event in acute coronary syndrome (ACS), is linked to disturbed blood flow (d-flow).
- The Hippo pathway kinases LATS1/2 are involved in mechanical stress responses, but their role in d-flow-induced atherothrombosis is unclear.
Purpose of the Study:
- To investigate the role of endothelial cell (EC)-specific LATS1/2 in atherothrombosis.
- To elucidate the molecular mechanisms by which disturbed blood flow induces atherothrombosis via ECs.
Main Methods:
- Utilized EC-specific knockout (EKO) mice in a partial left carotid ligation (PLCL) model.
- Performed plaque spatial multi-omics analysis integrating imaging mass cytometry, sequential immunofluorescence (COMET™), and spatial metabolomics.
Main Results:
- LATS1/2 deletion in ECs induced a senescence-associated stemness (SAS) phenotype, characterized by CD38 upregulation and reduced sulfite oxidase (SUOX) activity.
- Metabolic disturbances, including altered mitochondrial function and enhanced glutamate/TCA cycle metabolism, sustained EC proliferation despite ATP depletion.
- CD38 inhibition attenuated atherothrombotic plaque formation, and a similar EC phenotype was observed in human plaques.
Conclusions:
- Loss of LATS1/2 in ECs drives SAS, promoting EC proliferation, senescence, and fragile neovascularization, hallmarks of atherothrombosis.
- The LATS1/2-CD38-SUOX axis is identified as a novel pathway orchestrating SAS-driven atherothrombosis.
- Targeting CD38 presents a potential therapeutic strategy for vascular dysfunction and plaque instability in ACS.
Background:
Atherothrombosis, the main event leading to acute coronary syndrome (ACS), is strongly linked to disturbed blood flow (d-flow) regions. Although the involvement of the Hippo pathway and its kinases Large Tumor Suppressor Kinase 1and 2 (LATS 1 and 2) in mechanical stress responses is known, the mechanisms by which d-flow simultaneously induces senescence, proliferation, and atherothrombosis remain unclear.
Methods:
The role of endothelial cells (EC)-specific LATS1/2 was examined using EC specific knock-out (EKO) mice in a partial left carotid ligation (PLCL) model. Plaque spatial multi-omics analysis was performed by integrating imaging mass cytometry, sequential immunofluorescence (COMET™), and spatial metabolomics at the single-cell level in human and mouse atherosclerotic plaques.
Results:
In tamoxifen-inducible Lats1 homo( -/- ) / Lats2 homo( -/- ) EC-specific knockout (EKO) mice, deletion of LATS1/2 induced by tamoxifen led to fatal outcomes, characterized by severe systemic edema and markedly increased vascular permeability. In contrast, Lats1 het(+ /- ) / Lats2 homo( -/- ) -EKO mice survived and developed atherothrombotic plaques exhibiting neovascularization even without further additional dietary or genetic intervention. Spatial proteomics analysis revealed that LATS1/2 depletion in ECs triggered a senescence-associated stemness (SAS) phenotype, primarily driven by CD38 upregulation. Complementary spatial metabolomics profiling demonstrated a significant increase in sulfite and taurine within LATS1/2-deficient plaques, indicating lowered sulfite oxidase (SUOX) activity. Mechanistically, CD38 upregulation was found to suppress SUOX expression, induce the reverse mode of mitochondrial complex V, and increase succinate dehydrogenase (SDH) activity along with ATP consumption. Paradoxically, despite ATP depletion, this metabolic disturbance enhanced glutamate metabolism and the tricarboxylic acid (TCA) cycle, sustaining EC proliferation under energetically stressed conditions. The combined effect of LATS1/2 deletion and CD38 activation established a unique EC phenotype defined by increased SAS, leading to proliferation, senescence, and eventual cell death. These pathological processes culminated in the formation of atherothrombotic plaques, which were attenuated by inhibition of CD38. Notably, a similar phenotype-marked by metabolically active ECs-was observed in human atherothrombotic plaques, suggesting translational relevance.
Conclusion:
Loss of LATS1/2 in ECs induces SAS state that promotes excessive EC proliferation, senescent cell accumulation, and the development of structurally fragile, leaky neo vessels-hallmarks of atherothrombotic lesions. CD38-mediated SUOX deficiency further amplifies this pathological process by inducing mitochondrial dysfunction, depleting ATP, and triggering compensatory upregulation of glutamate and TCA cycle metabolism. These findings identify a novel LATS1/2-CD38-SUOX axis in ECs that orchestrates SAS-driven atherothrombosis. Targeting CD38 may represent a promising therapeutic strategy to mitigate vascular dysfunction and plaque instability in high-risk ACS patients.
Graphical Abstract:
Under normal physiological conditions, LATS1/2 and Lamin A work together to suppress CD38 expression. Lamin A binds directly to the CD38 promoter to repress transcription, and LATS1/2 interact with Lamin A to reinforce this suppression. This collaboration helps maintain low CD38 activity and preserves cellular NAD⁺ levels. However, under disturbed flow (d-flow), both LATS1/2 and Lamin A are downregulated. The loss of this dual repression leads to increased CD38 NADase expression. Elevated CD38 accelerates NAD⁺ consumption, causing NAD⁺ depletion-a hallmark of cellular senescence. Reduced NAD⁺ disrupts key metabolic and stress-response pathways, contributing to the onset of the senescent state. At the same time, CD38 suppresses sulfite oxidase (SUOX), leading to sulfite accumulation and mitochondrial redox imbalance. This shift activates the reverse mode of mitochondrial Complex V, which decreases ATP production and increases mitochondrial ROS, intensifying metabolic and oxidative stress in endothelial cells (ECs). In response, ECs compensate by upregulating succinate dehydrogenase (SDH), enhancing TCA cycle activity and glutamate metabolism. This metabolic adaptation provides the biosynthetic building blocks needed for cell growth and proliferation. As a result, ECs adopt a paradoxical phenotype: they show classical features of stress-induced senescence (such as NAD⁺ depletion, oxidative stress, and cell cycle arrest signals), while simultaneously undergoing metabolic activation and proliferation, also mediated by YAP. This defines a non-canonical endothelial program known as senescence-associated stemness (SAS), characterized by the formation of abnormal, proliferative, yet fragile neovessels. These dysfunctional vessels contribute to atherothrombosis, setting this process apart from the more stable lesions typical of conventional atherosclerosis.
More Related Videos
 11:00
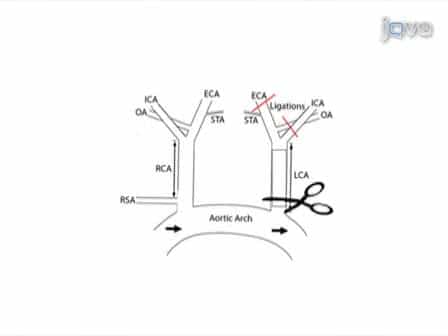
A Model of Disturbed Flow-Induced Atherosclerosis in Mouse Carotid Artery by Partial Ligation and a Simple Method of RNA Isolation from Carotid Endothelium
Published on: June 22, 2010
 08:41
Assessment of Vascular Tone Responsiveness using Isolated Mesenteric Arteries with a Focus on Modulation by Perivascular Adipose Tissues
Published on: June 3, 2019
Related Concept Videos
 01:24
Regulation of Angiogenesis and Blood Supply
 01:30
Atherosclerosis I: Introduction
 01:26
Coronary Artery Disease II: Pathophysiology
 01:30
Peripheral Artery Disease I: Introduction
 01:26
Atherosclerosis III: Management