Related Experiment Video
Updated: Sep 8, 2025

A Method for Screening and Validation of Resistant Mutations Against Kinase Inhibitors
Published on: December 7, 2014
Pharmacokinetics and Safety of Ripretinib in Participants with Hepatic Impairment: A Phase 1 Study
Anna Papinska1, Lakshmi Viswanathan2, Qiang Lu2
1Clinical Pharmacology, Deciphera Pharmaceuticals LLC, 200 Smith Street, Waltham, MA, 02451, USA. anna.papinska@deciphera.com.

Introduction:
Ripretinib, an oral switch-control inhibitor of KIT tyrosine kinase and platelet-derived growth factor receptor alpha kinase, is approved for adults with advanced gastrointestinal stromal tumor who received prior treatment with three or more kinase inhibitors, including imatinib. Ripretinib is metabolized into the equally active major metabolite DP-5439, a prominent component of total drug exposure in humans after oral administration. Ripretinib and DP-5439 undergo hepatic metabolism mainly via cytochrome P450 3A4. Therefore, exposure to ripretinib and/or DP-5439 may be affected in patients with hepatic impairment.
Methods:
This is a phase 1, open-label study evaluating the pharmacokinetics and safety of ripretinib and DP-5439 in participants with varying degrees of hepatic impairment compared with matched healthy participants after a single oral 50-mg ripretinib dose.
Results:
Mild hepatic impairment did not affect exposure to ripretinib and DP-5439. In participants with moderate and severe hepatic impairment, ripretinib exposure (area under the concentration-time curve) was 99% and 163% greater, respectively, compared with matched healthy participants, whereas DP-5439 exposure was approximately 20% greater in moderate and 44% lower in severe hepatic impairment. Exposure to combined ripretinib + DP-5439 was higher by approximately 51% and 37% in participants with moderate and severe hepatic impairment, respectively. No safety signals were identified.
Conclusion:
On the basis of the known safety profile of ripretinib, these increased ripretinib and combined ripretinib + DP-5439 exposures in participants with hepatic impairment are unlikely to be clinically relevant. Therefore, no dose adjustments are recommended for patients with gastrointestinal stromal tumor and hepatic impairment.
Insights
This study found that ripretinib (a KIT inhibitor) and its metabolite DP-5439 exposure increased in patients with moderate to severe hepatic impairment. However, these changes are unlikely to be clinically relevant, and no dose adjustments are recommended for gastrointestinal stromal tumor patients.
Area of Science:
- Pharmacology
- Oncology
- Clinical Pharmacology
Background:
- Ripretinib is an oral switch-control inhibitor of KIT and PDGFRA kinases, approved for advanced gastrointestinal stromal tumors (GIST) after prior treatments.
- Ripretinib is metabolized to DP-5439, an equally active metabolite, with both undergoing hepatic metabolism primarily via CYP3A4.
- Hepatic impairment may alter the exposure of ripretinib and DP-5439.
Purpose of the Study:
- To evaluate the pharmacokinetics and safety of ripretinib and its major metabolite DP-5439 in patients with hepatic impairment.
- To compare ripretinib and DP-5439 exposure in participants with mild, moderate, and severe hepatic impairment versus healthy controls.
Main Methods:
- Phase 1, open-label study design.
- Single oral 50-mg dose of ripretinib administered to participants with varying degrees of hepatic impairment and matched healthy controls.
- Pharmacokinetic assessment of ripretinib and DP-5439.
Main Results:
- Mild hepatic impairment showed no effect on ripretinib or DP-5439 exposure.
- Moderate hepatic impairment increased ripretinib exposure by 99% and DP-5439 by 20%.
- Severe hepatic impairment increased ripretinib exposure by 163% and decreased DP-5439 by 44%; combined exposure increased by 51% and 37% respectively. No safety signals were identified.
Conclusions:
- Increased ripretinib and combined ripretinib + DP-5439 exposures in hepatic impairment are unlikely to be clinically relevant.
- No dose adjustments are recommended for GIST patients with hepatic impairment.
- Ripretinib maintains a favorable safety profile in patients with hepatic impairment.
Related Concept Videos
 01:19
Phase I Oxidative Reactions: Overview
 01:16
Hepatic Drug Excretion: Influencing Factors
 01:11
Renal Failure: Dose Adjustments
Reduced renal clearance and elimination rate are common outcomes of renal impairment. These alterations lead to a prolonged elimination half-life and an altered apparent volume of distribution for drugs. As a result, dosage adjustments are typically necessary to maintain optimal drug levels in the body.
However, dosage adjustments...
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion,...
 01:17
Drug Metabolism: Phase I Reactions
 01:09
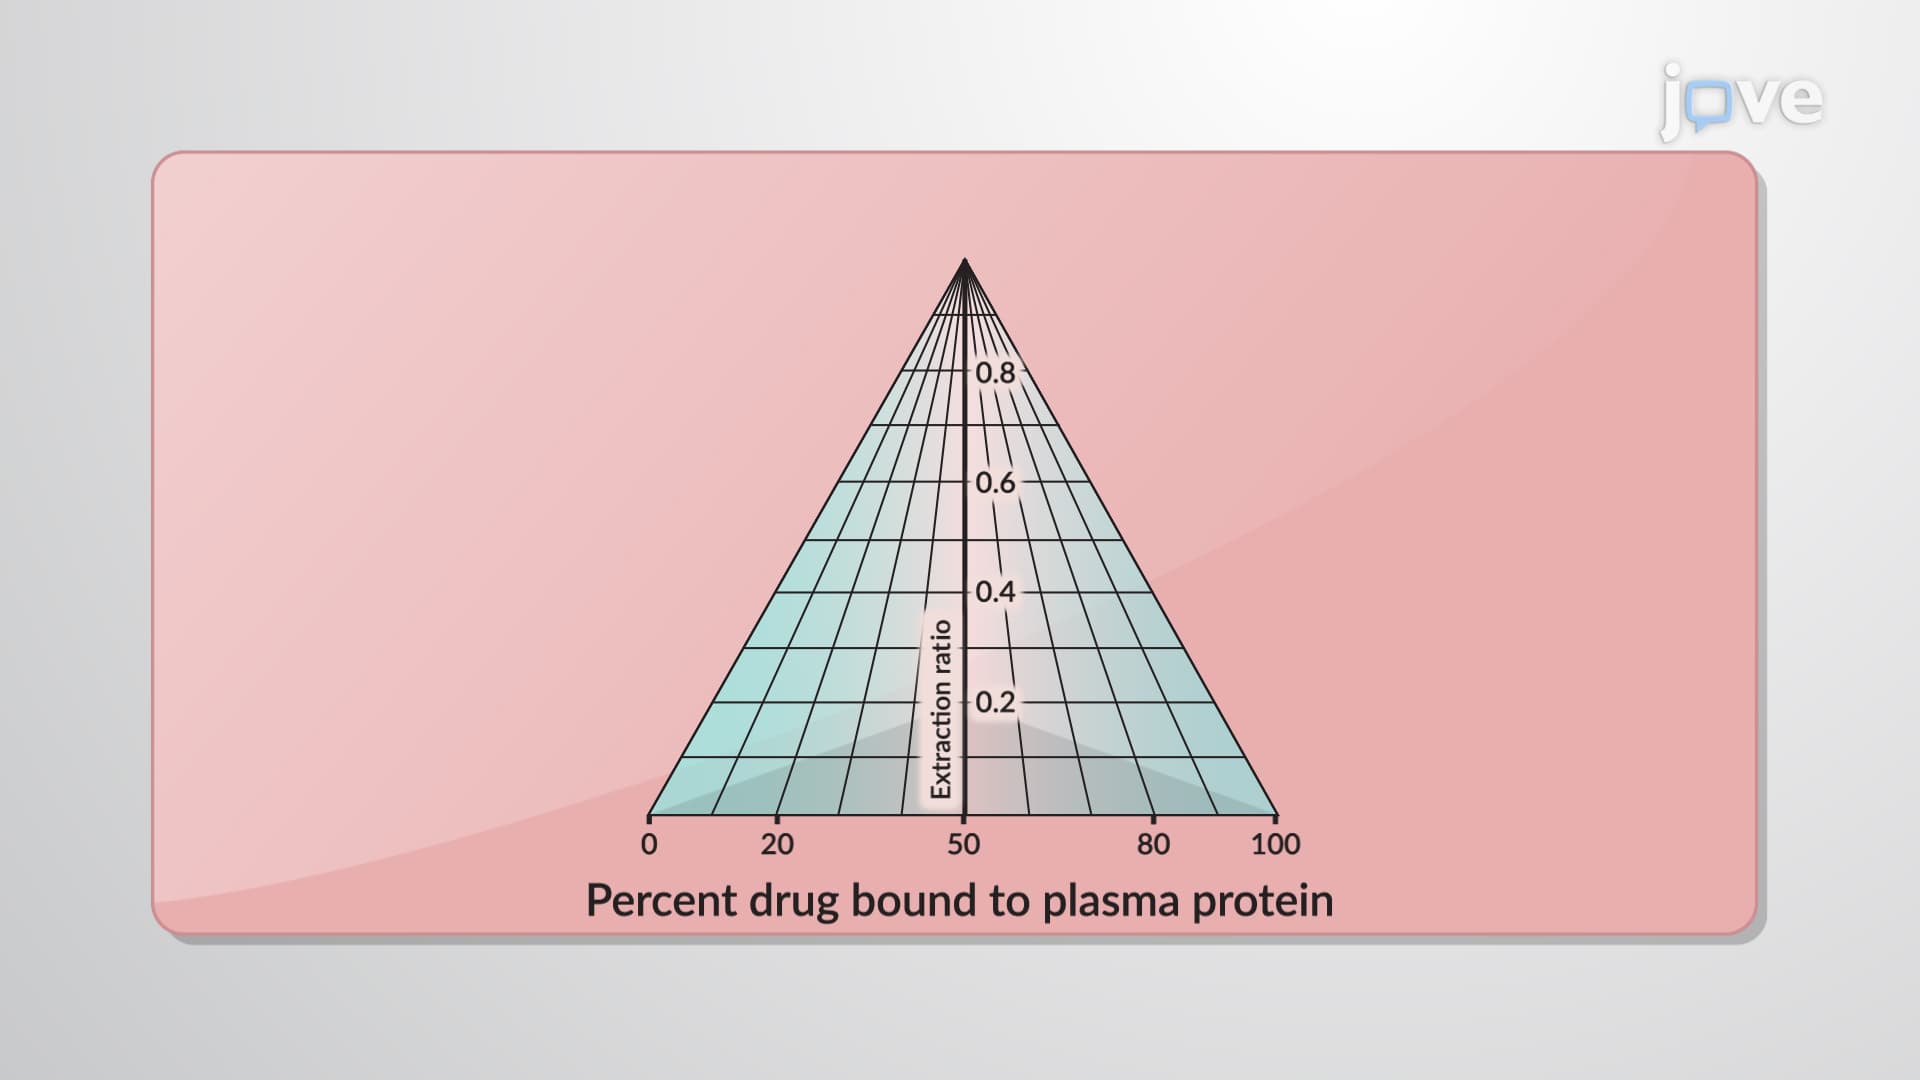
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...