Related Experiment Video
Updated: Sep 14, 2025


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
A Computationally Efficient Method to Generate Plausible Conformers for Ensemble Docking and Binding Free Energy
Ö Zeynep Güner Yılmaz1, Pemra Doruker2, Ozge Kurkcuoglu1
1Department of Chemical Engineering, Istanbul Technical University, Istanbul 34467, Turkey.

Abstract:
This study presents a computationally efficient approach to generate plausible protein conformers for ensemble docking to enable evaluations of interactions between ligand and protein for ranking the docked ligands according to their binding affinities. Two binding regions of triose phosphate isomerase (TIM), its catalytic site with DHAP (G. gallus TIM), and its dimer interface with 3PG (P. falciparumTIM) involving flexible loops were investigated as case studies. The binding sites of the apo and holo forms were modeled at the atomistic scale (high resolution) while the remaining structure was coarse-grained (low resolution) leading to a mixed-resolution description of the protein. The slowest three normal modes related to the functional dynamics of TIM were obtained using the Anisotropic Network Model and employed to derive 36 conformers of the truncated high-resolution regions by assessing six deformation parameters in both directions of the harmonic motions. Through energy minimization and docking calculations in Glide, optimal extents of deformation were identified. The docked truncated structures were then subjected to independent molecular dynamics (MD) simulations to confirm the interactions of the ligands in the binding sites. To prevent the disintegration of the truncated structure, different buffer zones and harmonic restraints were assessed to finally decide on four distinct zones with restraints of 0, 25, 35, and 50 kcal/mol·Å2. Each conformer underwent 900 ns-long simulations across three replicates reaching a total simulation time of 15.2 μs. Binding free energy calculations were conducted using the MM-GBSA approach using the first 50, first 100, first 200, and 300 ns intervals, which pointed out that 100 ns-long simulations were sufficient to estimate the binding affinities for TIM. Results consistently indicated comparable binding energies between the intact and truncated TIM structures underscoring the approach's reliability, where the truncated conformers also offered varying binding site geometries yielding favorable interactions. Comparative docking at the dimer interface of G. gallus and P. falciparum TIM further highlighted species-specific binding dynamics, affirming the methodology's applicability for diverse biological questions and establishing a computationally efficient approach to estimate binding free energy values even for supramolecular assemblages.


Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:18
The Equilibrium Binding Constant and Binding Strength
 02:27
Predicting Molecular Geometry
 02:00
Molecular Models
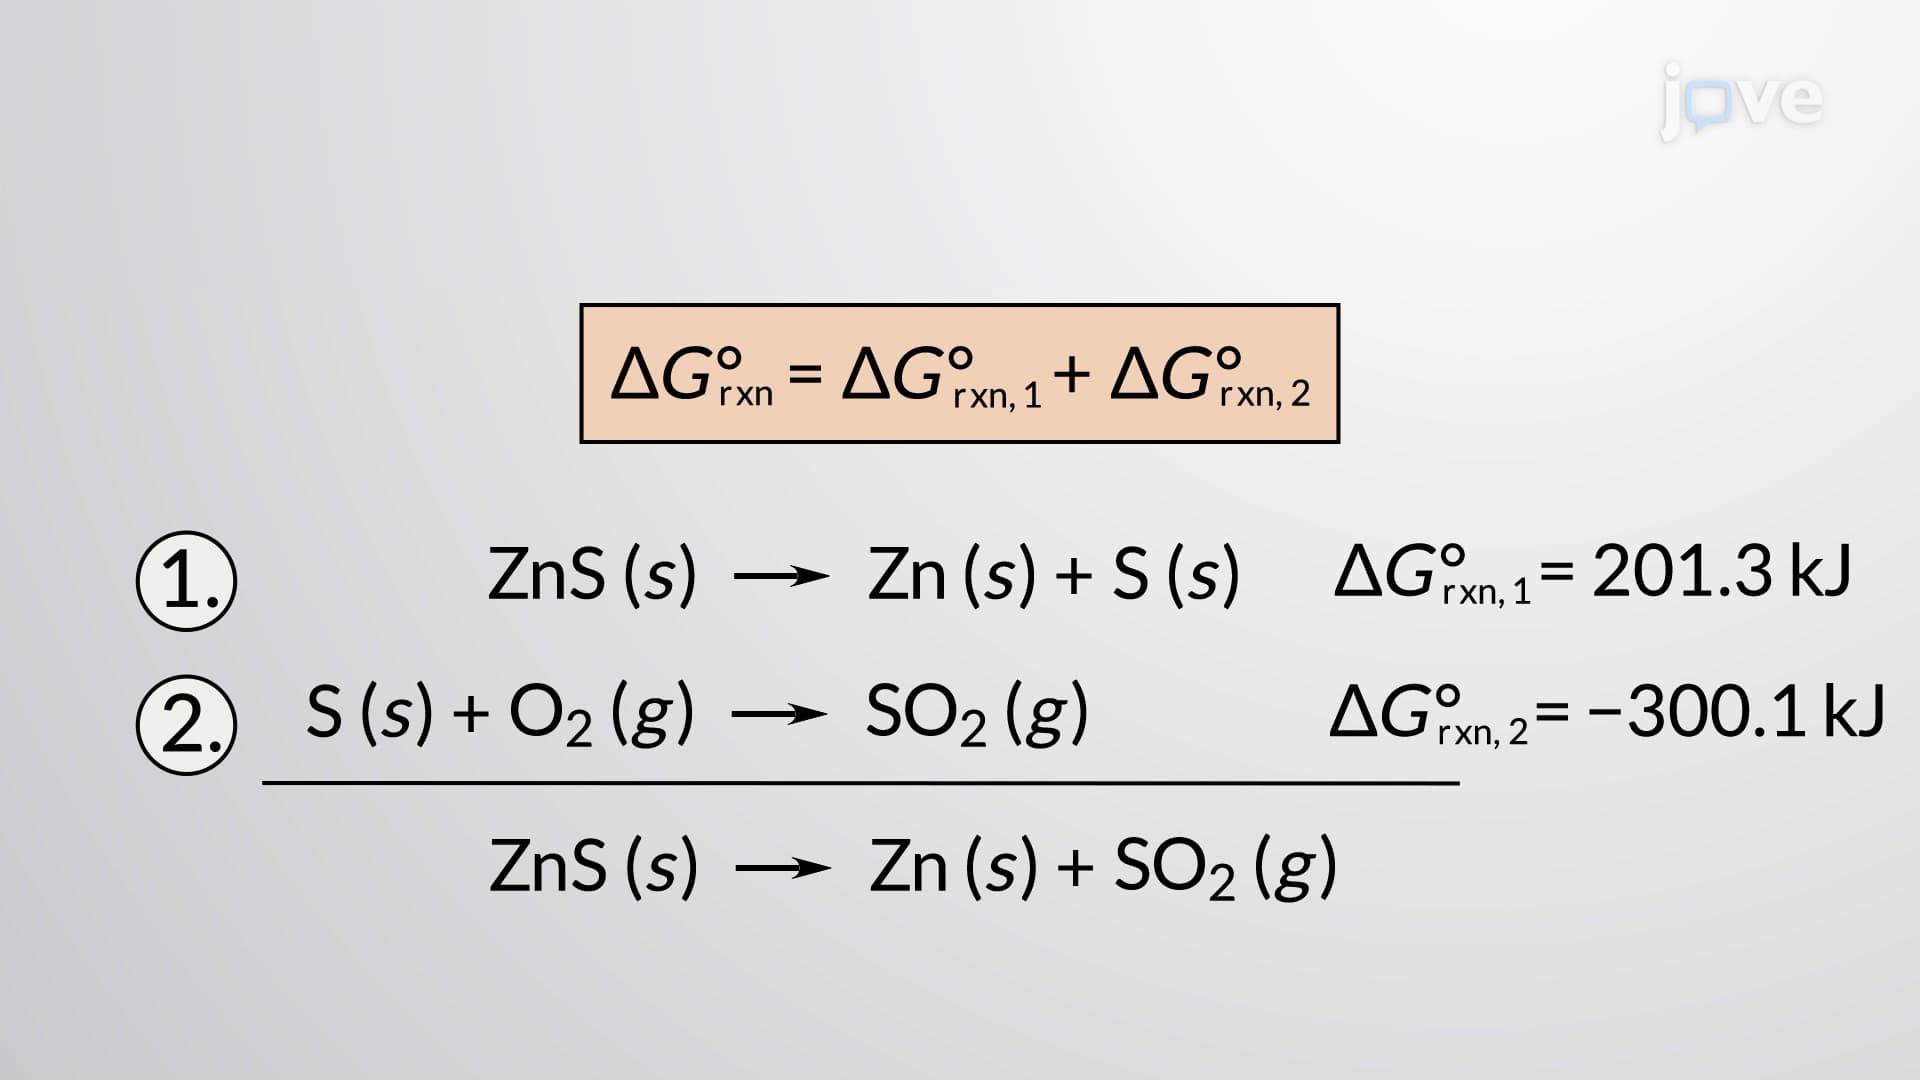 02:49
Calculating Standard Free Energy Changes
 02:20
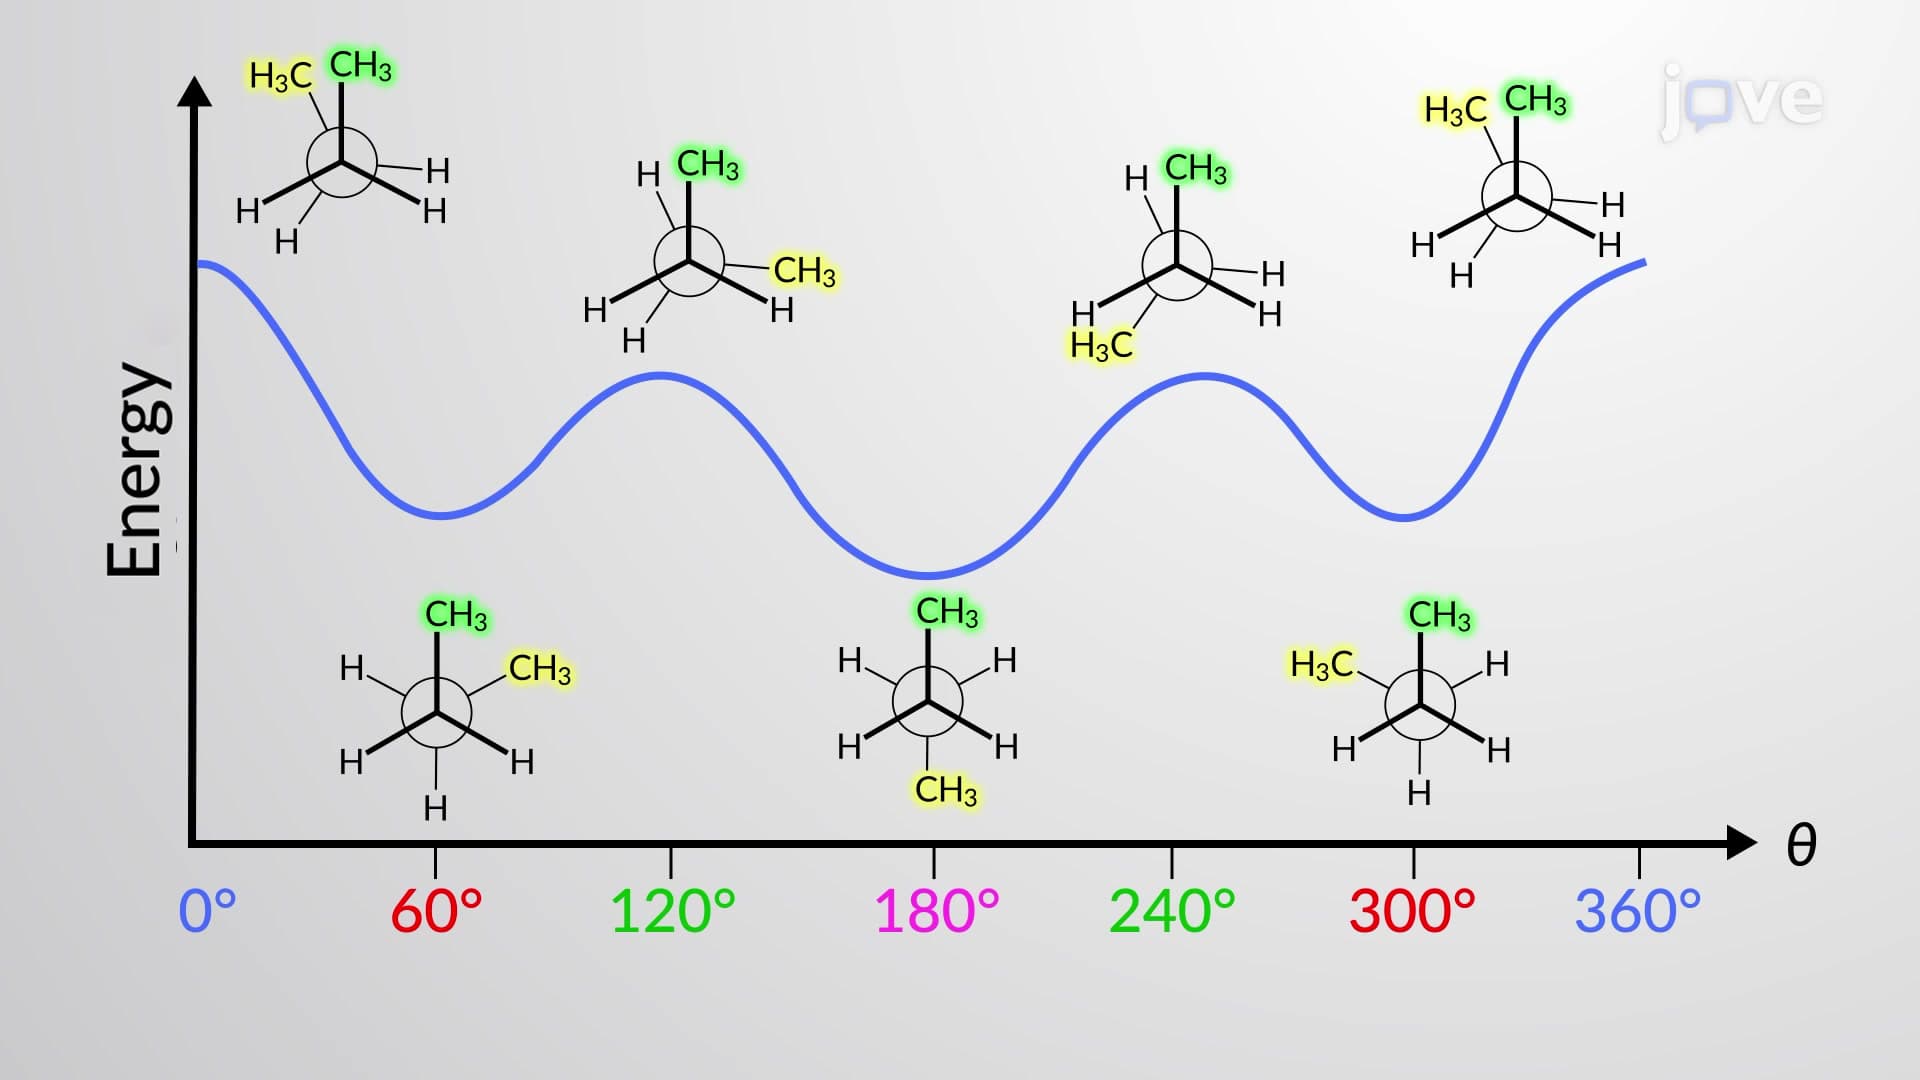
Conformations of Butane