Related Experiment Video
Updated: Sep 13, 2025

Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Apax: A Flexible and Performant Framework for the Development of Machine-Learned Interatomic Potentials
Moritz R Schäfer1, Nico Segreto1, Fabian Zills2
1Institute for Theoretical Chemistry, University of Stuttgart, Pfaffenwaldring 55, 70569 Stuttgart, Germany.

Abstract:
We introduce Atomistic learned potentials in JAX (apax), a flexible and efficient open source software package for training and inference of machine-learned interatomic potentials. Built on the JAX framework, apax supports GPU acceleration and implements flexible model abstractions for fast development. With features such as kernel-based data selection, well-calibrated uncertainty estimation, and enhanced sampling, it is tailored to active learning applications and ease of use. The features and design decisions made in apax are discussed before demonstrating some of its capabilities. First, a data set for the room-temperature ionic liquid EMIM+BF4- is created using active learning. It is highlighted how continuously learning models between iterations can reduce training times up to 85% with only a minor reduction of the models' accuracy. Second, we show good scalability in a data-parallel training setting. We report that a Gaussian Moment Neural Network model, as implemented in apax, achieves higher accuracy and up to 10 times faster inference times than a performance-optimized Allegro model. A recently published Li3PO4 data set, reported with comparable accuracy and inference performance metrics, is used as a point of comparison. Moreover, the inference speeds of the available simulation engines are compared. Finally, to highlight the modularity of apax, an equivariant message-passing model is trained as a shallow ensemble and used to perform uncertainty-driven dynamics.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:44
Atomic Orbitals
 02:27
Predicting Molecular Geometry
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 01:26
Thermodynamic Potentials
 01:13
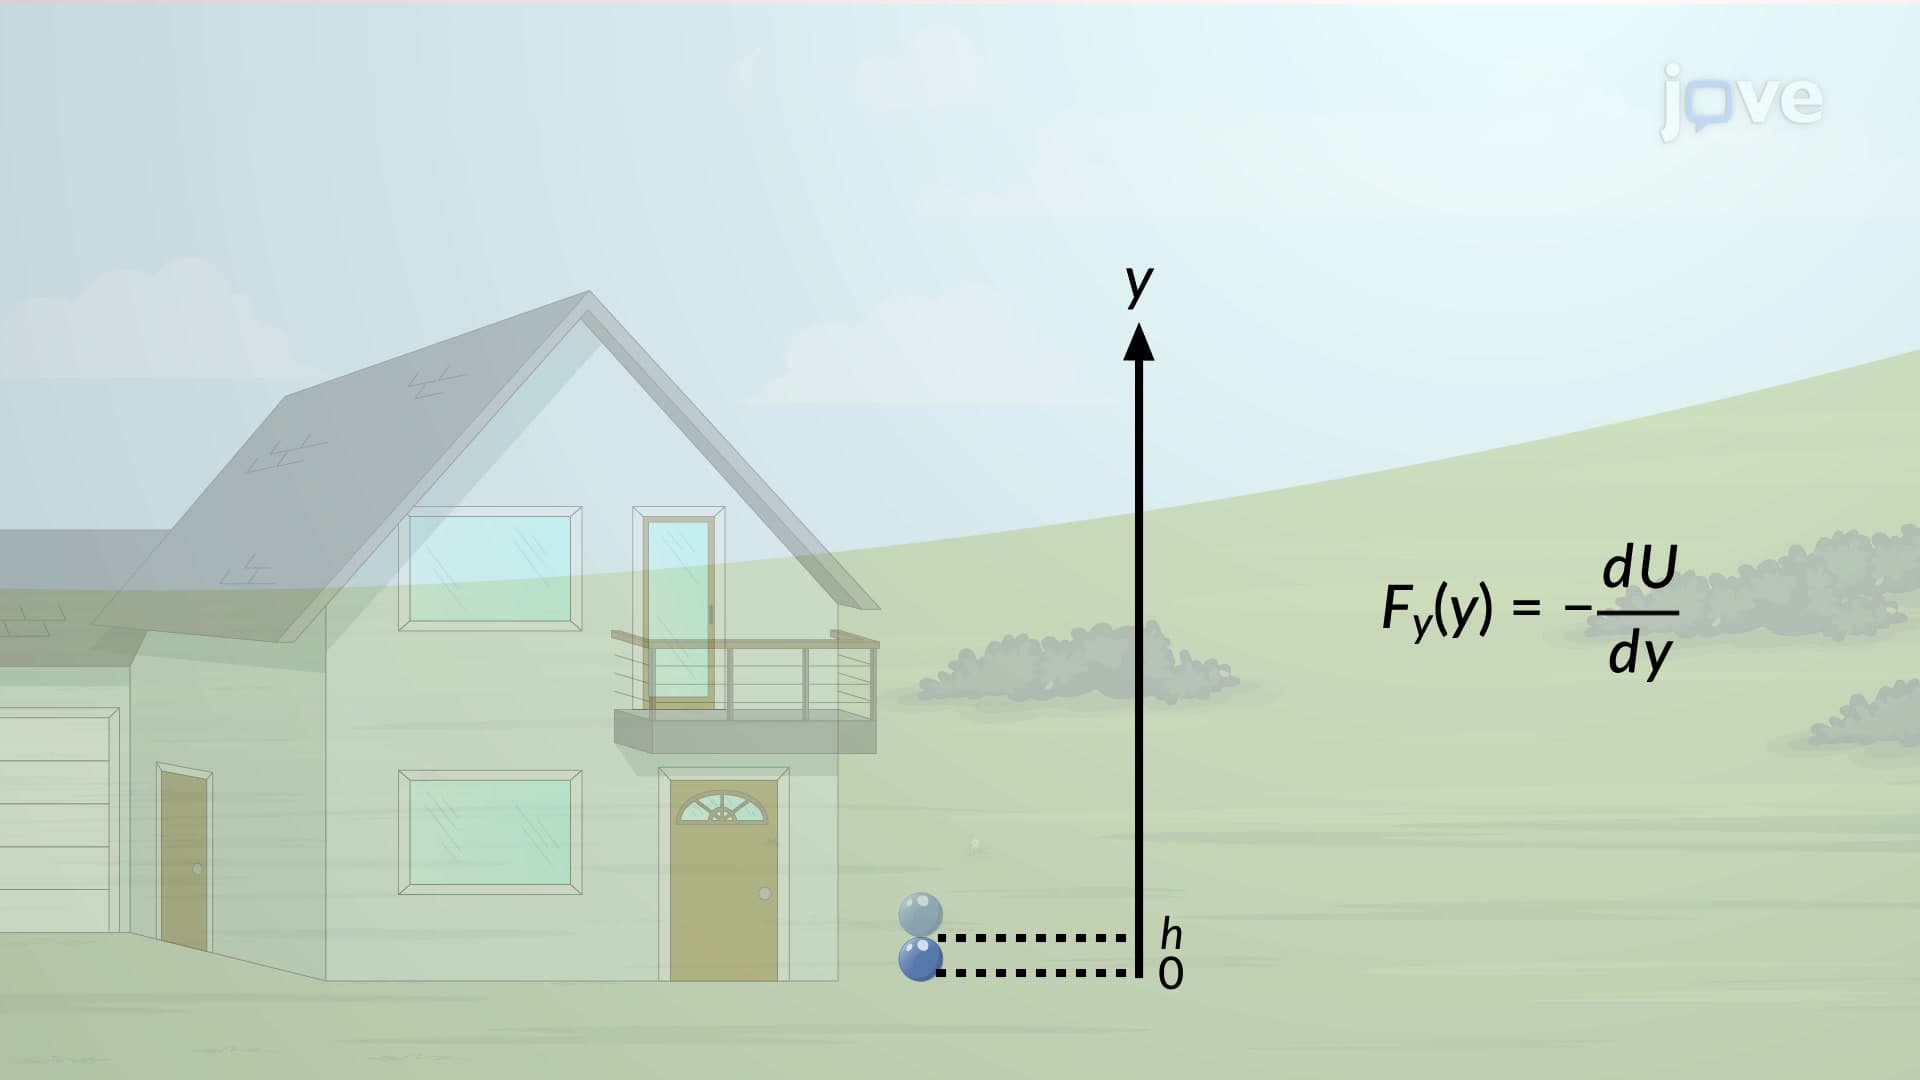
Force and Potential Energy in One Dimension
 02:35
Molecular Orbital Theory I