Related Experiment Video
Updated: Sep 12, 2025
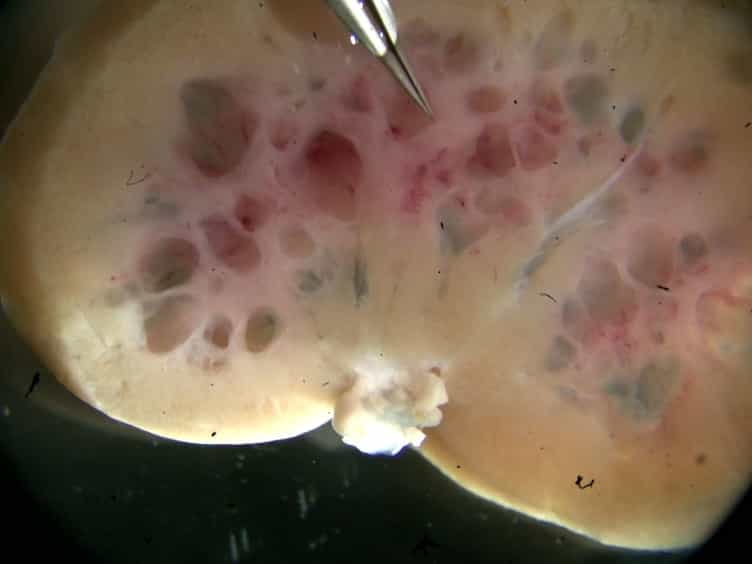

Implementing Patch Clamp and Live Fluorescence Microscopy to Monitor Functional Properties of Freshly Isolated PKD Epithelium
Published on: September 1, 2015
A novel gene therapy for ARPKD based on CFTR
Cristian Ciobanu1,2, Patricia Outeda3, William B Guggino2
1Department of Medicine, Johns Hopkins University, Baltimore, Maryland, United States.

Abstract:
Autosomal recessive polycystic kidney disease (ARPKD) is associated with cysts derived from abnormal bile ducts. We focused on targeting the cysts and show that a gene therapy for ARPKD that targets the abnormal bile ducts is feasible. We injected 1-mo-old, Pkhd1del3-4/del3-4 mice intraperitoneally with 2 × 10¹² particles/kg of adeno-associated virus (AAV1) containing either a GFP vector or a truncated cystic fibrosis transmembrane conductance regulator (CFTR) vector, Δ27-264-CFTR, or left them untreated. Two months after treatment, the cyst area and size in the liver were lower in the CFTR vector-treated mice than in mice receiving the GFP vector. We detected vector genomes and mRNA expression only in mice receiving the corresponding CFTR or GFP vector. We observed abundant GFP immunofluorescence in the cholangiocytes of the cysts and also saw expression of GFP and CFTR proteins above background levels in the corresponding treated mice. CFTR immunofluorescence was predominantly apically located in the ARPKD cholangiocytes, but after CFTR vector installation, it increased in the basolateral membrane. We stained mouse livers with Maackia amurensis lectin (MAL) or Sambucus nigra lectin (SNA), specific for α2,3- and α2,6-N-linked sialic acid, respectively, to detect the presence of sialic acid moieties contributing to AAV1 binding. Although immunofluorescent SNA was detected in the wild-type bile ducts, MAL 1 was not. MAL immunofluorescence was present in remarkably high levels on the apical surfaces of the cysts in cholangiocytes, offering a good target for AAV gene therapy. A gene therapy using an AAV1-based vector containing a truncated CFTR could be therapeutic in ARPKD.NEW & NOTEWORTHY Autosomal recessive polycystic kidney disease (ARPKD) causes severe disease in babies in the womb. Those who survive the neonatal period face chronic kidney and liver disease throughout their life. The overall goal of our study here is to develop a gene therapy to treat ARPKD.
More Related Videos
 07:04
Forskolin-induced Swelling in Intestinal Organoids: An In Vitro Assay for Assessing Drug Response in Cystic Fibrosis Patients
Published on: February 11, 2017
 08:00
Generation of Human Nasal Epithelial Cell Spheroids for Individualized Cystic Fibrosis Transmembrane Conductance Regulator Study
Published on: April 11, 2018
Related Concept Videos
 00:59
Gene Therapy
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...