Related Experiment Video
Updated: Sep 12, 2025

Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Investigating the electronic structure and adsorption of OCN- and CN- on Si12C12 Fullerene-like cages using DFT and
Abdulrahman Sumayli1, Saad M Alshahrani2
1Department of Mechanical Engineering, College of Engineering, Najran University, Najran, Saudi Arabia. aisumayli@nu.edu.sa.

Abstract:
The adsorption behavior of the toxic molecules OCN- and CN- onto Si12C12 fullerene-like cages was performed using density functional theory (DFT) calculations with the B3LYP and PBE functionals and a 6-311 + G** basis set, in both vacuum and aqueous phases (PCM model). Strong adsorption of OCN⁻ (nitrogen head) and CN⁻ (carbon head) to the cage surface results from covalent bonding and significant charge transfer from the fullerene-like cage to the anions in both phases. These adsorption processes, involving OCN- via its nitrogen head and CN- via its carbon head, were both spontaneous and exothermic, as indicated by the calculated adsorption energies and thermodynamic parameters. MD simulations show rapid OCN⁻ covalent bonding to the Si₁₂C₁₂ cage, followed by equilibration within 5 ps. The OCN⁻ exhibits higher mobility than the cage, as indicated by MSD analysis. This dynamic interaction, revealed by g(r) and MSD data, suggests a surface adsorption process. The impacts of OCN- and CN- interaction with the fullerene-like cage surface were assessed using infrared (IR) absorption spectroscopy. The photoelectron spectrum of OCN- and CN- adsorption suggests interactions between the fullerene-like cage and these molecules, emphasizing the involvement of strongly bonded electrons in facilitating these interactions. Significantly, during the adsorption process, there was a notable orbital hybridization between anions and fullerenes, as evidenced by the calculated total density of states (TDOS). DFT calculations show that Si12C12 fullerene-like cage exhibits strong binding energies for OCN⁻ and CN⁻, suggesting its potential for highly efficient removal in environmental samples.
More Related Videos
 11:38
In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
 08:18
Microscopic Visualization of Porous Nanographenes Synthesized through a Combination of Solution and On-Surface Chemistry
Published on: March 4, 2021
Related Concept Videos
 02:54
Lewis Structures of Molecular Compounds and Polyatomic Ions
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:18
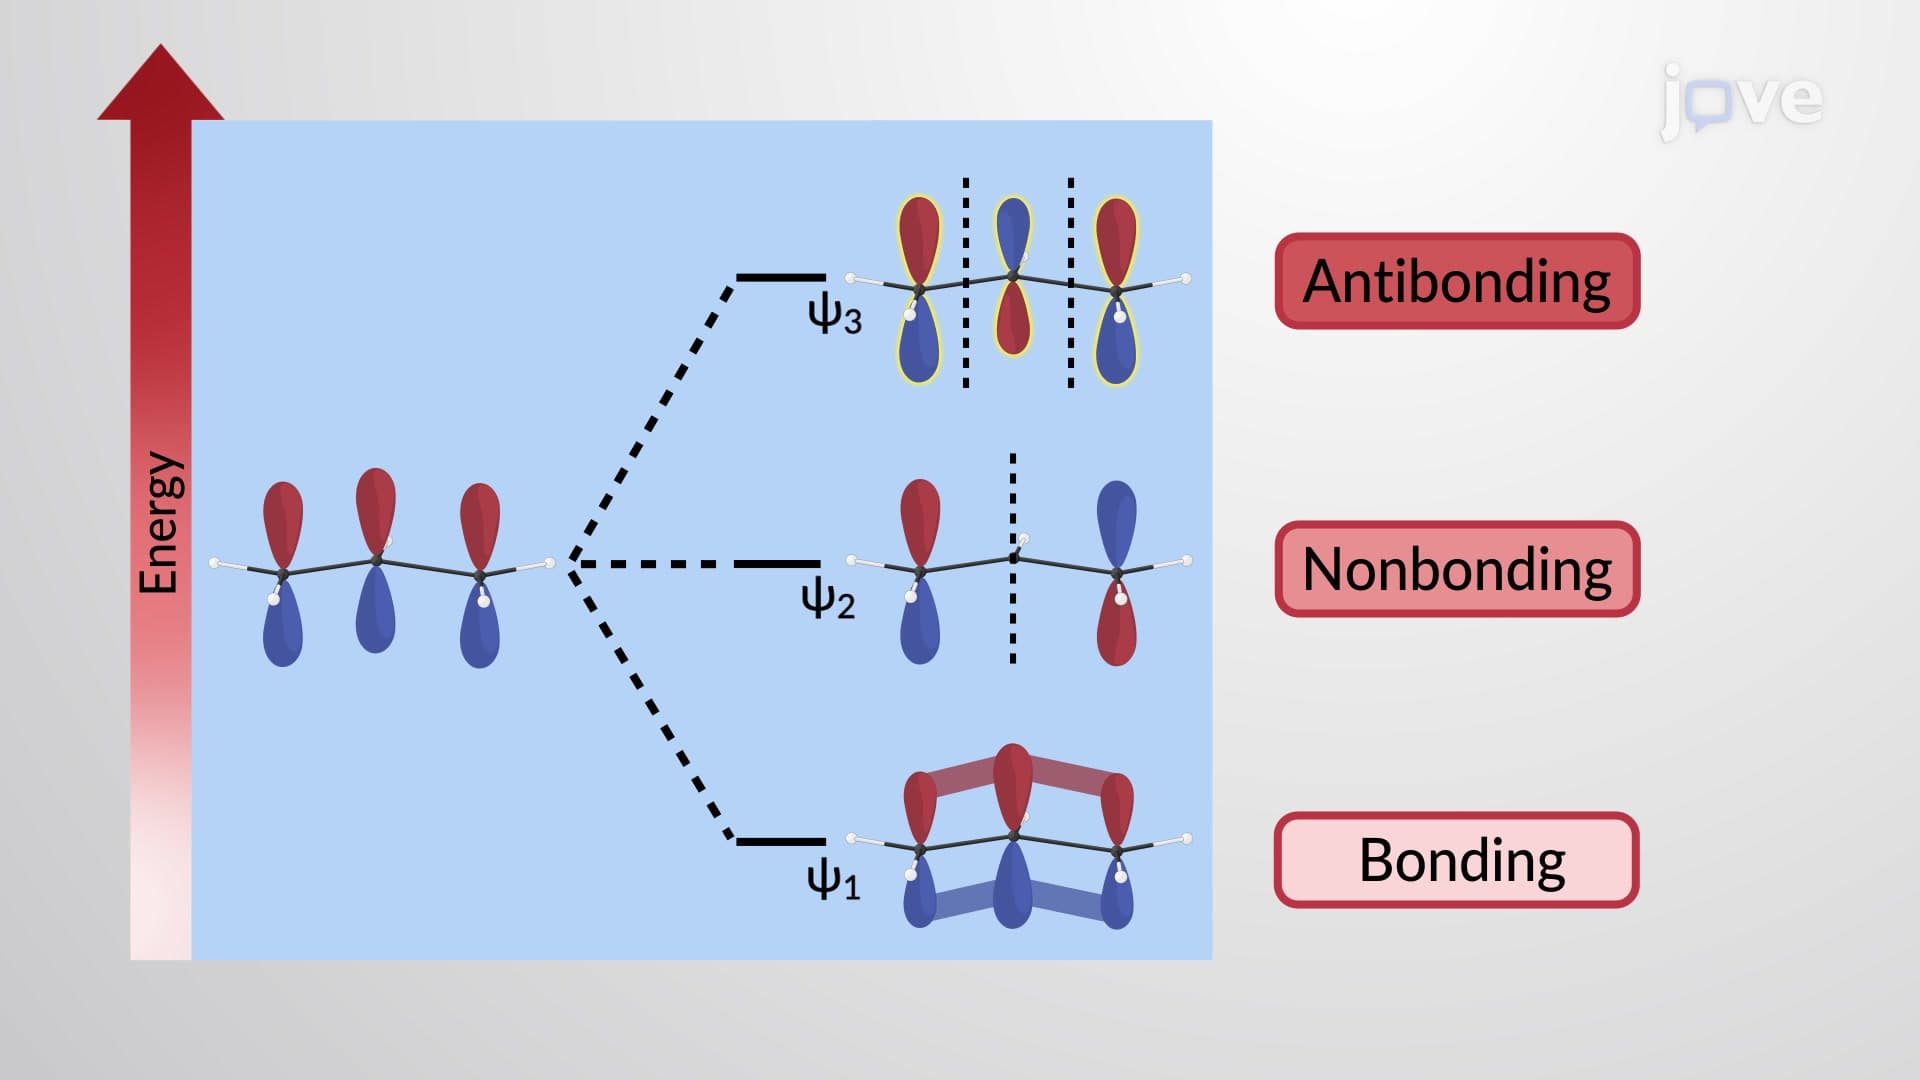
π Molecular Orbitals of the Allyl Cation and Anion
 02:46
Electron Configurations
The relative energies of the subshells determine the order in which atomic orbitals are filled (1s, 2s, 2p, 3s, 3p,...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...