Related Experiment Video
Updated: Sep 12, 2025

Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
Accurate prediction of drug-protein interactions by maintaining the original topological relationships among
Yanfei Li1, Xiran Chen2, Shuqin Wang3
1College of Computer Science, Nankai University, 300071, Tianjin, China.

Background:
Learning-based methods have recently demonstrated strong potential in predicting drug-protein interactions (DPIs). However, existing approaches often fail to achieve accurate predictions on real-world imbalanced datasets while maintaining high generalizability and scalability, limiting their practical applicability.
Results:
This study proposes a highly generalized model, GLDPI, aimed at improving prediction accuracy in imbalanced scenarios by preserving the topological relationships among initial molecular representations in the embedding space. Specifically, GLDPI employs dedicated encoders to transform one-dimensional sequence information of drugs and proteins into embedding representations and efficiently calculates the likelihood of DPIs using cosine similarity. Additionally, we introduce a prior loss function based on the guilt-by-association principle to ensure that the topology of the embedding space aligns with the structure of the initial drug-protein network. This design enables GLDPI to effectively capture network relationships and key features of molecular interactions, thereby significantly enhancing predictive performance.
Conclusions:
Experimental results highlight GLDPI's superior performance on multiple highly imbalanced benchmark datasets, achieving over a 100% improvement in the AUPR metric compared to state-of-the-art methods. Additionally, GLDPI demonstrates exceptional generalization capabilities in cold-start experiments, excelling in predicting novel drug-protein interactions. Furthermore, the model exhibits remarkable scalability, efficiently inferring approximately drug-protein pairs in less than 10 h.
More Related Videos
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
 08:31
Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
Related Concept Videos
 02:04
Protein-protein Interfaces
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:23
Factors Affecting Protein-Drug Binding: Drug Interactions
Displacement interactions can have varying outcomes, ranging from toxicity to virtually...
 01:16
Protein-Drug Binding: Mechanism and Kinetics
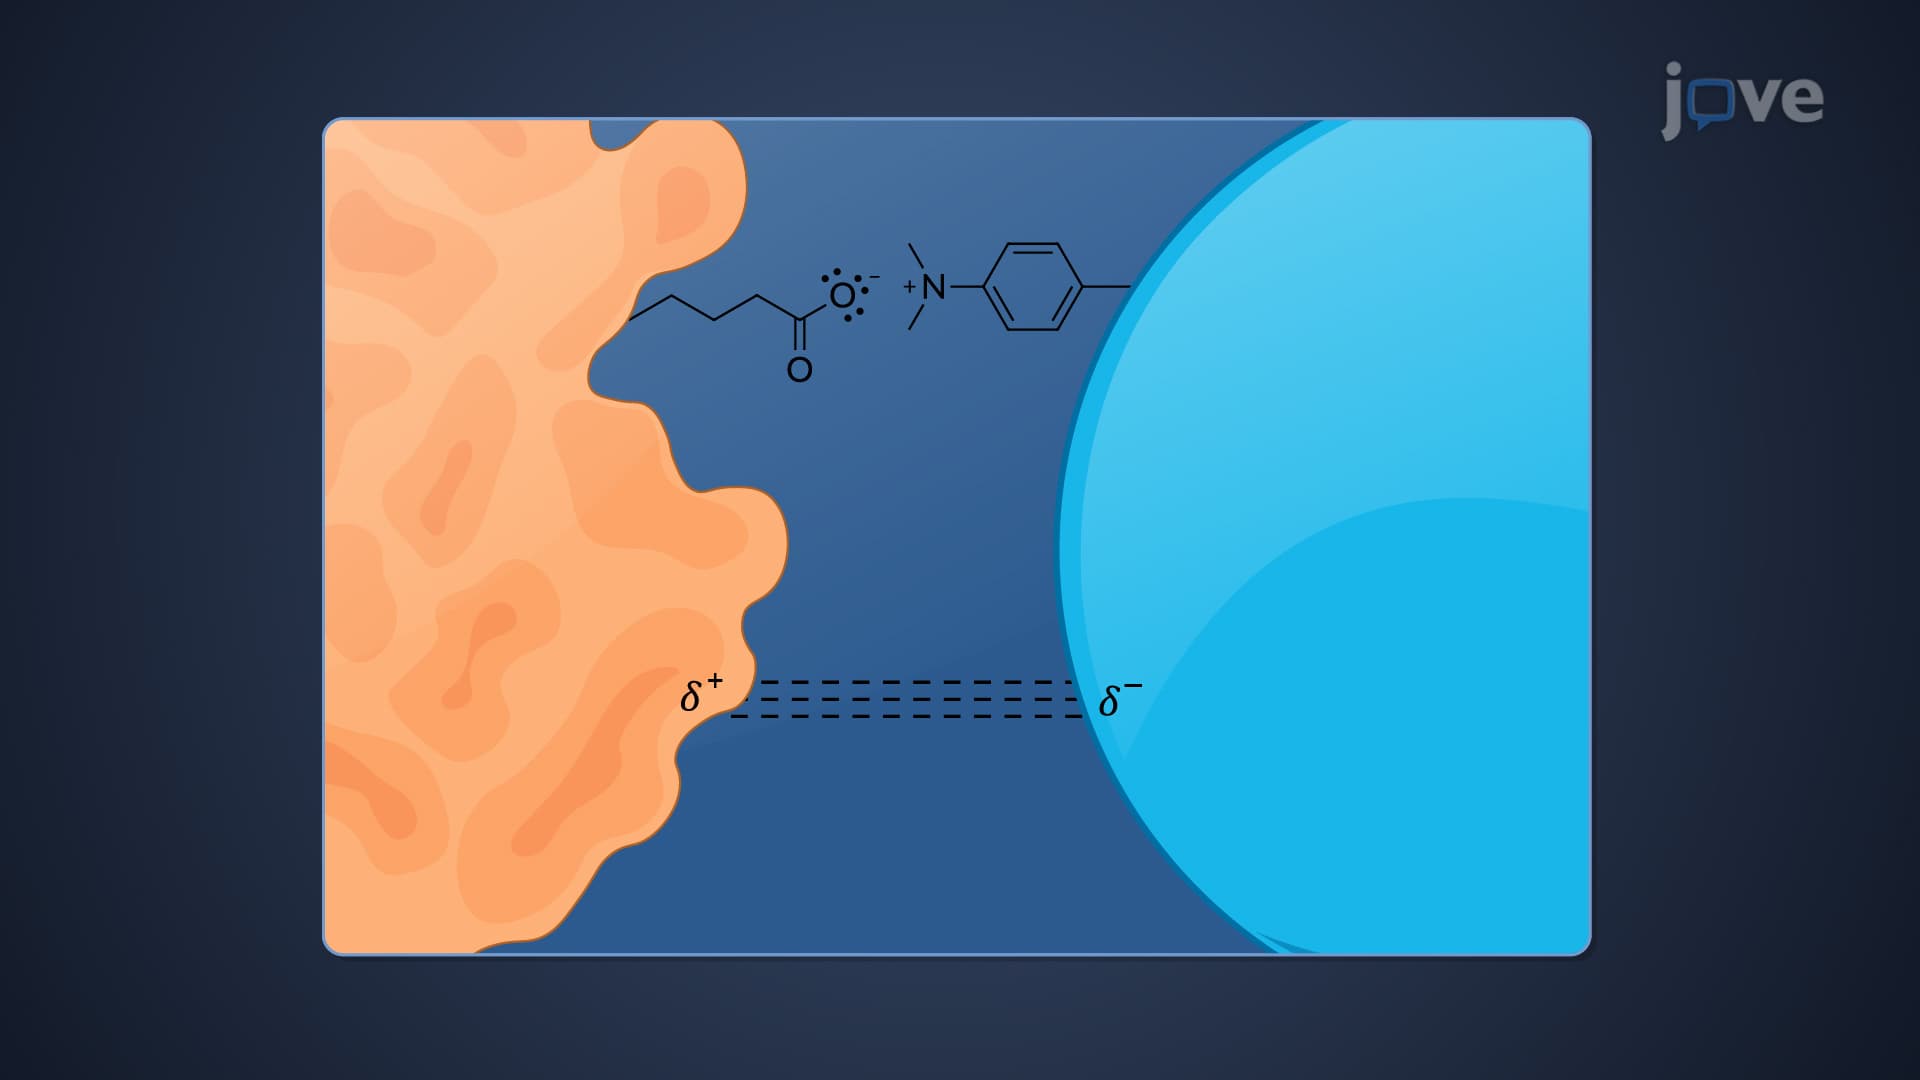
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:22
Protein-Drug Binding: Determination Methods
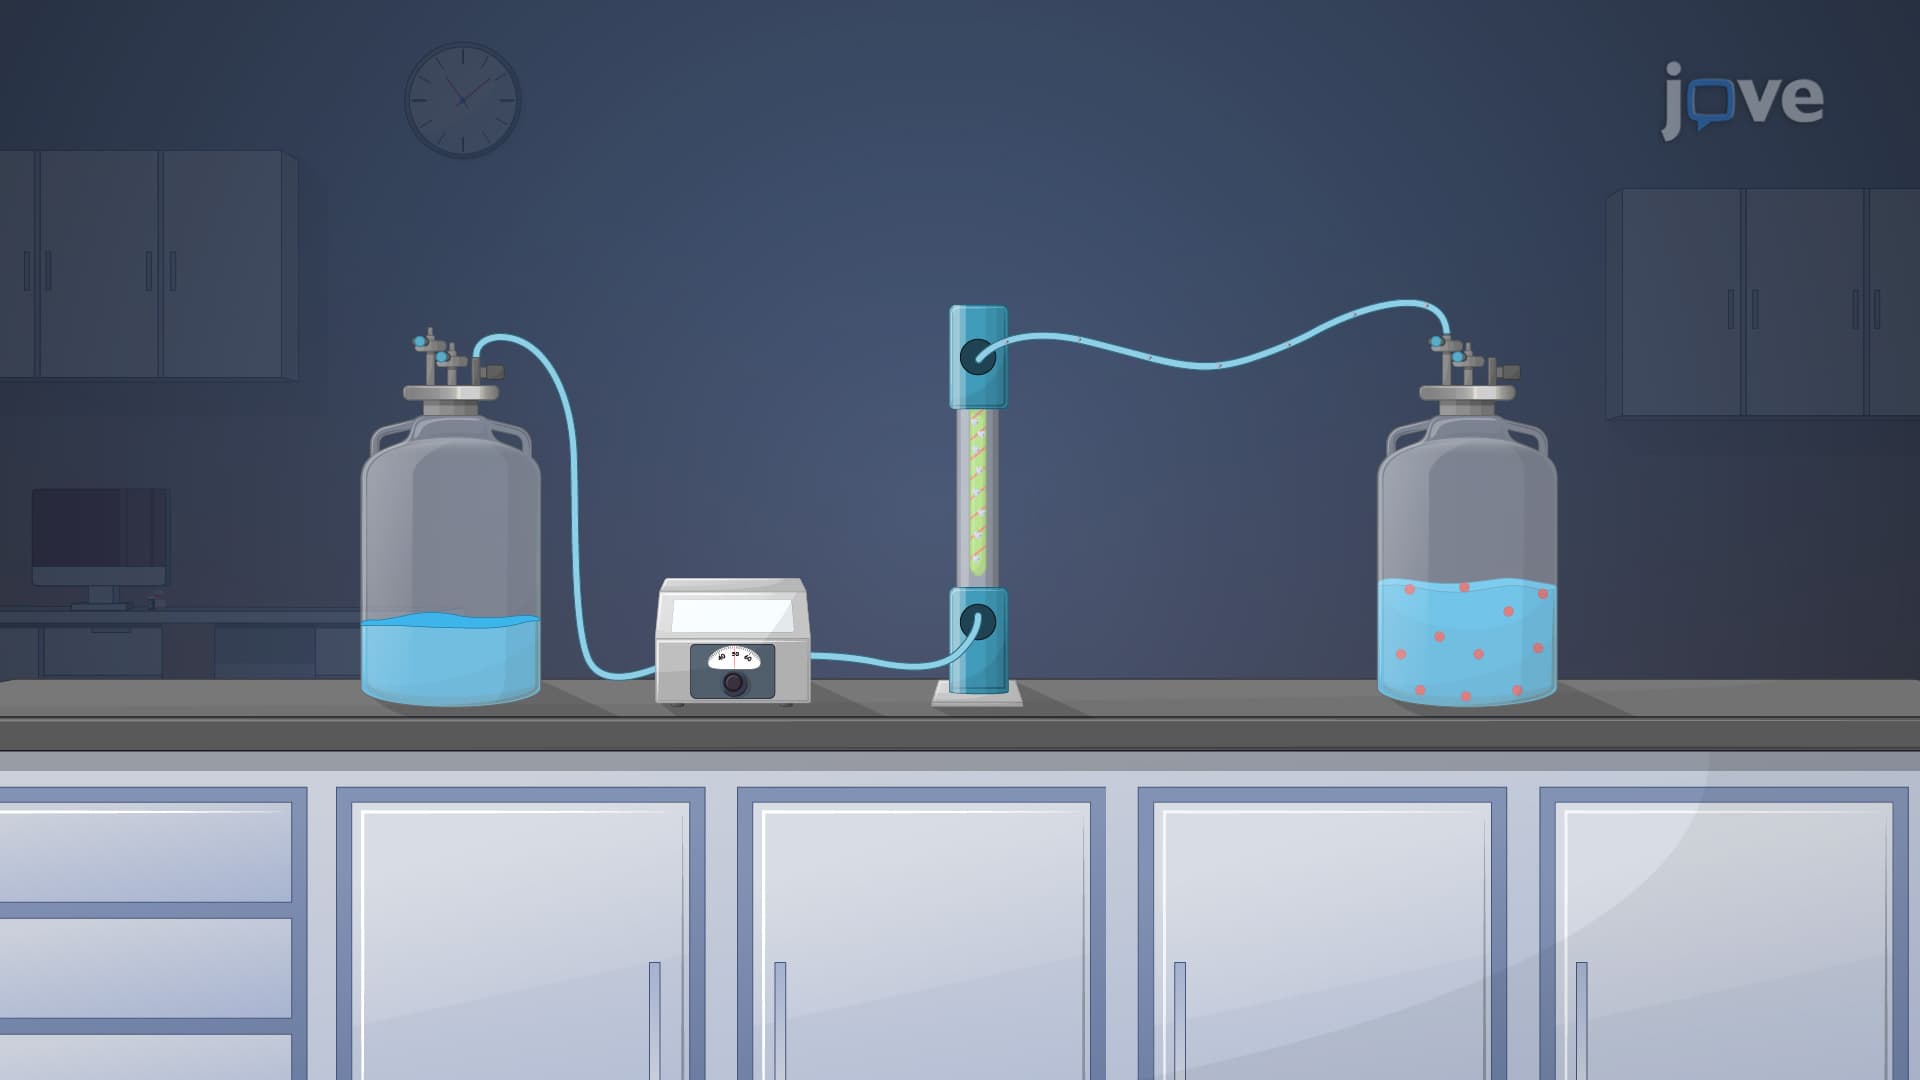
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...