Related Experiment Video
Updated: Sep 11, 2025

Methods of Ex Situ and In Situ Investigations of Structural Transformations: The Case of Crystallization of Metallic Glasses
Published on: June 7, 2018
Atomic scale structure and dynamical properties of (TeO2)1-x-(Na2O)x glasses through first-principles modeling and
Firas Shuaib1, Assil Bouzid1, Remi Piotrowski1
1Institut de Recherche sur les Céramiques (IRCER), UMR CNRS 7315, F-87068 Université de Limoges, Centre Européen de la Céramique, 12 rue Atlantis, Limoges, France. assil.bouzid@cnrs.fr.

Abstract:
We resort to first-principles molecular dynamics, in synergy with experiments, to study structural evolution and Na+ cation diffusion inside (TeO2)1-x-(Na2O)x (x = 0.10-0.40) glasses. Experimental and modeling results show a fair quantitative agreement in terms of total X-ray structure factors and pair distribution functions, thereby setting the ground for a comprehensive analysis of the glassy matrix evolution. We find that the structure of (TeO2)1-x-(Na2O)x glasses deviates drastically from that of pure TeO2 glass. Specifically, increasing the Na2O concentration leads to a reduction of the coordination number of Te atoms, reflecting the occurrence of a structural depolymerization upon introduction of the Na2O modifier oxide. The depolymerization phenomenon is ascribed to the transformation of Te-O-Te bridges into terminal Te-O non bridging oxygen atoms (NBO). Consequently, the concentration of NBO increases in these systems as the concentration of the modifier increases, accompanied by a concomitant reduction in the coordination number of Na atoms. The structure factors results show a prominent peak at ∼1.4 Å that becomes more and more pronounced as the Na2O concentration increases. The occurrence of this first sharp diffraction peak is attributed to the growth of Na-rich channels inside the amorphous network, acting as preferential routes for alkali-ion conduction inside the relatively stable Te-O matrix. These channels enhance the ion mobility.
Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:18
X-ray Crystallography
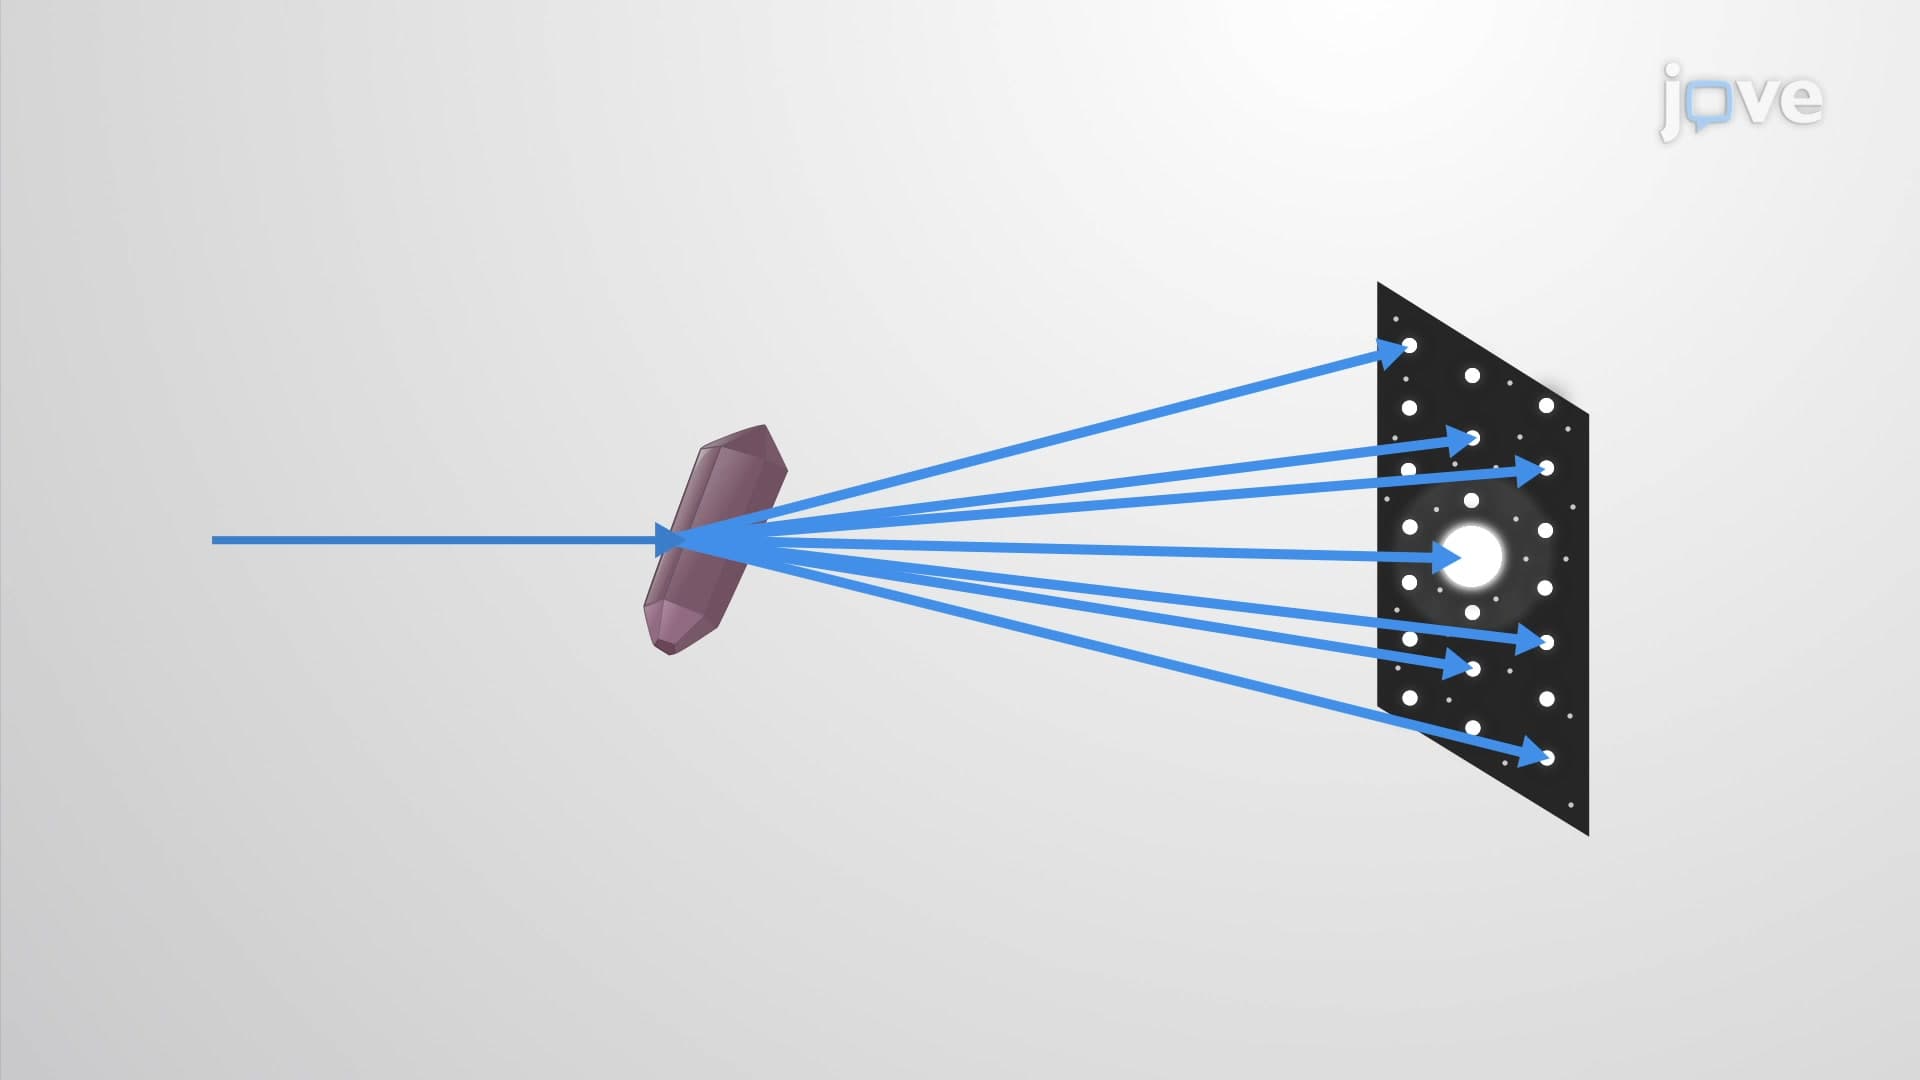
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 02:22
Structures of Solids