

Ligand-Based Pharmacophore Modeling and Quantum Computational Approaches for the Development of hCA XII Inhibitor

Abstract:
Globally, there is a threat to public health regarding the prevalence of cancer and its resistance problem. Among the various cancer therapy strategies, targeting Carbonic anhydrase (CA) has gained a lot of attention in recent times. CA XII was found to be a biomarker for the suppression of various hypoxic tumors in both the early and metastatic stages. This work aims to identify new lead CA XII inhibitors without inhibiting other isoforms and to replace the benzene sulfonamide scaffold. Existing lead molecules were computationally investigated, screened, and subjected to pharmacokinetic studies, molecular docking, DFT, and molecular dynamics simulation. A pharmacophore model was developed from the existing CA inhibitors, and 43 hit molecules were identified. All the hit molecules passed ADMET, and the molecular docking results revealed that among the identified novel CA inhibitors, compounds 3 and 26 showed promising leads with low energy conformation (-7.82 K/Cal and -7.36 K/Cal) and subsequently provided structural insights against target inhibition compared to the standard drug acetazolamide. RMSD and RMSF values followed by MMGBSA calculations revealed that compounds 3 and 26 are more stable than the standard. The study concludes that compounds 3 and 26 can be a new lead for CA XII inhibition.
Insights
New research identifies novel compounds 3 and 26 as potent inhibitors of Carbonic anhydrase XII (CA XII), a key target for hypoxic tumors. These promising leads offer a potential new therapeutic strategy for cancer treatment.
Area of Science:
- Biochemistry
- Medicinal Chemistry
- Computational Chemistry
Background:
- Cancer remains a significant global health threat, with drug resistance posing a major challenge.
- Targeting specific carbonic anhydrase (CA) isoforms, like CA XII, is a promising strategy for cancer therapy.
- CA XII is implicated as a biomarker in various hypoxic tumors, necessitating the development of selective inhibitors.
Purpose of the Study:
- To discover novel Carbonic anhydrase XII (CA XII) inhibitors.
- To move beyond the traditional benzene sulfonamide scaffold.
- To ensure selectivity against other CA isoforms.
Main Methods:
- Computational screening of existing lead molecules.
- Pharmacophore modeling and virtual screening to identify hit compounds.
- In silico pharmacokinetic (ADMET) and molecular dynamics simulations.
- Molecular docking, DFT, and MMGBSA calculations for stability and binding affinity assessment.
Main Results:
- A pharmacophore model yielded 43 hit molecules, all passing ADMET predictions.
- Compounds 3 and 26 demonstrated strong binding affinities and low energy conformations (-7.82 K/Cal and -7.36 K/Cal) via molecular docking.
- Molecular dynamics simulations indicated compounds 3 and 26 exhibit superior stability compared to acetazolamide.
- Structural insights were gained into the inhibition mechanism against CA XII.
Conclusions:
- Compounds 3 and 26 represent promising novel lead candidates for CA XII inhibition.
- These compounds offer a potential alternative scaffold to benzene sulfonamides.
- Further development of compounds 3 and 26 could lead to new anti-cancer therapeutics.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:15
G Protein-coupled Receptors
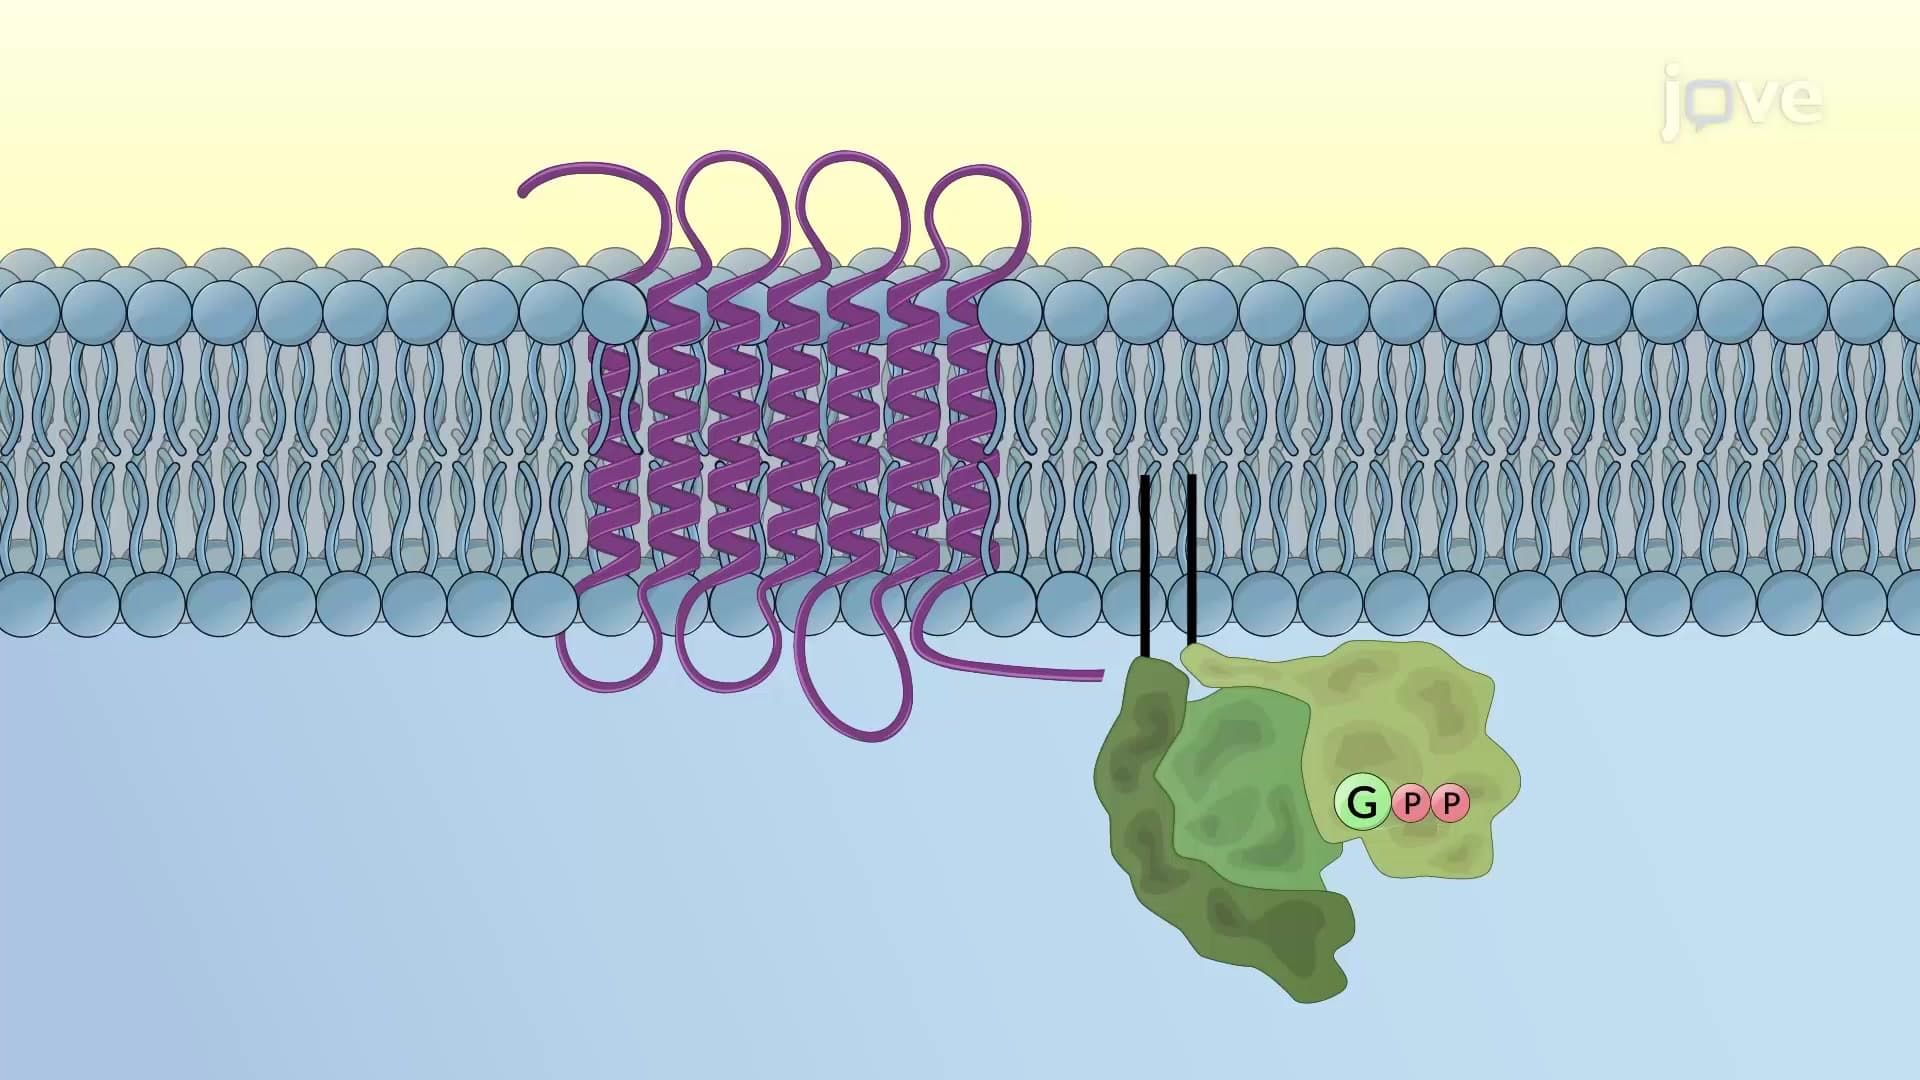
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...