Related Experiment Video
Updated: Sep 10, 2025


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
A statistical theory of reactivity based on molecular orbitals participation and its application to organic reactions
Javier Oller1,2, Paul Geerlings3, Frank De Proft3
1Departamento de Química Orgánica y Fisicoquímica, Facultad de Ciencias Químicas y Farmacéuticas, Universidad de Chile, Olivos 1007, Santiago, Chile. javier.oller@ciq.uchile.cl.

Abstract:
The FMO theory considers the highest occupied molecular orbital (HOMO) and lowest unoccupied molecular orbital (LUMO) as drivers of chemical reactivity. This theory has been successfully applied in many fields, but there are some cases where the reactivity is not well described by FMO even if orbitally controlled. In order to consider the participation of the molecular orbitals (MOs) beyond the frontier ones in the reactivity and also to explain the different reactivity possibilities that a same reagent can have (which can be correlated to experimental products e.g., major and minor products), we propose a theory that is based on two fundamental principles: the first one states that the participation/contribution of the MOs in the reactivity is not absolutely defined, therefore there are many possibilities involving just the reagent R1, that are related to many possibilities of reactivity. The second one states that the interaction between R1 and the second reagent R2 reduces these many possibilities of participation/contribution to a final option, which is related to just one type of reactivity, that can be correlated with experimental data. The search for these many possibilities of participation of MOs in the reactivity is based on an extension of the three-state grand canonical ensemble model and then because our theory focuses on non-ionization reactions, nucleophilic and electrophilic probabilities are derived. Electron densities are directly evaluated from the MOs, together with their orbital energies related to a microstate, and at the end an ensemble average electron density (average Fukui function) depending on the respective microstate probabilities at a constant kTp value is calculated. Upon varying kTp three reactivity zones: the FMO zone, the many-state zone (kTp ≈ ΔEij) and the no information zone appear in a natural way, each of them linked to their reactivity information content. We study the reactivity in electrophilic aromatic substitutions in substituted benzenes, hetero aromatic molecules and in polycyclic aromatic hydrocarbons. Furthermore, the nucleophilic addition reaction on acrolein and benzaldehyde, and the nucleophilic substitution on quinoline is scrutinized. With our theory it was possible to describe the reactivity correctly when the FMO theory is incomplete or incorrect, and also when FMO is correct according to experiment.


Related Concept Videos
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding
 02:35
Molecular Orbital Theory I
 01:11
Radical Reactivity: Overview
 02:43
E1 Reaction: Stereochemistry and Regiochemistry
 02:43
E2 Reaction: Stereochemistry and Regiochemistry
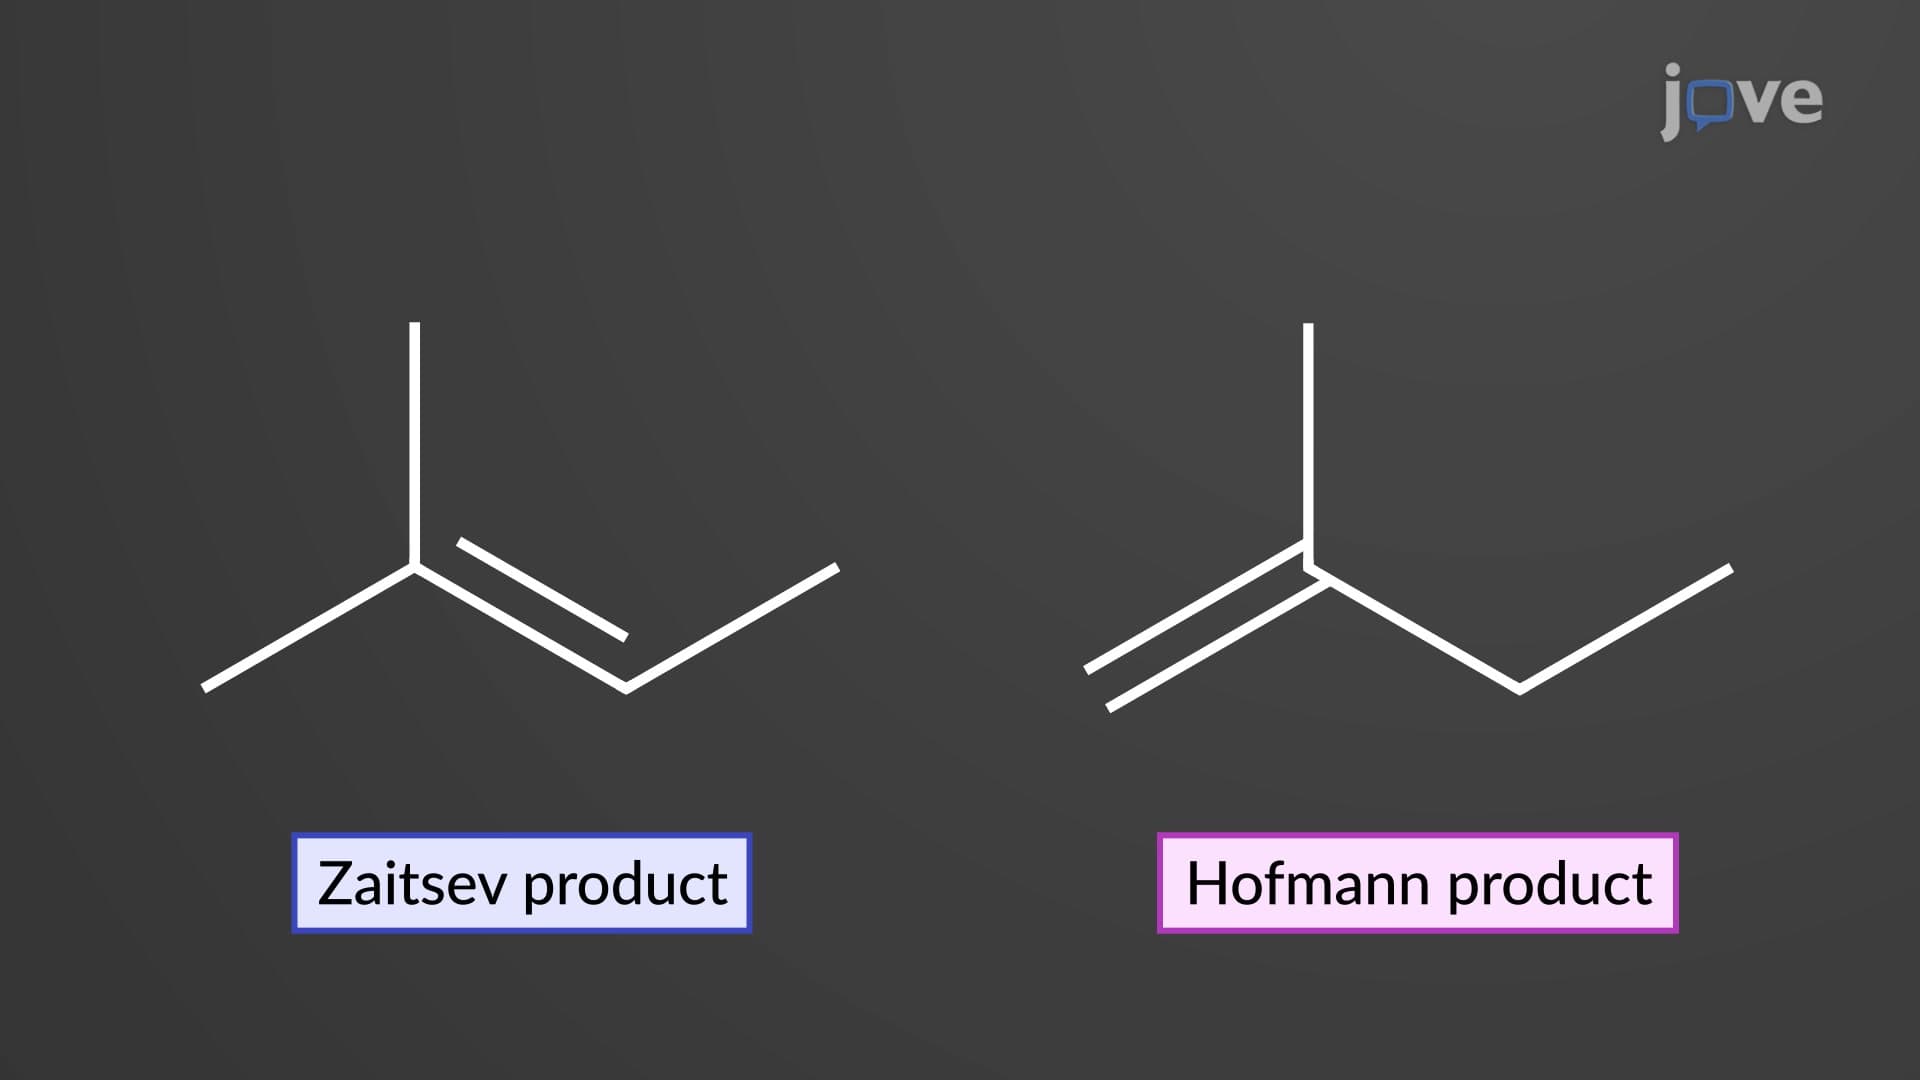
When a substrate with two different β hydrogens undergoes an E2 elimination, the presence of a strong base can yield two regioisomeric alkenes. The more-substituted alkene is the major...