Related Experiment Video
Updated: Sep 10, 2025
Author Spotlight: Advancing Therapeutics to Treat Vibriosis in Humans and Aquatic Organisms
Published on: May 31, 2024
Synthesis, Absorption and Emission Behaviour, Solvation, AIM, Topology, and Docking Studies on 4-(((perfluorophenyl)
V Tamilselvi1,2, Aamal A Al-Mutairi3, M Arivazhagan4
1Department of Physics, Pioneer Kumaraswamy College, Kanyakumari District, Nagercoil, 629 003, Tamil Nadu, India.

Abstract:
This work aims to synthesize 4-(((perfluorophenyl) methylene)amino)-N-(thiazol-2-yl) benzene sulfonamide (PFTH) and characterize it using various instrumental techniques such as UV-visible (UV-vis), fluorescence, Fourier-transform-infrared (FTIR), Raman, and nuclear magnetic resonance (NMR) spectroscopy analyses, respectively. The density functional theory (DFT) calculations have been performed using correlation-consistent polarized valence double-zeta (cc-pVDZ) basis sets and Becke, 3-parameter Lee-Yang-Parr (B3LYP) functionals, respectively. In this study, we used Gauge-Including Atomic Orbitals (GIAO) to determine the computational techniques for nuclear magnetic resonance (13C NMR and 1H NMR) spectroscopy. The C-S bond lengths have been measured as follows: C24-S8 = 1.75 Å, C27-S28 = 1.73 Å, and C17-S20 = 1.77 Å, respectively. A higher absorption wavelength (307 nm) has been observed in the gas phase due to the solvent interaction. The synthesized PFTH displays two emission peaks at the excitation wavelengths of 356 nm and 648 nm, respectively, in DMSO solvent. The calculated energy gap of about ~ 7.6047 eV in the gas phase has been determined using frontier molecular orbital (FMO) analysis. The estimated value is comparatively lower than that in the solvent environments, such as water, chloroform, and DMSO. Electrophilic and nucleophilic regions are revealed by the molecular electrostatic potential (MEP) of the PFTH. The natural bond orbital (NBO) analysis of the PFTH compound at LP(2)S28→π∗(C24-N25) had the highest stabilization energy (E(2)) value of about 47.87 kcal/mol. Topological analyses such as electron localization function (ELF), localized orbital locator (LOL), and average localized ionization energy (ALIE), respectively, were conducted in this study. In addition, non-covalent interaction (NCI) and atoms in molecules (AIM) studies revealed that hydrogen bonds formed between C15-H31 and N25. A molecular docking study was also performed on the molecules, and the lowest binding energy was observed at - 6.67 kcal/mol.
More Related Videos
Related Concept Videos
 01:22
Electrophilic Aromatic Substitution: Sulfonation of Benzene
 01:13
Electrophilic Aromatic Substitution: Fluorination and Iodination of Benzene
 02:26
Preparation and Reactions of Sulfides
 01:34
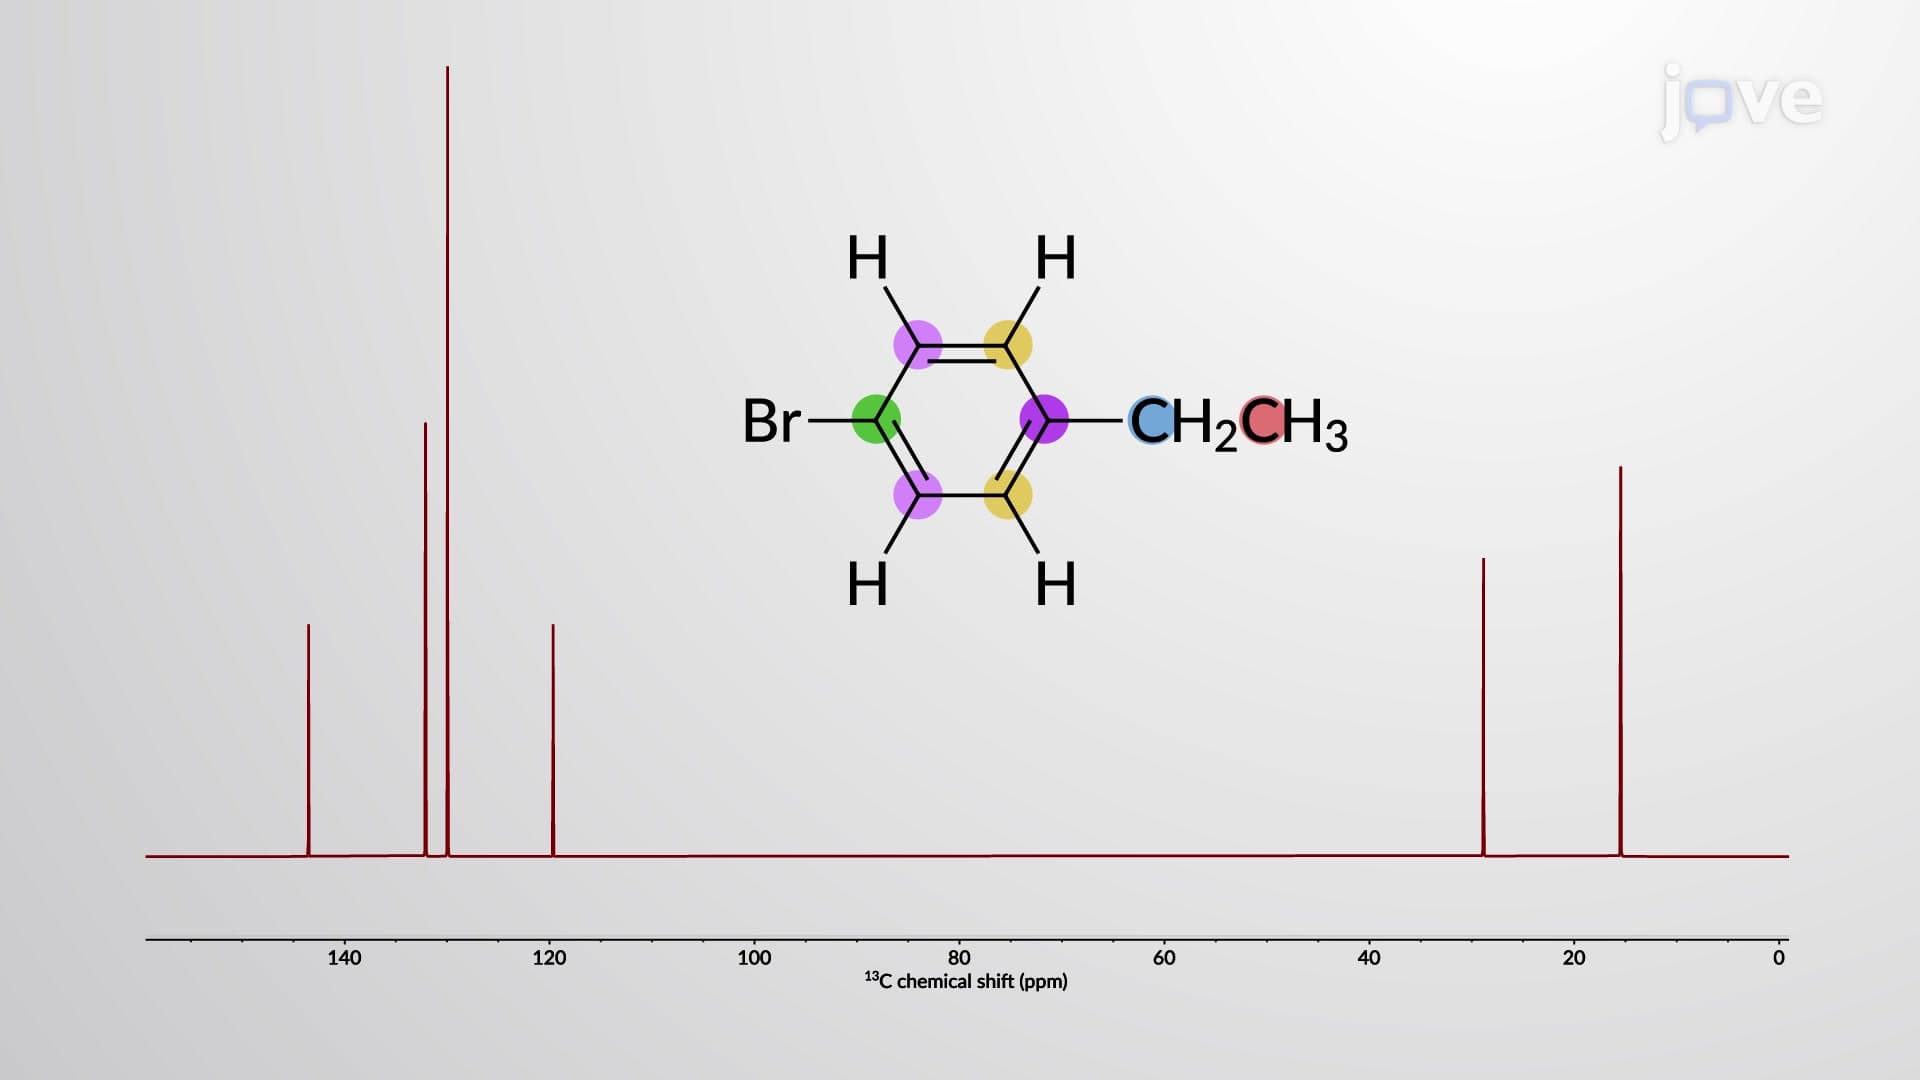
NMR Spectroscopy of Benzene Derivatives
 01:11
Aryldiazonium Salts to Azo Dyes: Diazo Coupling
 01:20
Diazonium Group Substitution with Halogens and Cyanide: Sandmeyer and Schiemann Reactions