Related Experiment Video
Updated: Sep 10, 2025
Methods of Ex Situ and In Situ Investigations of Structural Transformations: The Case of Crystallization of Metallic Glasses
Published on: June 7, 2018
Density functional theory analysis of the structural, electronic, elastic, phonon dispersion and AIMD properties of
Mazhar Haleem Awan1, Sehrish Munsif2, Huma Habib3
1Department of Physics, Abbottabad University of Science and Technology Abbottabad KPK Pakistan mazharhaleem7@gmail.com +923149888807.

Abstract:
Perovskite chalcogenides have attracted significant interest due to their potential applications in optoelectronics, catalysis, and renewable energy systems. This paper examines the structural, electronic, elastic, and phononic properties of KMgX3 (X = O, S, Se) using density functional theory (DFT) in the context of the full-potential linearized augmented plane wave plus local orbital (FP-LAPW + lo) approach. Their stability in the cubic phase (Pm3̄m symmetry) is confirmed by the computed lattice parameters for KMgO3 (4.1325 Å), KMgS3 (5.0008 Å), and KMgSe3 (5.2070 Å). KMgO3 exhibits semiconducting behavior with a direct bandgap of 7.323 eV in the spin-up state, according to electronic band structure studies, whereas KMgS3 and KMgSe3 show metallic properties. Elastic constants (C11, C12, and C44) meet the requirements for mechanical stability, which is evaluated using the Born criterion. Upon further examination of mechanical characteristics such as Bulk modulus, Shear modulus, Young's modulus, and Poisson's ratio. Materials such as KMgO3 and KMgS3 exhibit ductile behavior, whereas KMgSe3 exhibits brittleness. Phonon dispersion curves and ab initio molecular dynamics simulations confirm the dynamical and thermal stability of these compounds. The results show that KMgX3 perovskites have potential uses in optoelectronic devices and spintronics.
More Related Videos
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:40
MO Theory and Covalent Bonding
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 02:35
Molecular Orbital Theory I
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:18
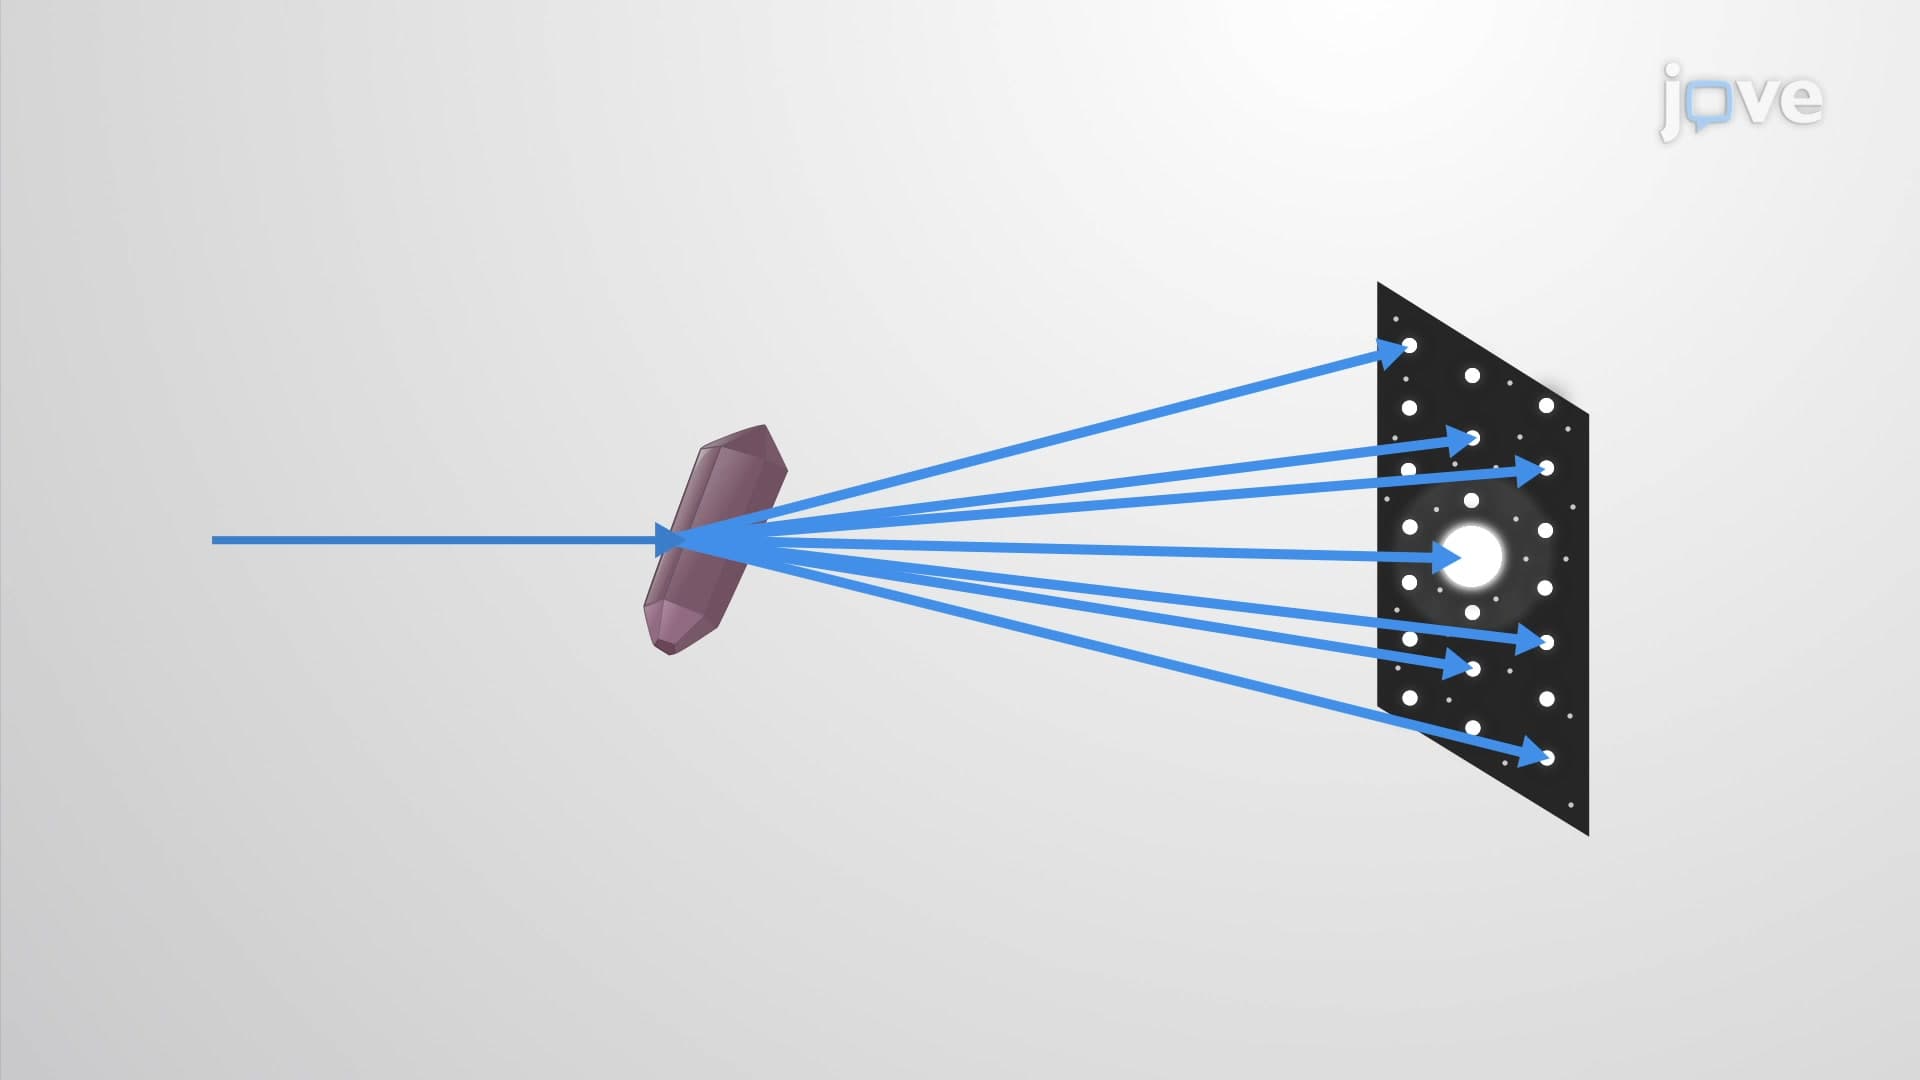
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...