Related Experiment Video
Updated: Sep 9, 2025


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
An Interpretable Deep Learning and Molecular Docking Framework for Repurposing Existing Drugs as Inhibitors of
Juan Huang1, Jialong Gao1, Qu Chen1
1School of Biological and Chemical Engineering, Zhejiang University of Science and Technology, Hangzhou 310023, China.

Abstract:
Despite the widespread use of vaccines against SARS-CoV-2, COVID-19 continues to pose global health challenges, requiring efficient drug screening and repurposing strategies. This study presents a novel hybrid framework that integrates deep learning (DL) with molecular docking to accelerate the identification of potential therapeutics. The framework comprises three crucial steps: (1) a previously developed DL model is employed to rapidly screen candidate compounds, selecting those with predicted interaction scores above a cut-off value of 0.8; (2) AutoDock Vina version 1.5.6 and LeDock version 1.0 are used to evaluate binding affinities, with a threshold of <-7.0 kcal·mol-1; and (3) predicted drug-protein binding sites are evaluated to determine their overlap with known active residues of the target protein. We first validated the framework using four experimentally confirmed COVID-19 drug-target pairs and then applied it to identify potential inhibitors of the SARS-CoV-2 main protease (MPro). Among 29 drug candidates selected based on antiviral, anti-inflammatory, or anti-cancer properties, only Enasidenib met all three selection criteria, showing promise as an MPro inhibitor. However, further experimental and clinical studies are required to confirm its efficacy against SARS-CoV-2. This work provides an interpretable strategy for virtual screening and drug repurposing, which can be readily adapted to other DL models and docking tools.
More Related Videos
 10:21
Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 02:04
Protein-protein Interfaces
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:26
Drug Discovery: Overview
 01:16
Protein-Drug Binding: Mechanism and Kinetics
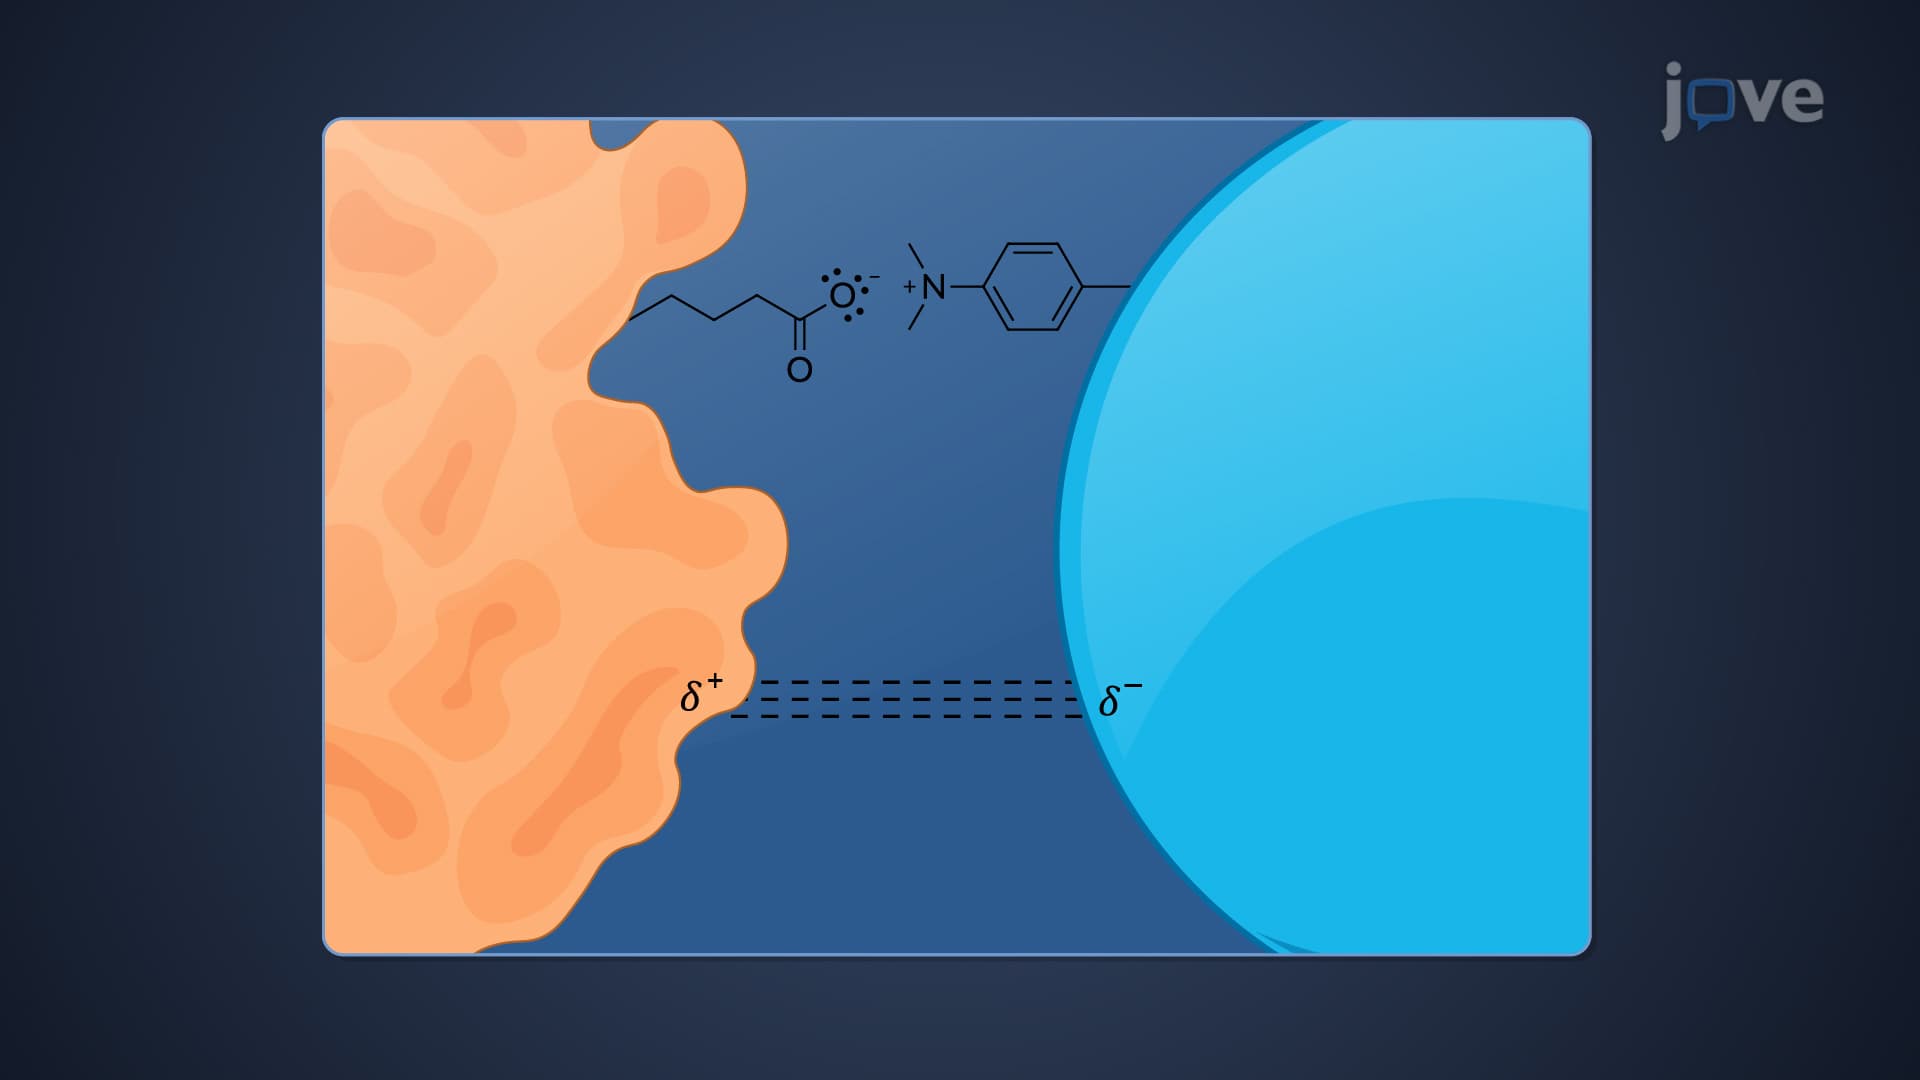
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:26
Targets for Drug Action: Overview
Receptors are either membrane-spanning or intracellular proteins, which upon binding a ligand, get activated and transmit the signal downstream to elicit a response. Drugs bind receptors, either mimicking the action of endogenous ligands or blocking the receptor activity to bring about a modified response. Nearly 35% of approved drugs target the G...