Related Experiment Video
Updated: May 6, 2026
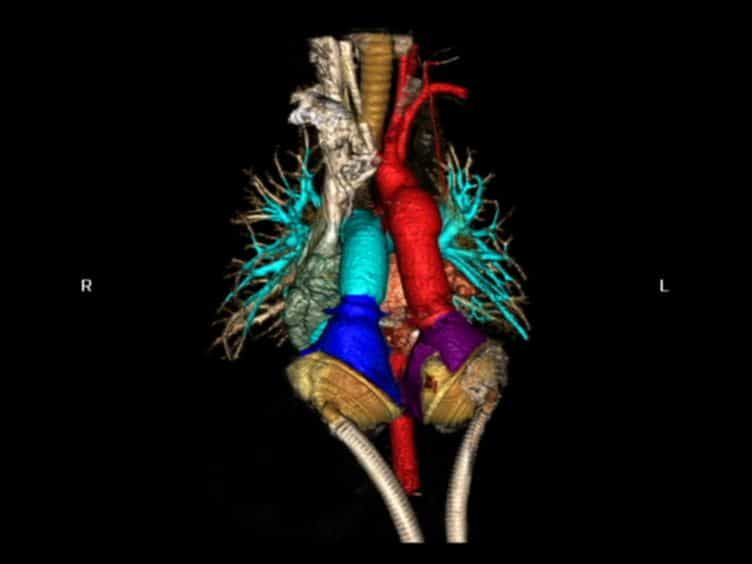

Implantation of Total Artificial Heart in Congenital Heart Disease
Published on: July 18, 2014
A novel TAFAZZIN gene variant c.525_533del causing Barth syndrome and leading to heart transplantation: a case report
Michał Krawiec1, Joanna Śliwka2, Szymon Pawlak2
1Faculty of Medical Sciences in Zabrze, Medical University of Silesia in Katowice, Zabrze, Poland.

Introduction:
Barth syndrome (BTHS) is an ultra-rare genetic disease caused by a mutation in the TAFAZZIN gene, located on the X chromosome. This gene codes for the protein tafazzin, which is involved in the metabolism of the mitochondrial phospholipid - cardiolipin. Symptoms of this genetic defect include dilated cardiomyopathy (DCM), skeletal myopathy, neutropenia, growth retardation, reduced cholesterol levels, increased serum lactic acid levels, and hypoglycemia in the neonatal period.
Case Description:
A Caucasian boy with DCM and left ventricular non-compaction associated with BTHS, caused by a previously unreported variant in the TAFAZZIN gene: NM_000116.4:c.525_533del; NP_000107.1:p.(His176_Phe178del) at NC_000023.11:g.154419607_154419615del, in the exon 6. Due to the patient's heart failure, a mechanical circulatory support (MCS) system was required, followed by orthotopic heart transplantation (OHT). Because of the presence of neutropenia, standard immunosuppressive therapy had to be modified in the postoperative period.
Conclusions:
A previously unreported mutation is presented, leading to BTHS. This disease can have severe cardiovascular manifestations, requiring MCS and OHT.

Related Concept Videos
 01:24
Tissue Transplantation
The Biology of Tissue Transplantation
The biology of tissue transplantation hinges on the Major Histocompatibility Complex (MHC) molecules. These molecules...
 01:30
Cardiomyopathy II: Dilated Cardiomyopathy
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy
 01:29
Cardiomyopathy IV: Restrictive Cardiomyopathy