Related Experiment Video
Updated: Sep 9, 2025


Author Spotlight: Streamlining Protein Target Prediction and Validation via Molecular Docking and CETSA
Published on: February 23, 2024
Decoding the limits of deep learning in molecular docking for drug discovery
1Xiangya School of Pharmaceutical Sciences, Central South University Changsha 410013 Hunan P.R. China jiang_dj@zju.edu.cn oriental-cds@163.com.

Abstract:
Structure-based molecular docking, a cornerstone of computational drug design, is undergoing a paradigm shift fueled by deep learning (DL) innovations. However, the rapid proliferation of DL-driven docking methods has created uncharted challenges in translating in silico predictions to biomedical reality. Here, we delve into the performance and prospects of traditional methods and state-of-the-art DL docking paradigms-encompassing generative diffusion models, regression-based architectures, and hybrid frameworks-across five critical dimensions: pose prediction accuracy, physical plausibility, interaction recovery, virtual screening (VS) efficacy, and generalization across diverse protein-ligand landscapes. We reveal that generative diffusion models achieve superior pose accuracy, while hybrid methods offer the best balance. Regression models, however, often fail to product physically valid poses, and most DL methods exhibit high steric tolerance. Furthermore, our analysis reveals significant challenges in generalization, particularly when encountering novel protein binding pockets, limiting the current applicability of DL methods. Finally, we explore failure mechanisms from a model perspective and propose optimization strategies, offering actionable insights to guide docking tool selection and advance robust, generalizable DL frameworks for molecular docking.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:22
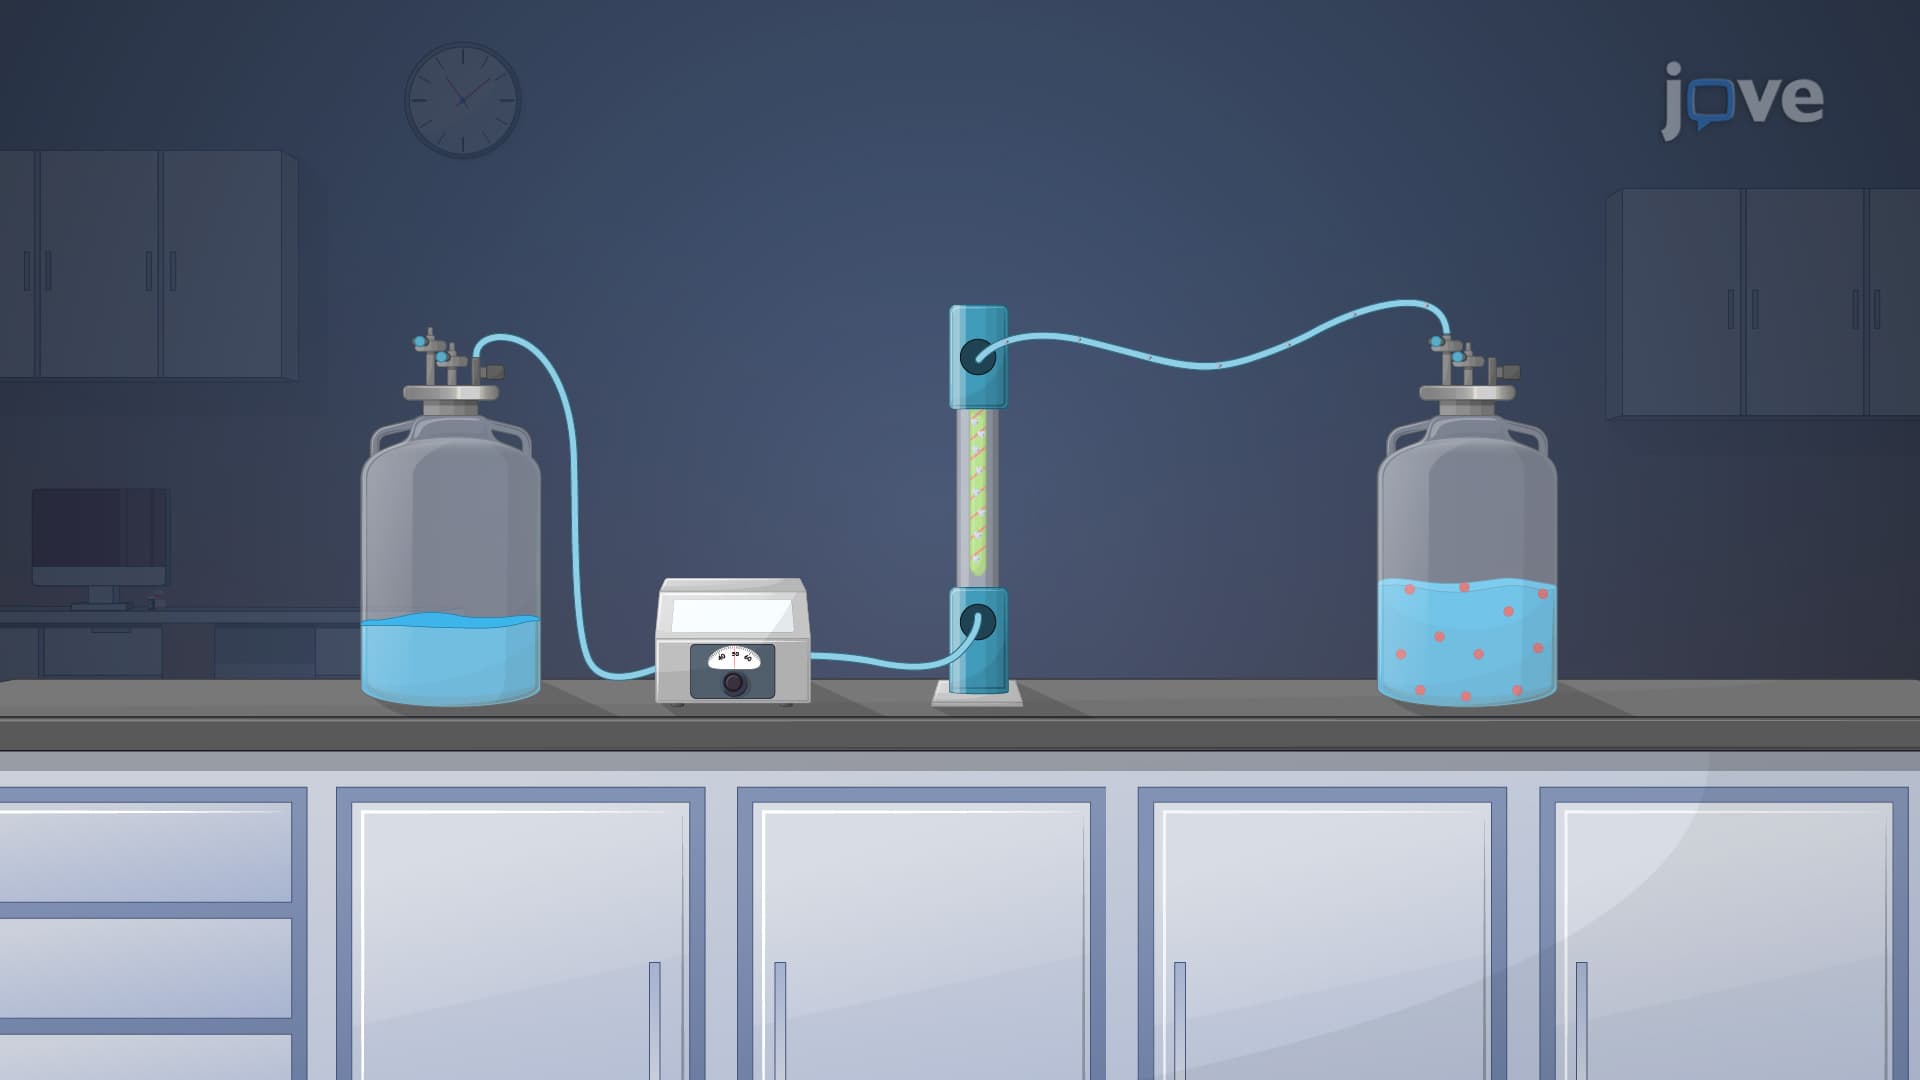
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:26
Drug Discovery: Overview
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:04
Protein-protein Interfaces