Related Experiment Video
Updated: Jan 18, 2026


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Compositional design rules for tuning functionalities in CuInP2X6 (X = S, Se) van der Waals semiconductor
Mona Layegh1, Joseph W Bennett1
1Department of Chemistry & Biochemistry, University of Maryland Baltimore County, Baltimore, MD 21250, USA. bennettj@umbc.edu.

Abstract:
Two-dimensional van der Waals (2D-vdW) semiconducting ferroelectrics, such as CuInP2Se6 (CIPSe) and CuInP2S6 (CIPS), offer unique opportunities for lightweight, scalable, low-power nanoscale electronic devices. However, the limited pool of functional 2D-vdW ferroics highlights the need for clear design principles that can be used to guide experiments. Here, we use first-principles density functional theory (DFT) to study how isovalent atomistic substitution at In and P sites modifies structure, polarization, and electronic properties in CIPSe and CIPS. When substituting In with Sb and Bi, and P with As and Sb, we reveal how ionic radius mismatch, electronegativity differences, and stereochemical lone pair activity shape the ferroelectric and semiconducting response in both rigid (S-based) and soft (Se-based) 2D lattices. In CIPSe, Bi doping at In sites widens the band gap to ∼1.07 eV without reducing polarization or switching performance, provided the Bi atoms are placed in a balanced zigzag arrangement that limits local strain. In CIPS, polarization values drop more noticeably than in CIPSe, and dopant location has a stronger effect on switching behavior. P-site substitution leads to stronger distortions in CIPSe, where larger dopants destabilize the P-P dimer network and in many instances, we predict it to be a metal. However, with As doping in CIPS, polarization remains stable, and the band gap decreases without major structural disruption. Our results establish structure-property design rules based on dopant size and location, host lattice stiffness, and chemical compatibility, offering a route to engineer new 2D ferroelectrics in which polarization and band gap can be tuned together through specific compositional changes.
Related Concept Videos
 02:42
Valence Bond Theory
 01:22
Semiconductors
Metals such as copper (Cu), zinc (Zn), or lead (Pb) have low resistivity and feature conduction bands that are either not fully occupied or overlap with the valence band, making a bandgap non-existent. This allows electrons in the highest energy levels of the valence band to easily transition to the conduction band upon gaining...
 01:27
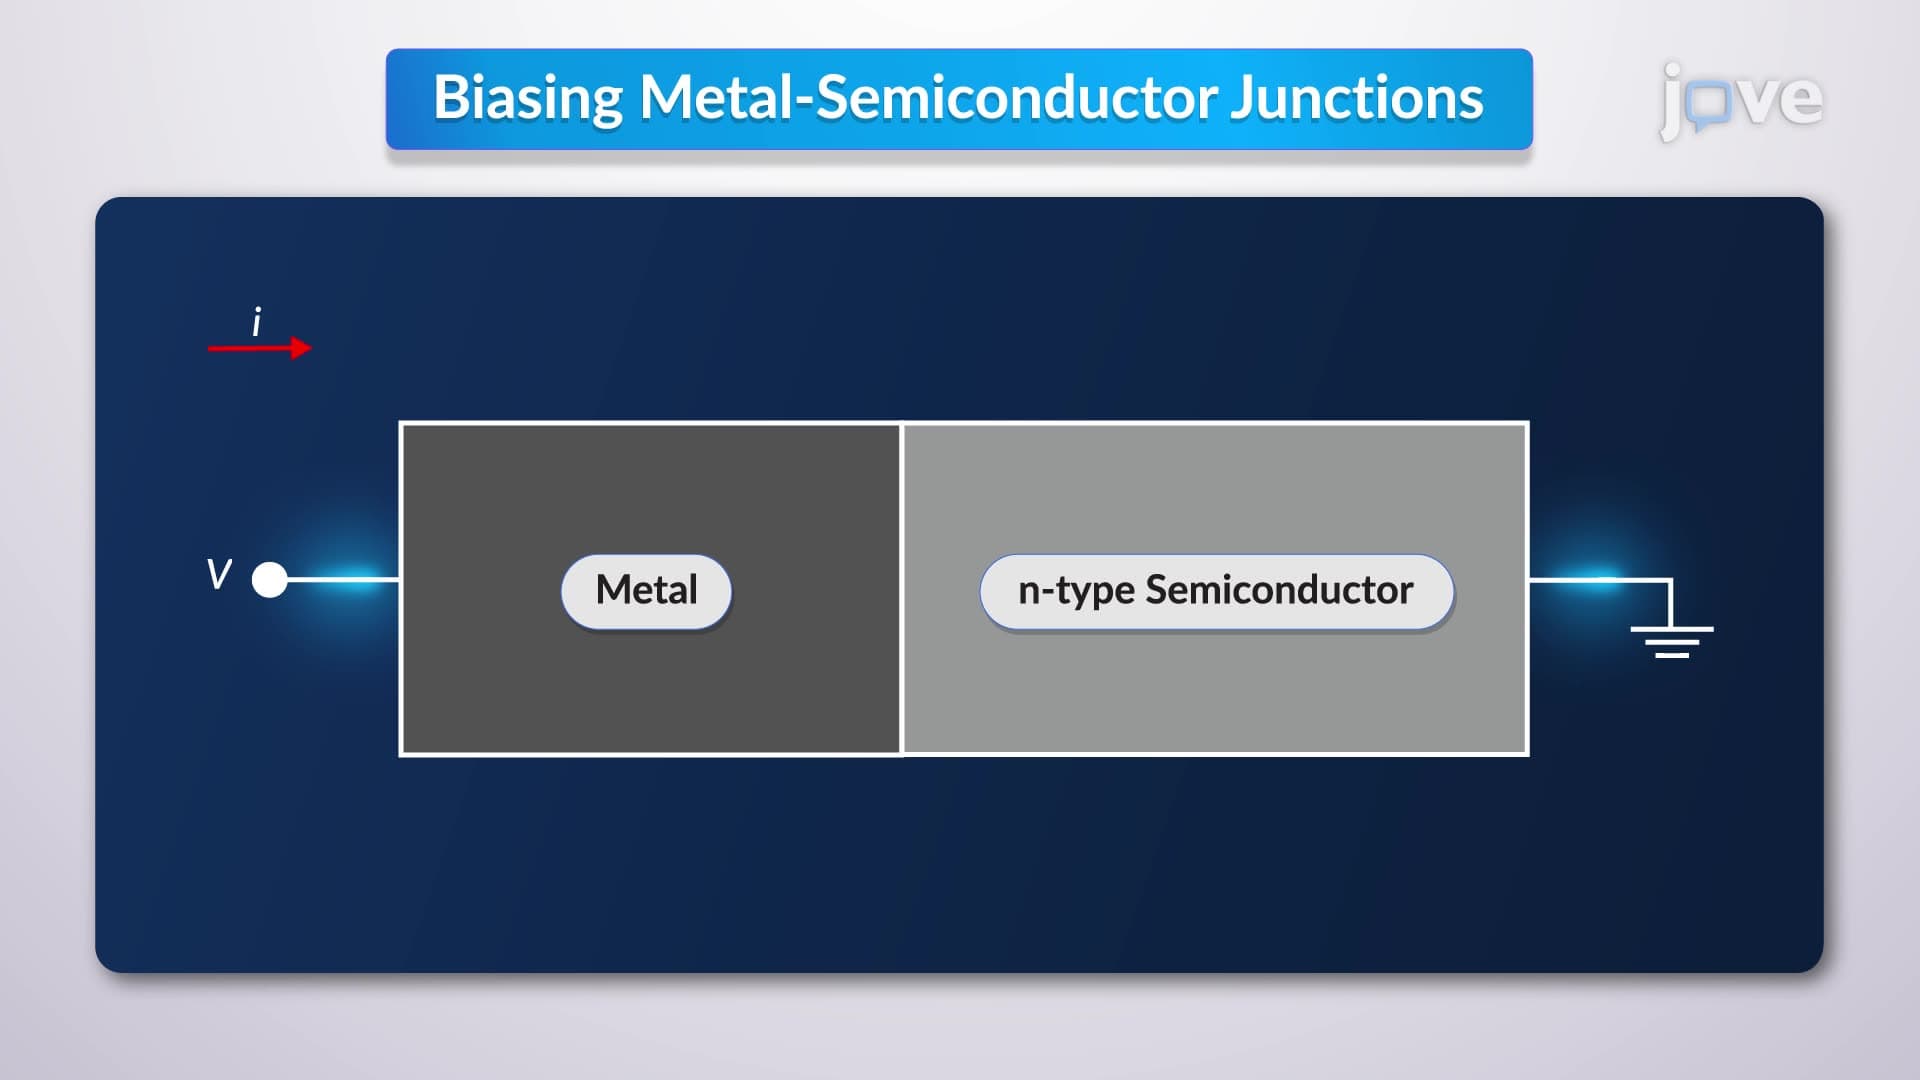
Biasing of Metal-Semiconductor Junctions
In Schottky junctions, where the semiconductor is n-type, applying a positive voltage to the metal relative to the semiconductor reduces its Fermi...
 01:12
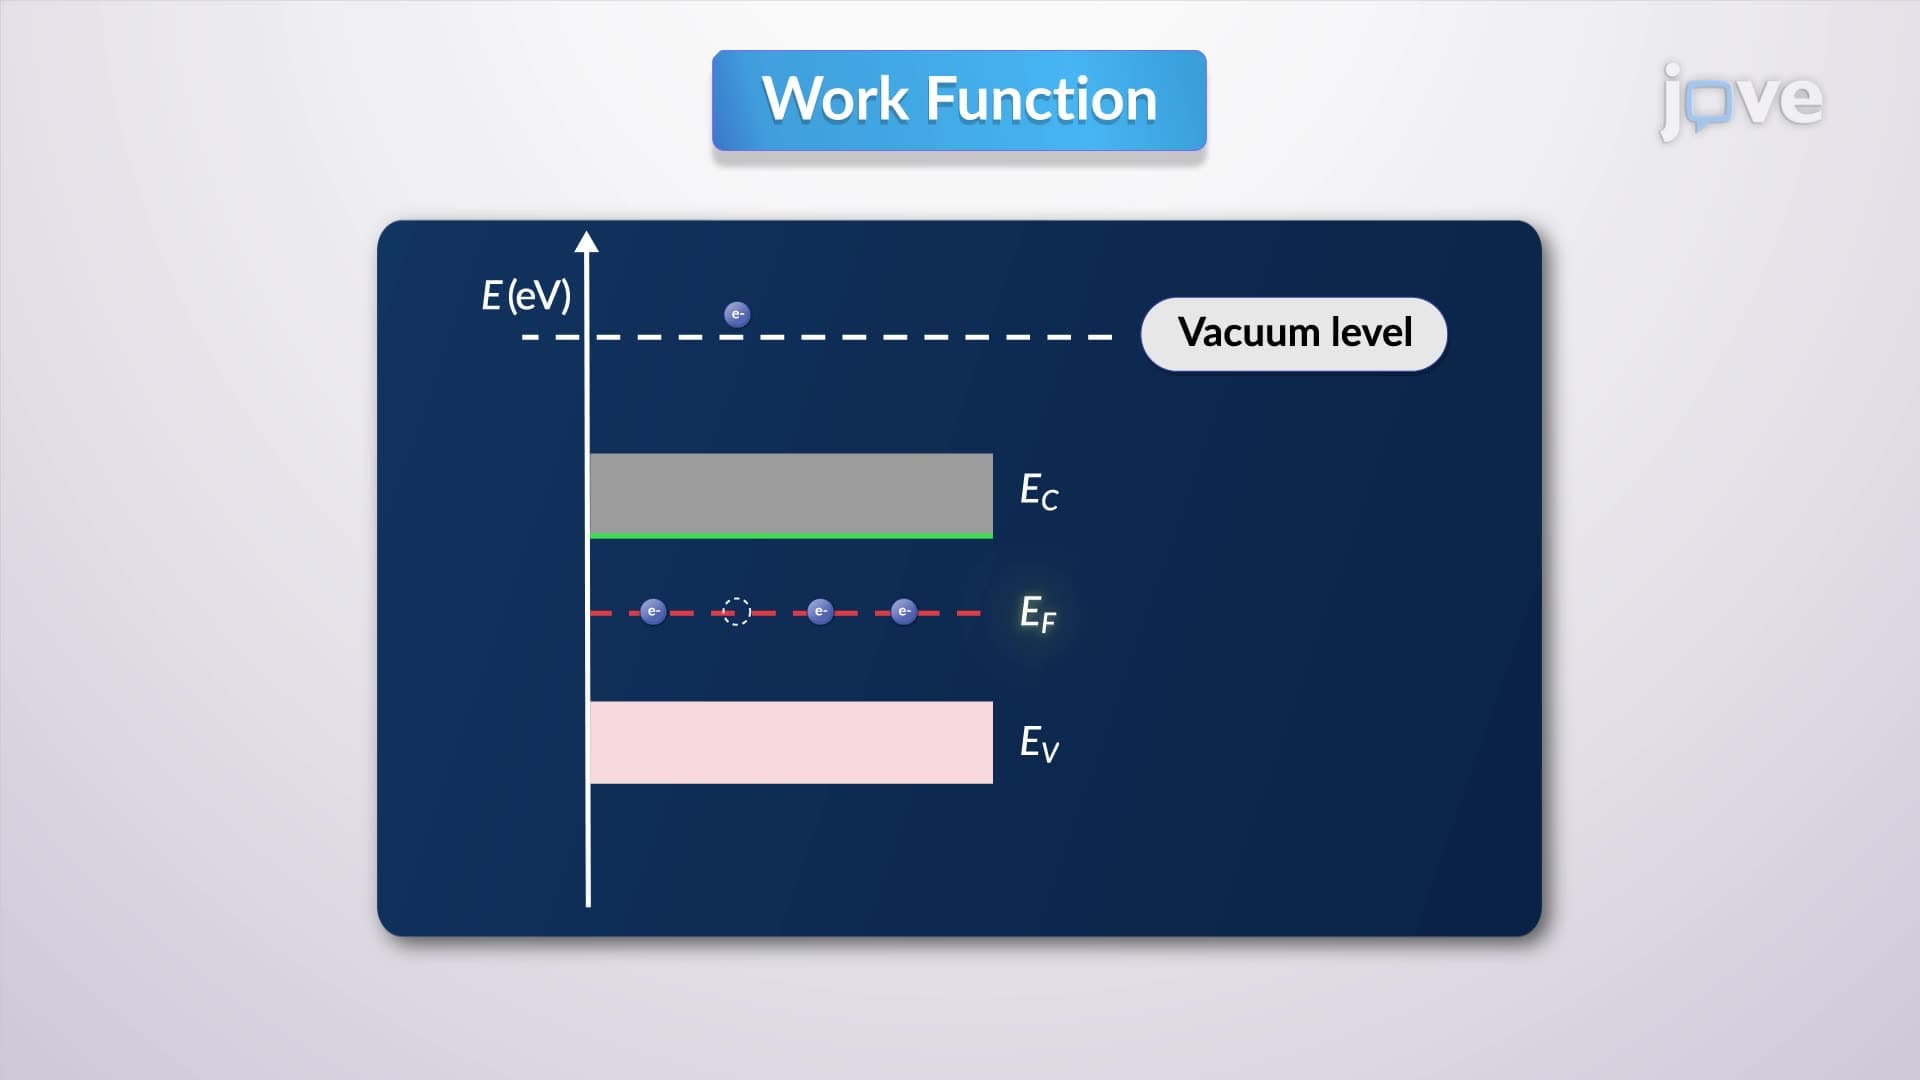
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 01:20
Types of Semiconductors
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...