Related Experiment Video
Updated: Jan 18, 2026

Author Spotlight: Streamlining Visual Dynamics to Simplify Molecular Dynamics Simulations Using Gromacs
Published on: August 9, 2024
Efficient, Hierarchical, and Object-Oriented Electronic Structure Interfaces for Direct Nonadiabatic Dynamics
Sascha Mausenberger1,2, Severin Polonius1,2, Sebastian Mai1
1Institute of Theoretical Chemistry, Faculty of Chemistry, University of Vienna, Währinger Straße 17, 1090 Vienna, Austria.

Abstract:
We present a novel, flexible framework for electronic structure interfaces designed for nonadiabatic dynamics simulations, implemented in Python 3 using concepts of object-oriented programming. This framework streamlines the development of new interfaces by providing a reusable and extendable code base. It supports the computation of energies, gradients, various couplings─like spin-orbit couplings, nonadiabatic couplings, and transition dipole moments─and other properties for an arbitrary number of states with any multiplicities and charges. A key innovation within this framework is the introduction of hybrid interfaces, which can use other interfaces in a general hierarchical manner. Hybrid interfaces are capable of using one or more child interfaces to implement multiscale approaches, such as quantum mechanics/molecular mechanics where different child interfaces are assigned to different regions of a system. The concept of hybrid interfaces can be extended through nesting, where hybrid parent interfaces use hybrid child interfaces to easily setup complex workflows without the need for additional coding. We demonstrate the versatility of hybrid interfaces with two examples: one at the method level and one at the workflow level. The first example showcases the numerical differentiation of wave function overlaps, implemented as a hybrid interface and used to optimize a minimum-energy conical intersection with numerical nonadiabatic couplings. The second example presents an adaptive learning workflow, where nested hybrid interfaces are used to iteratively refine a machine learning model. This work lays the groundwork for more modular, flexible, and scalable software design in excited-state dynamics.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Related Concept Videos
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 01:06
Interfacial Electrochemical Methods: Overview
 02:00
Molecular Models
 01:18
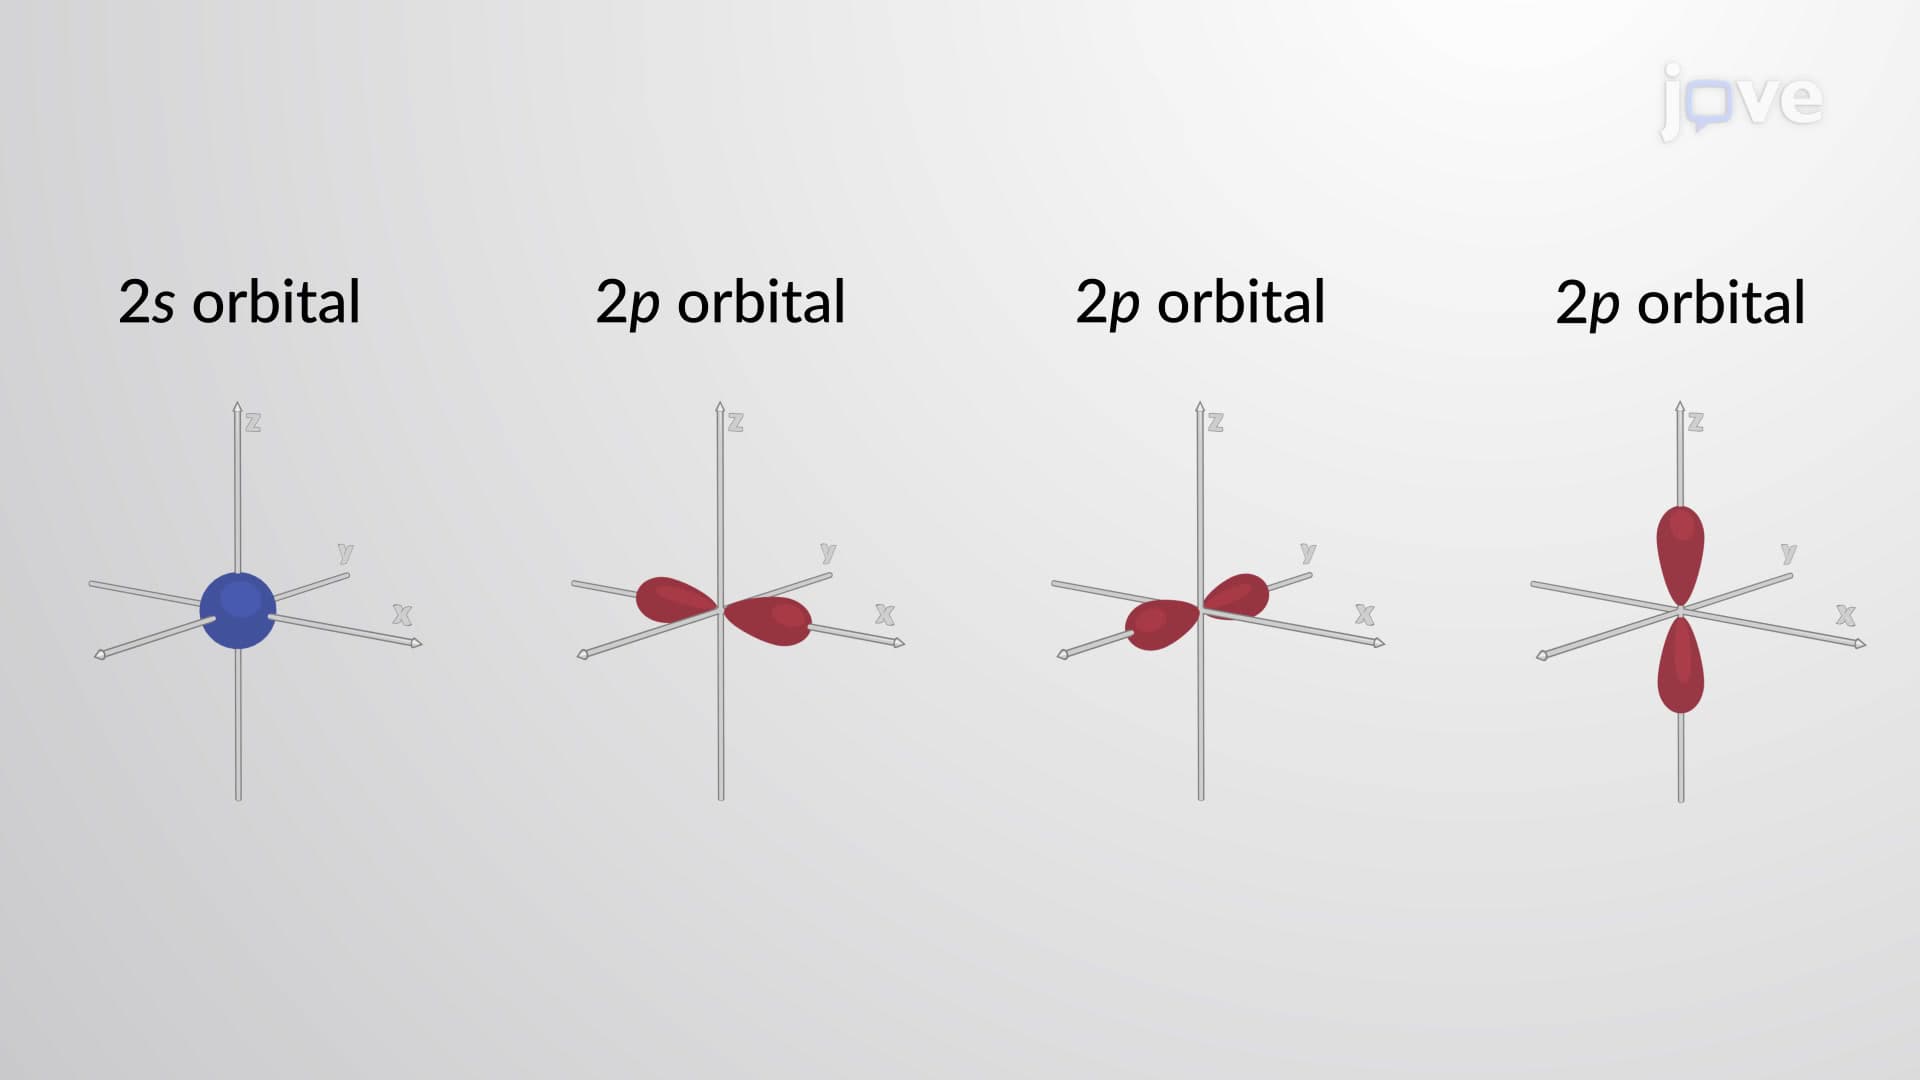
Electron Orbital Model
The first shell is closest to the nucleus, and it has only one subshell with a single spherical orbital called the...