Related Experiment Video
Updated: Jan 18, 2026


Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Leveraging Language Model, Crystal Structure Prediction and First-Principles Calculation for Material Design
Lei Zhang1,2, Ben Ni1, Kaiyang Xu2
1Department of Internet Engineering, School of Software, Nanjing University of Information Science & Technology, Nanjing 210044, China.

Abstract:
Large language models (LLMs) have demonstrated transformative potential for materials discovery in condensed matter systems, but their full utility requires both broader application scenarios and integration with ab initio crystal structure prediction (CSP), density functional theory (DFT) methods and domain knowledge to benefit future inverse material design. Here, we develop an integrated computational framework combining language model-guided materials screening with genetic algorithm (GA) and graph neural network (GNN)-based CSP methods to predict new photovoltaic material. This LLM + CSP + DFT approach successfully identifies a previously overlooked oxide material with unexpected photovoltaic potential. Through transformer-based vector similarity analysis coupled with unsupervised clustering and first-principles calculations, we demonstrate that this material exhibits a direct band gap and high theoretical efficiencies that are suitable for photovoltaic application. Our work highlights a hierarchical computational inverse design pipeline that can efficiently navigate the material space to identify nonintuitive functional materials with tailored optoelectronic properties.
Related Concept Videos
 02:00
Molecular Models
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:27
Predicting Molecular Geometry
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:18
X-ray Crystallography
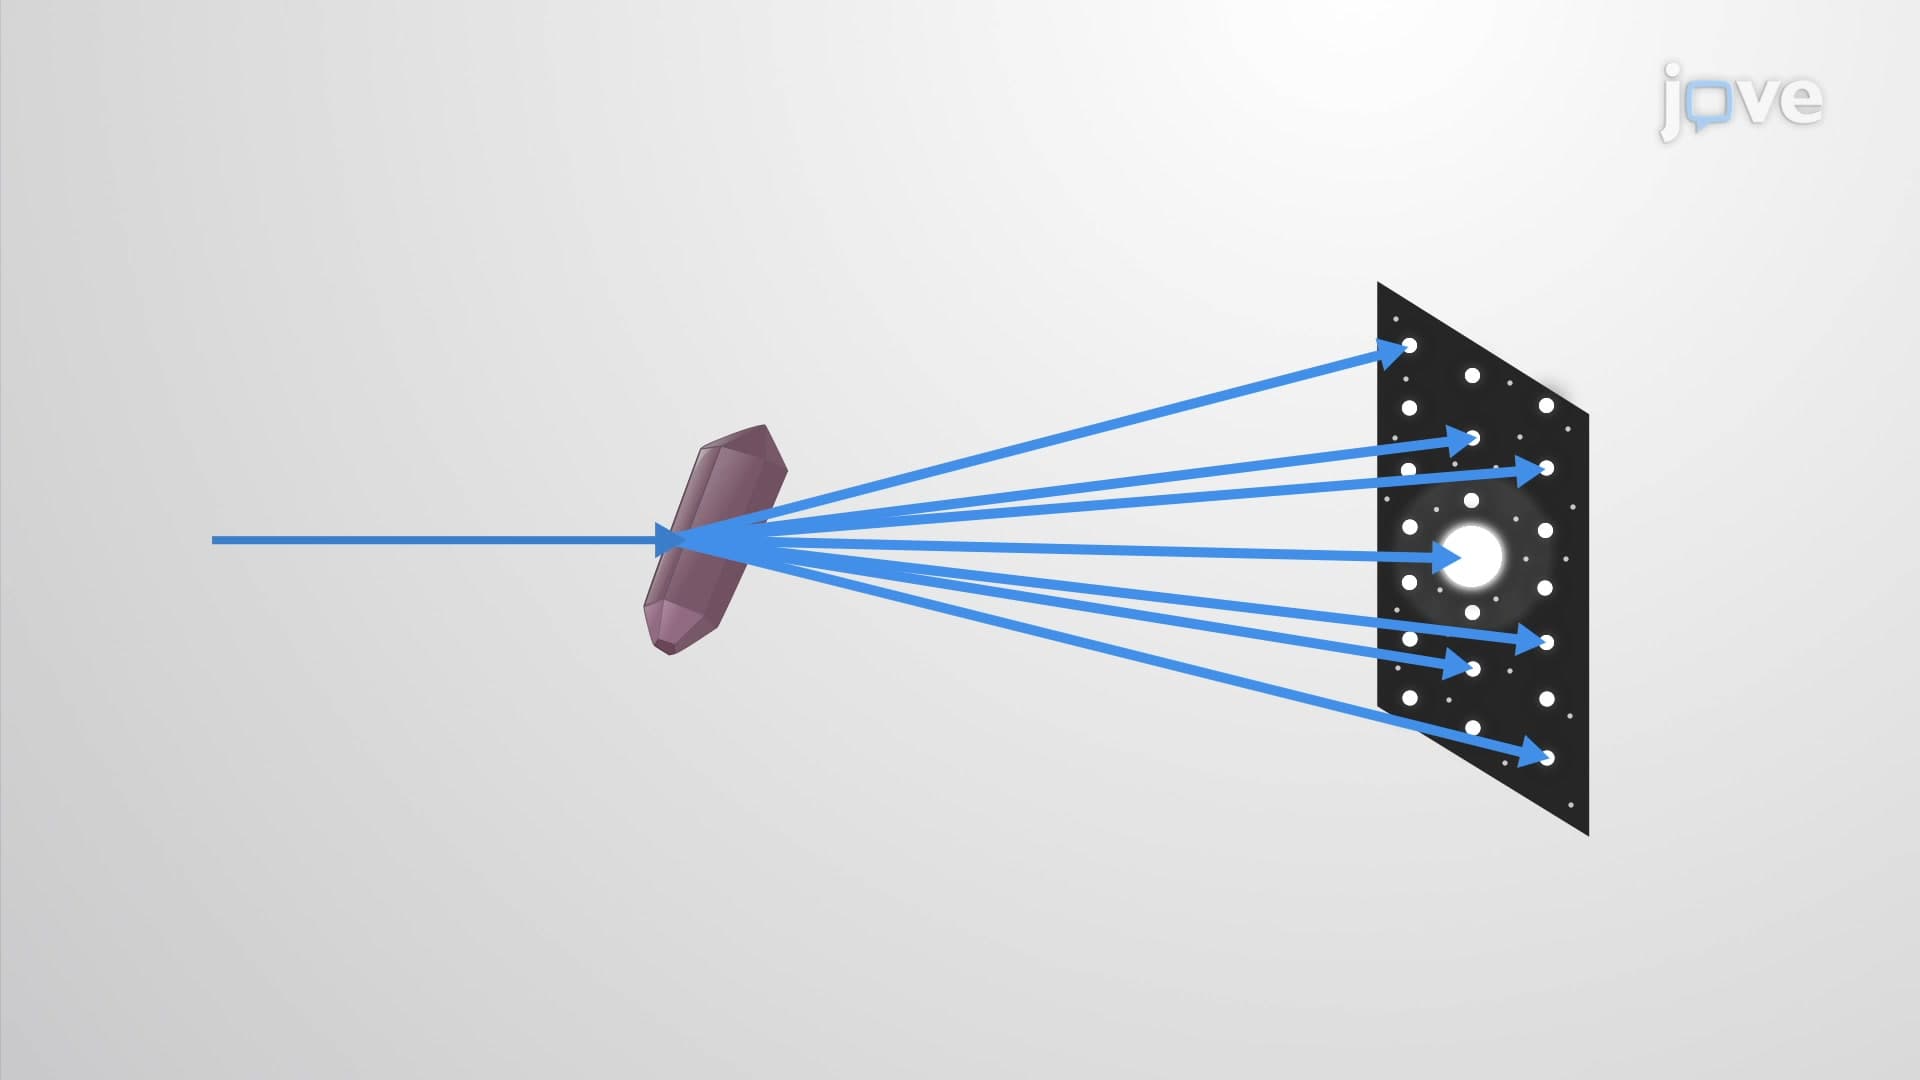
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...