Related Experiment Video
Updated: Jan 18, 2026

Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Enabling accurate modelling of materials for a solid electrolyte interphase in lithium-ion batteries using effective
Wen-Qing Li1, Gang Wu1, Juan Manuel Arce-Ramos1
1Institute of High Performance Computing (IHPC), Agency for Science, Technology and Research (A*STAR), 1 Fusionopolis Way, #16-16 Connexis, Singapore 138632, Republic of Singapore. ngmf@a-star.edu.sg.

Abstract:
Accurate modelling of the structural and dynamic properties of the solid electrolyte interphase (SEI) in lithium-ion batteries remains a longstanding challenge due to the high complexity of the SEI structure and the lack of structural information. Atomistic simulations using molecular dynamics (MD) can provide insights into the structure of the SEI but require large models and accurate interatomic potentials; however, existing computational tools struggle to evaluate these potentials in mixed-material systems efficiently and reliably. Here, we demonstrate the effectiveness of machine learning interatomic potentials (MLIPs) defined using amorphous structures as reference data, specifically the moment tensor potential (MTP), combined with density functional theory (DFT) calculations and active learning loops that enable rapid sampling of MD trajectories. For SEI relevant materials (e.g., Li2CO3, bulk Li, LiPF6, and Li2EDC), our trained MTP models accurately capture the key structural properties (e.g., lattice parameters, elastic constants, or phonon spectra). For the dynamical properties of monoclinic Li2CO3 and amorphous Li2EDC, the models are validated against previous theoretical predictions in the literature. Particularly, we illustrate the finite temperature effects on computing energy barriers. The determined mechanism of dominant diffusion carriers (Li vacancy, interstitial Li, and Li Frenkel pair) in Li2CO3 is highly consistent with DFT calculations. Furthermore, we show that the generated training datasets can be applied to train graph-neural-network (GNN)-based interatomic potentials that can further improve accuracy. The developed machine learning workflow provides a scalable approach for SEI modelling, enabling simulations at larger time and length scales to tackle the limitations of conventional DFT methods.
Related Concept Videos
 02:54
Trends in Lattice Energy: Ion Size and Charge
 01:27
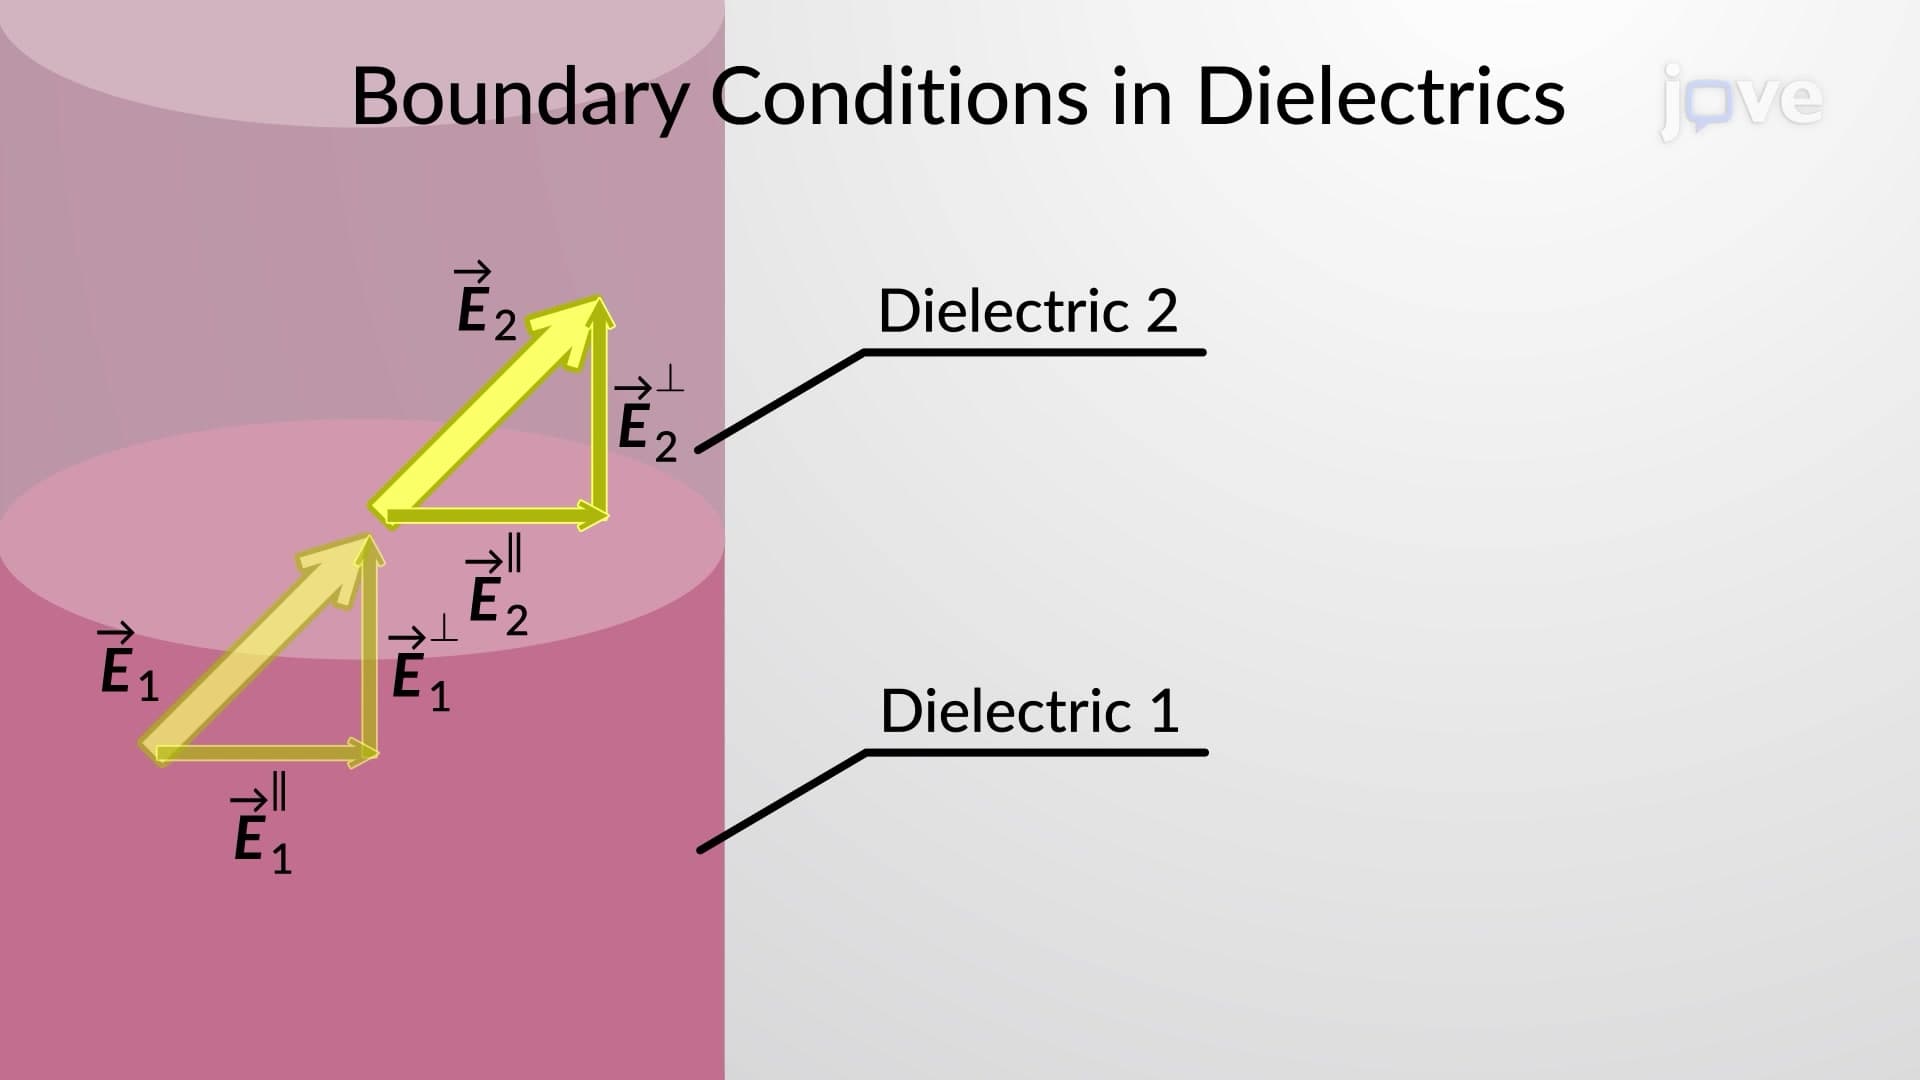
Electrostatic Boundary Conditions in Dielectrics
Consider a case where both the mediums across a boundary are two different dielectric materials. Recall that the electric field and electric displacement are proportional and related through the material's permittivity....
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...
 03:02
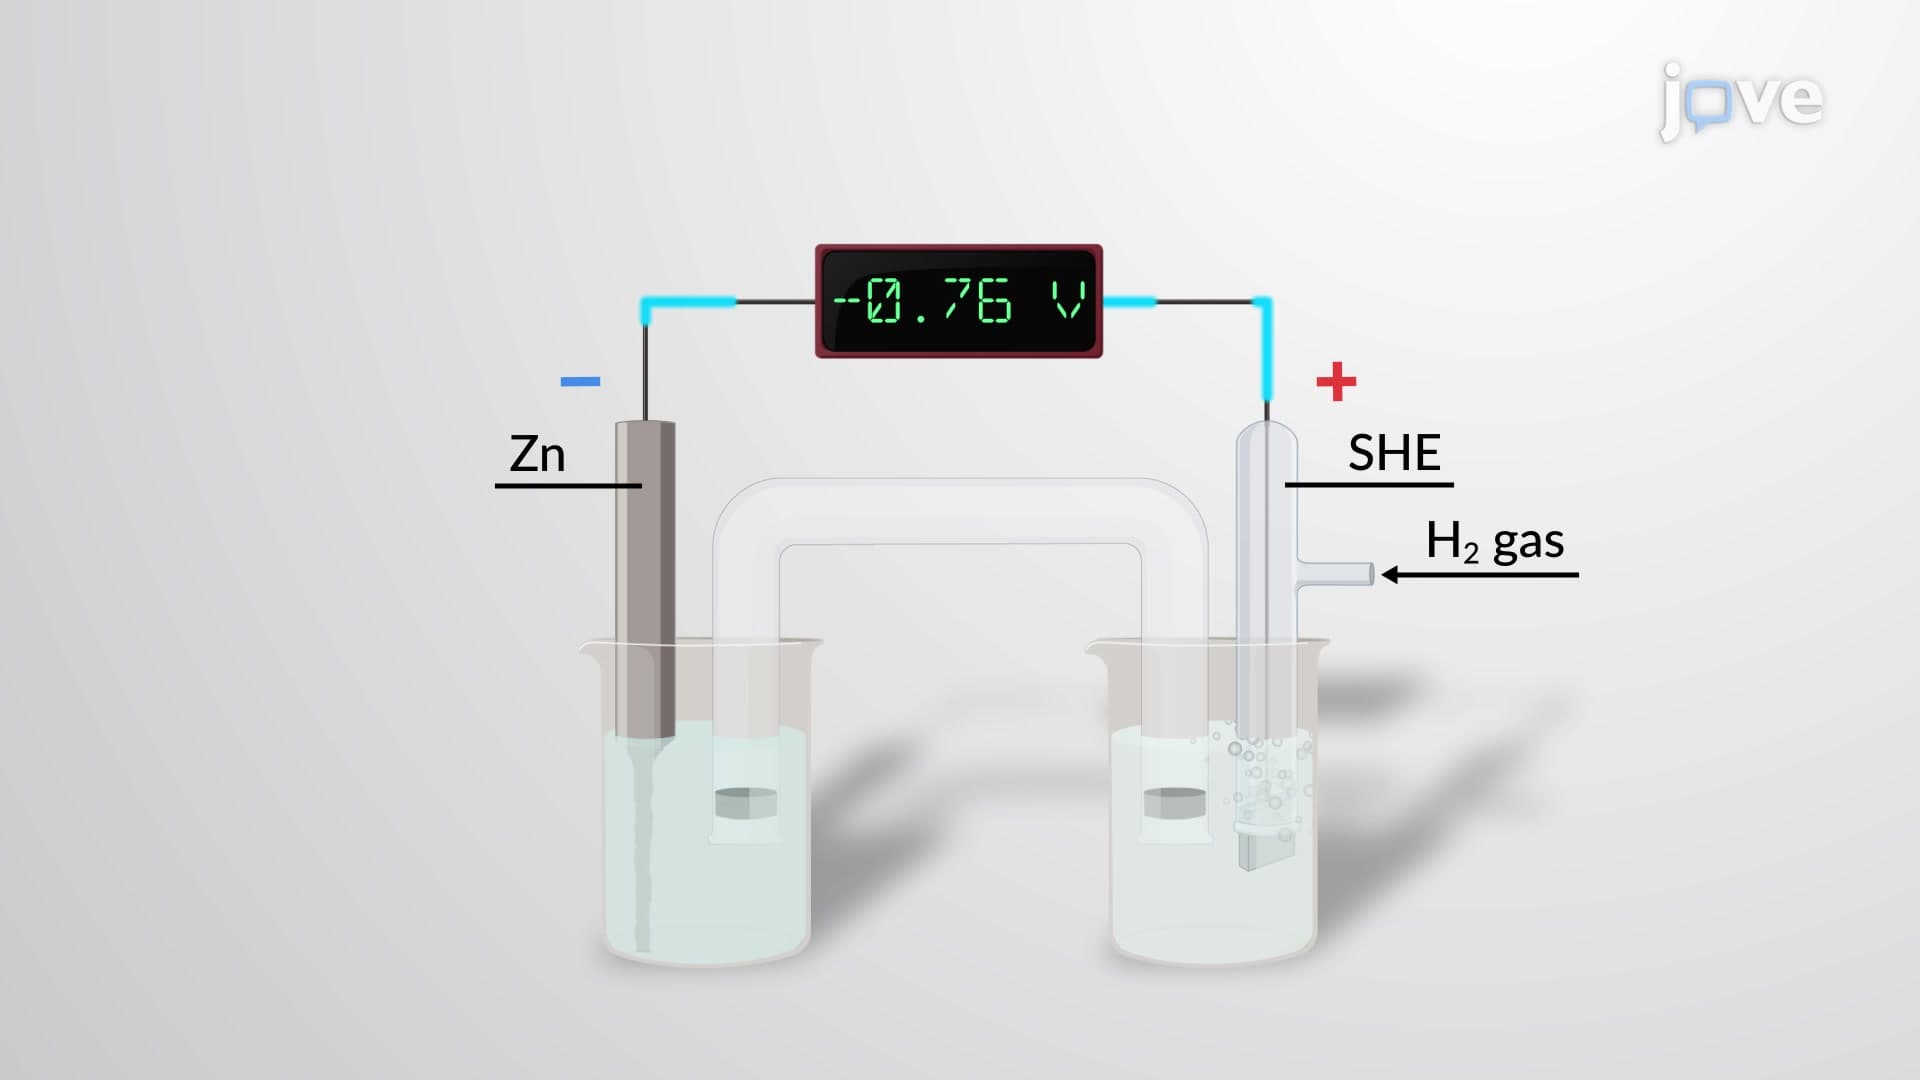
Standard Electrode Potentials
 01:15
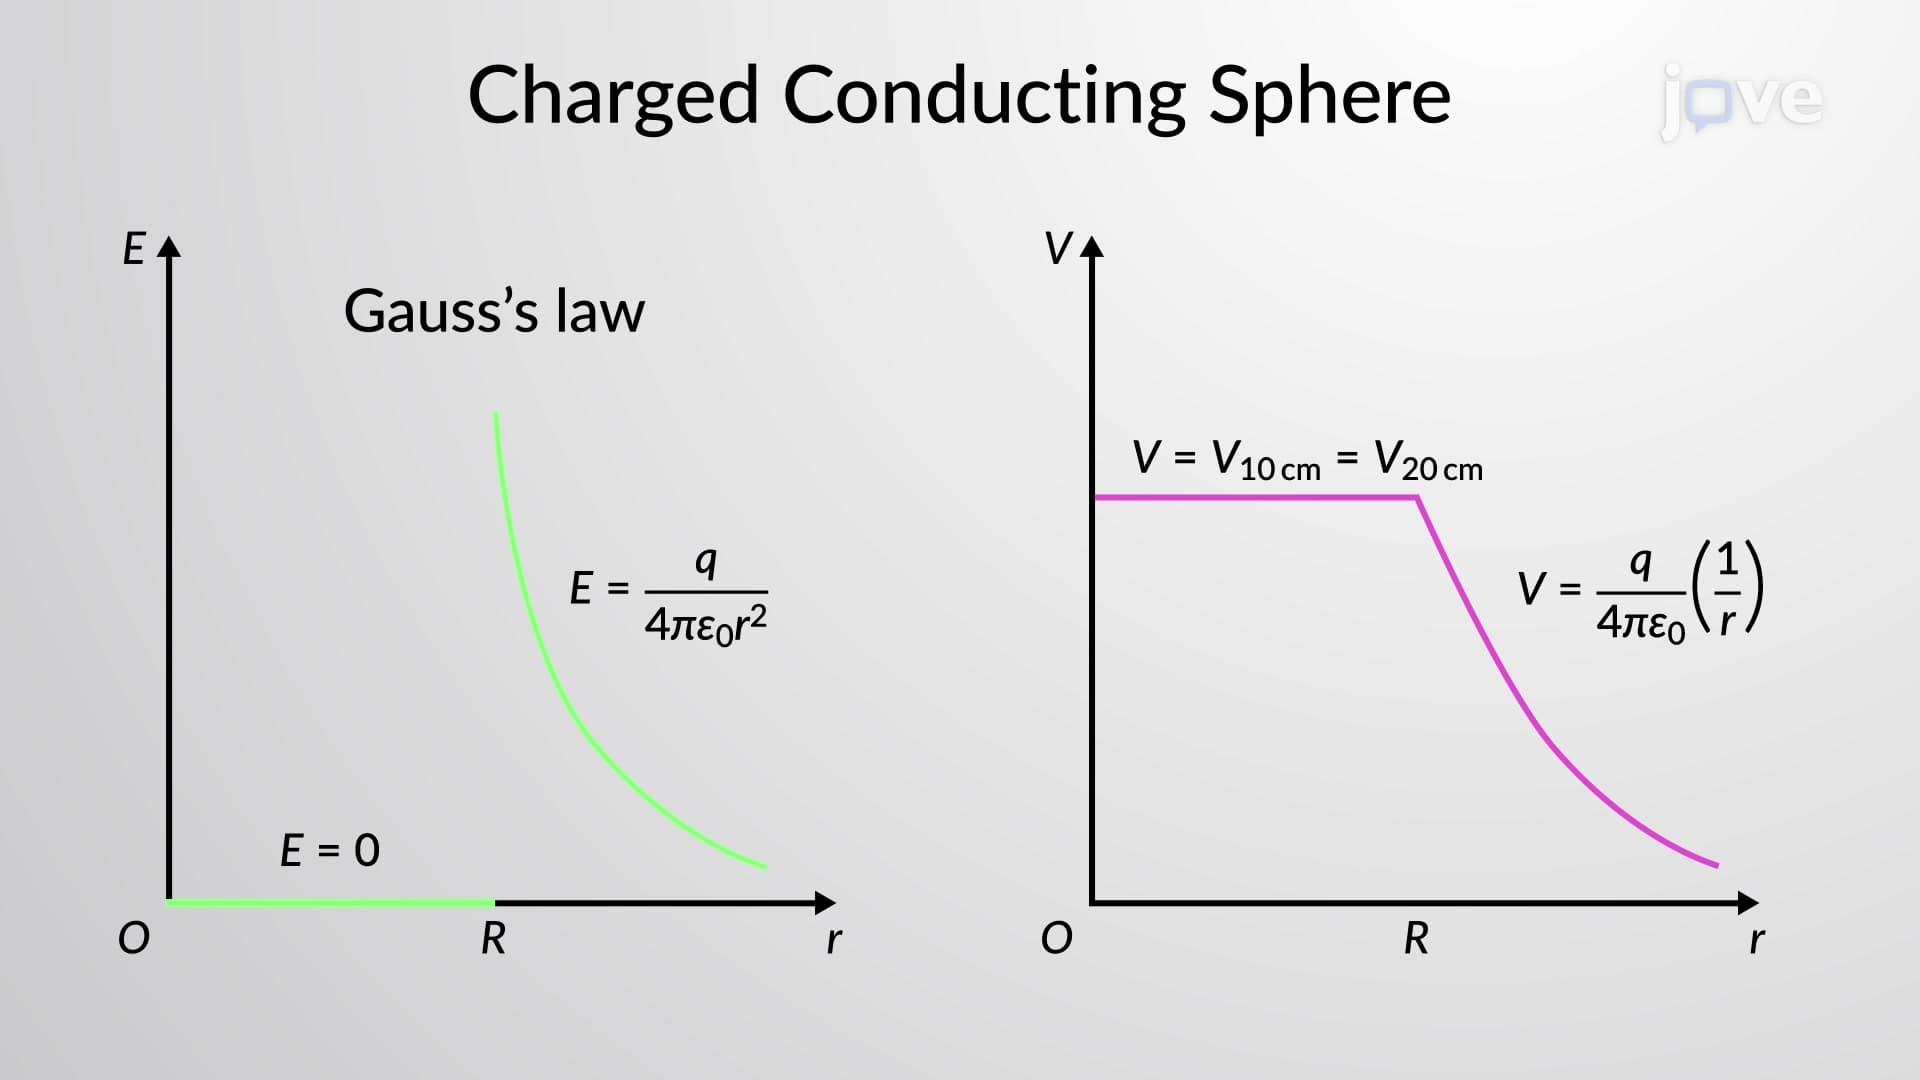
Calculations of Electric Potential I
The ring is divided into infinitesimal small arcs such that point M is equidistant from all the arcs. Here, the cylindrical coordinate system is used to calculate the electric potential at point M. A general element of the arc between angles θ and θ + dθ is of the length Rdθ and has a charge of...