Related Experiment Video
Updated: Jan 17, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Leveraging the Bias-Variance Tradeoff in Quantum Chemistry for Accurate Negative Singlet-Triplet Gap Predictions: A
Atreyee Majumdar1, Raghunathan Ramakrishnan1
1Tata Institute of Fundamental Research, Hyderabad, India.

Abstract:
Molecules that have been suggested to violate the Hund's rule, having a first excited singlet state ( ) energetically below the triplet state ( ), are rare. Yet, they hold the promise to be efficient light emitters. Their high-throughput identification demands exceptionally accurate excited-state modeling to minimize qualitatively wrong predictions. We benchmark twelve energy gaps to find that the local-correlated versions of ADC(2) and CC2 excited state methods deliver excellent accuracy and speed for screening medium-sized molecules. Notably, we find that double-hybrid DFT approximations (e.g., B2GP-PLYP and PBE-QIDH) exhibit high mean absolute errors ( ) despite very low standard deviations ( ). Exploring their parameter space reveals that a configuration with 75% exchange and 55% correlation, which reduces the mean absolute error to below 5 meV, but with an increased variance. Using this low-bias parameterization as an internal reference, we correct the systematic error while maintaining low variance, effectively combining the strengths of both low-bias and low-variance DFT parameterizations to enhance overall accuracy. Our findings suggest that low-variance DFT methods, often overlooked due to their high bias, can serve as reliable tools for predictive modeling in first-principles molecular design. The bias-correction data-fitting procedure can be applied to any general problem where two flavors of a method, one with low bias and another with low variance, have been identified a priori.

Related Concept Videos
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...
 03:35
Hybridization of Atomic Orbitals II
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:24
Hybridization of Atomic Orbitals I
 01:22
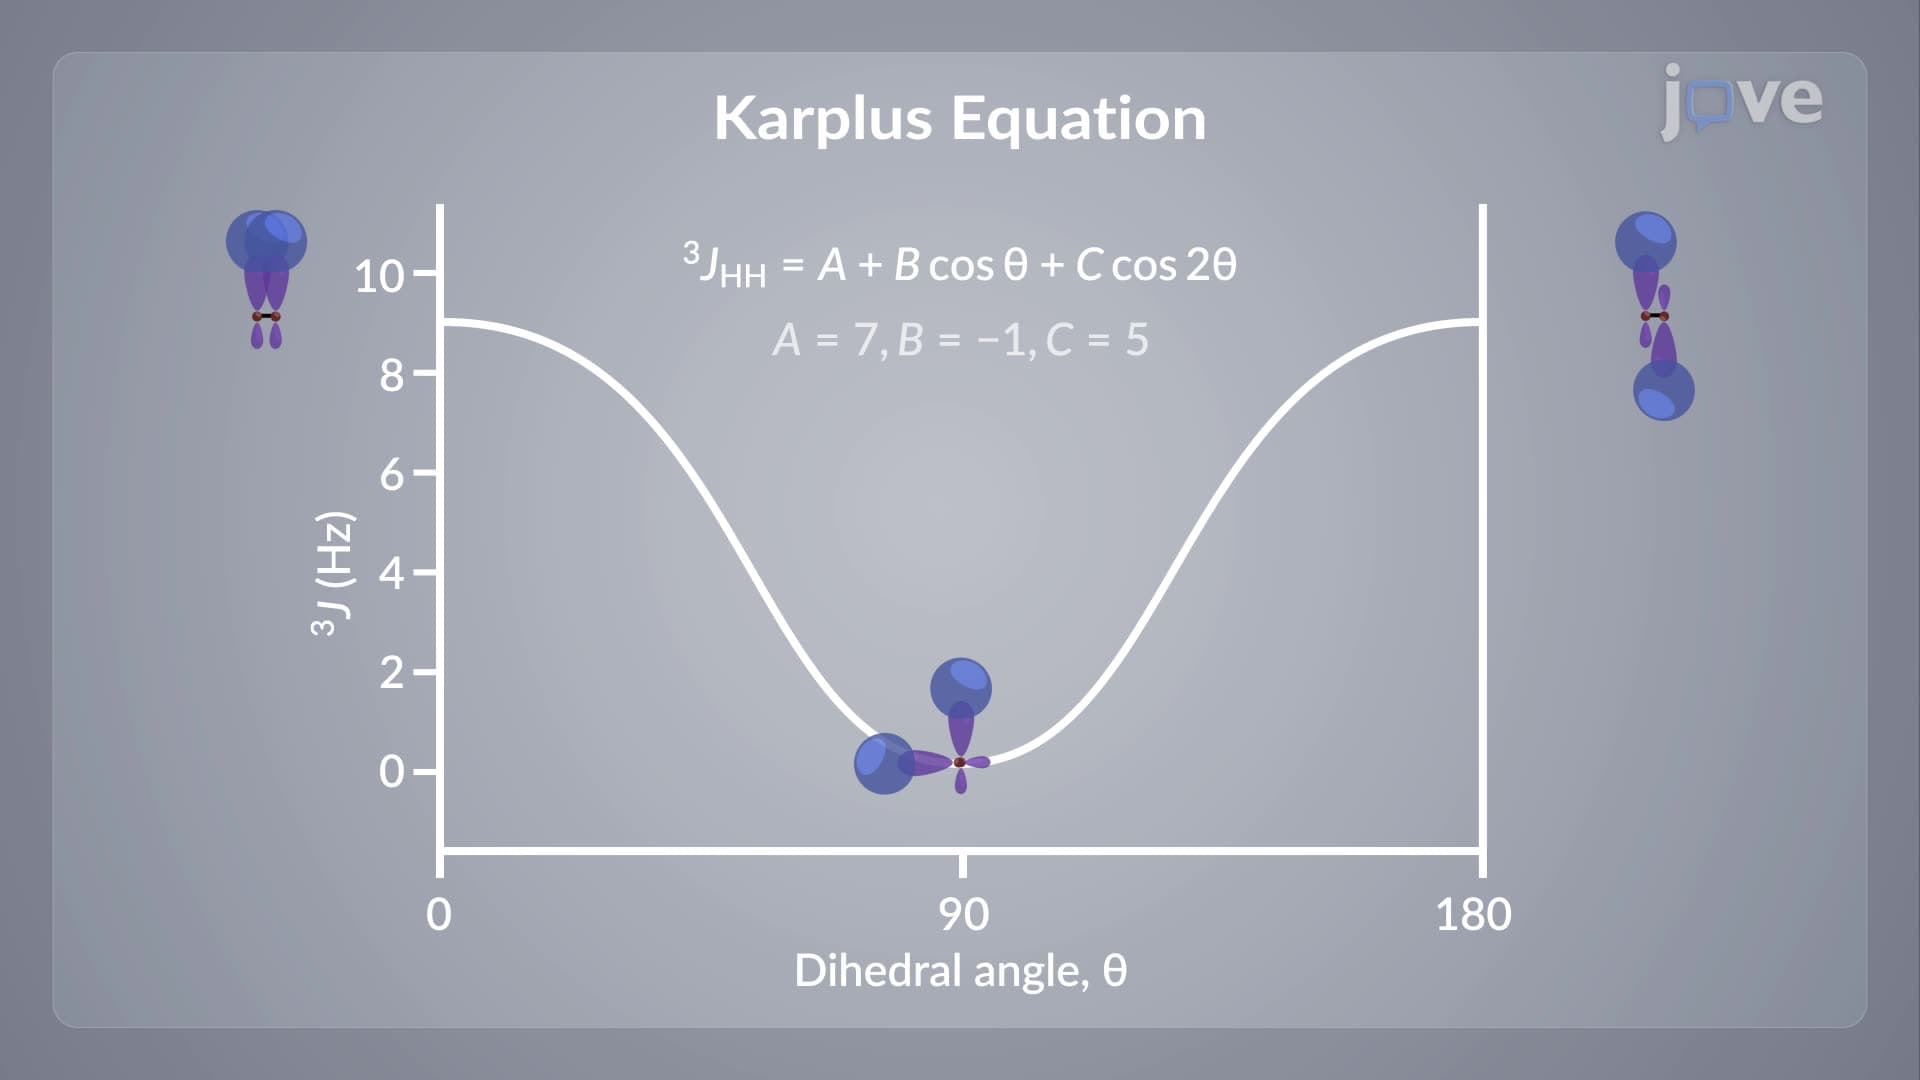
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:20
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...