Related Experiment Video
Updated: Jan 16, 2026
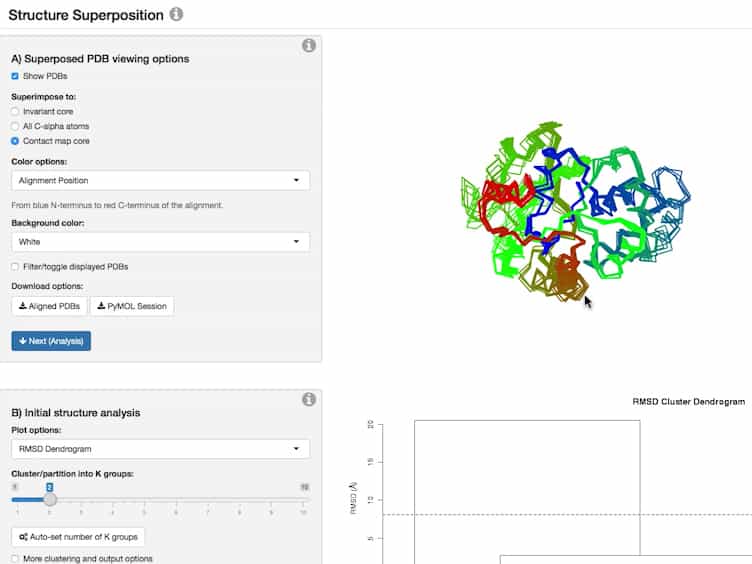

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Robust Conformational Space Exploration of Cyclic Peptides by Combining Different MD Protocols and Force Fields
Samuel Murail1, Jaysen Sawmynaden2, Akli Zemirli1
1Université Paris Cité, CNRS UMR 8251, INSERM ERL U1133, Unité de Biologie Fonctionnelle et Adaptative, BFA, F-75013 Paris, France.

Abstract:
Cyclic peptides are an important class of pharmaceutical drugs. We used replica-exchange molecular dynamics (REMD) and simulated tempering (ST) simulations to explore the conformational landscape of a set of nine cyclic peptides. The N-ter to C-ter backbone-cyclized peptides of 7-10 residues were previously designed for high conformational stability with a mixture of l- and d-amino acids. Their experimental NMR structures are available in the protein data bank (PDB). For each peptide, we tested several force fields, namely, Amber96, Amber14, RSFF2C, and Charmm36m in implicit and explicit solvents. We find that the variability of the free energy maps obtained from several protocols is larger than the variability obtained by just repeating the same protocol. Running multiple protocols is therefore important for the convergence assessment of REMD or ST simulations. The majority of the free energy maps showed clusters with a high RMSD compared to the native structures, revealing the residual flexibility of this set of cyclic peptides. The high RMSD clusters had in some cases the lowest free energy, rendering the prediction of the native structure more difficult with a single protocol. Fortunately, the combination of four implicit solvent REMD and ST simulations, mixing the Amber96 and Amber14 force fields, predicted robustly the native structure. As implicit solvent simulations in the REMD or ST setup are up to one hundred times faster than explicit solvent simulations, running four implicit solvent simulations is a good practical choice. We checked that the use of an explicit solvent REMD or ST simulation, taken alone or combined with implicit solvent simulations, did not significantly improve our results. It results in our combination of four implicit solvent simulations being tied in terms of success rate with much more expensive combinations that include explicit solvent simulations. This may be used as a guideline for further studies of cyclic peptide conformations.
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...