Related Experiment Video
Updated: Jan 16, 2026
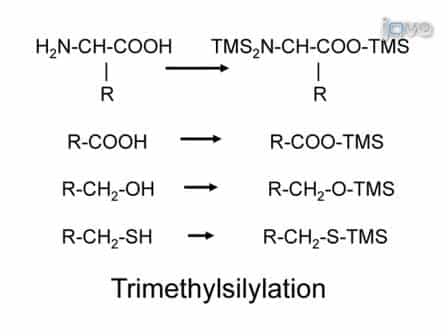

One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
A Deep Clinical and Biochemical Characterization of a Patient With Combined Malonic and Methylmalonic Aciduria
Vincenza Gragnaniello1, Alfonso Galderisi2, Sara Tucci3
1Department of Women's and Children's Health University of Padua Padua Italy.

Abstract:
Combined malonic and methylmalonic aciduria (CMAMMA) is an inborn error of metabolism caused by a deficiency in mitochondrial malonyl-CoA synthetase, the enzyme responsible for activating malonic acid (MA) to malonyl-CoA, a precursor of lipoic acid. The clinical phenotype is highly heterogeneous, ranging from asymptomatic cases to severe neurological impairment. We describe a patient affected by CMAMMA. The patient presented in the neonatal period with hyperinsulinemic hypoglycemia. Urinary organic acid analysis revealed elevated levels of both malonic and methylmalonic acids. The diagnosis of CMAMMA was confirmed through molecular testing of the ACSF3 gene. Levels of both lipoylated pyruvate dehydrogenase (PDH) and α-ketoglutarate dehydrogenase (αKGDH) were decreased. Given the role of lipoic acid in regulating insulin secretion, the involvement of impaired mitochondrial lipoic acid biosynthesis in the clinical presentation of hyperinsulinemic hypoglycemia cannot be excluded. We describe a case of CMAMMA associated with hyperinsulinemic hypoglycemia. While a definitive association between CMAMMA and hyperinsulinism cannot be established based on a single case, the observed reduction in lipoic acid levels may suggest a mechanistic connection between the two disorders. We suggest considering urinary organic acid testing in patients with hyperinsulinism, especially when the cause is unknown.
Related Concept Videos
 01:20
Inborn Errors of Metabolism
 01:35
Loss of Carboxy Group as CO2: Decarboxylation of Malonic Acid Derivatives
 01:21
Overview of Protein Metabolism
Amino acids play various roles in the body once they are absorbed into cells. They are restructured...
 01:22
Lysosomal Hydrolases
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 02:59
Animal Mitochondrial Genetics