Related Experiment Video
Updated: Jan 16, 2026
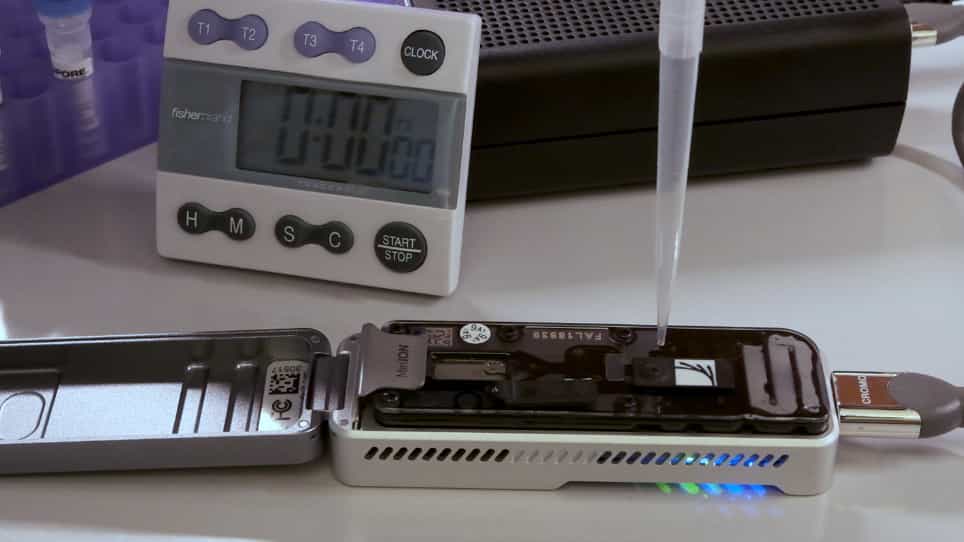

Hybrid De Novo Genome Assembly for the Generation of Complete Genomes of Urinary Bacteria using Short- and Long-read Sequencing Technologies
Published on: August 20, 2021
Evaluating urine volume and host depletion methods to enable genome-resolved metagenomics of the urobiome
Zachary J Lewis1,2, Angela Scott1, Christopher Madden1
1Department of Veterinary Preventive Medicine, The Ohio State University, Columbus, OH, USA.

Background:
The gut microbiome has emerged as a clear player in health and disease, in part by mediating host response to environment and lifestyle. The urobiome (microbiota of the urinary tract) likely functions similarly. However, efforts to characterize the urobiome and assess its functional potential have been limited due to technical challenges including low microbial biomass and high host cell shedding in urine. Here, to begin addressing these challenges, we evaluate urine sample volume (100 ml-5 mL) and host DNA depletion methods and their effects on urobiome profiles in healthy dogs, which are a robust large animal model for the human urobiome. We collected urine from seven dogs and fractionated samples into aliquots. One set of samples was spiked with host (canine) cells to model a biologically relevant host cell burden in urine. Samples then underwent DNA extraction followed by 16S rRNA gene and shotgun metagenomic sequencing. We then assembled metagenome-assembled genomes (MAGs) and compared microbial composition and diversity across groups. We tested six methods of DNA extraction: QIAamp BiOstic Bacteremia (no host depletion), QIAamp DNA Microbiome, Molzym MolYsis, NEBNext Microbiome DNA Enrichment, Zymo HostZERO, and propidium monoazide.
Results:
In relation to urine sample volume, ≥ 3.0 mL resulted in the most consistent urobiome profiling. In relation to host depletion, individual (dog) but not extraction method drove overall differences in microbial composition. DNA Microbiome yielded the greatest microbial diversity in 16S rRNA sequencing data and shotgun metagenomic sequencing data and maximized MAG recovery while effectively depleting host DNA in host-spiked urine samples. As proof-of-principle, we then mined MAGs for select metabolic functions including central metabolism pathways and environmental chemical degradation.
Conclusions:
Our findings provide guidelines for studying the urobiome in relation to sample volume and host depletion and lay the foundation for future evaluation of urobiome function in relation to health and disease. Video Abstract.