Related Experiment Video
Updated: Jan 15, 2026


Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
NO-modulated triplet ground state and two-state antiaromaticity in BN-doped cyclobutadienes: a combined DFT and
Sajid Imran1, Wenhao Wang1, Yuan Gao1
1Guangdong Basic Research Center of Excellence for Aggregate Science, School of Science and Engineering, The Chinese University of Hong Kong, Shenzhen, Guangdong 518172, China. jun.zhu@cuhk.edu.cn.

Abstract:
Controlling aromaticity across electronic states is crucial for designing novel species. While aromaticity typically could be achieved in either the lowest singlet state (S0) or the lowest triplet state (T1), dual-state aromaticity or antiaromaticity remains less developed. Herein, we demonstrate that NO-substitution uniquely induces antiaromaticity in both S0 and T1 states of 1,2-BN-doped cyclobutadiene (1,2-BN-CBD), initially nonaromatic in S0 and weakly aromatic in T1. Unlike attachment to nitrogen or carbon, NO bonding to boron (2) induces two-state antiaromaticity, as confirmed by Nucleus-Independent Chemical Shift (NICS), Electron Localization Function (ELFπ), NICS-grid, and Isomerization Stabilization Energy (ISE) analyses. Furthermore, compounds with NO at boron (2 and 10) exhibit triplet ground states. Spin density mainly localizes on NO, driving antiaromaticity in T1. Principal Interaction Orbital (PIO) and Principal Interacting Spin Orbital (PISO) analyses reveal that exocyclic BN double bond formation enforces planarization and enables localization in the S0 and T1 states, leading to two-state antiaromaticity in 2. K-means clustering combined with principal component analysis (one of the most commonly used unsupervised machine learning algorithms) classified BN-doped CBDs based on their electronic and structural properties, uniquely isolating 2 due to its distinct substituent positions and aromaticity behaviors. These findings highlight an important role of the substituent position in tuning electronic and aromatic properties.
Related Concept Videos
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group...
 01:20
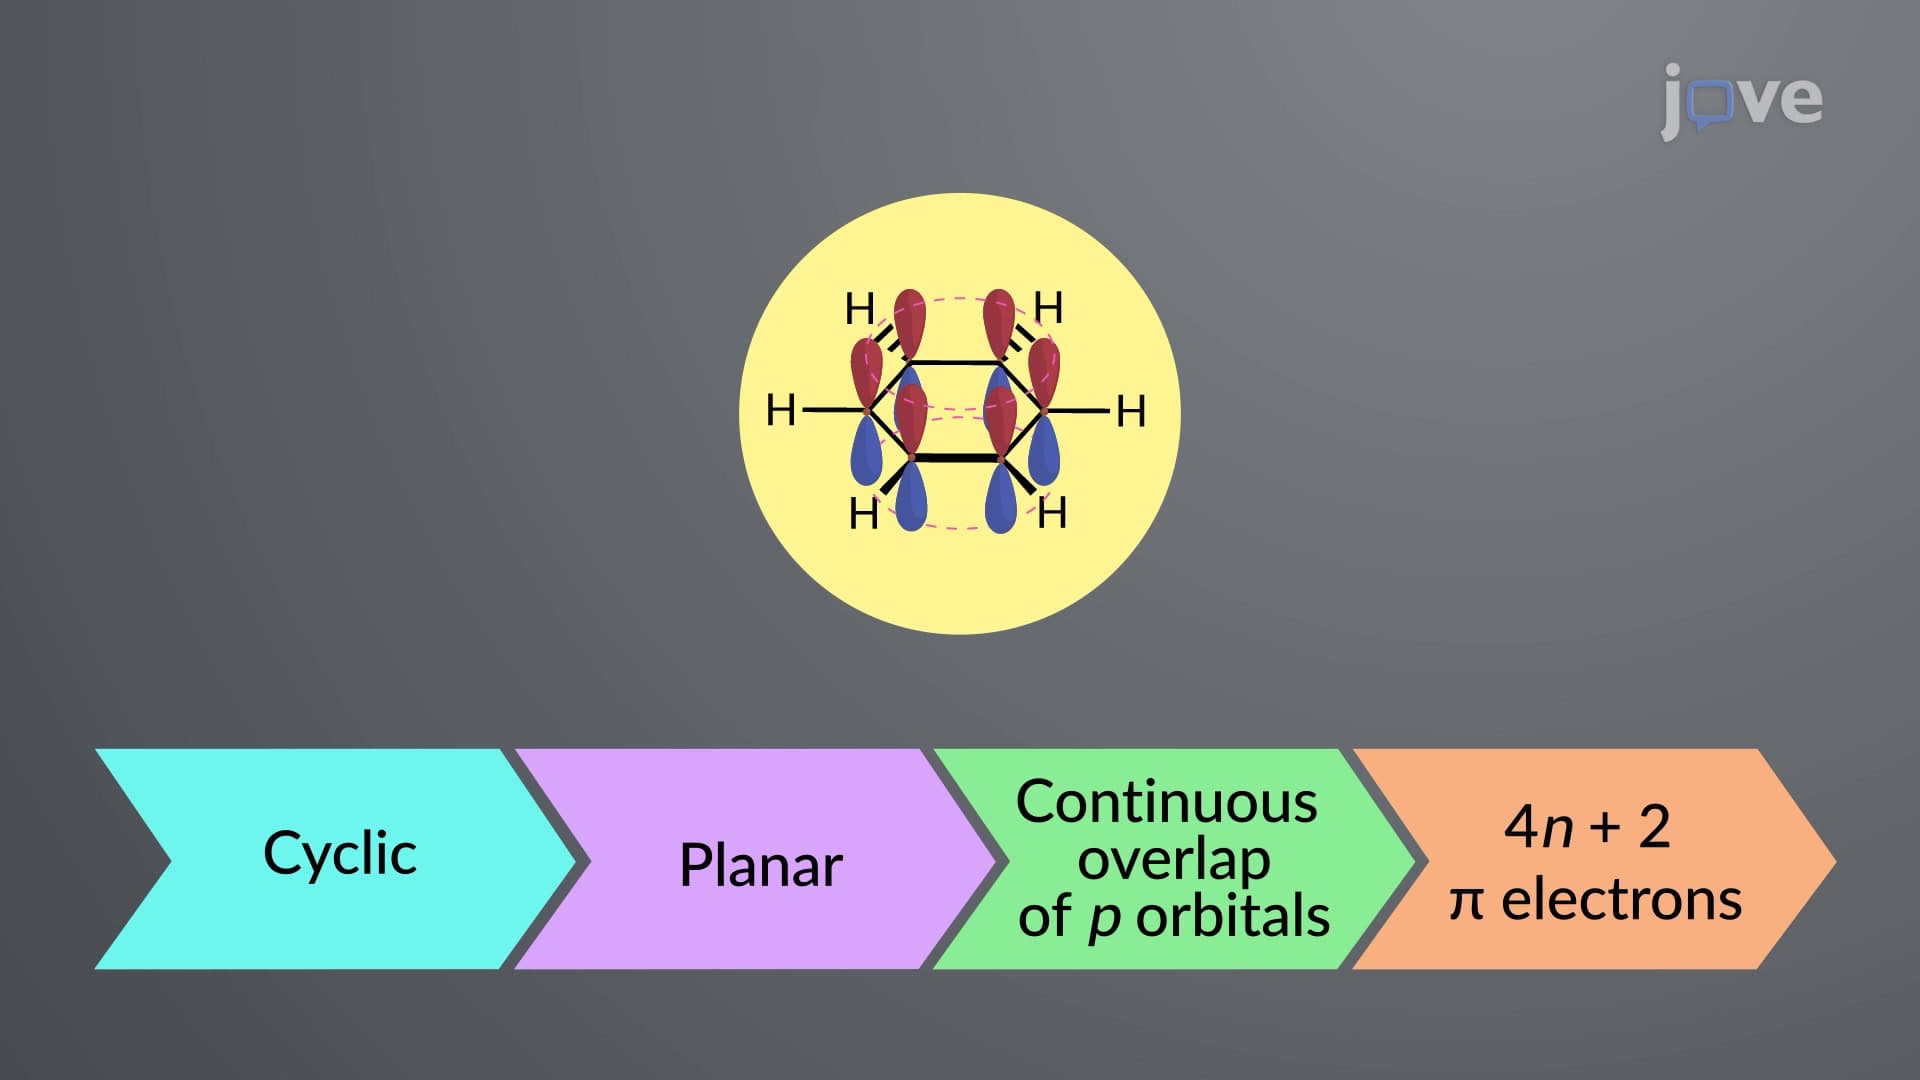
Criteria for Aromaticity and the Hückel 4n + 2 Rule
For the first time, Eric Hückel, a German chemical physicist, derived a set of structural features for a compound to be classified as aromatic. This is now known as Hückel’s rule or the 4n +...
 01:18
Frost Circles for Different Conjugated Systems
 01:18
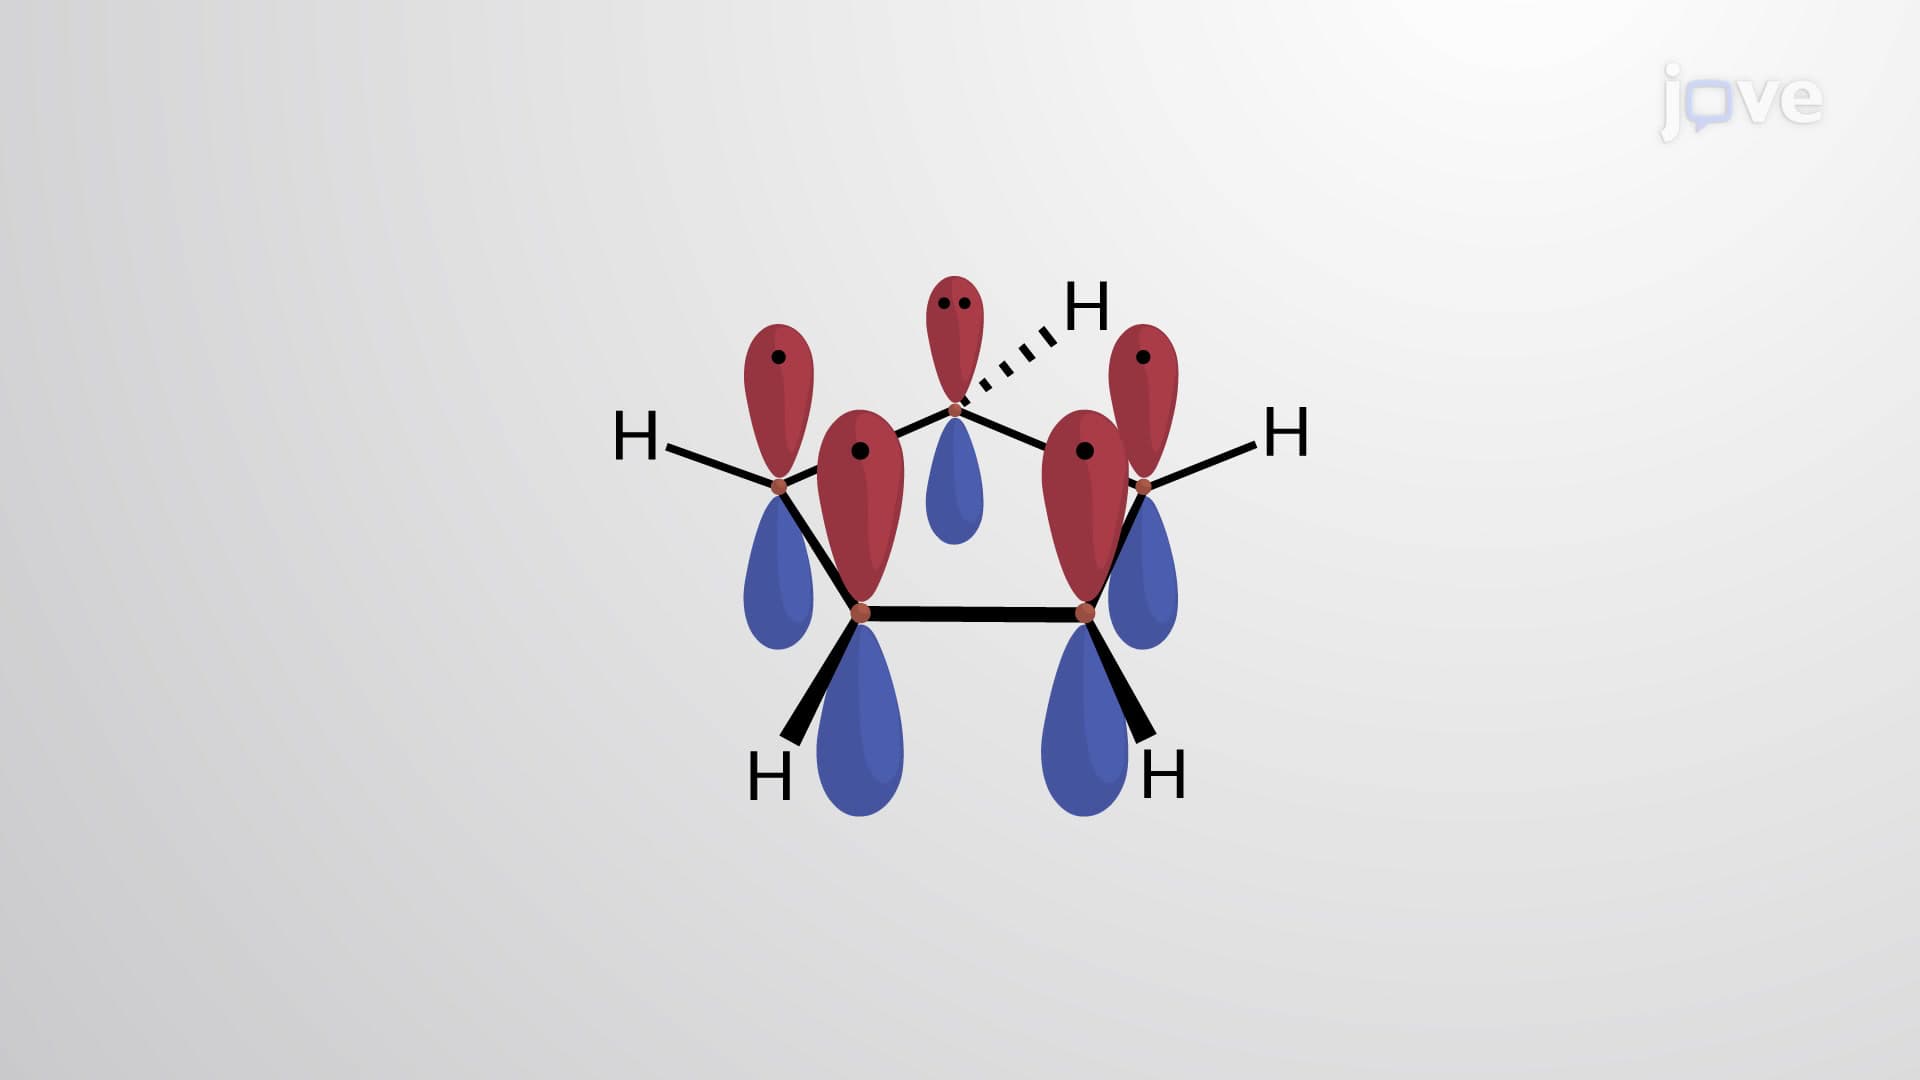
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds