Related Experiment Video
Updated: Jan 14, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Evaluation of Density Functionals for Si-O-C-H Molecule Thermochemistry
Ingeborg-Helene Svenum1, Francesca Lønstad Bleken2, Stefan Andersson1
1SINTEF Industry, P.O. Box 4760, Torgarden, Trondheim 7465, Norway.

Abstract:
The energies and vibrational frequencies of molecular species with Si-O-C-H compositions have been calculated at the CCSD(T) level (coupled cluster with single and double excitations and a perturbative treatment of triple excitations). The CCSD(T) results compare well with experimental data where the difference in enthalpy of formation is typically only about 1-2 kJ/mol in most cases. In addition, the same molecules have been calculated with density functional theory (DFT) calculations using nine commonly used density functionals and two different basis sets. The performance of the DFT calculations is compared with the CCSD(T) benchmark values in terms of enthalpy of formation, reaction energy, vibrational frequencies, and zero-point energies. The results show that the M06-2X functional provides the lowest mean absolute error (MAE) in terms of the enthalpy of formation, whereas, for the vibrational frequencies and zero-point energies, the SCAN functional gives the lowest MAE values. The results were also grouped according to the types of bonds that are present in the molecules. Moreover, an elaborate set of possible reactions within the molecular species in the Si-O-C-H system is included to evaluate the performance of the different DFT functionals with respect to the relative stability of species within the same reaction system. In this case, the B2GP-PLYP functional shows the smallest errors. PW6B95 is the functional that most consistently performs well for the studied properties of the included molecules. The coupled cluster data sets provide new benchmark data, several of which are not previously available for silicon chemistry.


Related Concept Videos
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding
 03:40
Hess's Law
 01:24
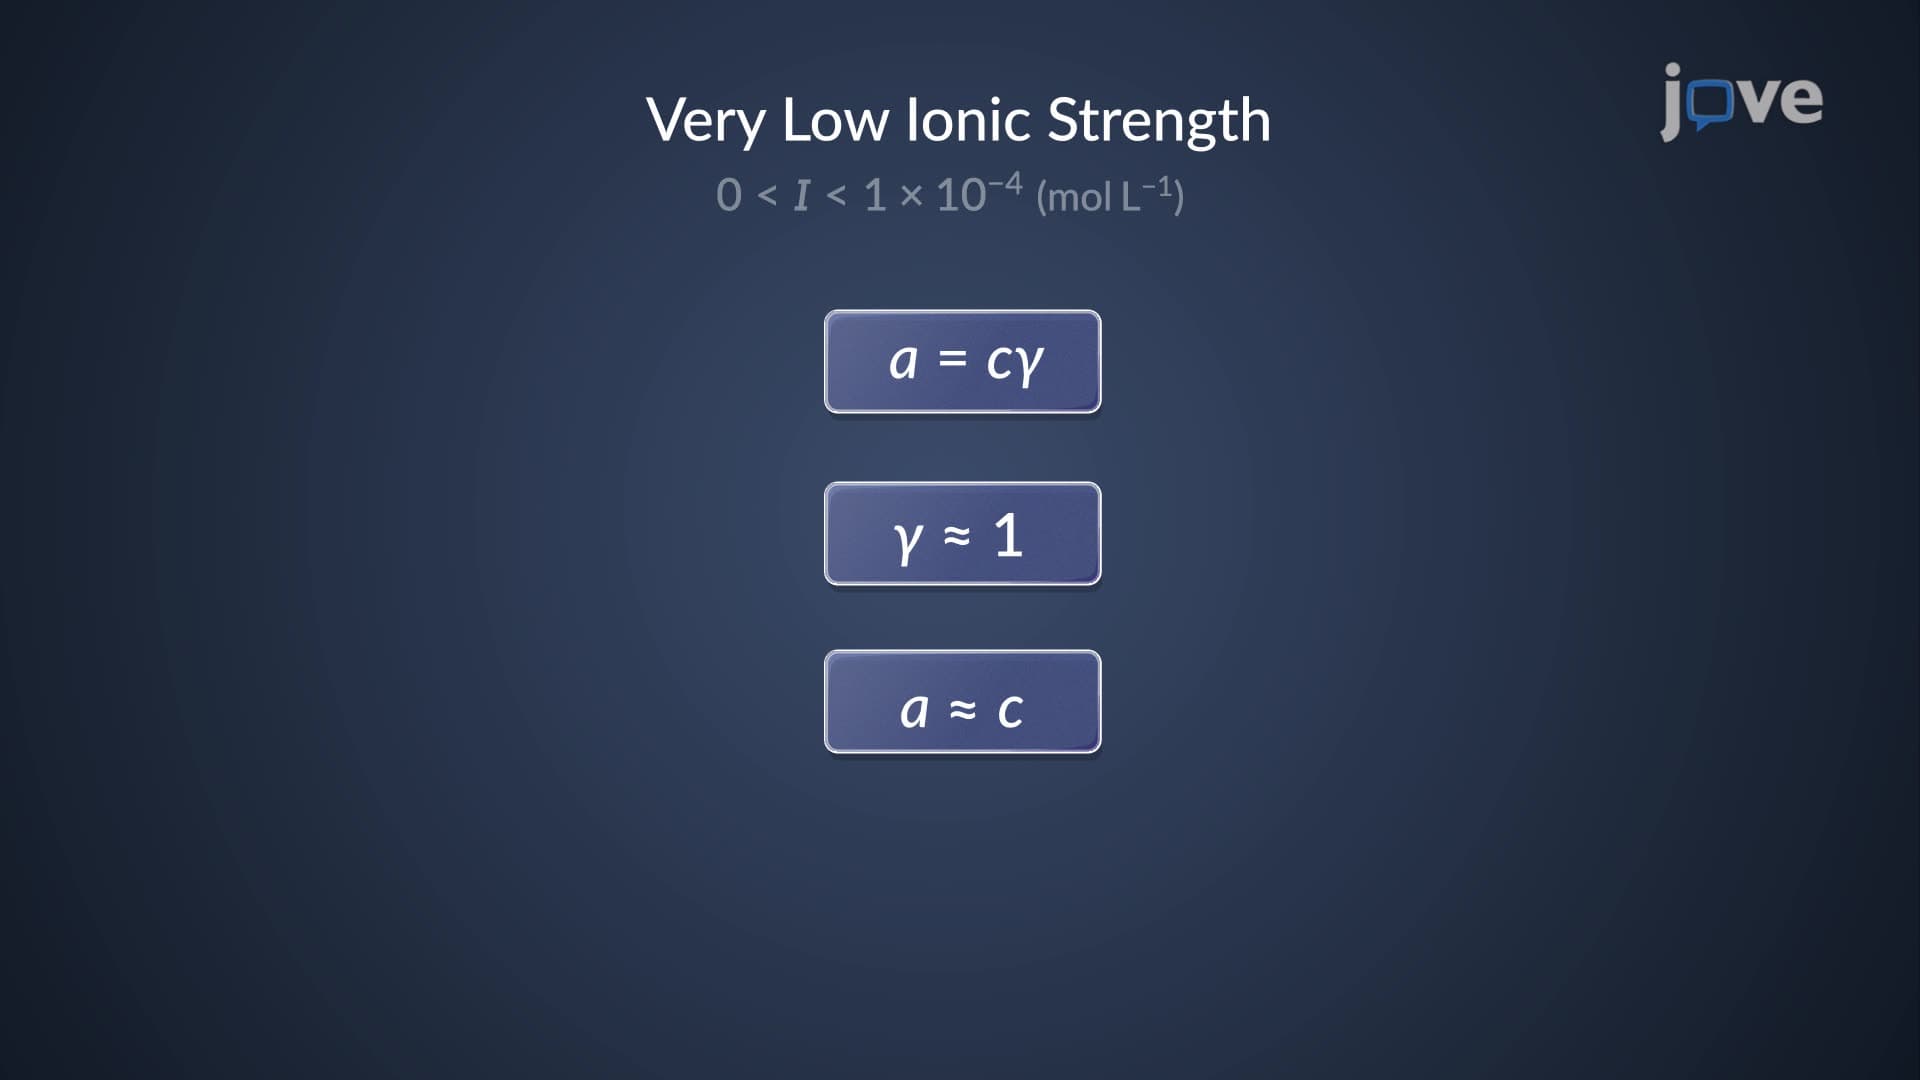
Thermodynamics: Activity Coefficient
The activity coefficient is a measure of the deviation from ideal behavior. When the ionic strength of the solution is minimal, the activity coefficient of an ionic species is close to unity, making...
 02:36
Molecular Geometry and Dipole Moments
 01:26
Thermodynamic Potentials