Related Experiment Video
Updated: Jan 12, 2026


Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Machine learning workflow for analysis of high-dimensional order parameter space: A case study of polymer
Elyar Tourani1, Brian J Edwards1, Bamin Khomami1
1Materials Research and Innovation Laboratory, Department of Chemical and Biomolecular Engineering, University of Tennessee, Knoxville, Tennessee 37996, USA.

Abstract:
Currently, identification of crystallization pathways in polymers is being carried out using molecular simulation-based data on a preset cutoff point on a single order parameter (OP) to define nucleated or crystallized regions. Aside from sensitivity to the cutoff, each of these OPs introduces its own systematic biases. In this study, an integrated machine learning workflow is presented to quantify accurately crystallinity in polymeric systems using atomistic molecular dynamics simulation data. Each atom is represented by a high-dimensional feature vector that combines geometric, thermodynamic-like, and symmetry-based descriptors. Low-dimensional embeddings are employed to expose latent structural fingerprints within atomic environments. Subsequently, unsupervised clustering on the embeddings is used to identify crystalline and amorphous atoms with high fidelity. After generating high-quality labels with multidimensional data, we use supervised learning techniques to identify a minimal set of order parameters that can fully capture this label. Various tests were conducted to reduce the feature set, and it is shown that using only three order parameters, namely q6, S̄i, and p2, is sufficient to recreate the crystallization labels with great accuracy. Based on these observed OPs, the crystallinity index (C-index) is introduced as the logistic regression model's probability of crystallinity. This measure remains bimodal at all stages of the process and achieves >98% classification performance. Notably, a model trained on one or a few snapshots enables efficient on-the-fly computation of crystallinity. Lastly, we demonstrate how the optimal C-index fit evolves during various stages of crystallization, supporting the hypothesis that entropy dominates early nucleation, while q6 gains relevance in the later stages. This workflow yields a data-driven strategy for OP selection and provides a generalizable metric to monitor structural transformations in large-scale polymer simulations.
Related Concept Videos
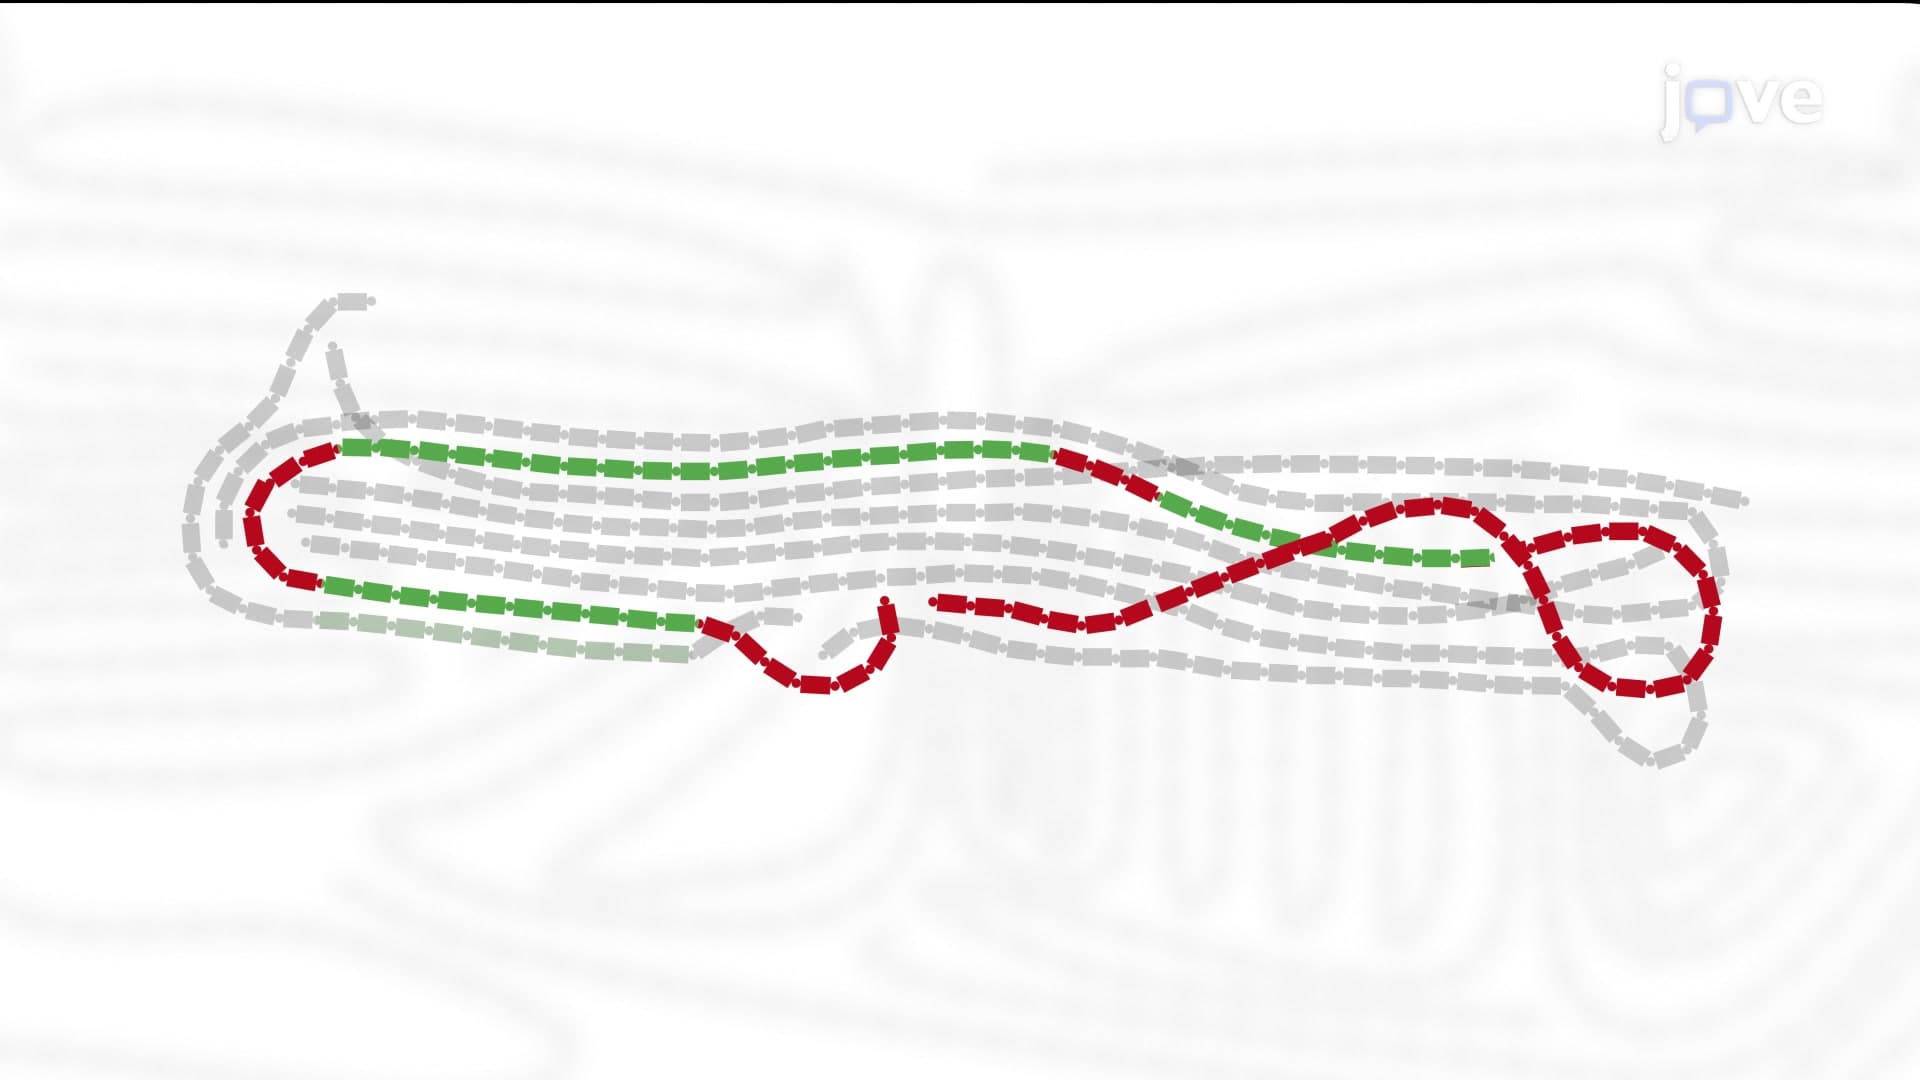 01:21
Polymer Classification: Crystallinity
Crystalline domains are the regions where polymer chains are aligned in an orderly manner and held together in proximity by intermolecular forces. For example, chains in the crystalline domains of polyethylene and nylon are bound together by van der Waals...
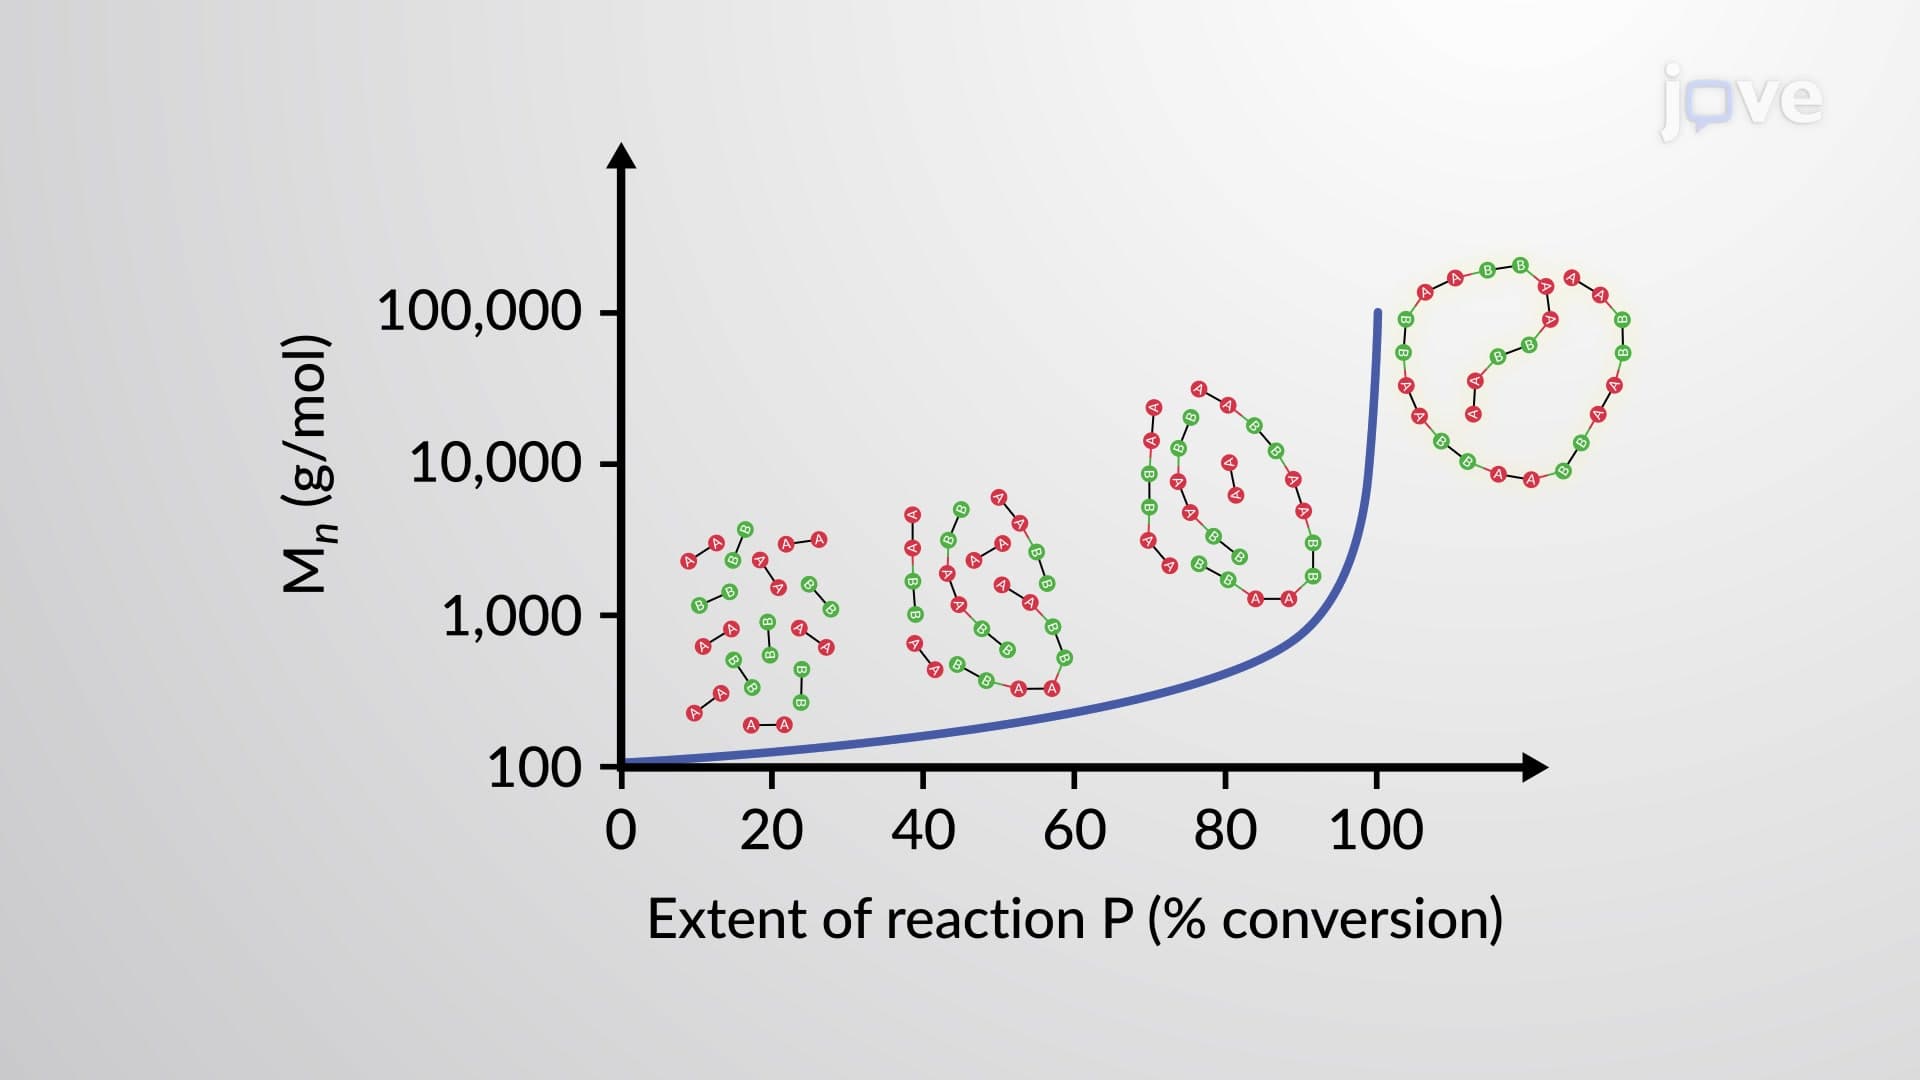 01:08
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...
 01:03
Step-Growth Polymerization: Overview
Many natural and synthetic polymers are produced by...