Related Experiment Video
Updated: Jan 11, 2026


Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Machine learning-enhanced multiple time-step ab initio molecular dynamics
François Mouvet1, Nicholas J Browning1, Pablo Baudin1
1Laboratory of Computational Chemistry and Biochemistry, Ecole Polytechnique Fédérale de Lausanne, Lausanne, Switzerland.

Abstract:
The efficiency of molecular dynamics is limited by the time step that can be used to integrate equations of motion, which is dictated by the highest frequency motion in the system. Multiple time step (MTS) integrators alleviate this issue by decomposing the forces acting on the particles into "fast" and "slow" components, which can then be integrated using different time steps. In ab initio MTS, an inexpensive, low level electronic structure method can be used to integrate the fast components, while its difference with an expensive but accurate high level method is used for the slow components. In this work, we present a machine learning-enhanced multiple time step (ML-MTS) method for performing accurate Born-Oppenheimer molecular dynamics at significantly reduced computational cost. We propose two alternative ML-MTS schemes, which invoke different timescale separations and result in stable and accurate trajectories. In the first scheme, ML force estimates bypass the need for a high level calculation, resulting in speedups of two orders of magnitude over standard velocity Verlet (VV) integration using a hybrid exchange-correlation functional. In the second scheme, we keep the high level calculation for the slow component and an ML correction is applied to the fast component, allowing a fourfold increase in time step compared to modern ab initio MTS algorithms without any loss of stability, thus yielding speedups up to almost an order of magnitude over straightforward VV.
More Related Videos
 05:00
Author Spotlight: Streamlining Visual Dynamics to Simplify Molecular Dynamics Simulations Using Gromacs
Published on: August 9, 2024
 07:31
Author Spotlight: Advancing Cell Membrane Biophysics - Exploring Interactions and Challenges Through Experimental and Computational Approaches
Published on: September 1, 2023
Related Concept Videos
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...
 02:31
Multi-Step Reactions
 01:22
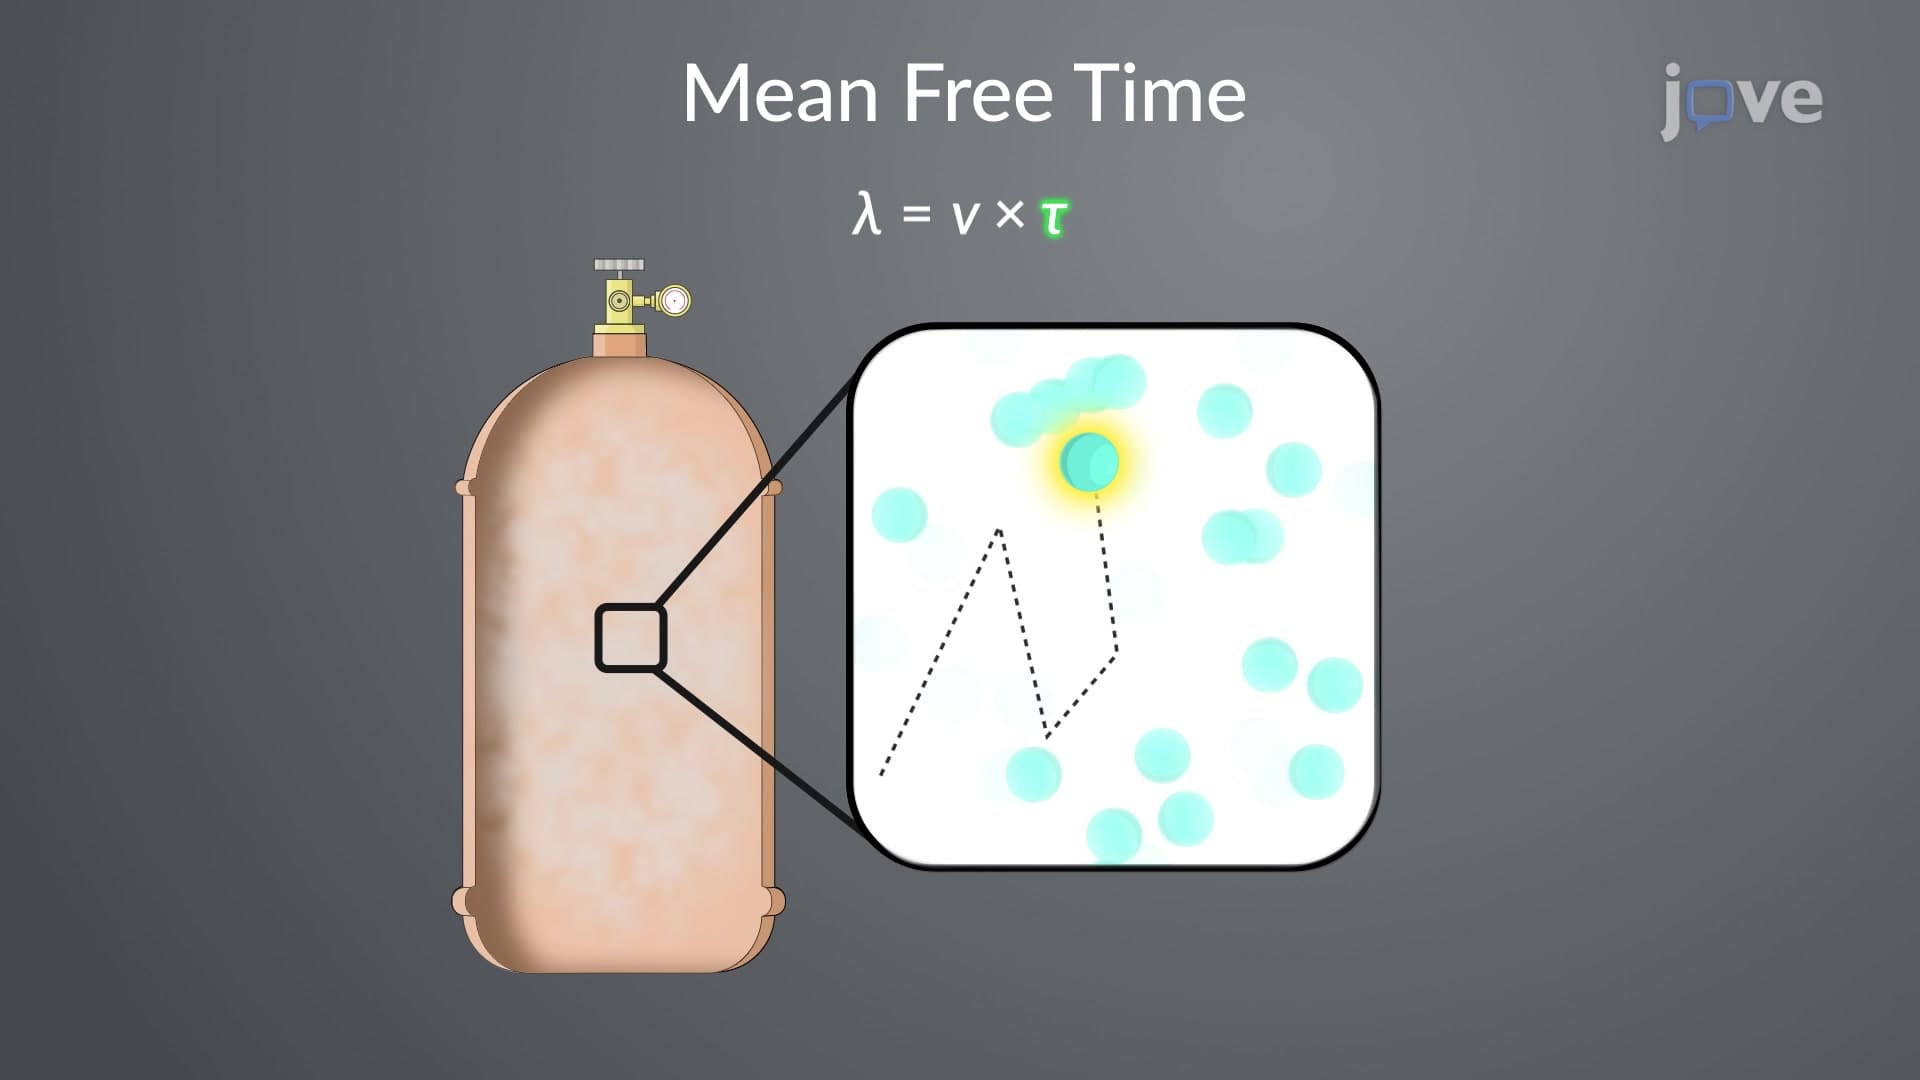
Mean free path and Mean free time
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:08
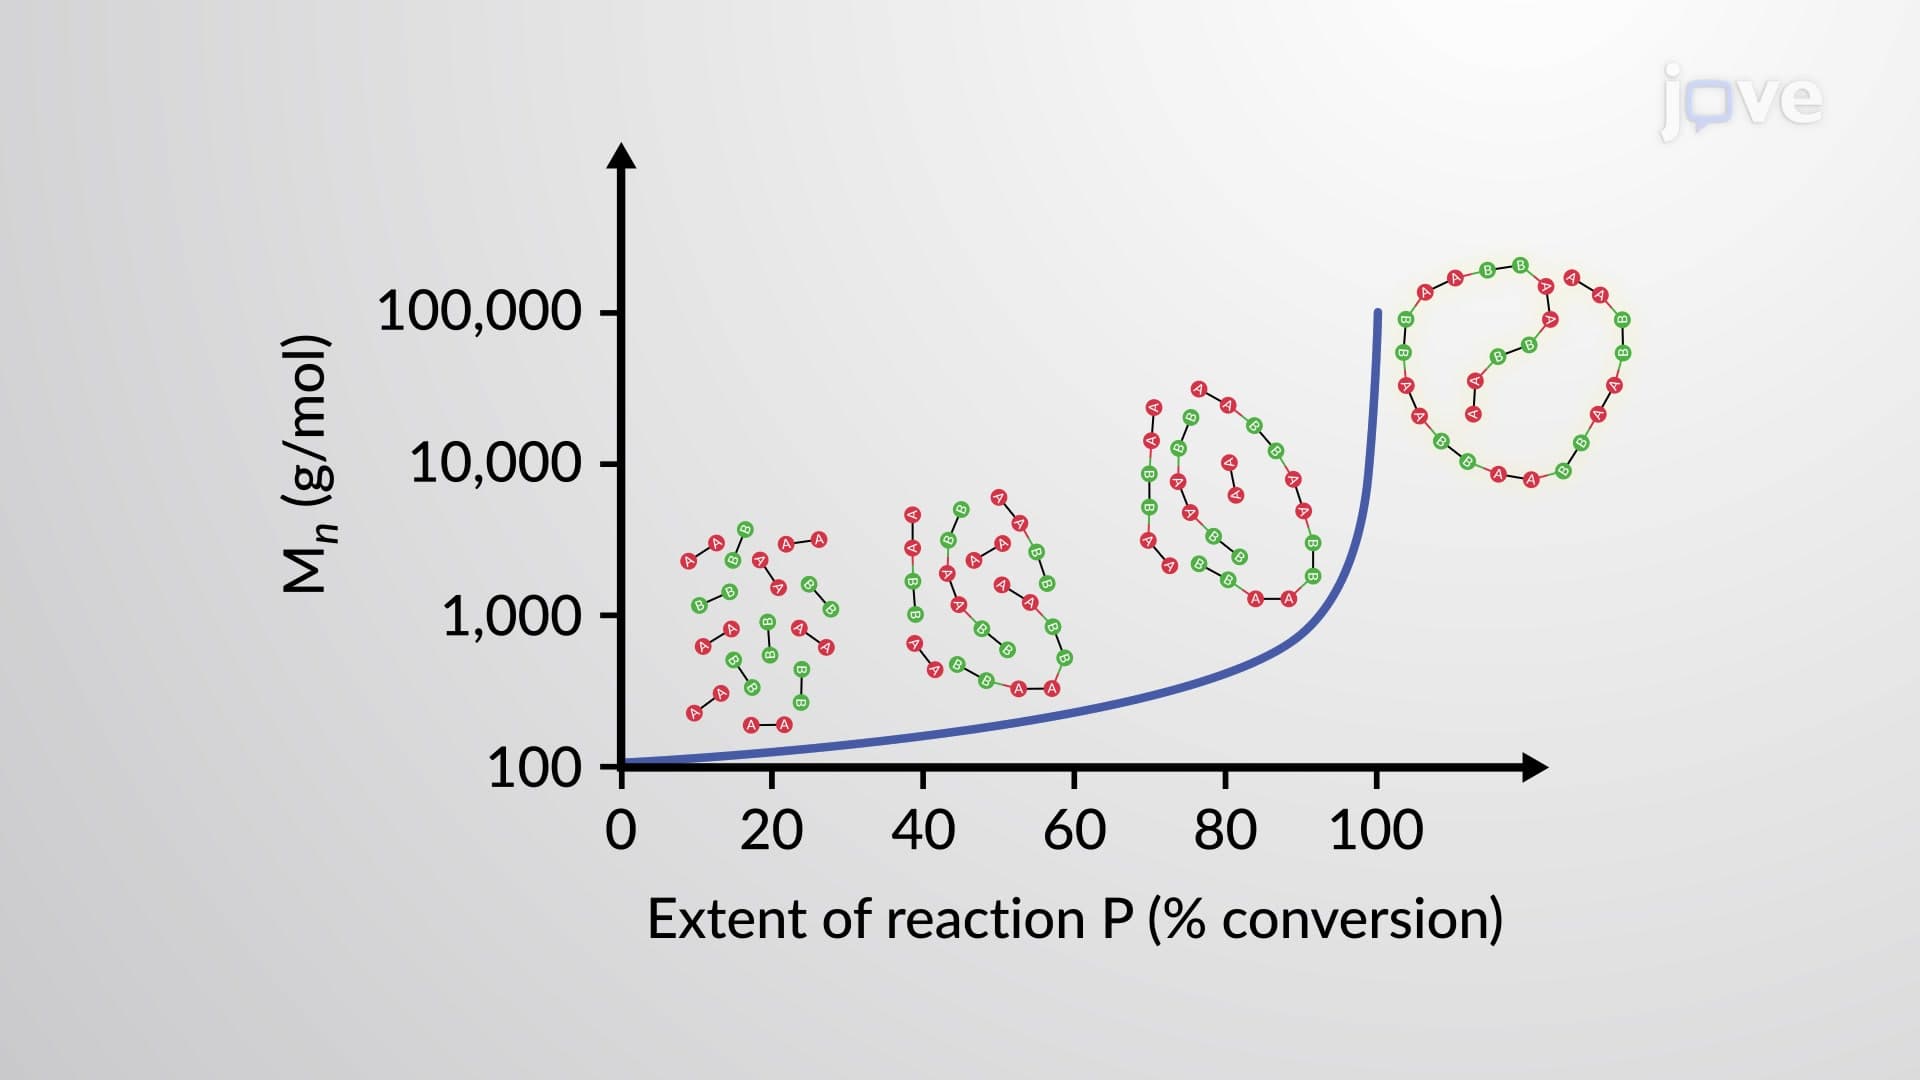
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...