Related Experiment Video
Updated: Jun 12, 2026
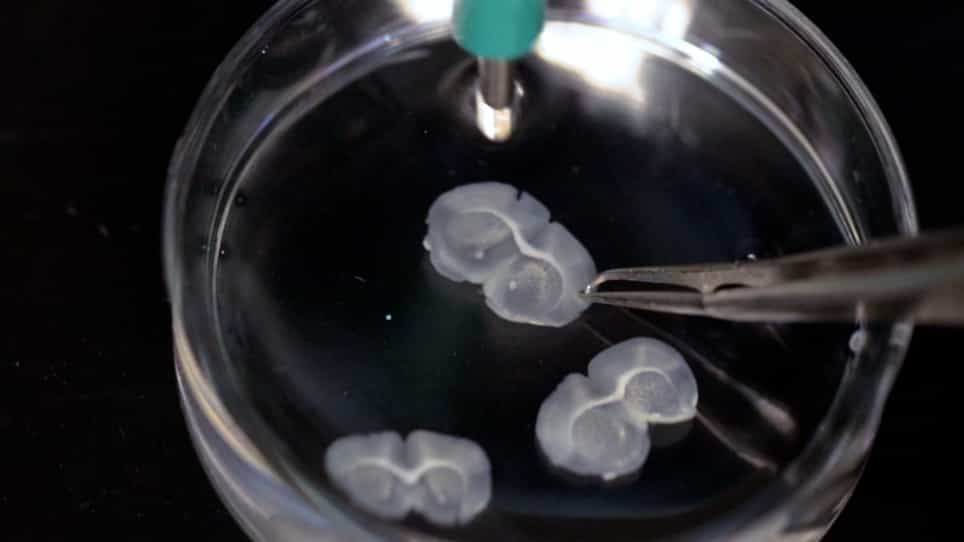

Measurement of Oxygen Consumption Rate in Acute Striatal Slices from Adult Mice
Published on: June 8, 2022
CHCHD2 mutant mice link mitochondrial deficits to PD pathophysiology
Szu-Chi Liao1,2,3,4, Kohei Kano1,4, Sadhna Phanse5
1Gladstone Institute of Neurological Disease, Gladstone Institutes, San Francisco, CA, USA.

Abstract:
Mitochondrial dysfunction is a hallmark of Parkinson's disease (PD), but the mechanisms by which it drives autosomal dominant and idiopathic forms of PD remain unclear. To investigate this, we generated and performed a comprehensive phenotypic analysis of a knock-in mouse model carrying the T61I mutation in the mitochondrial protein CHCHD2 (coiled-coil-helix-coiled-coil-helix domain-containing 2), which causes late-onset symptoms indistinguishable from idiopathic PD. We observed pronounced mitochondrial disruption in substantia nigra dopaminergic neurons, including distorted ultrastructure and CHCHD2 aggregation, as well as disrupted mitochondrial protein-protein interactions in brain lysates. These abnormalities were associated with a whole-body metabolic shift toward glycolysis, elevated mitochondrial reactive oxygen species (ROS), and progressive accumulation of aggregated α-synuclein. In idiopathic PD, CHCHD2 gene expression also correlated with α-synuclein levels in vulnerable dopaminergic neurons, and CHCHD2 protein accumulated in early Lewy aggregates. These findings delineate a pathogenic cascade in which CHCHD2 accumulation impairs mitochondrial respiration and increases ROS production, driving α-synuclein aggregation and neurodegeneration.
Related Concept Videos
 01:24
Parkinson Disease ll: Pathophysiology
 01:21
Huntington Disease l: Introduction