Related Experiment Video
Updated: Jan 11, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Learned free-energy functionals from pair-correlation matching for dynamical density functional theory
Karnik Ram1,2, Jacobus Dijkman3,4, René van Roij5
1TU Munich, School of Computation, Information and Technology, Munich, Germany.

Abstract:
Classical density functional theory (cDFT) and dynamical density functional theory (DDFT) are modern statistical mechanical theories for modeling many-body colloidal systems at the one-body density level. The theories hinge on knowing the excess free-energy accurately, which is, however, not feasible for most practical applications. Dijkman et al. [Phys. Rev. Lett. 134, 056103 (2025)0031-900710.1103/PhysRevLett.134.056103] recently showed how a neural excess free-energy functional for cDFT can be learned from bulk simulations via pair-correlation matching. In this article, we demonstrate how this same functional can be applied to DDFT, without any retraining, to simulate nonequilibrium overdamped dynamics of inhomogeneous densities. We evaluate this on a three-dimensional Lennard-Jones system with planar geometry under various complex external potentials, and we observe good agreement of the dynamical densities with those from expensive Brownian dynamic simulations, up to the limit of the adiabatic approximation. We further develop and apply an extension of DDFT based on gradient flows to a grand-canonical system modeled after breakthrough gas adsorption studies, finding similarly good agreement. Our results demonstrate a practical route for leveraging learned free-energy functionals in DDFT, paving the way for accurate and efficient modeling of many-body nonequilibrium systems.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 01:26
Thermodynamic Potentials
 03:25
Free Energy Changes for Nonstandard States
 01:10
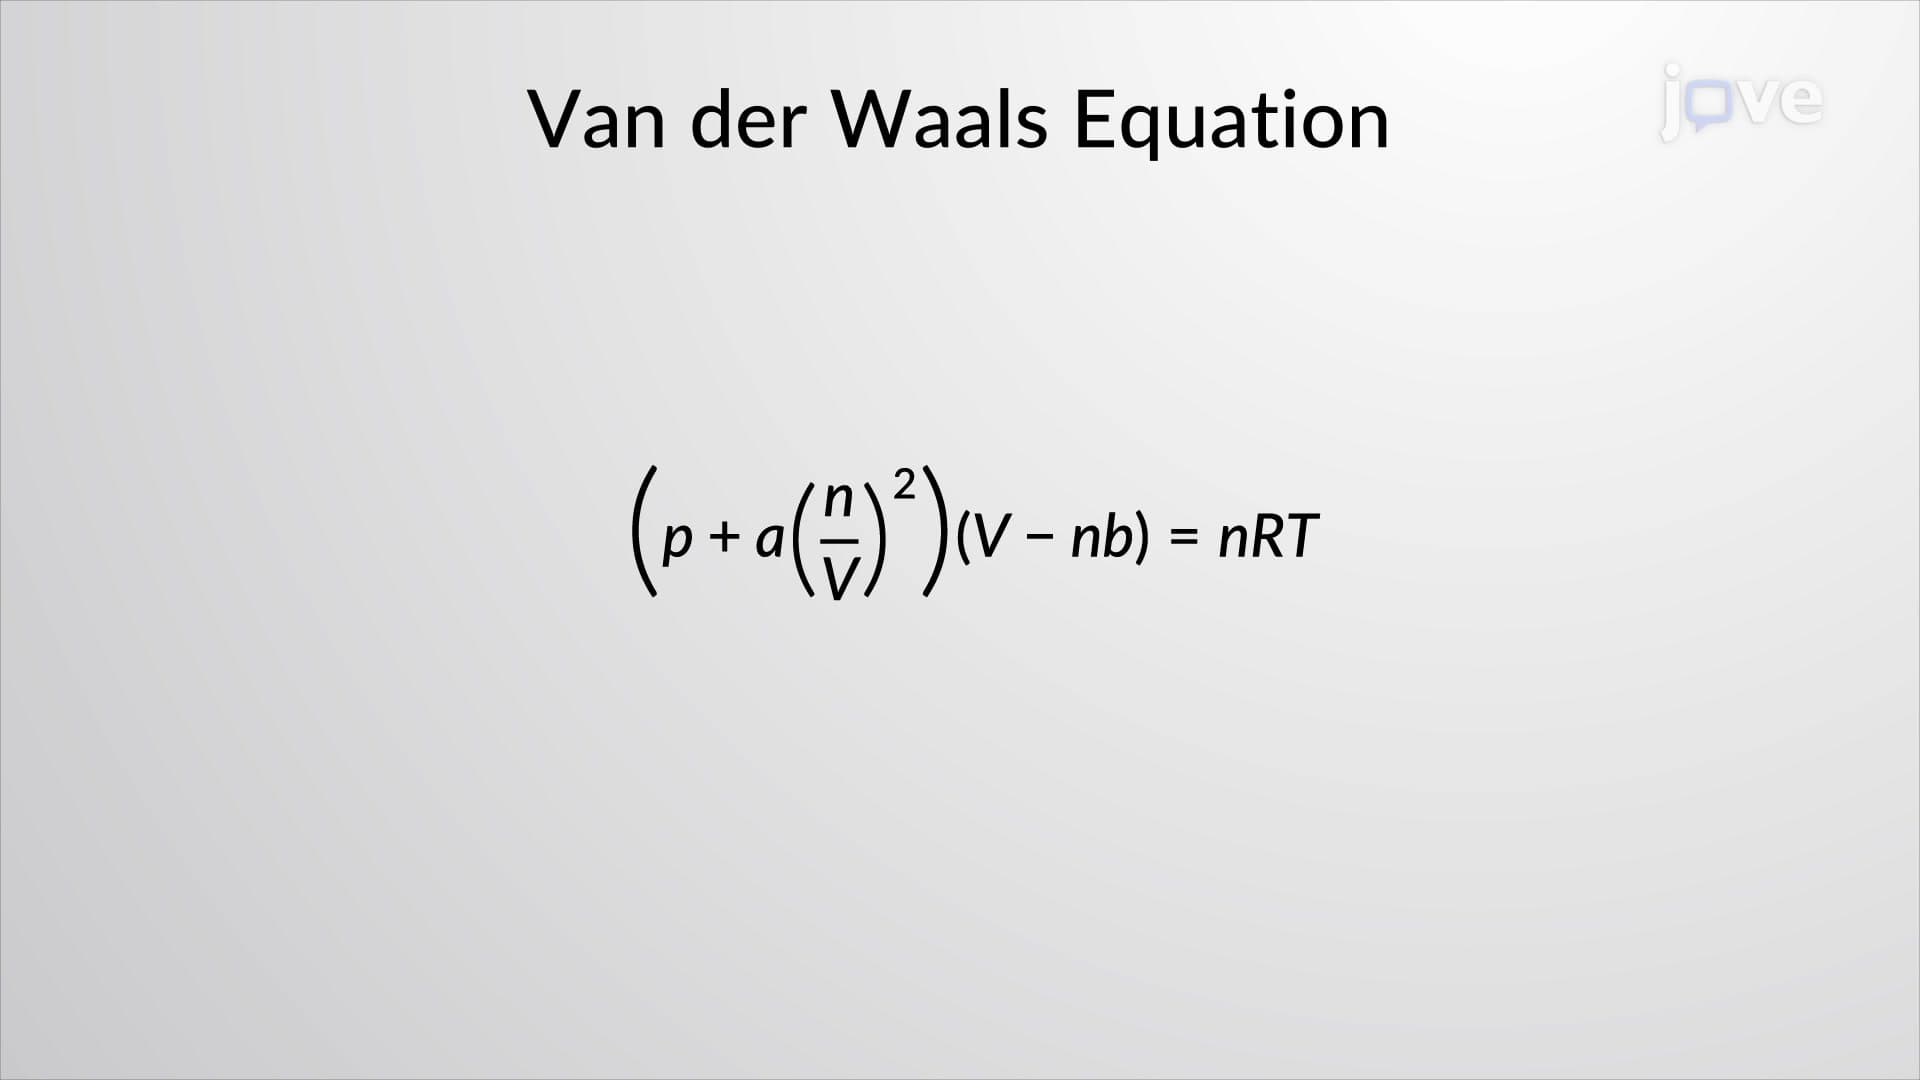
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
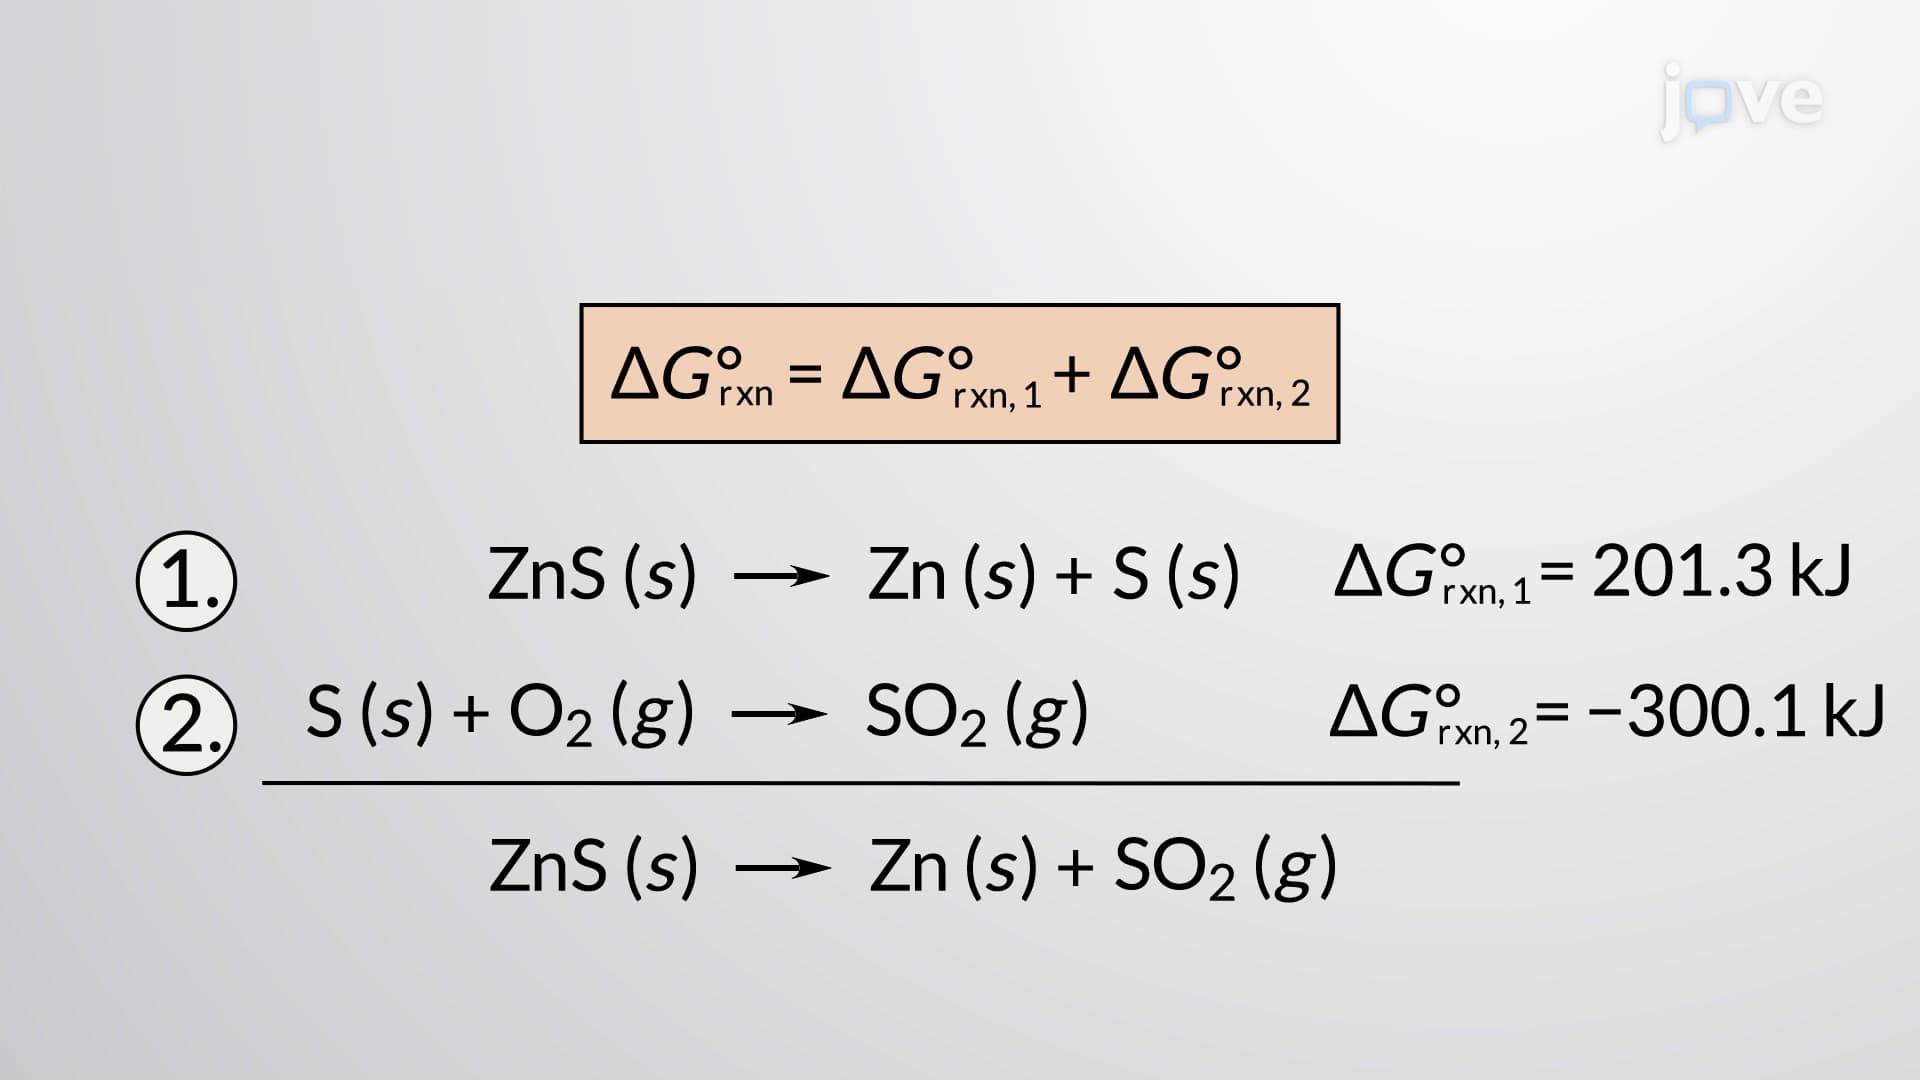 02:49
Calculating Standard Free Energy Changes
 01:24
Van der Waals Interactions
 01:16
Potential-Energy Criterion for Equilibrium