Related Experiment Video
Updated: Jan 10, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Structure-activity relationship analysis of 5'-substituted C2-hexynyl-4'-thioadenosine analogs as dual A2A/A3
Seung Woo Kim1, Jahyun Bok1, Hongseok Choi1
1Research Institute of Pharmaceutical Sciences, College of Pharmacy, Seoul National University, Seoul, 08826, Republic of Korea.

Abstract:
Adenosine receptors (ARs), particularly the A2A and A3 receptors are complementary regulators of inflammation, motivating a dual-agonist strategy. We designed, synthesized, and evaluated 2-hexynyl-6-amino 4'-thioadenosine derivatives and established a 5'-uronamide structure-activity relationship (SAR) guided by comparative docking. Small, hydrophobic 5' substituents were favored across both A2A and A3AR, mapping a shared hydrophobic pocket and clarifying the donor/acceptor interactions between the ligands and key residues (Thr88/His250 in A2AAR; Thr94 in A3AR). Critically, steric and electronic control at the 5'-uronamide enabled the identification of A2A- and A3-selective ligands, demonstrating tunable A2A/A3 selectivity via 5'-substituent design. The lead compound 2j (SHENECA) showed high-affinity dual binding (hA2AKi = 2.9 ± 0.28 nM; hA3Ki = 0.8 ± 0.1 nM) and subnanomolar functional potency (hA2A EC50 = 0.51 ± 0.17 nM; hA3 EC50 < 0.10 nM). In LPS-stimulated bone-marrow-derived dendritic cells, 2j (and 2i) produced greater suppression of IL-6 and IL-12/IL-23p40 than selective A2A or A3 agonists under our assay conditions, findings compatible with synergistic effects of dual A2A/A3 activation. In vitro ADMET profiling indicated favorable stability in human liver microsomes and plasma and no meaningful hERG inhibition. Collectively, the study demonstrates tunable A2A/A3 selectivity via 5'-uronamide design and the successful discovery of dual agonist 2j, and supports 2j as a strong lead for further optimization toward anti-inflammatory applications.
More Related Videos
 04:17
Author Spotlight: Advancing Cellular and Protein Engineering to Control Biological Functions and Develop Novel Therapies
Published on: September 27, 2024
 10:24
NMR-Based Activity Assays for Determining Compound Inhibition, IC50 Values, Artifactual Activity, and Whole-Cell Activity of Nucleoside Ribohydrolases
Published on: June 30, 2019
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...
 01:17
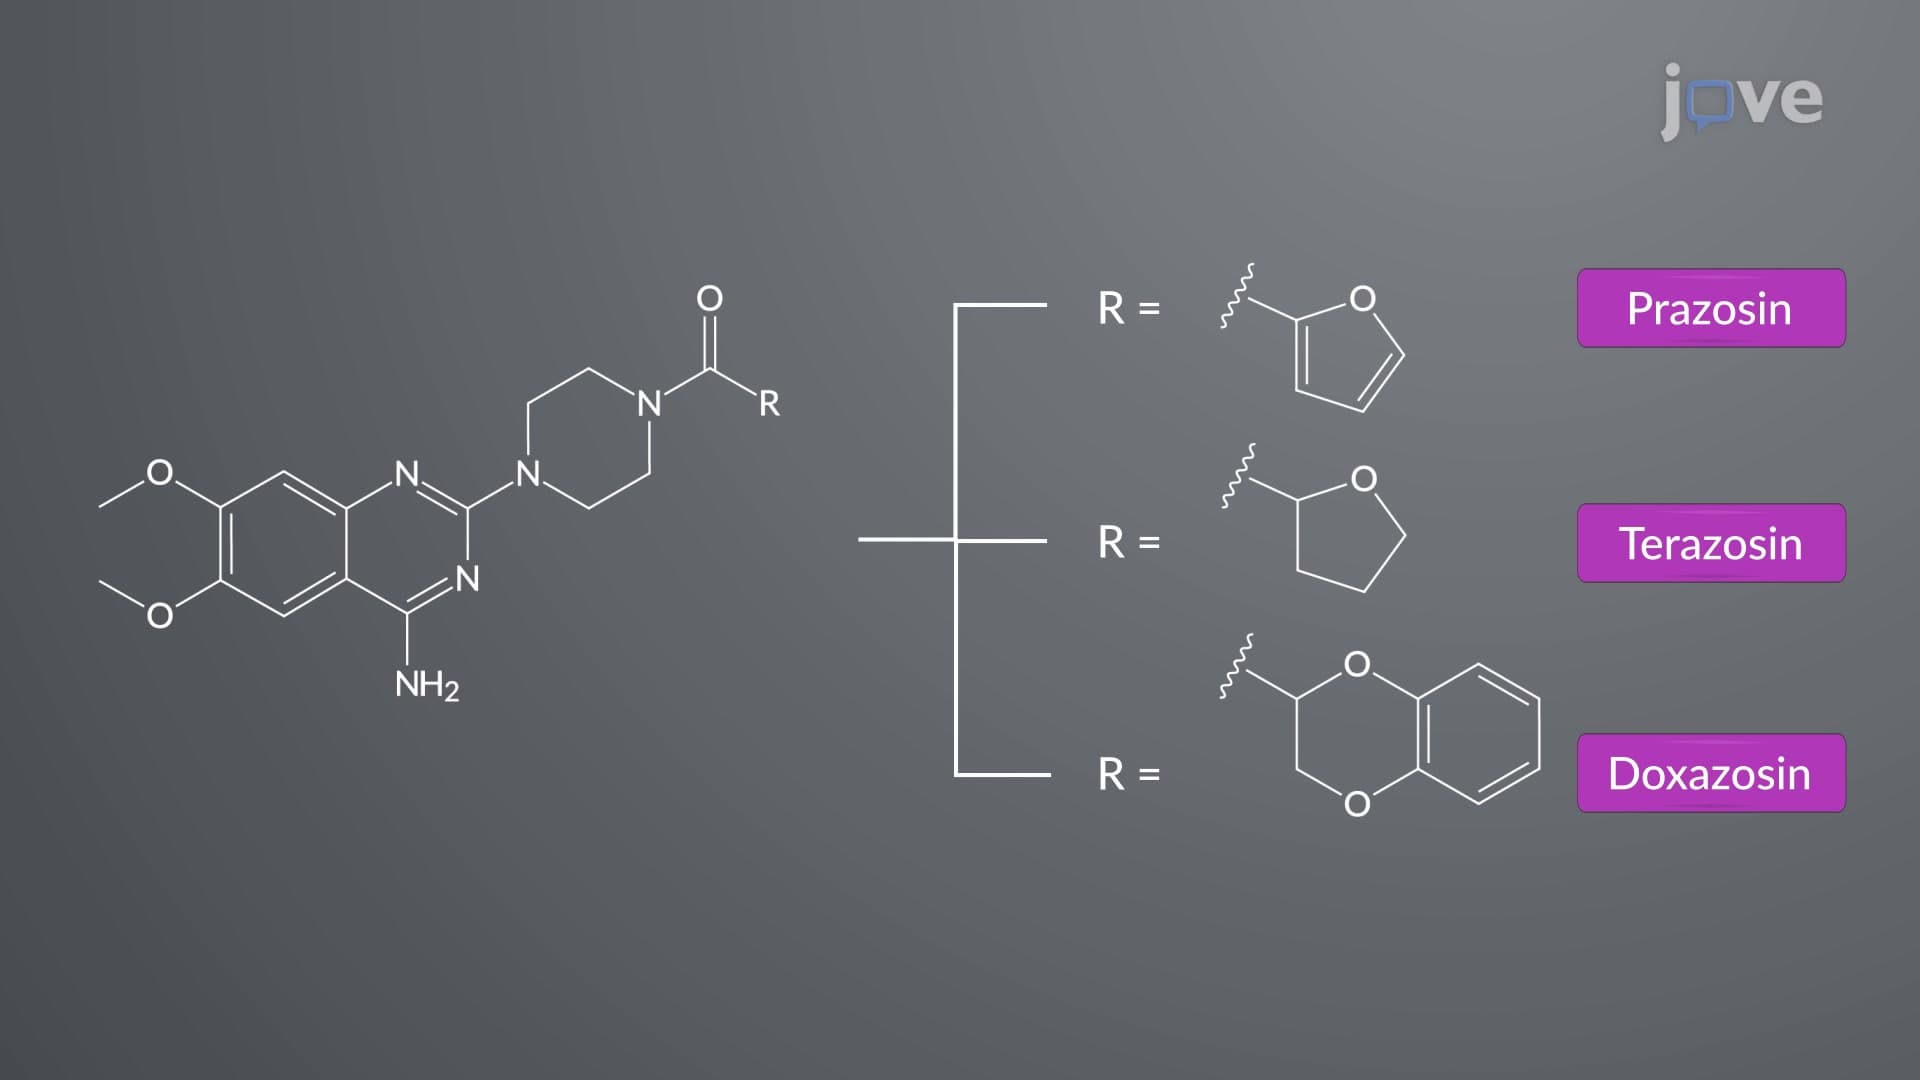
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline...
 01:31
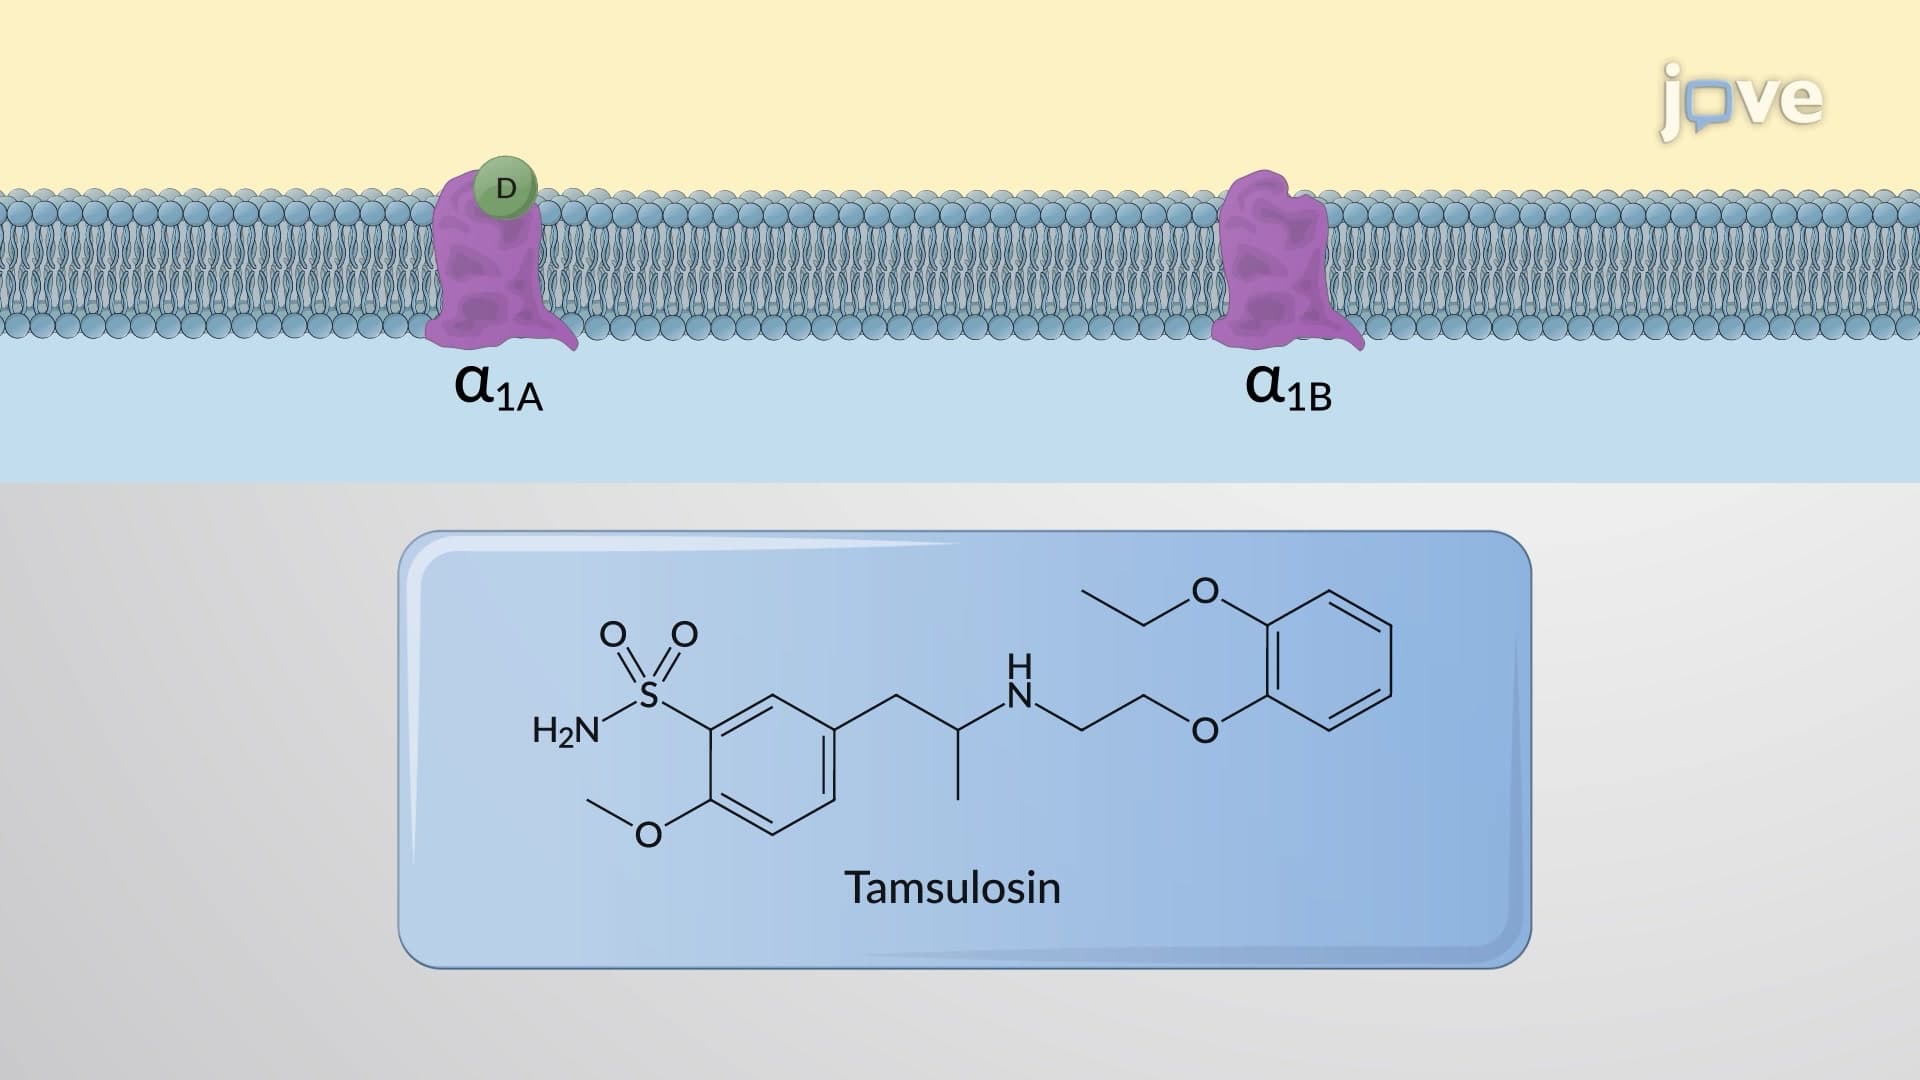
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:29
Indirect-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
Reversible inhibitors display short to medium durations of action. Short-acting agents include simple alcohols with...
 01:29
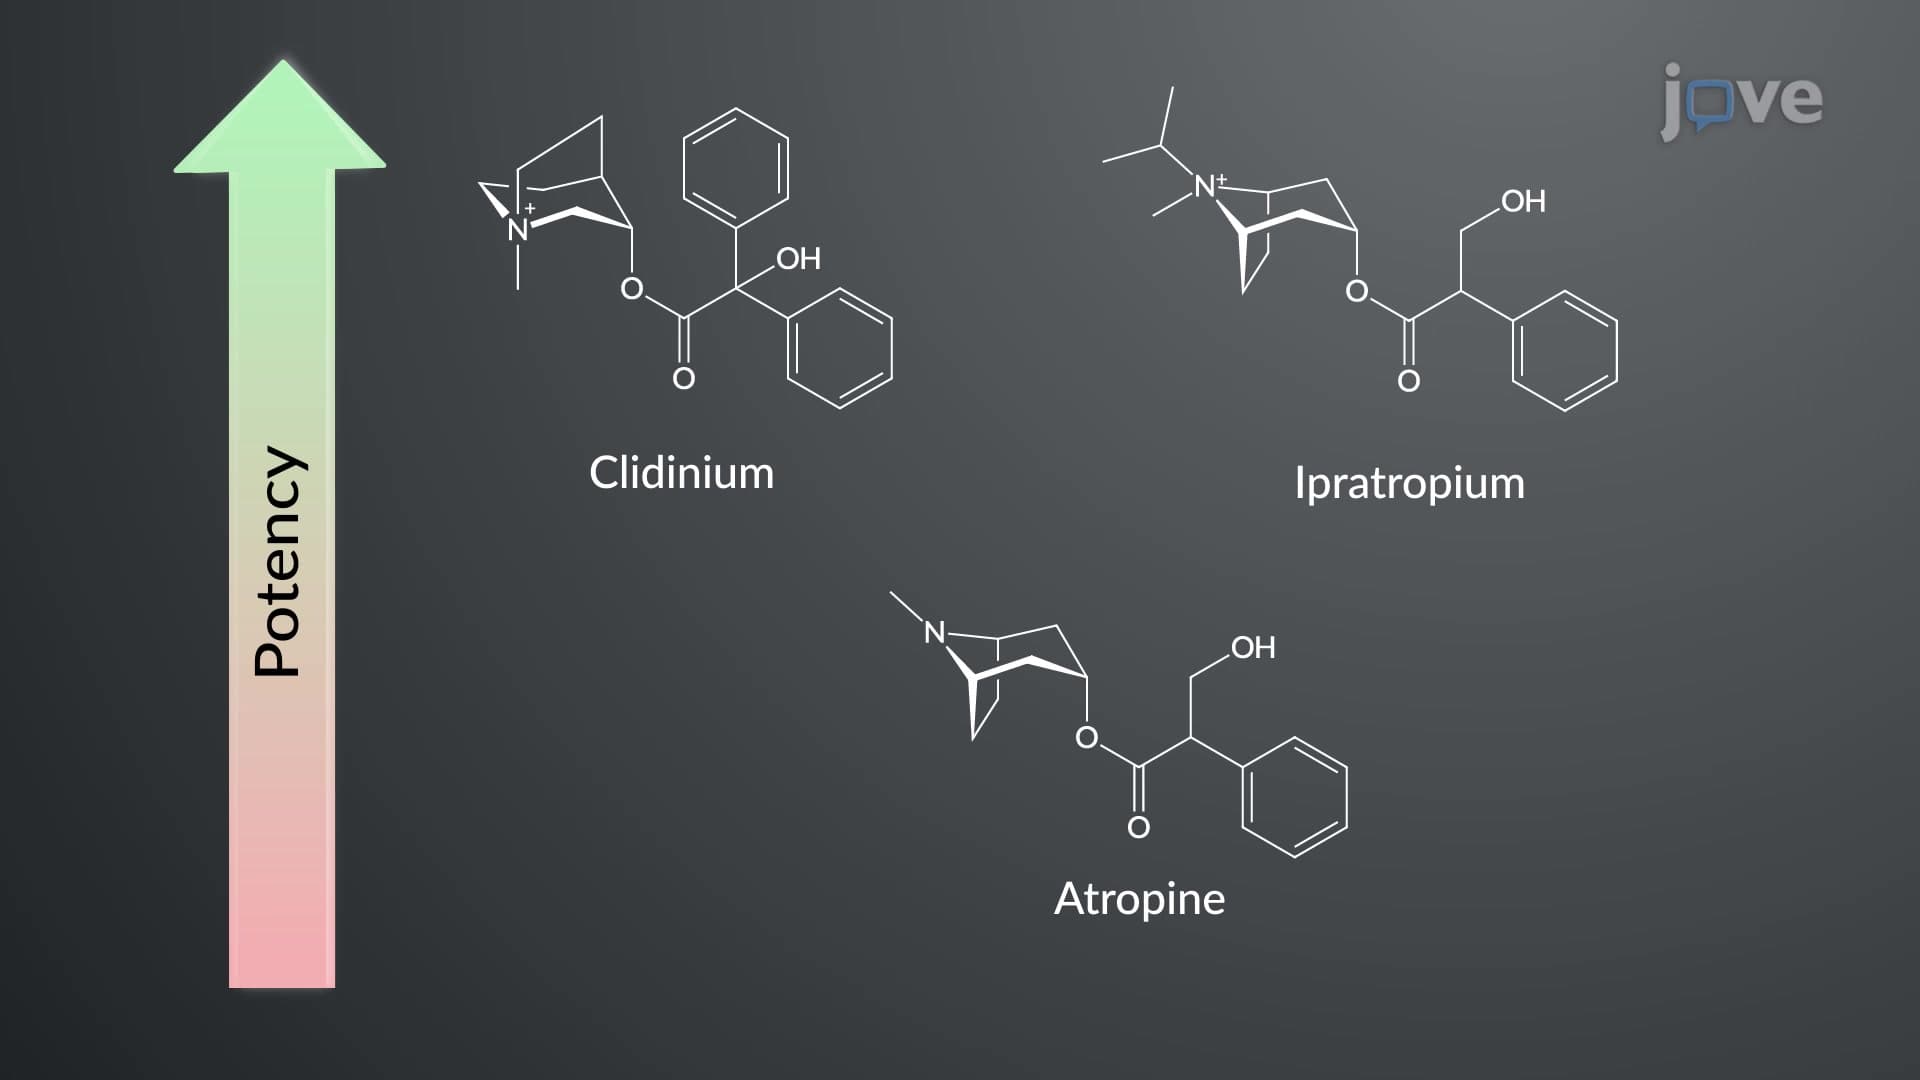
Cholinergic Antagonists: Chemistry and Structure-Activity Relationship