Related Experiment Video
Updated: Jan 10, 2026

Combining X-Ray Crystallography with Small Angle X-Ray Scattering to Model Unstructured Regions of Nsa1 from S. Cerevisiae
Published on: January 10, 2018
Bayesian multi-state multi-condition modeling of a protein structure based on X-ray crystallography data
Matthew Hancock1, James Holton2,3,4, James S Fraser1
1Department of Bioengineering and Therapeutic Sciences, University of California San Francisco, San Francisco, CA, USA.

Abstract:
An atomic structure model of a protein can be computed from a diffraction pattern of its crystal. While most crystallographic studies produce a single set of atomic coordinates, the billions of protein molecules in a crystal sample many conformational modes during data collection. As a result, a 'multi-state' model that depicts these conformations could reproduce the X-ray data better than a single conformation, and thus likely be more accurate. Computing such a multi-state model is challenging due to a lower data-to-parameter ratio than that for single-state modeling. To address this challenge, additional information could be considered, such as X-ray datasets collected for the same system under distinct experimental conditions (eg, temperature, ligands, mutations, and pressure). Here, we develop, benchmark, and illustrate MultiXray: Bayesian multi-state multi-condition modeling for X-ray crystallography. The input information is several X-ray datasets collected under distinct conditions and a molecular mechanics force field. The model consists of an independent coordinate set for each of several states and the weight of each state under each condition. A Bayesian posterior model density quantifies the match of the model with all X-ray datasets and the force field. A sample of models is drawn from the posterior model density using biased molecular dynamics (MD) simulations. We benchmark MultiXray on simulated CypA X-ray data. Using a second X-ray dataset improves the from 0.105 to 0.089. We then demonstrate MultiXray on experimental temperature-dependent data for SARS-CoV-2 Mpro. Using multiple X-ray datasets improves of the PDB-deposited structure from 0.253 to 0.237. MultiXray is implemented in our open-source Integrative Modeling Platform (IMP) software, relying on integration with Phenix, thus making it easily applicable to many studies.
More Related Videos
Related Concept Videos
 01:24
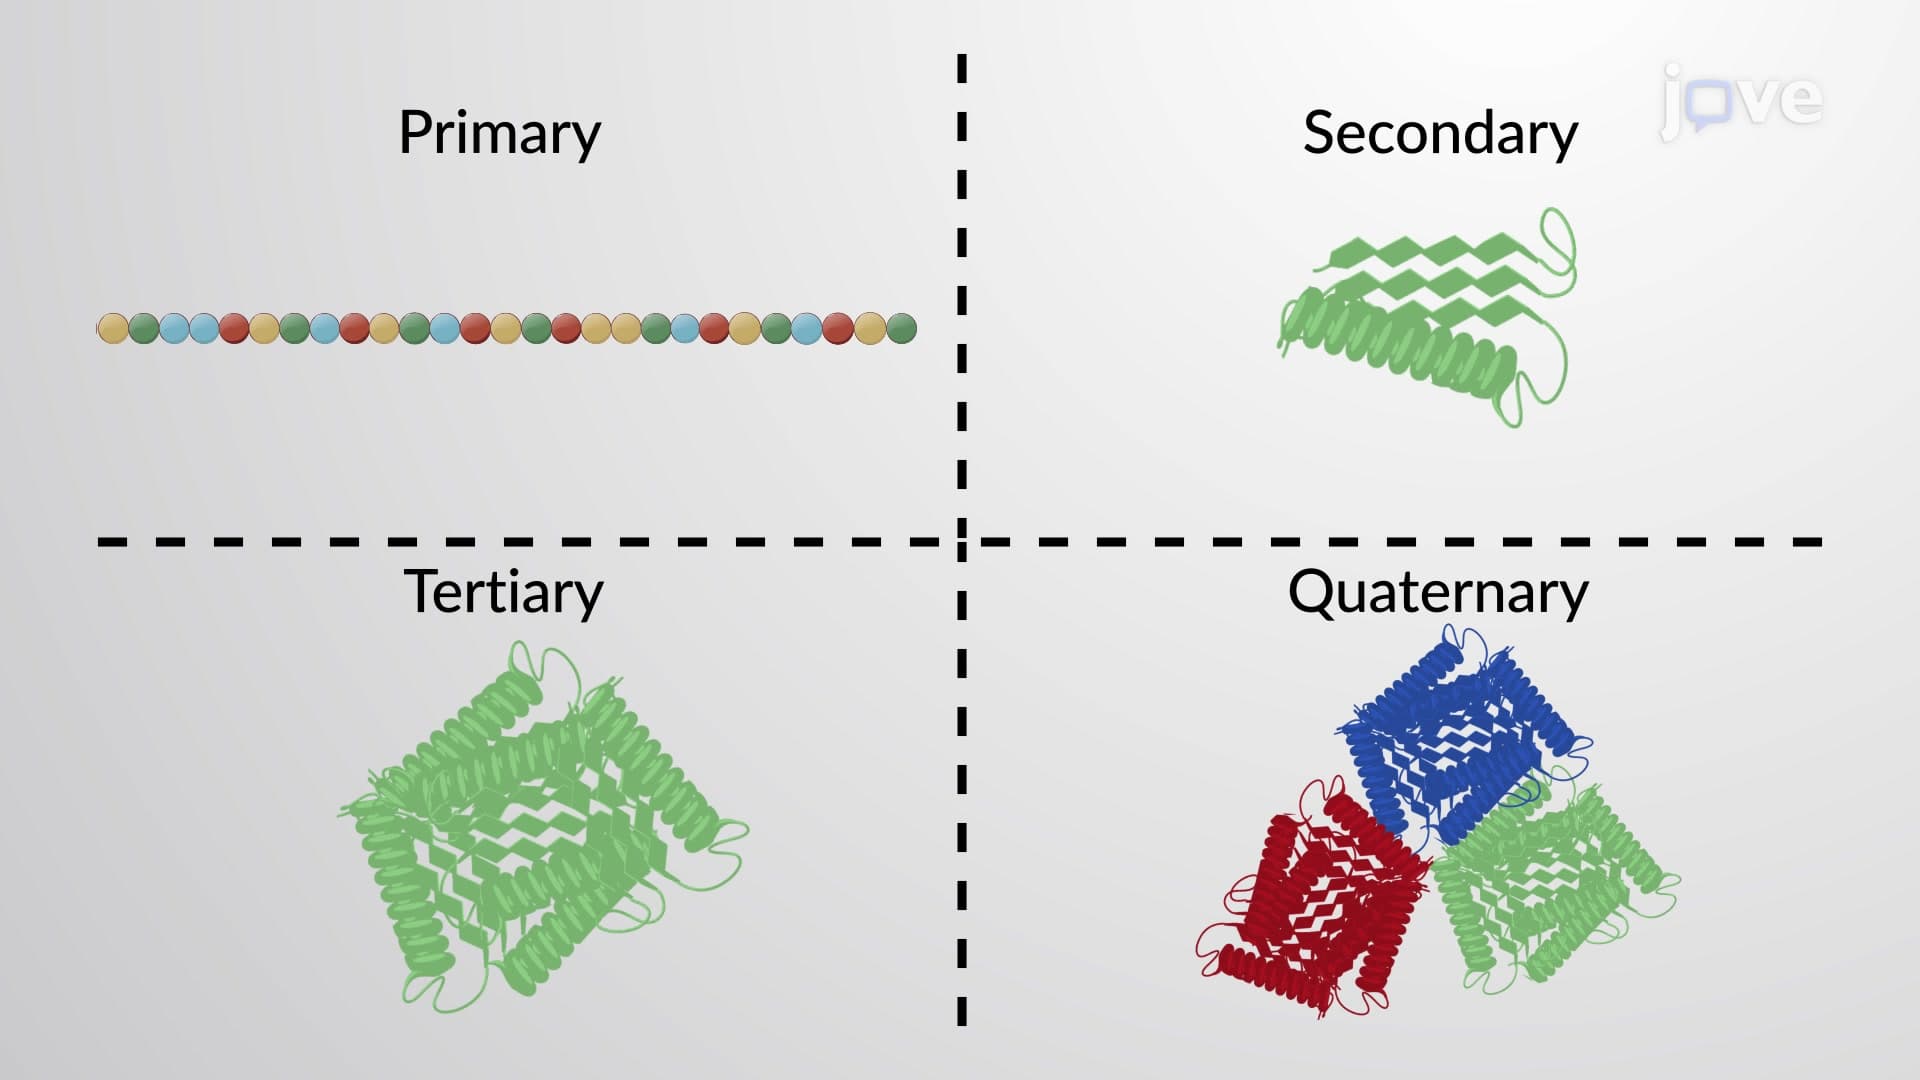
Protein Organization
The primary structure of a protein is its amino acid sequence....
 01:57
Gene Families
Occasionally these regions can be adapted to take on new roles within the organism, becoming novel genes...
 02:15
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...
 02:18
X-ray Crystallography
Diffraction
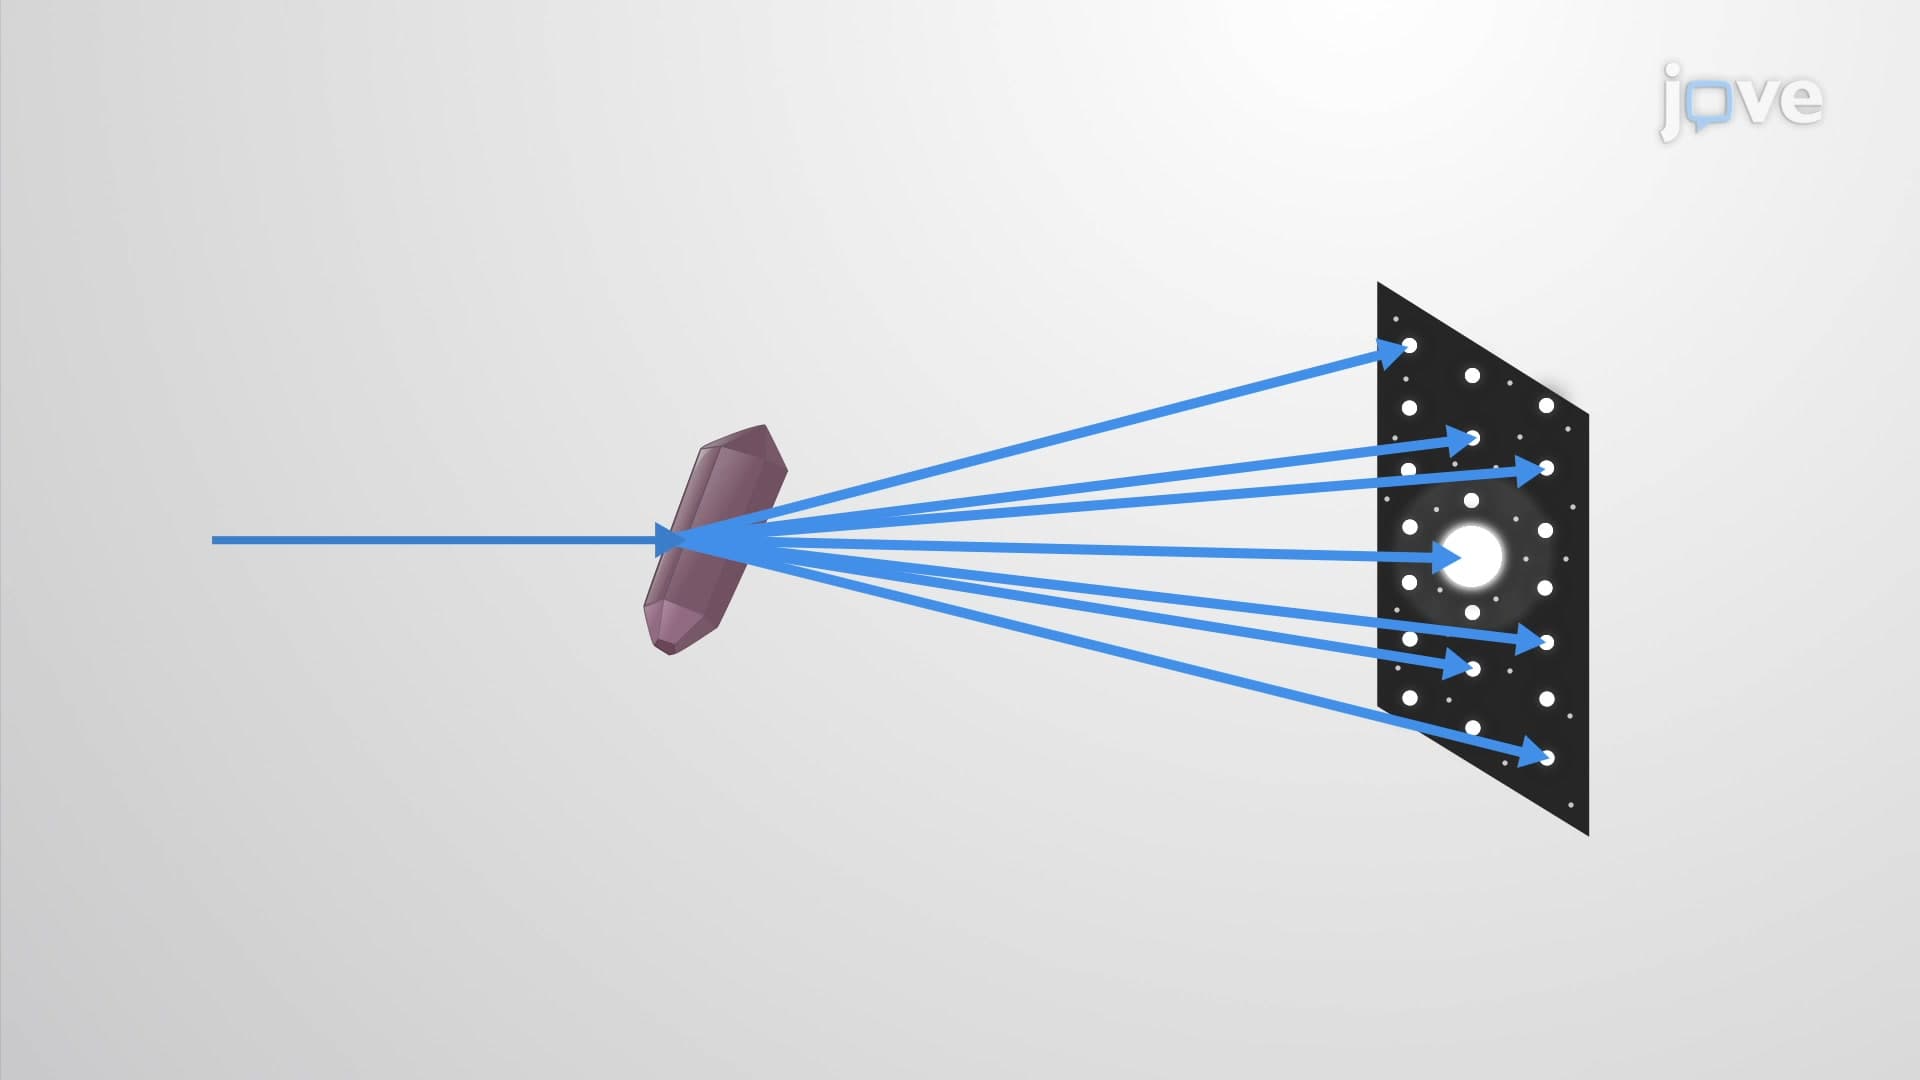
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 01:22
Protein Folding