Related Experiment Video
Updated: Jan 10, 2026
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Molecular simulations of the early structural development of normal alkanes with different chain lengths and
Patcharaporn Teeranontmongkol1, Go Matsuba2, Visit Vao-Soongnern3
1Laboratory of Computational and Applied Polymer Science (LCAPS), School of Chemistry, Suranaree University of Technology, Nakhon Ratchasima, 30000, Thailand.

Context:
The development of ordered structures upon immediately quenching from 473 K to different crystallization temperatures (Tc = 300 K, 320 K, 340 K, and 360 K) of three n-alkanes (n-eicosane (C20H42), n-tetracontane (C40H82) and n-octacontane (C80H162) to accommodate the formation of bilayer, monolayer and the folded-chain configurations, respectively), was quantitatively analyzed by evaluating chain/bond order parameters, interaction energies, conformational statistics, local dynamics, structural pair correlation function, and X-ray scattering profiles. The formation of ordered structures seems to be best at Tc = 340 K. Generally, systems that tend to form the monolayer and folded-chain configurations exhibit the most and the least ordered structures, respectively. Chains in monolayer structures tend to have a larger fraction of trans state, higher anisotropy of bond orientation, slower monomer dynamics, and more densely packed structures than chains in bilayer structures, while longer alkanes with chain-folded structures exhibit the least characteristics of these properties. The nucleation temperature (Tn) can be affected by their structural configuration, with the highest Tn for the monolayer structure. Nevertheless, the melting temperature (Tm) tends to depend solely on the molecular weights of these alkanes, not on their structural configuration.
Methods:
Monte Carlo (MC) simulation of the coarse-grained (CG) models of three n-alkanes (one CG bead equivalent to an ethylene unit) on the second nearest neighbor diamond (2nnd) lattice with the same periodic box dimension of 5 nm in each size. The energetics of CG models were composed of the Rotational Isomeric State (RIS) model and the Lennard-Jones (LJ) potential energies to represent their intra- and intermolecular interactions, respectively. Structure developments within 100 million Monte Carlo steps (MCS) trajectories were monitored, and data analysis was based on snapshots collected at intervals of 10,000 MCS. All simulations and data analysis were performed using in-house FORTRAN codes with the g77 compiler.
Related Concept Videos
 02:33
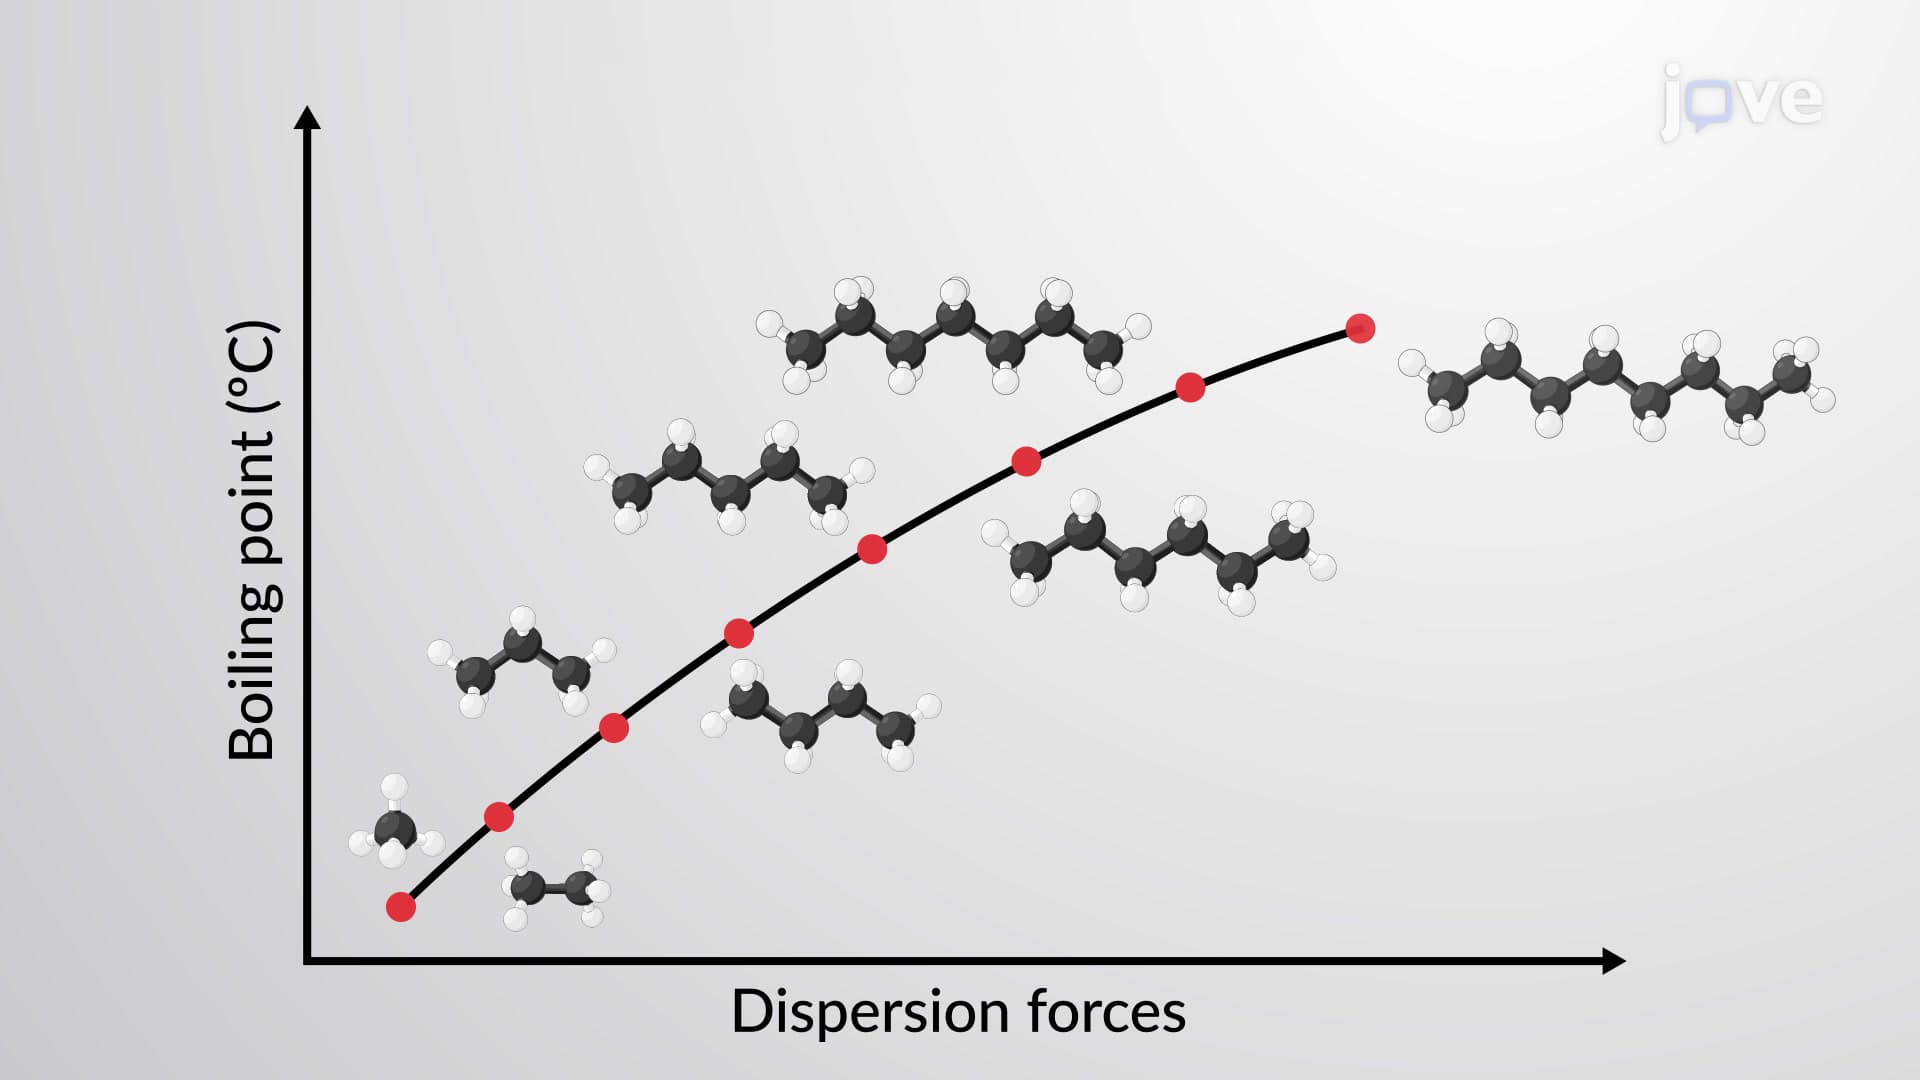
Physical Properties of Alkanes
 02:23
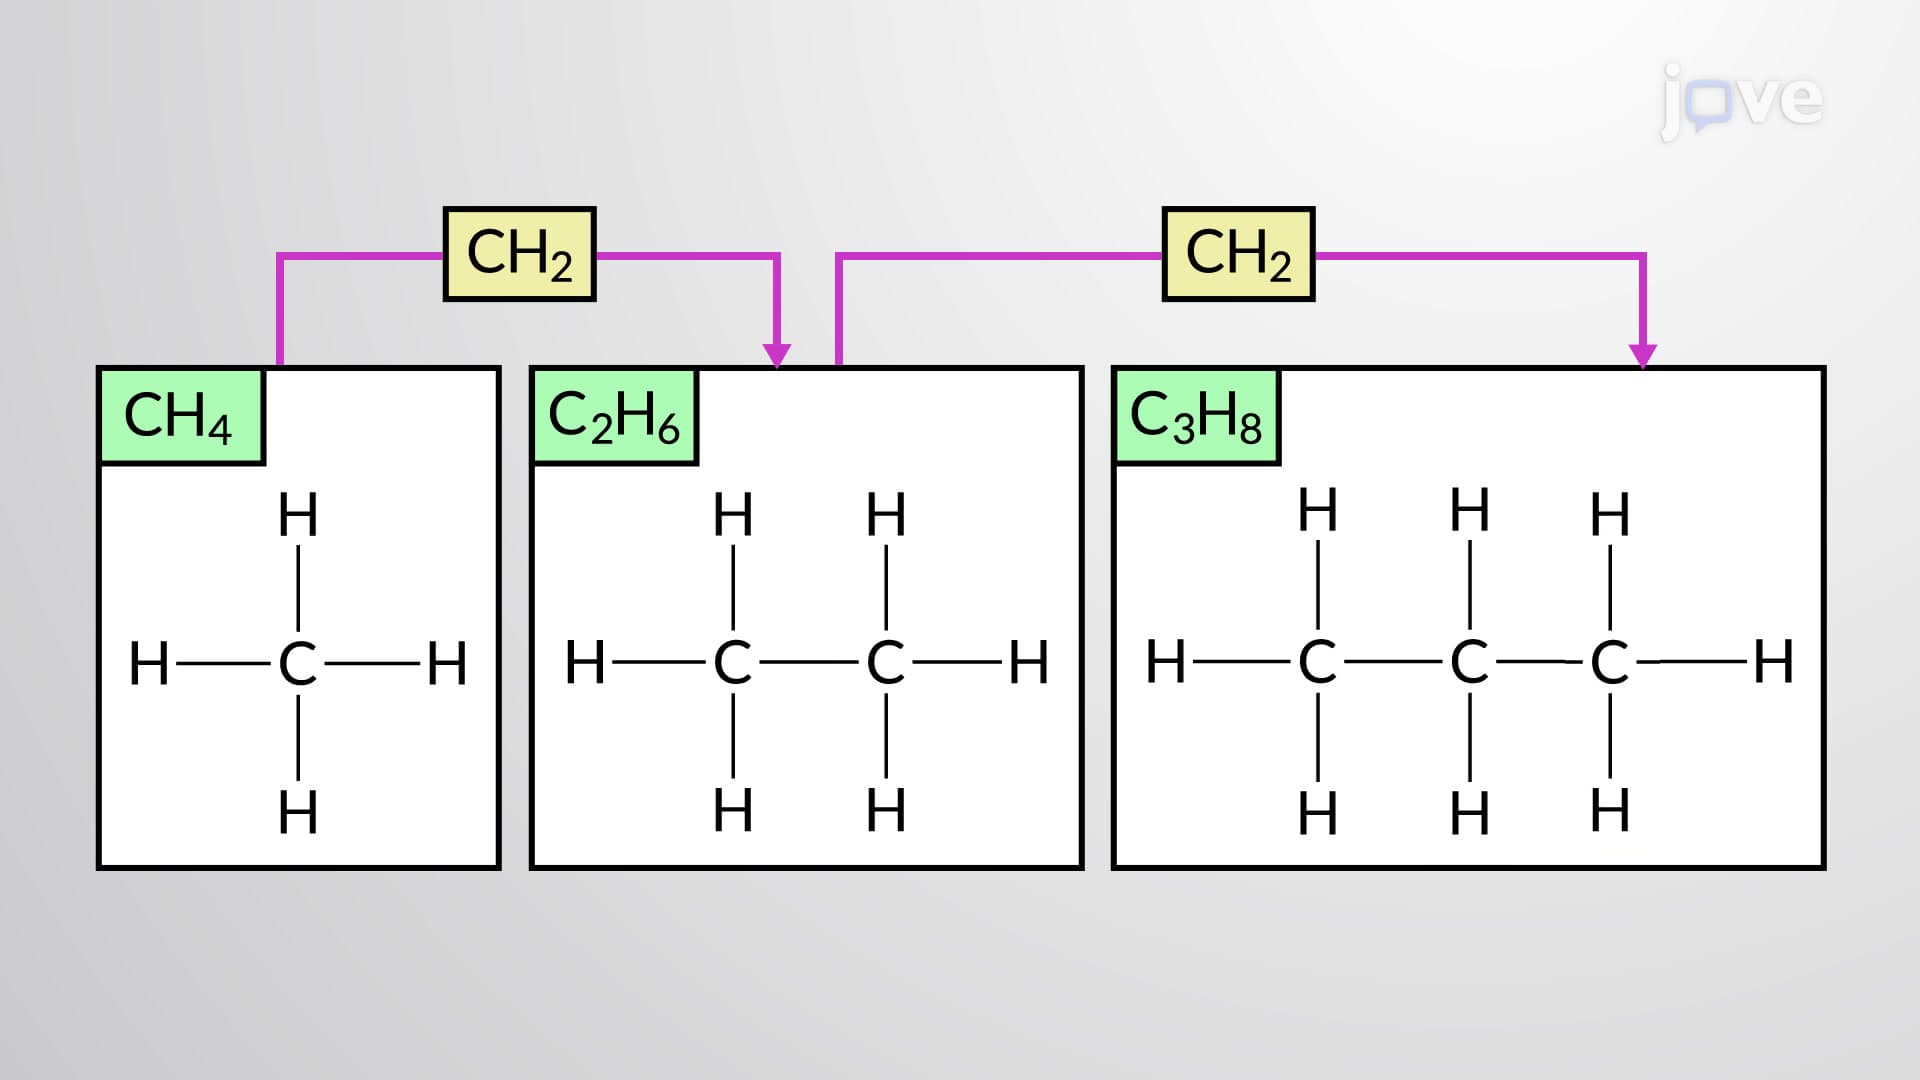
Structure of Alkanes
Hydrocarbons are the simplest organic compounds composed of carbons and hydrogens. Based on the bond order between carbons, the hydrocarbons are further classified into alkanes, alkenes, and alkynes.
Alkanes are the simplest hydrocarbons with sp3 hybrid carbon atoms....
 02:14
Combustion Energy: A Measure of Stability in Alkanes and Cycloalkanes
Alkanes undergo combustion in the presence of excess oxygen and high-temperature conditions to give carbon dioxide and water. A combustion reaction is the energy source in natural gas, liquified...
 02:18
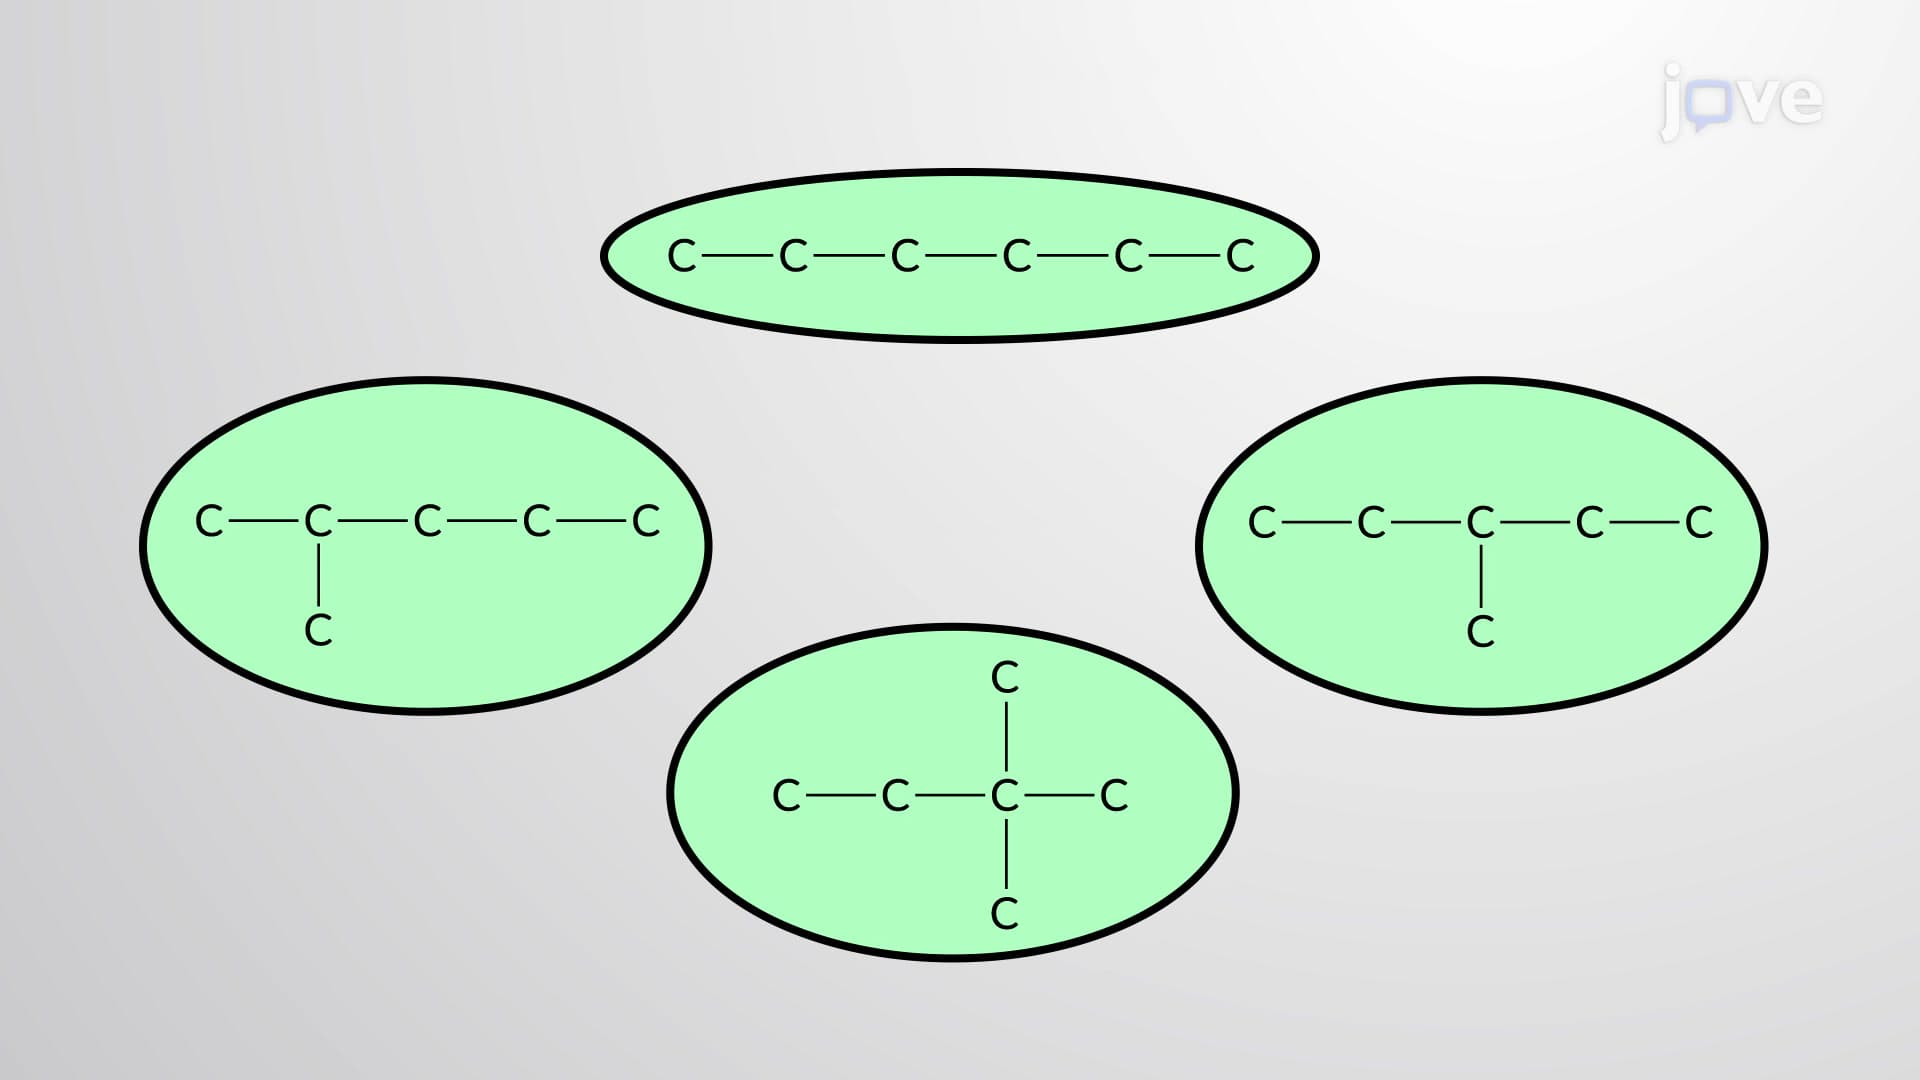
Constitutional Isomers of Alkanes
The linear isomer of an alkane is prefixed by the term “n”; hence a linear isomer of pentane is known as n-pentane. Based on the type of branching, some of the...
 02:22
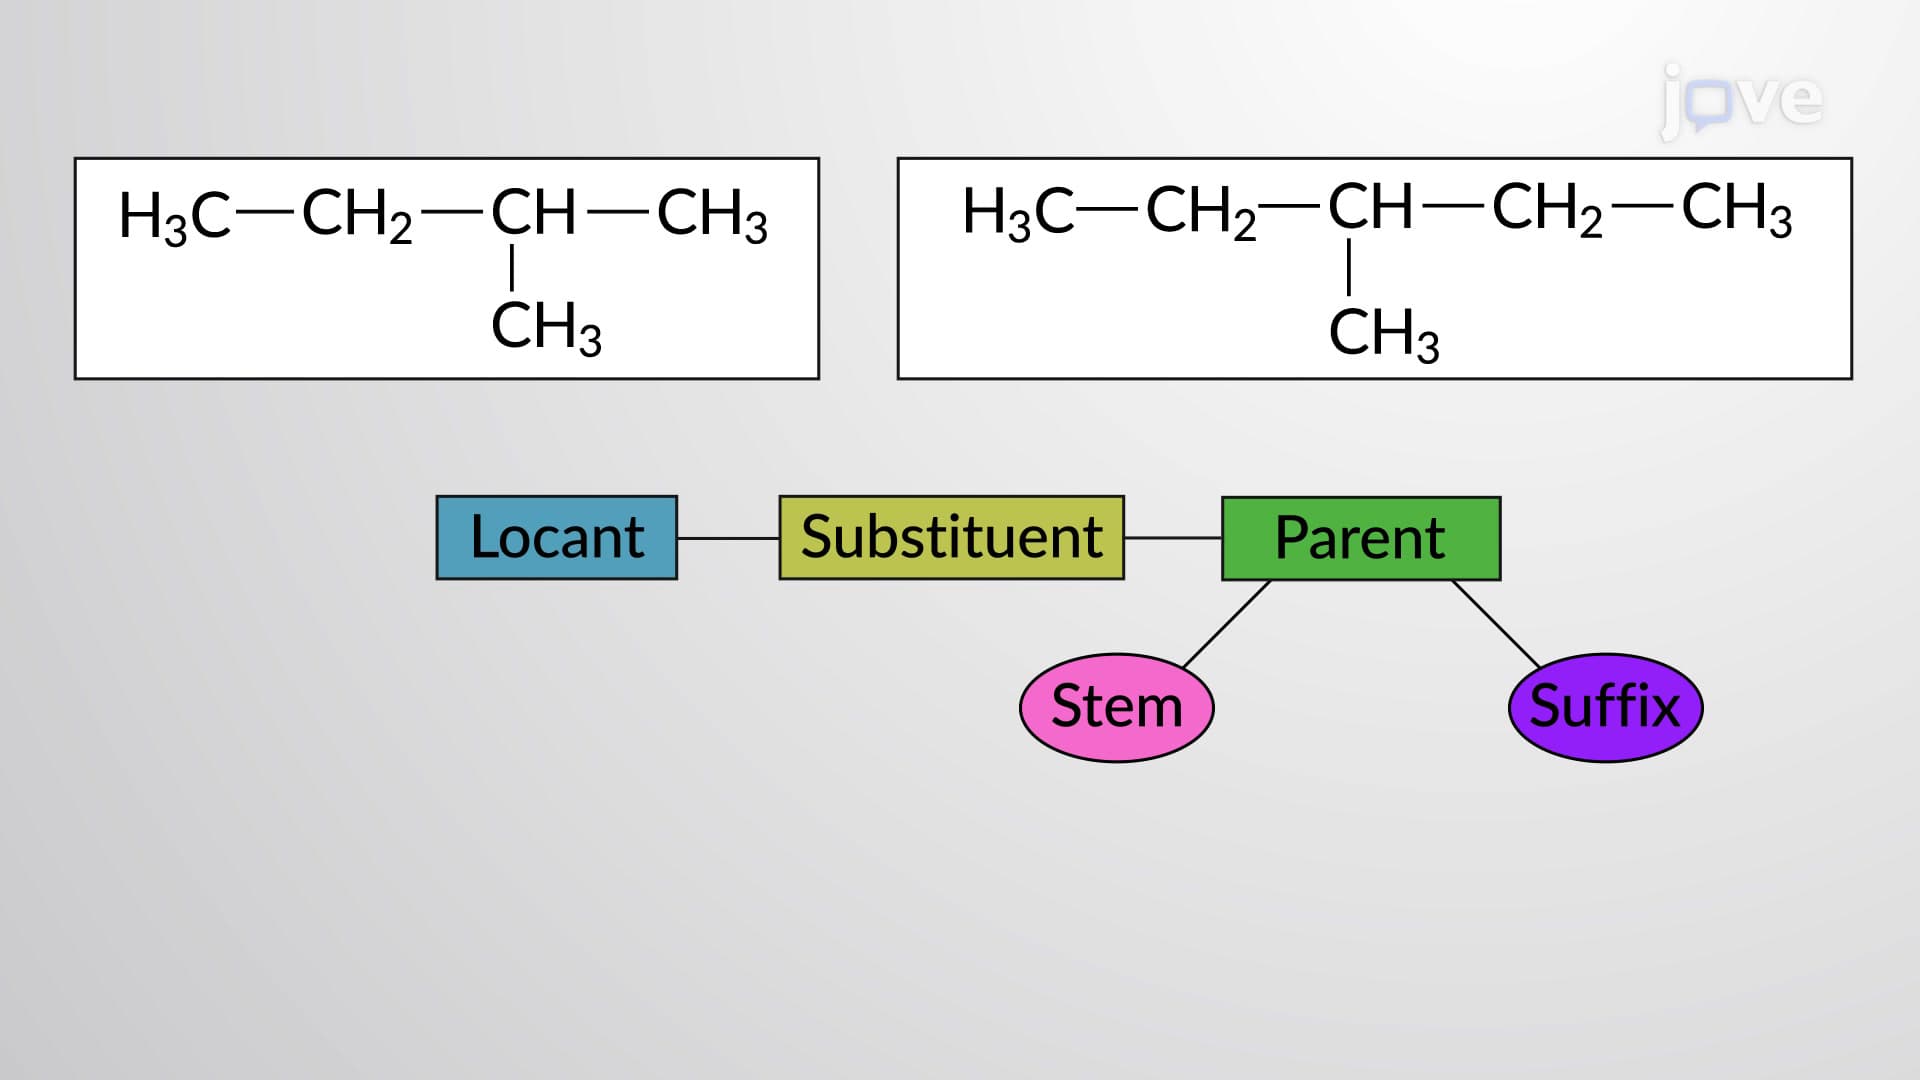
Nomenclature of Alkanes
The alkane nomenclature considers the length of the carbon chain, the number, and the location of the substituent to arrive at its systematic name. The IUPAC...
 01:18
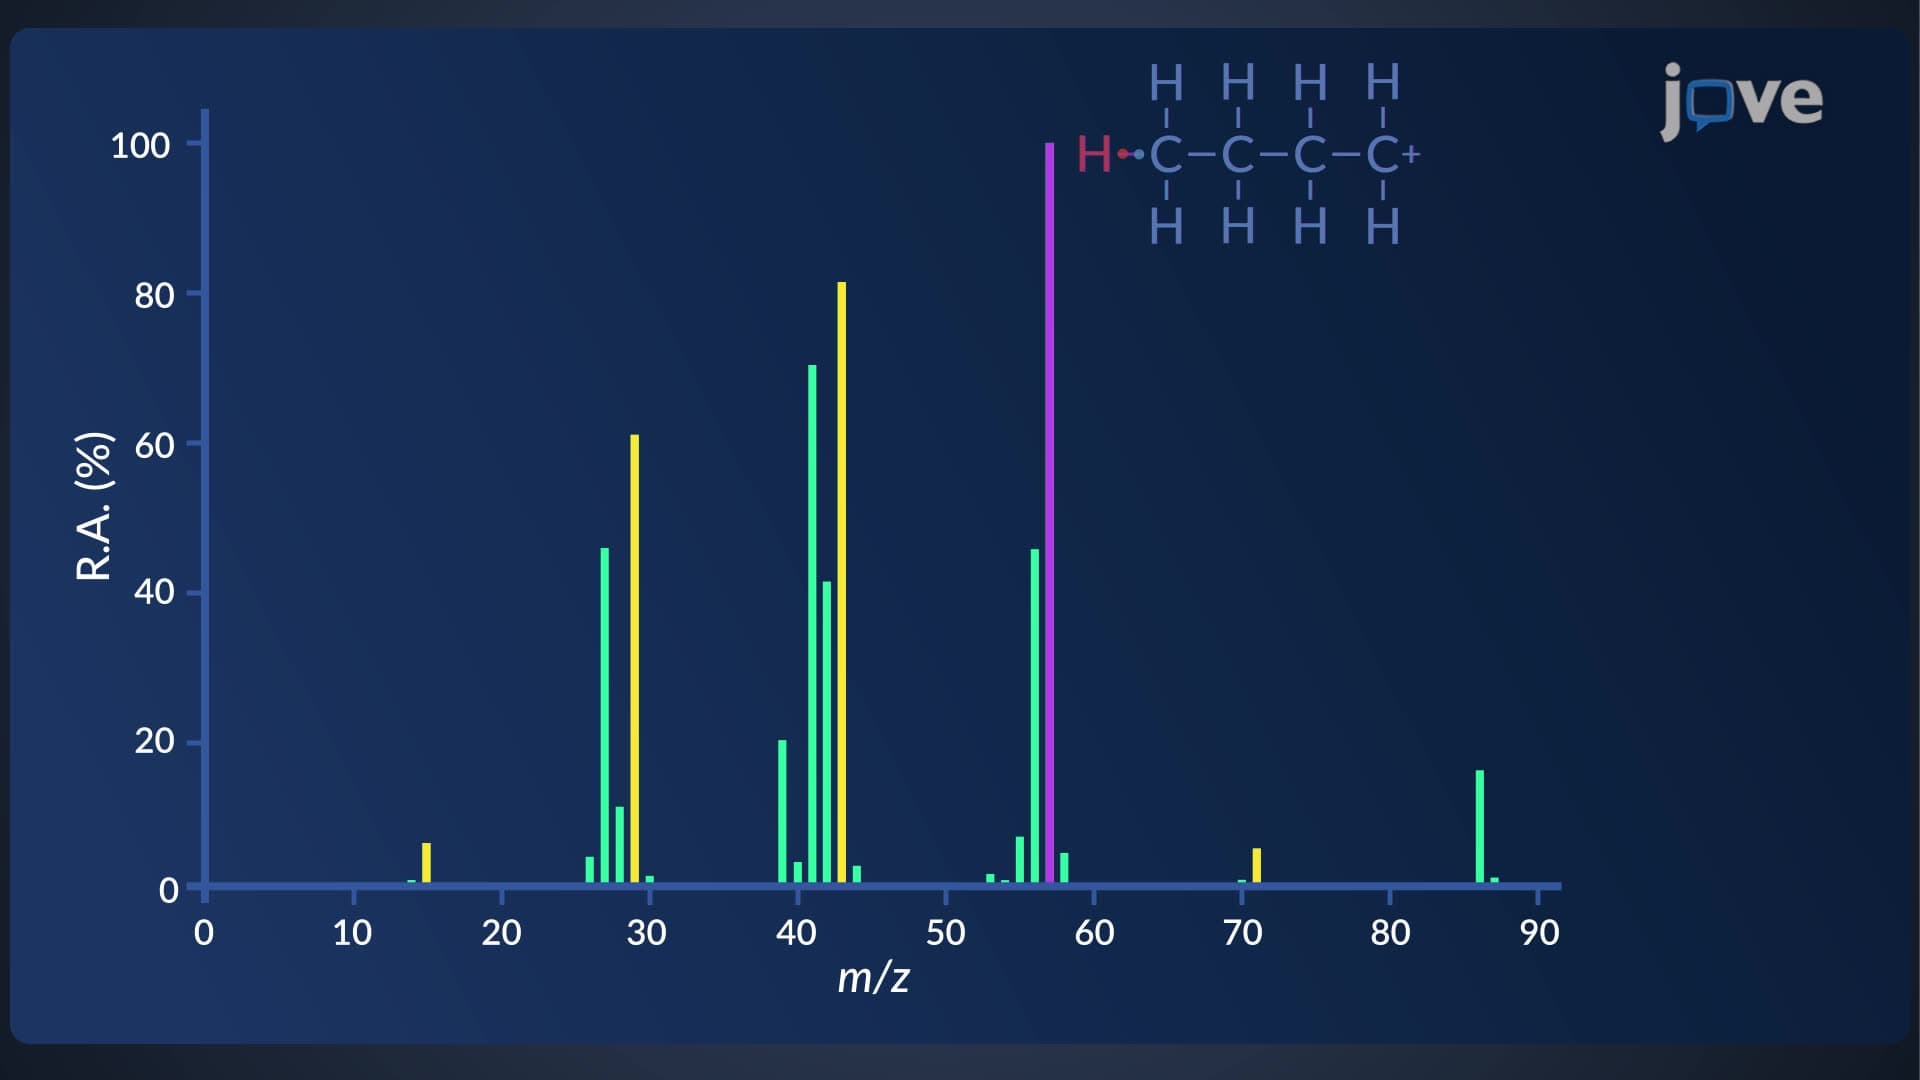
Mass Spectrometry: Long-Chain Alkane Fragmentation