Related Experiment Video
Updated: Jan 10, 2026


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Engineering drug-protein kinetics via intrinsic quantum Stark fields in binding motifs
Moses Udoisoh1, Mujahid Usman Yahuza2, Patience Folashade Olabinri3
1Department of Physics, Ignatius Ajuru University of Education, Rivers State, Rumuolumeni, Nigeria. moses.udoisoh@iaue.edu.ng.

Context:
Intrinsic, oriented electric fields inside protein active sites, quantified by vibrational Stark effect (VSE) measurements and electrostatic calculations have been implicated in catalytic preorganization, yet their systematic use to program drug-target lifetimes has remained underexplored. We advance a field-first paradigm in which the component of the protein field projected along a dominant reaction coordinate, denoted as (the component of the protein's electric field along the reaction coordinate), serves as a tunable design variable for off-rate and residence time (τ). Focusing on two mechanistic archetypes proton sharing (Ketosteroid Isomerase, KSI-like) and bond polarization/dissociation (human aldose reductase, hALR2-like) we show that realistic changes in can tilt barriers, alter curvatures, and modulate tunneling, yielding exponential leverage on kinetics. This reframes pharmacodynamic lifetime as an electrostatic, geometry-addressable property, complementary to affinity optimization and accessible through protein mutations or ligand substituents that re-orient local dipoles.
Method:
We developed a predictive, quantum-mechanical framework grounded in analytical solutions to the one-dimensional time-independent Schrödinger equation. Two experimentally validated systems were modeled viz the proton-transfer dynamics in ketosteroid isomerase (KSI) using an asymmetric double-well potential, and carbonyl polarization/dissociation in human aldose reductase (hALR2) using a modified Morse potential. The intrinsic Stark field was incorporated via its projection onto the reaction coordinate ( ), coupling through molecular dipole and polarizability terms to tilt and reshape the potential energy landscape. The resulting eigenvalues and wavefunctions provided the parameters for a Grote-Hynes-corrected transition-state theory model with explicit quantum tunneling corrections. This approach quantitatively connects field strength to activation barriers, vibrational frequencies, and ultimately the off-rate ( ), enabling the prediction of how perturbations via mutagenesis or ligand design alter residence time.
More Related Videos
 10:50
Using In Vitro Fluorescence Resonance Energy Transfer to Study the Dynamics Of Protein Complexes at a Millisecond Time Scale
Published on: March 14, 2019
 08:31
Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
Related Concept Videos
 01:16
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 02:18
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:25
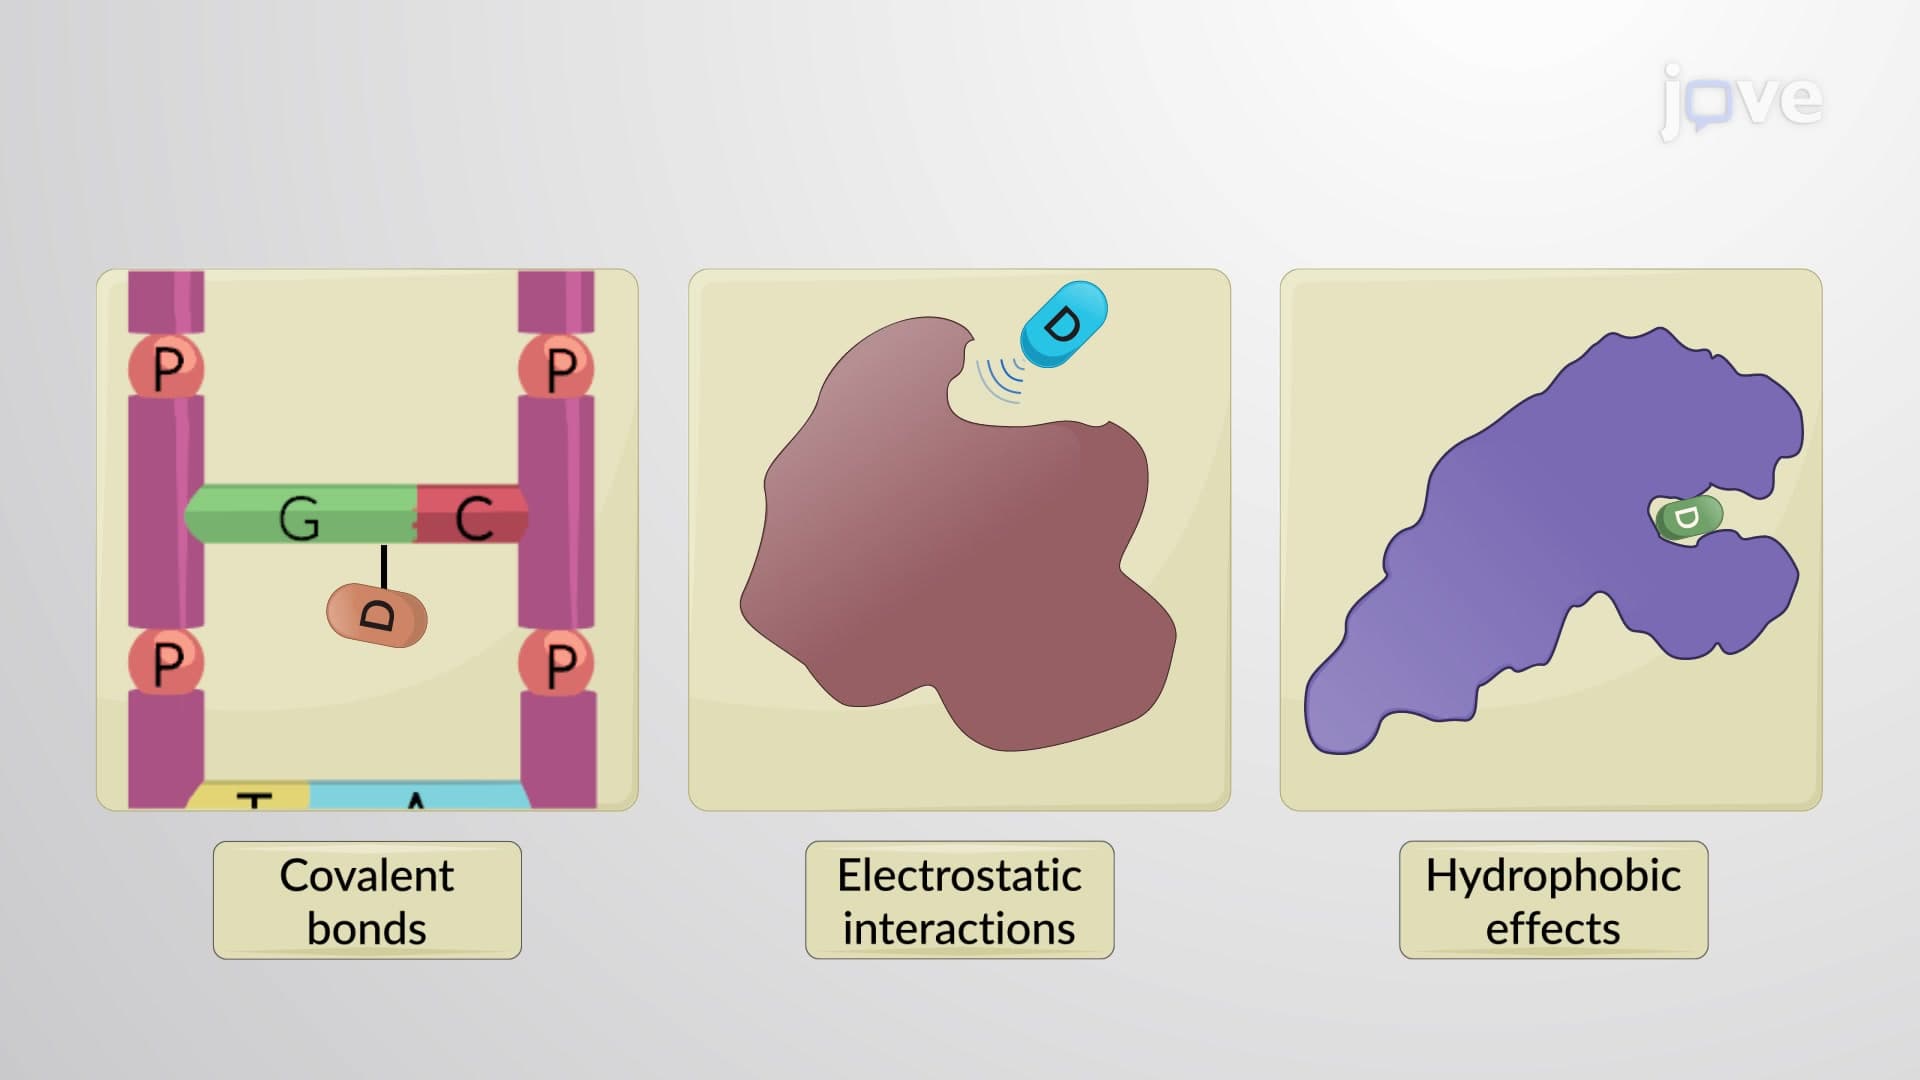
Drug-Receptor Bonds
In...
 00:49
Ligand Binding and Linkage