Related Experiment Video
Updated: Jan 10, 2026

Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Data-Efficient Equivariant NNPs Enable DFT-Accurate Simulations and Implicit Solvation Free Energies
Esma Mutlu1, Selonou G Kankinou1, Omer Tayfuroglu1,2
1Department of Chemistry, Gebze Technical University, Kocaeli 41400, Turkey.

Abstract:
The use of machine learning (ML) potentials has emerged as a powerful approach in computational chemistry, particularly in computer-aided drug design studies. Neural network potentials (NNPs) provide a more physics-informed estimation of binding and solvation events, bridging the accuracy gap between classical force fields and quantum mechanical methods. Current universal neural network potentials (NNPs) have not yet achieved consistent chemical accuracy required for reliable molecular dynamics simulations. Accurate yet data efficient representation of potential energy surfaces and prediction of solvation free energies are essential for large-scale molecular simulations and drug-design workflows. Here, we utilize data efficient E(3)-equivariant graph neural network potentials that are capable of estimating the solvation free energies (SFEs) of small compounds with density functional theory (DFT)-level accuracy and significantly reduced computational cost. Leveraging the data efficiency of equivariant architectures, our models achieve chemical accuracy with a relatively small training data set. We demonstrate that the method is data-efficient in constructing ML potentials. Our focus is on hydration free energy changes of small compounds from the FreeSolv database. We develop and test two distinct NNPs─one for the gas phase and one for an implicit water model─that can be applied in molecular simulations and SFE calculations using implicit solvation. The Solvation Model based on Density (SMD)-based implicit NNP model achieves an accuracy of 89% while offering a substantial computational speed-up compared to its DFT counterpart, which attains 90% accuracy.
More Related Videos
 05:00
Author Spotlight: Streamlining Visual Dynamics to Simplify Molecular Dynamics Simulations Using Gromacs
Published on: August 9, 2024
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:26
Thermodynamic Potentials
 01:17
Ampere-Maxwell's Law: Problem-Solving
To solve the problem, we can use the equations from the analysis of an RC circuit and Maxwell's version of Ampère's law.
For the first part of the...
 01:13
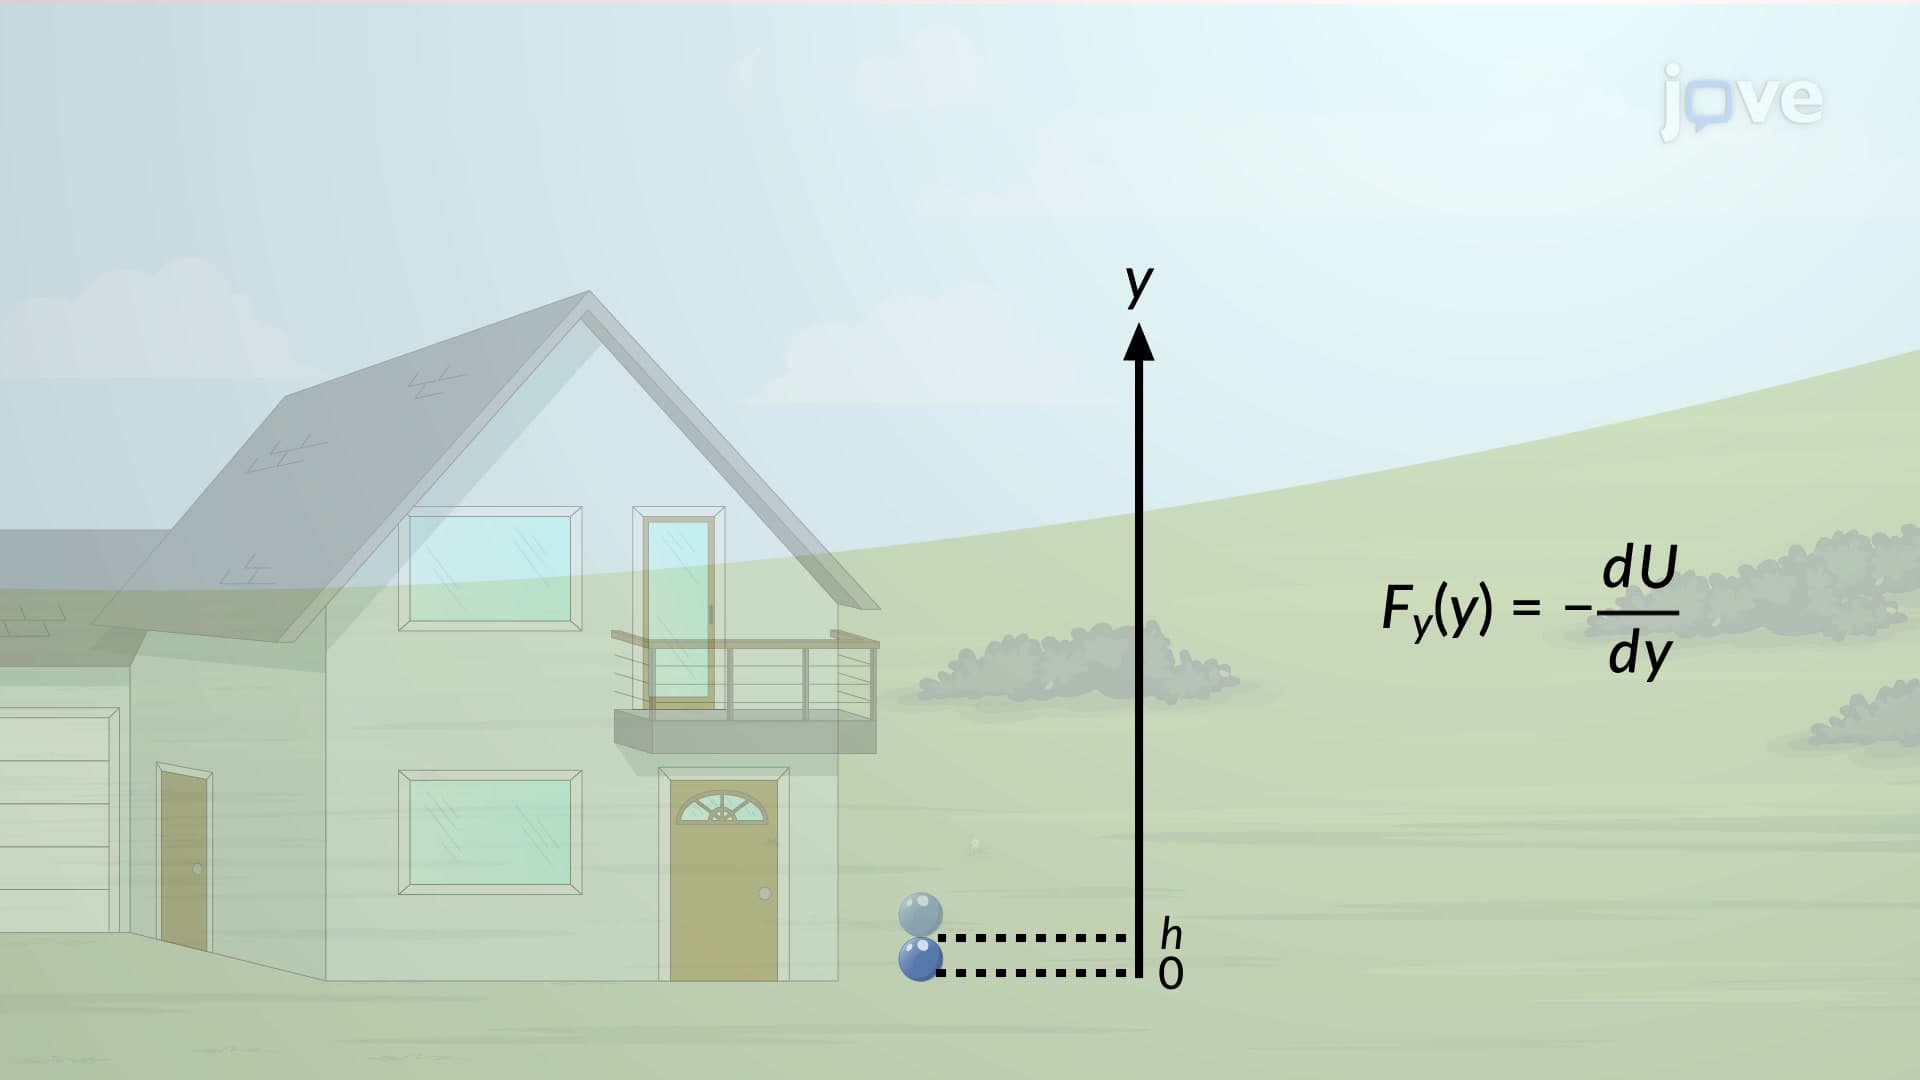
Force and Potential Energy in One Dimension
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...
 02:05
Entropy and Solvation
 03:25
Free Energy Changes for Nonstandard States